Related Experiment Video
Updated: Aug 15, 2026

A Strategy for Sensitive, Large Scale Quantitative Metabolomics
Published on: May 27, 2014
Quantitative prediction of biodegradability, metabolite distribution and toxicity of stable metabolites
S Dimitrov1, R Breton, D Macdonald
1Laboratory of Mathematical Chemistry, University Prof As. Zlatarov, Bourgas, Bulgaria.

Abstract:
An evaluation of the capability of organic chemicals to mineralize is an important factor to consider when assessing their fate in the environment. Microbial degradation can convert a toxic chemical into an innocuous one, and vice versa, or alter the toxicity of a chemical. Moreover, primary biodegradation can convert chemicals into stable products that can be difficult to mineralize. In this paper, we present some new results obtained on the basis of a recently developed probabilistic approach to modeling biodegradation based on microbial transformation pathways. The metabolic transformations and their hierarchy were calibrated by making use of the ready biodegradability data from the MITI-I test and expert knowledge for the most probable transformation pathways. A model was developed and integrated into an expert software system named CATABOL that is able to predict the probability of biodegradation of organic chemicals directly from their structure. CATABOL simulates the effects of microbial enzyme systems, generates the most plausible transformation pathways, and quantitatively predicts the persistence and toxicity of the biodegradation products. A subset of 300 organic chemicals were selected from Canada's Domestic Substances List and subjected to CATABOL to compare predicted properties of the parent chemicals with their respective first stable metabolite. The results show that most of the stable metabolites have a lower acute toxicity to fish and a lower bioaccumulation potential compared to the parent chemicals. In contrast, the metabolites appear to be generally more estrogenic than the parent chemicals.
Related Concept Videos
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:07
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:30
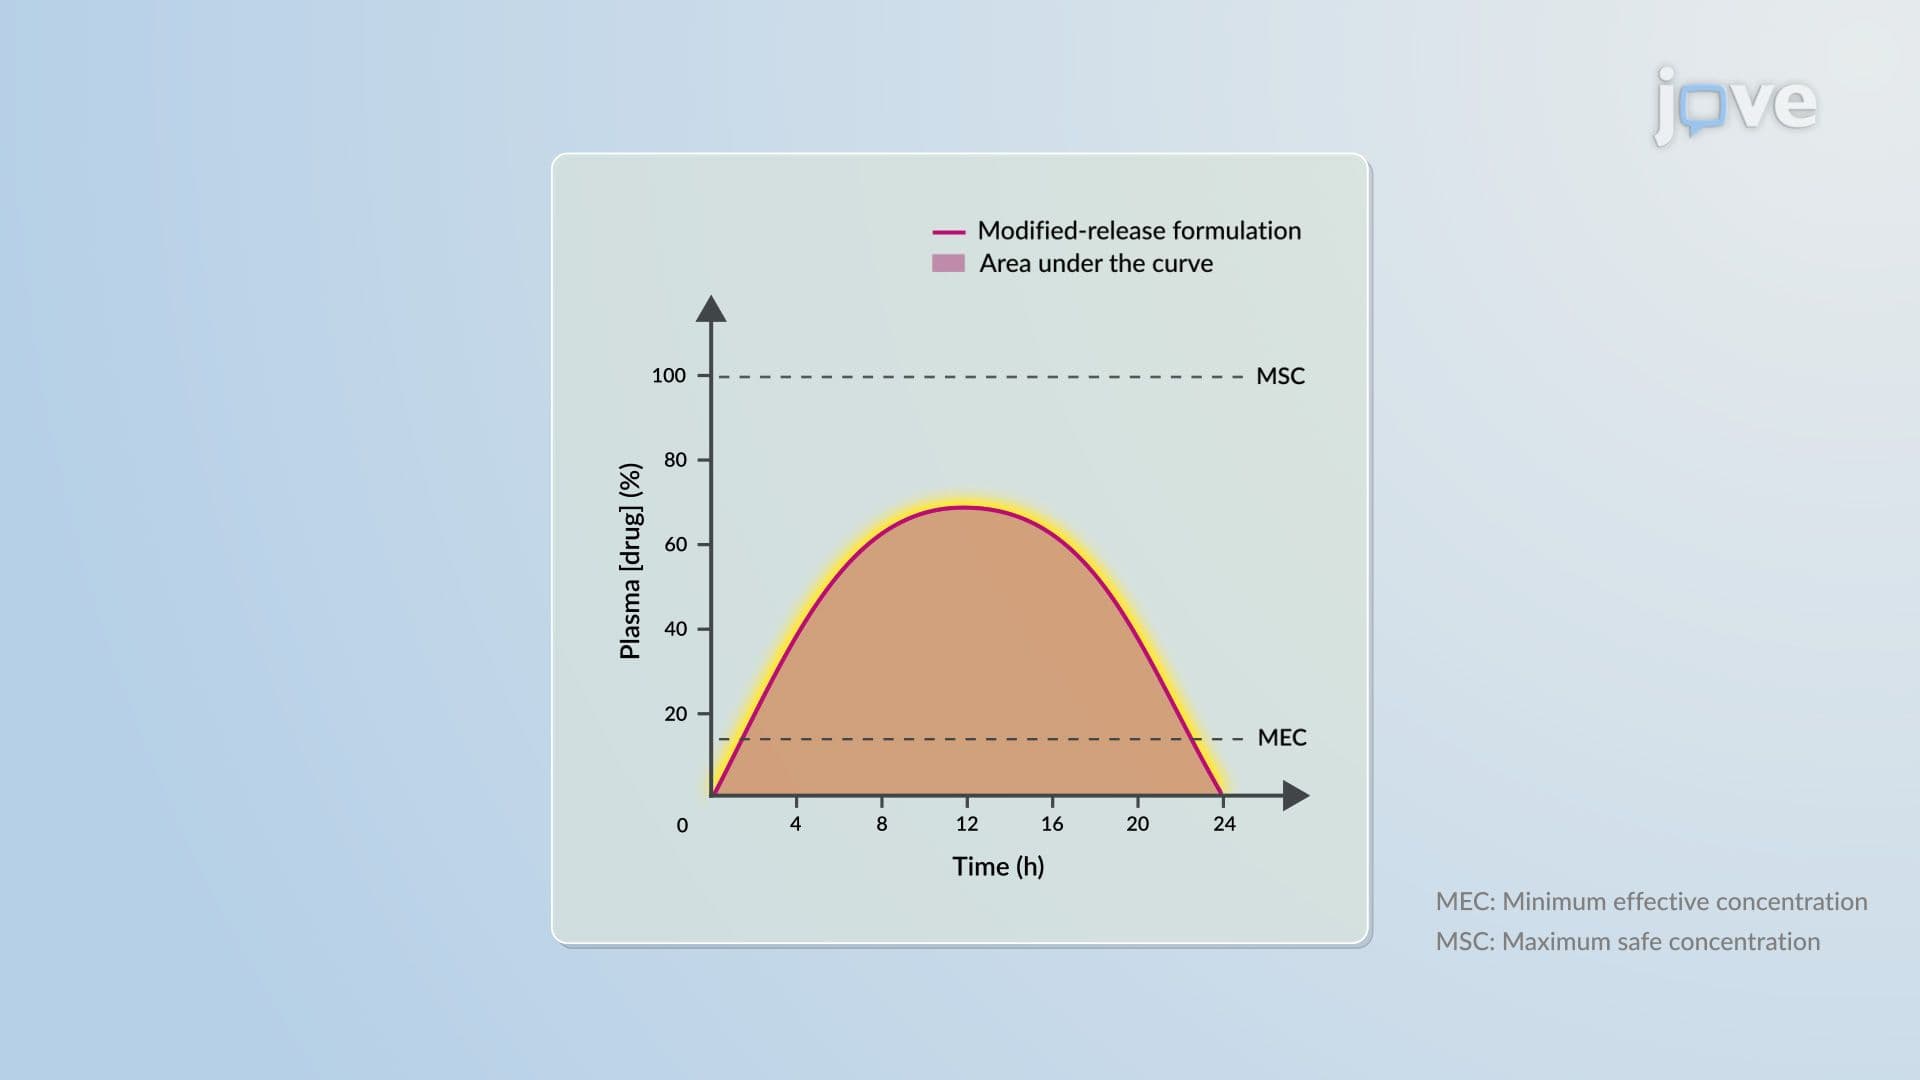
Modified-Release Drug Delivery Systems: Bioavailability
 01:21
Toxicokinetics: Overview
 01:23
Toxicity Testing in Animals