Normal mode analysis of macromolecular motions in a database framework: developing mode concentration as a useful classifying statistic
Related Experiment Video
Updated: Sep 29, 2026
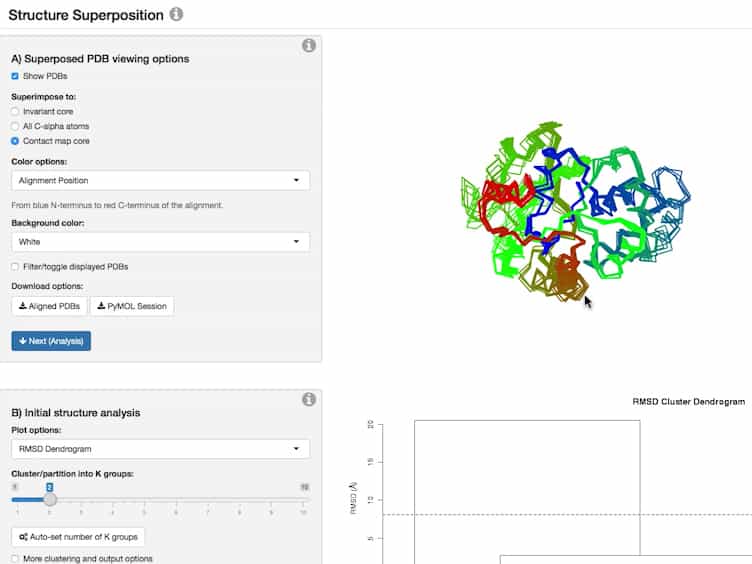
Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
W G Krebs1, Vadim Alexandrov, Cyrus A Wilson
1Department of Molecular Biophysics and Biochemistry, Yale University, New Haven, Connecticut 06520, USA.

Abstract:
We investigated protein motions using normal modes within a database framework, determining on a large sample the degree to which normal modes anticipate the direction of the observed motion and were useful for motions classification. As a starting point for our analysis, we identified a large number of examples of protein flexibility from a comprehensive set of structural alignments of the proteins in the PDB. Each example consisted of a pair of proteins that were considerably different in structure given their sequence similarity. On each pair, we performed geometric comparisons and adiabatic-mapping interpolations in a high-throughput pipeline, arriving at a final list of 3,814 putative motions and standardized statistics for each. We then computed the normal modes of each motion in this list, determining the linear combination of modes that best approximated the direction of the observed motion. We integrated our new motions and normal mode calculations in the Macromolecular Motions Database, through a new ranking interface at http://molmovdb.org. Based on the normal mode calculations and the interpolations, we identified a new statistic, mode concentration, related to the mathematical concept of information content, which describes the degree to which the direction of the observed motion can be summarized by a few modes. Using this statistic, we were able to determine the fraction of the 3,814 motions where one could anticipate the direction of the actual motion from only a few modes. We also investigated mode concentration in comparison to related statistics on combinations of normal modes and correlated it with quantities characterizing protein flexibility (e.g., maximum backbone displacement or number of mobile atoms). Finally, we evaluated the ability of mode concentration to automatically classify motions into a variety of simple categories (e.g., whether or not they are "fragment-like"), in comparison to motion statistics. This involved the application of decision trees and feature selection (particular machine-learning techniques) to training and testing sets derived from merging the "list" of motions with manually classified ones.
More Related Videos
 09:17
Structure-Based Simulation and Sampling of Transcription Factor Protein Movements along DNA from Atomic-Scale Stepping to Coarse-Grained Diffusion
Published on: March 1, 2022
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Related Concept Videos
 01:07
What is a Mode?
There can be more than one mode in a data set if multiple values have the same highest frequency. For instance, suppose that the Statistics exam scores of 20 students are: 50; 53; 59; 59; 63; 63; 72; 72; 72; 72; 72; 76; 78; 81; 83; 84; 84; 84; 90; 93. Here, the mode is 72, as it occurs most frequently, five times.
A data set with two modes is called bimodal. For example,...
 02:00
Molecular Models
 01:29
Mechanistic Models: Compartment Models in Algorithms for Numerical Problem Solving
In individual population analyses, different algorithms are employed, such as Cauchy's method, which uses a...