Related Experiment Video
Updated: Sep 29, 2026
An Electrochemiluminescence-Based Assay for MeCP2 Protein Variants
Published on: May 22, 2020
MECP2 mutations in Israel: implications for molecular analysis, genetic counseling, and prenatal diagnosis in Rett
Yuval Yaron1, Bruria Ben Zeev, Ruth Shomrat
1Genetic Institute & Prenatal Diagnosis Unit, Tel Aviv Sourasky Medical Center, Israel.

Abstract:
This report describes molecular analysis of the MECP2 gene in 37 Israeli patients suspected of having Rett syndrome (RTT). The patients were from various Jewish ethnic groups and from Arabic origin. Of the 17 patients with classical RTT, bi-directional sequencing of the coding exons revealed MECP2 mutations in 14 patients. About 66% of the mutations were located in previously described hot-spots. One case presented a novel mutation (141insA). Mutation-negative patients were further analyzed by Southern blot, which detected a novel gross rearrangement in another classical case. Altogether, detection rate in classical cases was 88%. In a non-classical case, a novel missense mutation (1451G>C) was detected in an affected girl and in her normal father, suggesting that this is a non-pathogenic alteration. Another variant (1461 + 96insA) was detected in an affected girl and in her healthy mother and also in another affected girl and her healthy father, suggesting that this variant too, is non-pathogenic. No significant difference in mutation type was noted among the different ethnic groups. In one familial case, the same mutation was detected in two sibs but not in their mother, suggesting germ-line mosaicism. Our results suggest that mutation-negative cases should be further assessed for gross rearrangements and that molecular analysis of the parents is often required when previously undescribed sequence alterations are detected.
More Related Videos
 08:27
A Non-random Mouse Model for Pharmacological Reactivation of Mecp2 on the Inactive X Chromosome
Published on: May 22, 2019
 08:22
A Novel Strategy Combining Array-CGH, Whole-exome Sequencing and In Utero Electroporation in Rodents to Identify Causative Genes for Brain Malformations
Published on: December 1, 2017
Related Concept Videos
 03:09
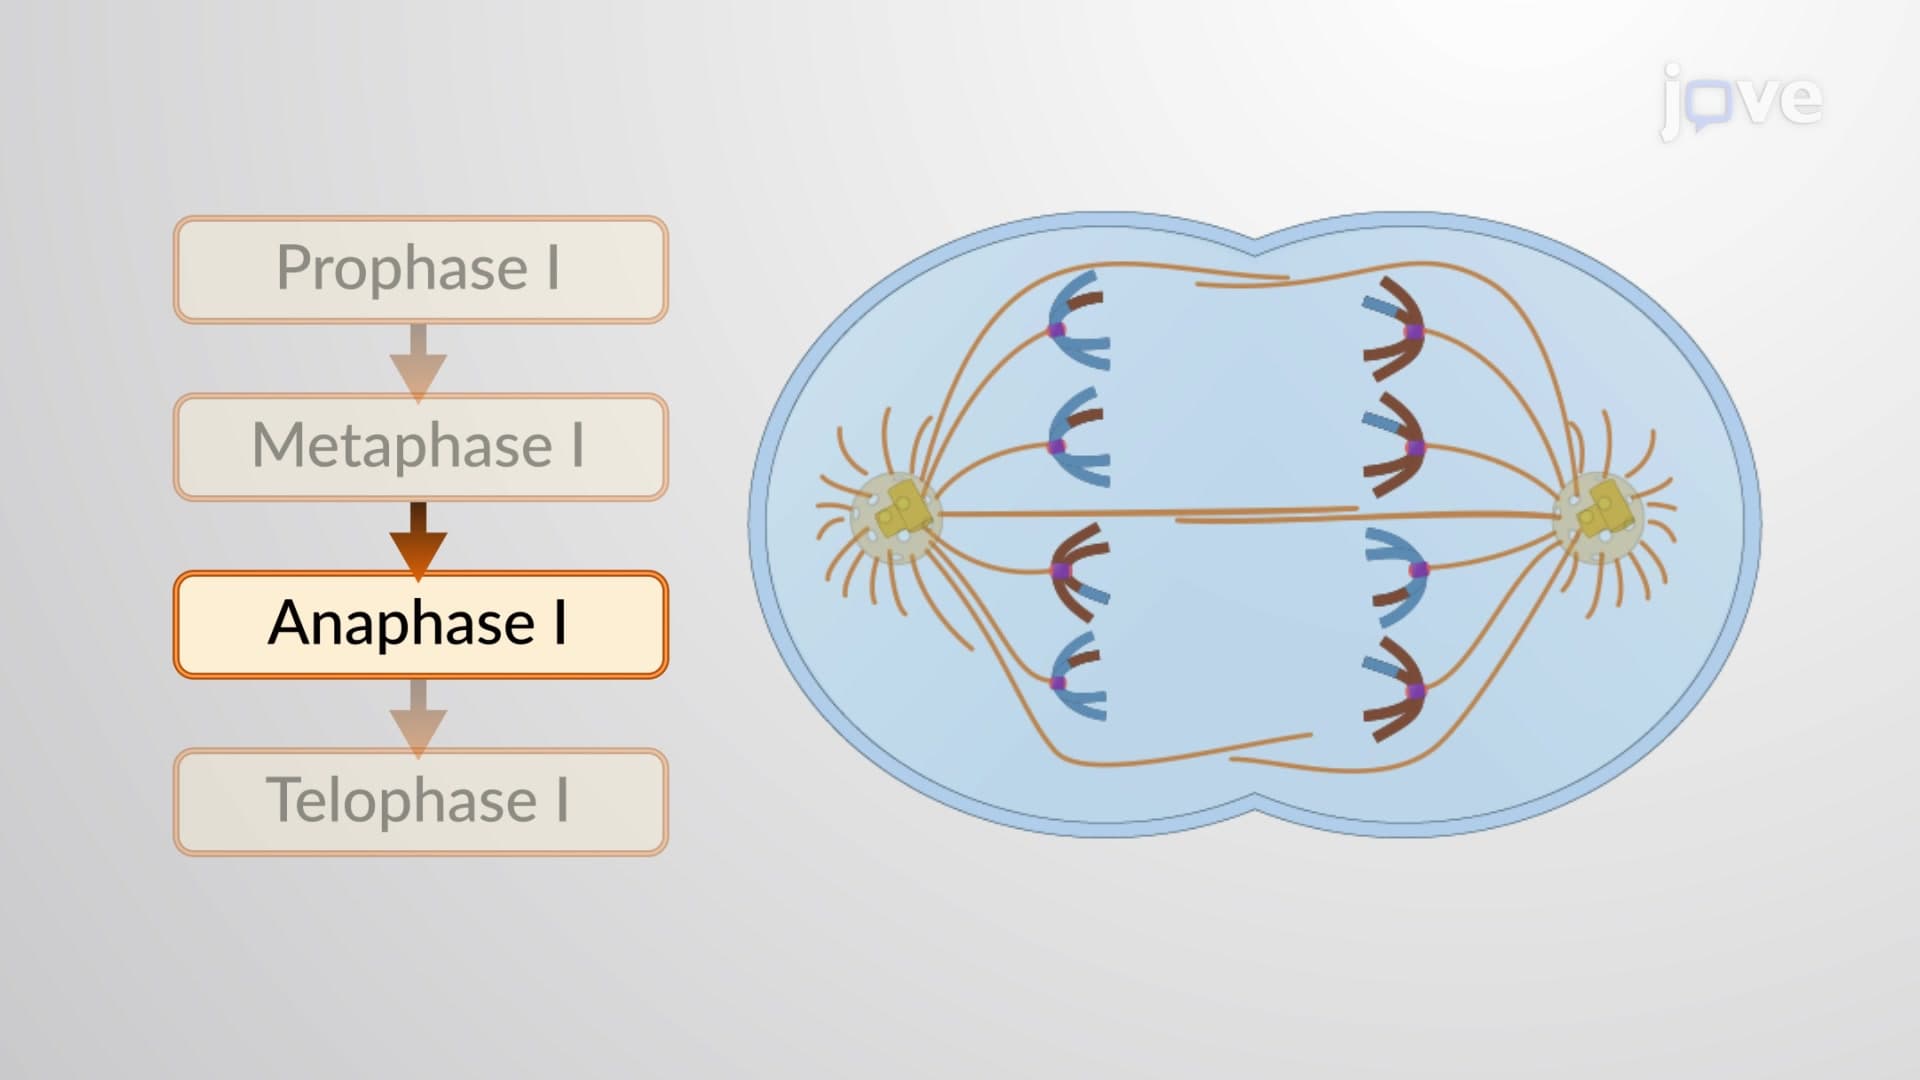
Meiosis I
Prophase I is the most extended and complex step of meiosis I characterized by synapsis, chromosome pairing, and recombination of the homologous chromosomes. This process is facilitated by a proteinaceous structure called the...
 01:20
Mismatch Repair
The Mutator Protein Family Plays a Key Role in DNA Mismatch Repair
The human genome has more than 3 billion base pairs of DNA per cell. Prior to cell division, that vast amount of genetic...
 03:18
Non-LTR Retrotransposons
 01:20
The Retinoblastoma Gene
The first-ever tumor suppressor gene called Rb was identified in retinoblastoma - a rare eye tumor in children. In inherited forms of the disease, a child inherits one defective copy of the Rb gene, which predisposes them to retinoblastoma. However,...