Related Experiment Video
Updated: Sep 29, 2026

Characterizing Histone Post-translational Modification Alterations in Yeast Neurodegenerative Proteinopathy Models
Published on: March 24, 2019
Heterogeneity of presenile dementia with bone cysts (Nasu-Hakola disease): three genetic forms
T Kondo1, K Takahashi, N Kohara
1Department of Immunology, National Institute of Neuroscience, National Center for Neurology and Psychiatry, Kodairo, Tokyo, Japan.

Insights
Nasu-Hakola disease (NHD) in Japanese patients is often caused by mutations in the DAP12 gene, leading to presenile dementia and bone cysts. Genetic analysis revealed loss-of-function mutations in five out of six cases studied.
Area of Science:
- Genetics
- Neurology
- Bone Biology
Background:
- Nasu-Hakola disease (NHD) is a rare autosomal recessive disorder.
- NHD presents with presenile dementia and bone cysts.
- Previous studies identified DAP12 gene deletions in Finnish NHD patients.
Purpose of the Study:
- To investigate the genetic basis of NHD in Japanese patients.
- To identify mutations in the DAP12 gene in Japanese NHD cases.
- To understand the genetic heterogeneity of NHD in Japan.
Main Methods:
- Genetic analysis of DAP12 alleles in six Japanese NHD patients.
- Mutation screening including deletion and point mutation analysis.
- Assessment of DAP12 protein expression in a patient without mutation.
Main Results:
- Five of the six Japanese NHD patients had loss-of-function mutations in DAP12.
- Identified mutations included a single-base deletion and a novel point mutation.
- One patient without DAP12 mutation showed normal protein expression, suggesting other genetic forms.
Conclusions:
- The DAP12 gene is a significant cause of NHD in Japanese populations.
- Japanese NHD exhibits genetic heterogeneity, with at least three forms related to DAP12.
- Further research is needed to elucidate the genetic basis in all Japanese NHD cases.
Abstract:
Nasu-Hakola disease (NHD) is an autosomal recessive disorder characterized by presenile dementia and bone cysts. Finnish patients revealed a large deletion in DAP12 gene encoding a key element for transducing activation signal. The authors examined six Japanese cases for DAP12 alleles. Five of the six had loss-of-function mutation, either a single-base deletion or a novel point mutation. The single patient without mutation normally expressed DAP12 protein. Japanese NHD has at least three genetic forms regarding DAP12.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:22
Lysosomal Hydrolases
 01:29
Alzheimer Disease l: Introduction
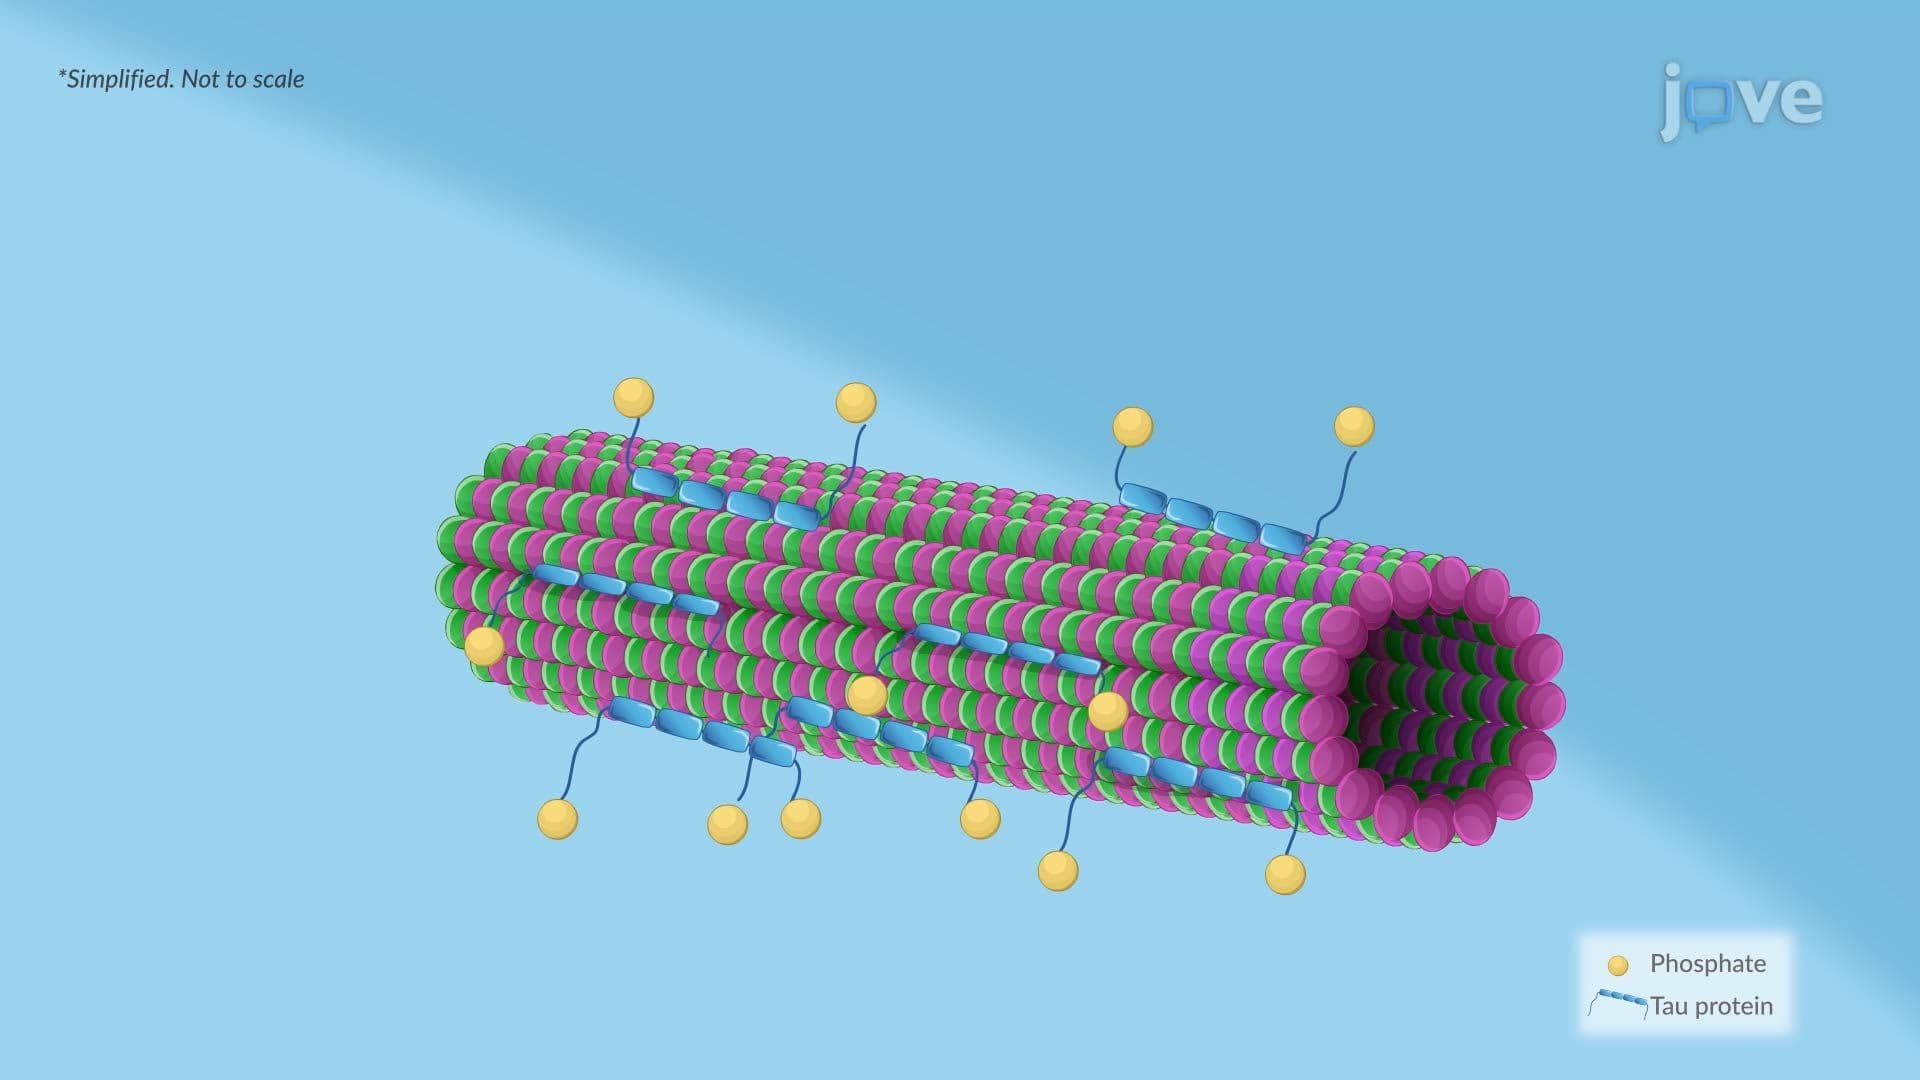 01:23
Alzheimer Disease ll: Pathophysiology
 01:22
Dementia l: Introduction
 01:24
Parkinson Disease ll: Pathophysiology