Related Experiment Video
Updated: Jul 28, 2026
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
Terminal glycosylation in cystic fibrosis (CF): a review emphasizing the airway epithelial cell
A D Rhim1, L Stoykova, M C Glick
1The Cystic Fibrosis Center and Department of Pediatrics, University of Pennsylvania School of Medicine, The Children's Hospital of Philadelphia, 3516 Civic Center Boulevard, Philadelphia, PA 19104, USA.

Abstract:
Altered terminal glycosylation, with increased fucosylation and decreased sialylation is a hallmark of the cystic fibrosis (CF) glycosylation phenotype. Oligosaccharides purified from the surface membrane glycoconjugates of CF airway epithelial cells have the Lewis x, selectin ligand in terminal positions. This review is focused on the investigations of the glycoconjugates of the CF airway epithelial cell surface. Two of the major bacterial pathogens in CF, Pseudomonas aeruginosa and Haemophilus influenzae, have binding proteins which recognize fucose in alpha-1,3 linkage and asialoglycoconjugates. Therefore, consideration has been given to the possibility that the altered terminal glycosylation of airway epithelial glycoproteins in CF contributes to both the chronic infection and the robust, but ineffective, inflammatory response in the CF lung. Since the glycosylation phenotype of CF airway epithelial cells have been modulated by the expression of wtCFTR, the hypotheses which have been proposed to relate altered function of CFTR to the regulation of the glycosyltransferases are discussed. Understanding the effects of mutant CFTR on glycosylation may provide further insight into the regulation of glycoconjugate processing as well as new approaches to the therapy of CF.
More Related Videos
 10:18
Real-Time, Semi-Automated Fluorescent Measurement of the Airway Surface Liquid pH of Primary Human Airway Epithelial Cells
Published on: June 13, 2019
 05:56
Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
Related Concept Videos
 01:25
Protein Glycosylation
Glycosylation occurs in...
 01:24
Oligosaccharide Assembly
Multiple sugar molecules that may or may...
 01:05
Proteoglycans
 01:14
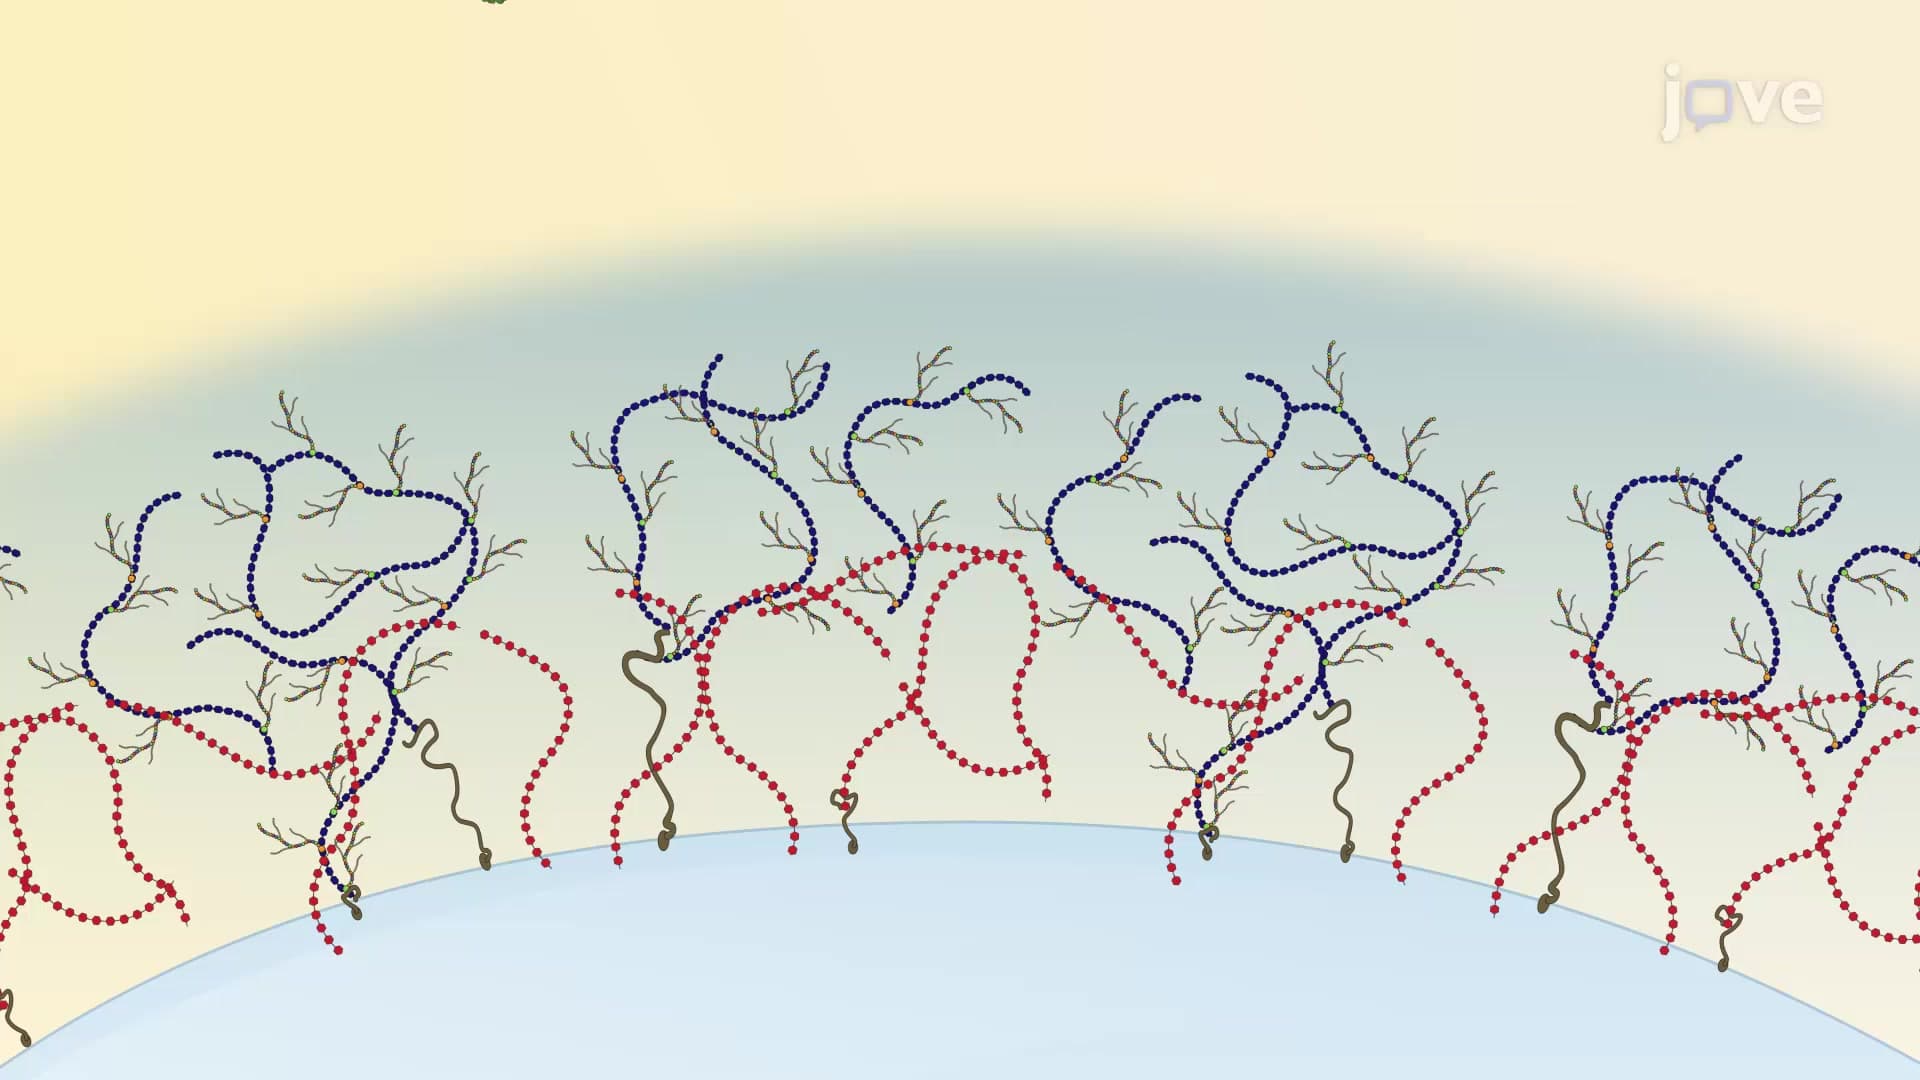
Glycocalyx and its Functions
Components of...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:19
Overview of Carbohydrate Metabolism
Glucose transport into cells is facilitated by a family of transport proteins called GLUT (Glucose Transporters). GLUT4 is the primary glucose transporter for insulin-stimulated glucose...