Related Experiment Video
Updated: Sep 28, 2026
An Intestine/Liver Microphysiological System for Drug Pharmacokinetic and Toxicological Assessment
Published on: December 3, 2020
The influence of nonspecific microsomal binding on apparent intrinsic clearance, and its prediction from
Rupert P Austin1, Patrick Barton, Scott L Cockroft
1Department of Physical and Metabolic Science, AstraZeneca R&D Charnwood, Loughborough, United Kingdom. rupert.austin@astrazeneca.com

Abstract:
The apparent intrinsic clearance of 13 drugs has been determined using rat liver microsomes at three different concentrations of microsomal protein. The kinetics was studied using the in vitro half-life method. The nonspecific binding of these drugs to the microsomes was also studied under the same conditions, except for cofactor removal, using equilibrium dialysis. The intrinsic clearances are shown to be dependent on the microsomal concentration, but are approximately constant when corrected for the extent of nonspecific binding to the microsomes. The large difference between observed intrinsic clearance and unbound intrinsic clearance that exists for some compounds, particularly lipophilic bases, is highlighted. A simple model has been developed for understanding the binding of compounds to microsomes and is demonstrated to accurately predict the extent of microsomal binding at one concentration of microsomes from measurement at another. The binding of a further 25 drugs to rat liver microsomes at a microsomal concentration of 1 mg/ml was also studied, along with measurements of lipophilicity using octanol-water partition coefficients. It is shown that the extent of microsomal binding is correlated with lipophilicity, but that basic compounds show a different behavior to acidic and neutral compounds. Microsomal binding is shown to be best predicted using a model where log P is used for basic compounds, and log D(7.4) is used for acidic and neutral compounds. This model has been developed further so that the extent of binding to microsomes of any given concentration can be estimated purely from a knowledge of lipophilicity and ionization.
Related Concept Videos
 01:09
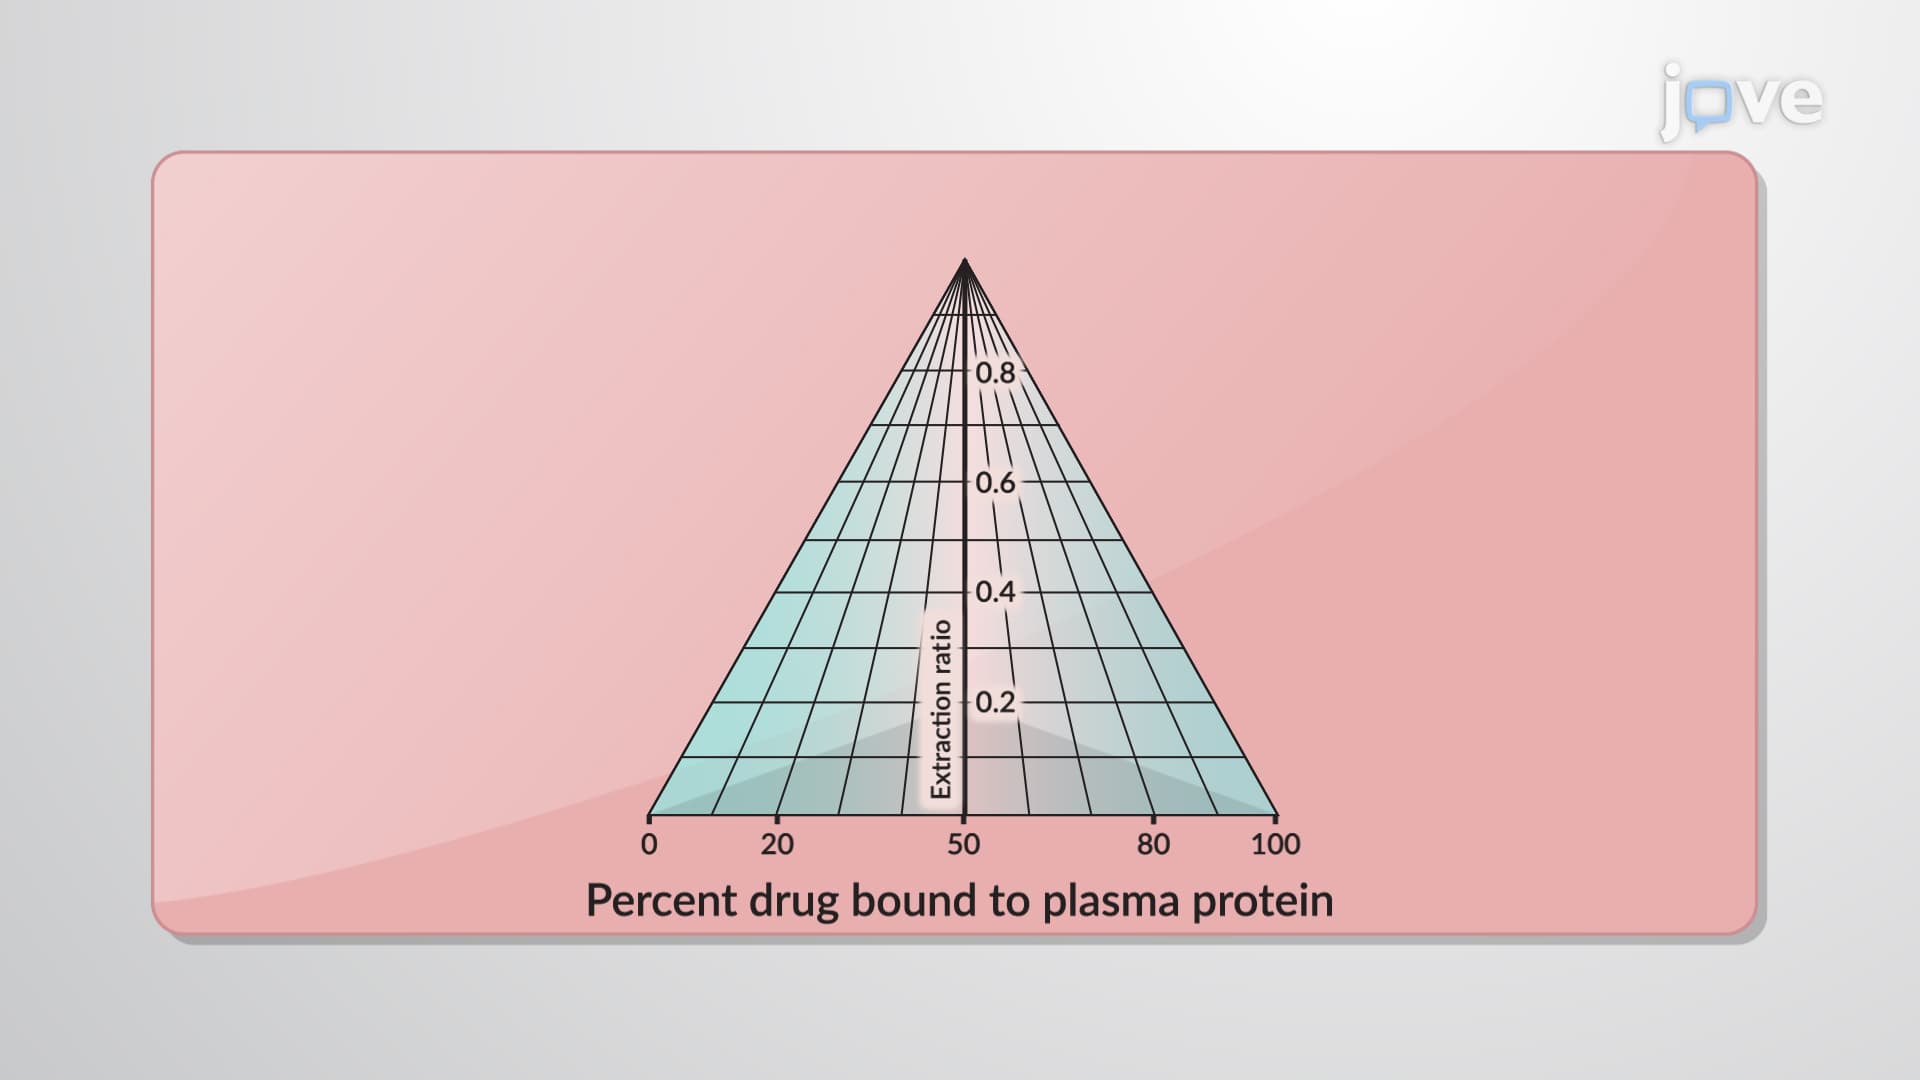
Hepatic Drug Clearance: Effect of Protein Binding
For low-extraction-ratio drugs that are less than 80% protein-bound, minor changes in protein binding...
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:07
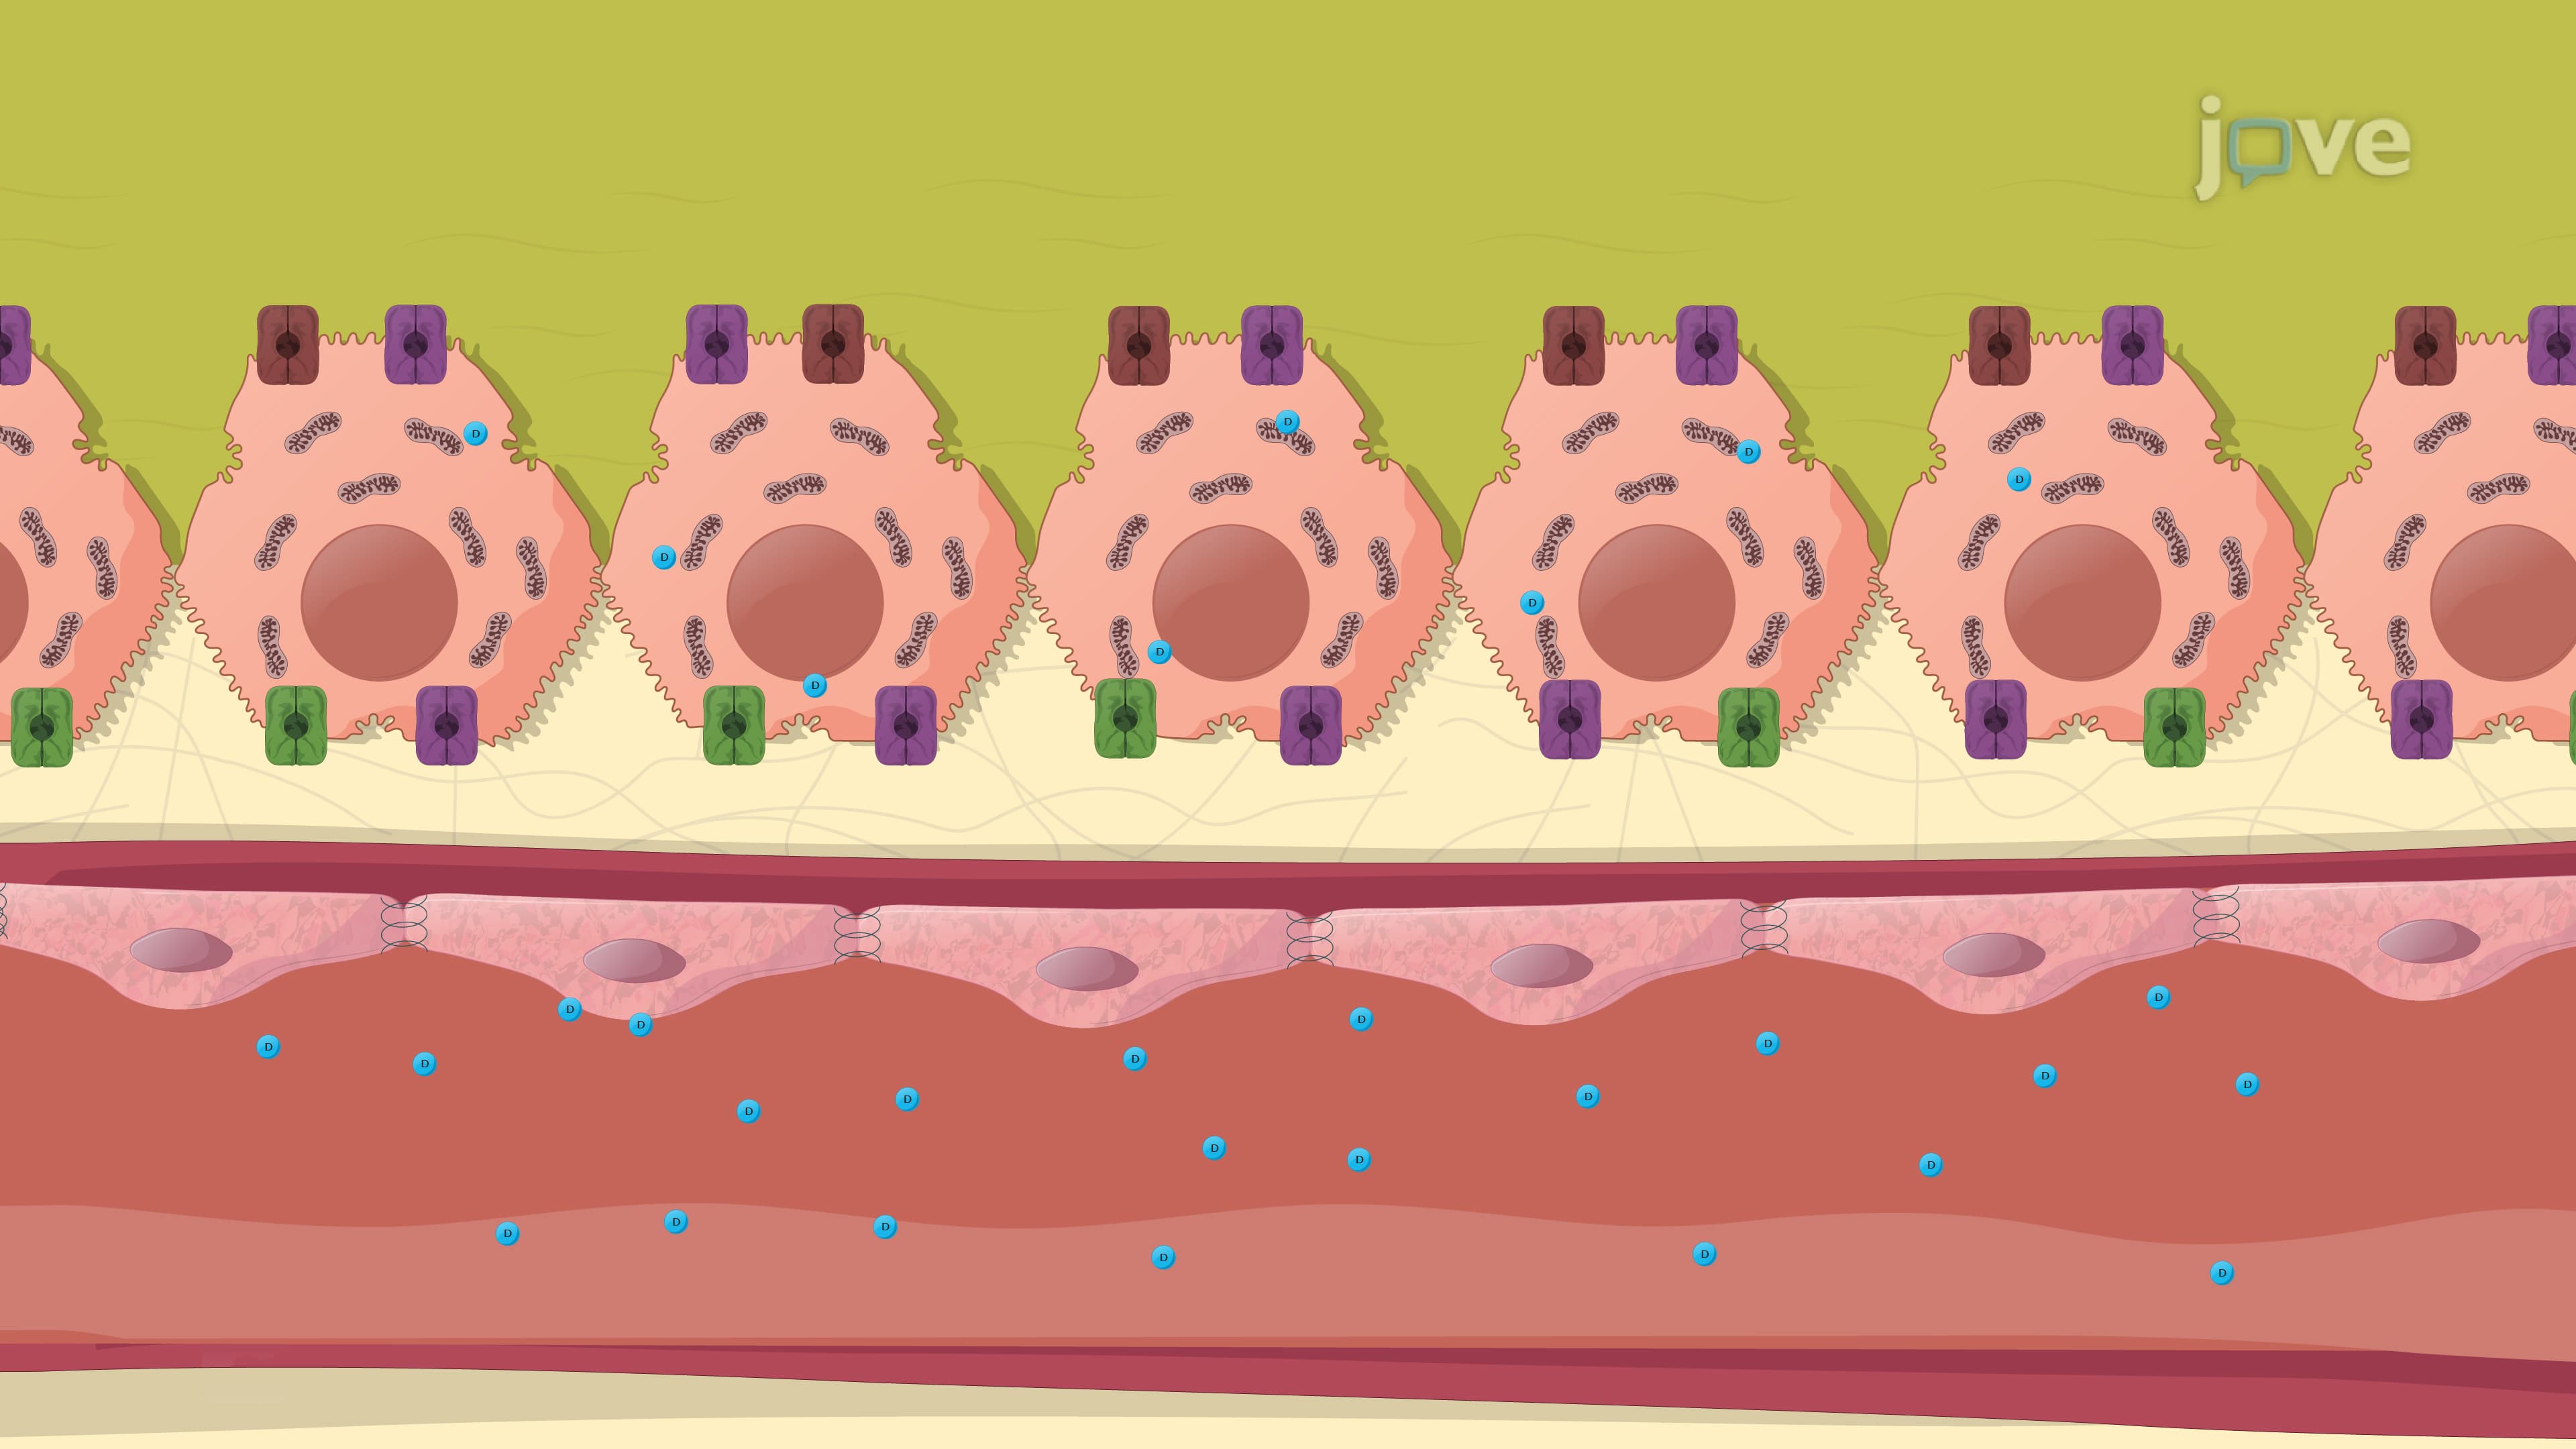
Physiological Pharmacokinetic Models: Incorporating Hepatic Transporter-Mediated Clearance
A recent model describes pravastatin's hepatobiliary excretion, mediated...
 01:31
Factors Affecting Renal Clearance: Drug's Physicochemical Properties and Plasma Levels
One important factor is the drug's molecular size. The kidneys readily excrete smaller molecules below 300 Daltons (Da). On the other hand, molecules weighing between 300 and 500 Da are excreted through both urine and bile. Larger molecules above 500 Da tend to be excreted less...
 00:54
Noncompartmental Analysis: Miscellaneous Pharmacokinetic Parameters
One key aspect of the noncompartmental approach is determining a drug's total clearance. This can be done by dividing the drug dose by the area under the concentration-time curve from zero to infinity. The area under the concentration-time curve represents the drug's overall...
 01:14
Hepatic Drug Clearance: Role of Transporters