Related Experiment Video
Updated: Sep 28, 2026
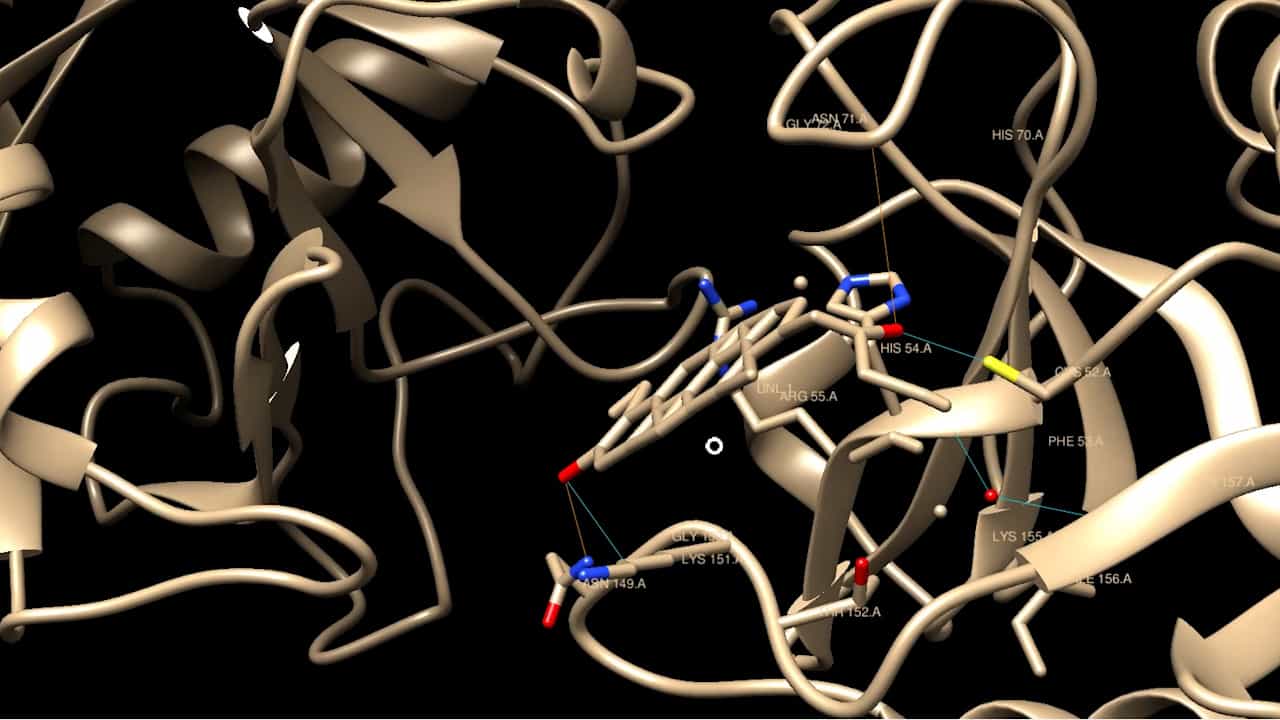
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Methodology and problems of protein-ligand docking: case study of dihydroorotate dehydrogenase, thymidine kinase, and
Pavel Pospisil1, Thomas Kuoni, Leonardo Scapozza
1Department of Applied Biosciences, Swiss Federal Institute of Technology (ETH) Zürich, Winterthurerstrasse 190, CH-8057 Zürich, Switzerland.

Abstract:
The docking methodology was applied to three different therapeutically interesting enzymes: human dihydroorotate dehydrogenase (DHODH), Herpes simplex virus type I thymidine kinase (HSV1 TK) and human phosphodiesterase 4 (PDE4). Programs FlexX, AutoDock and DOCK where used. The three targets represent three distinct cases. For DHODH and HSV1 TK, the binding modes of substrate and inhibitors within the active site are known, while the binding orientation of cAMP within PDE4 has been solely hypothesized. Active site of DHODH is mainly hydrophobic and the binding mode of the inhibitor brequinar was used as a template for evaluating the docking strategies. The presence of cofactors revealed to be crucial for the definition of the docking site. The HSV1 TK active site is small and polar and contains crystal water molecules and ATP. Docking of thymidine and aciclovir (ACV) within the active site was analyzed by keeping or removing water molecules. It showed the crucial role of water in predicting the binding of pyrimidines and purines. The crystal structure of PDE4 contains magnesium and zinc cations as well as catalytic water molecule but no ligand. Several docking experiments of cAMP and rolipram were performed and the results showed clear-cut dependence between the ligand orientation and the presence of metals in the active site. All three cases show specific problems of the docking methodology, depending on the character of the active site.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 00:49
Ligand Binding and Linkage
 01:19
Allosteric Proteins-ATCase
Aspartate transcarbamoylase (ATCase) is a cytosolic enzyme that catalyzes the condensation of L-aspartate and carbamoyl phosphate to N-carbamoyl-L-aspartate. This reaction is the first step in pyrimidine biosynthesis. UTP and CTP, the end products of the pyrimidine synthesis pathway,...
 02:18
The Equilibrium Binding Constant and Binding Strength
 02:04
Protein-protein Interfaces
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...