Related Experiment Video
Updated: Sep 27, 2026


Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
Progressive familial intrahepatic cholestasis
Giulia Martina Cavestro1, Luca Frulloni, Elena Cerati
1Dept. of Clinical Science, Chair of Gastroenterology, University of Parma, Parma. giuliamartina@yahoo.it

Insights
Progressive familial intrahepatic cholestasis (PFIC) encompasses genetic liver diseases in children. Mutations in ATP8B1, BSEP, and MDR genes cause PFIC types 1, 2, and 3, respectively, leading to liver failure.
Area of Science:
- Hepatology
- Genetics
- Pediatric Gastroenterology
Background:
- Progressive familial intrahepatic cholestasis (PFIC) is a group of inherited liver diseases in children.
- PFIC presents with varying genetic causes and clinical manifestations, including cholestasis and liver failure.
Purpose of the Study:
- To review the genetic basis and clinical features of PFIC types 1, 2, and 3.
- To discuss current therapeutic strategies for PFIC.
Main Methods:
- Review of literature on PFIC genetics and clinical presentations.
- Analysis of therapeutic interventions for different PFIC subtypes.
Main Results:
- PFIC 1 (Byler disease) involves ATP8B1 gene mutations, presenting early with cholestasis.
- PFIC 2 is linked to Bile Salt Export Pump (BSEP) defects, causing similar symptoms.
- PFIC 3, associated with MDR gene mutations, appears later and carries risks of portal hypertension and liver failure.
Conclusions:
- PFIC subtypes have distinct genetic origins and clinical trajectories.
- While external biliary diversion and ursodeoxycholic acid are initial options, liver transplantation remains a primary treatment for many PFIC patients.
Abstract:
Progressive familial intrahepatic cholestasis (PFIC) is a heterogeneous group of autosomal recessive childhood cholestasis of hepatocellular origin. PFIC 1, also known as Byler disease, was first described in Amish kindred. It is characterized by cholestasis often arising in the neonatal period and it leads to death due to liver failure. PFIC 1, like Benign Recurrent Intrahepatic Cholestasis (BRIC) which is the benign form of the same disease, recognizes mutations in the ATP8B1 gene. PFIC 2 disease is clinically similar to PFIC 1 but it has a different gene mutation causing a defect in the Bile Salt Export Pump (BSEP), exclusively expressed in the liver and involved in the canalicular secretion of bile acids. PFIC 3 usually appears later in life and it has a higher risk of portal hypertension, gastrointestinal bleeding and liver failure. This particular form of disease (the only one with high serum values of g-glutamil transpeptidase), is associated to a genetic defect in the class III multidrug resistance protein (MDR). External biliary diversion and ursodeoxycholic acid therapy, should be considered as the initial therapy in these patients, even if liver transplantation still seems to be the only solution for most patients.
Related Concept Videos
 01:25
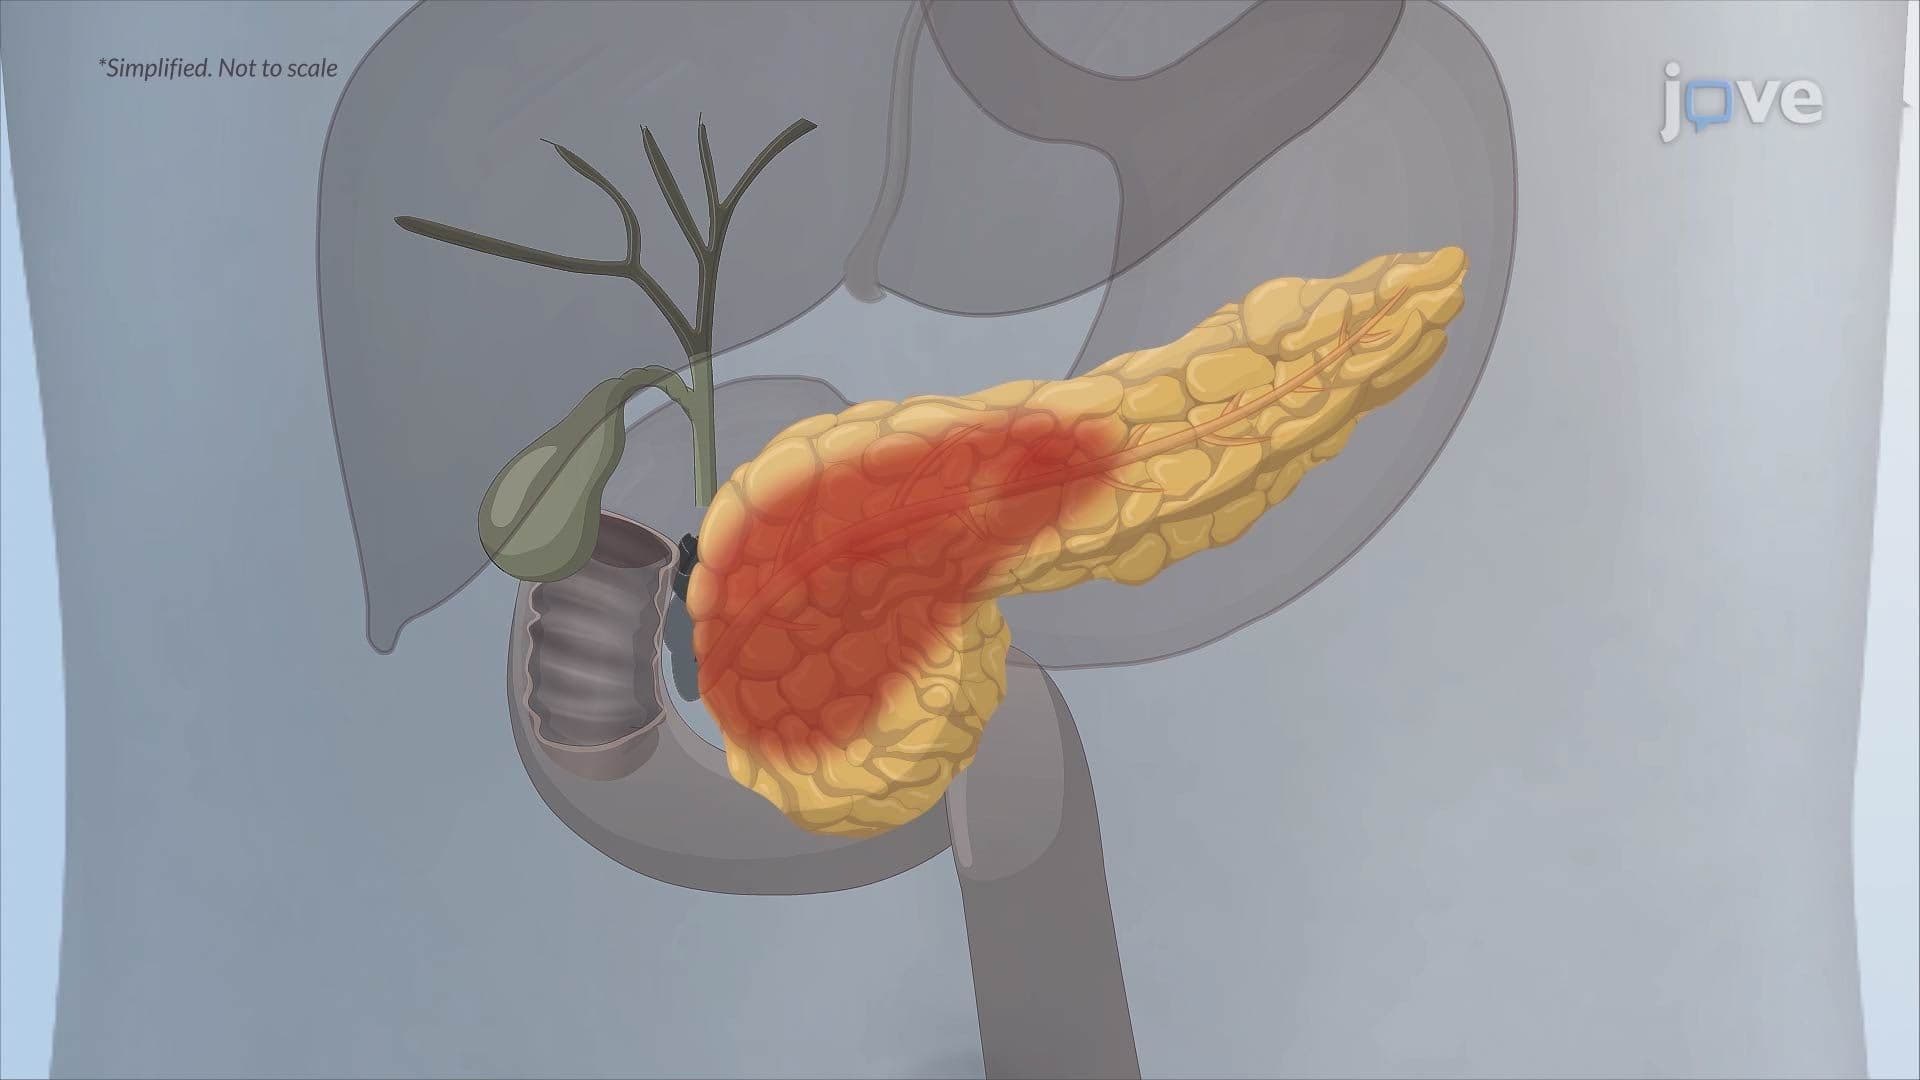
Chronic Pancreatitis I: Introduction
 01:24
Chronic Pancreatitis I: Introduction
Pancreatitis is the inflammation of the pancreas, which occurs when the immune system becomes active and causes swelling, pain, and disruptions in organ function. Pancreatitis can manifest as either an acute or chronic condition.
Acute pancreatitis arises suddenly and lasts for a brief duration, while chronic pancreatitis is a long-term affliction...
 01:21
Chronic Pancreatitis II: Pathophysiology
 01:24
Cirrhosis II: Pathophysiology
 01:26
Diseases of the Liver and Gallbladder
Cirrhosis is characterized by the scarring of hepatic lobules in the liver, which are replaced by fibrous tissue, affecting the liver's normal functioning. NAFLD, on the other hand, is caused by an excessive build-up of fat in the liver, not related to...
 01:20
Cholecystitis