Related Experiment Video
Updated: Sep 27, 2026

Using Phylogenetic Analysis to Investigate Eukaryotic Gene Origin
Published on: August 14, 2018
Simultaneous sequence alignment and tree construction using hidden Markov models
Robert C Edgar1, Kimmen Sjölander
1Department of Bioengineering, University of California, Berkeley, California 94720, USA. bob@drive5.com

Abstract:
We present a new algorithm (SATCHMO) that simultaneously estimates a tree and generates a set of multiple sequence alignments given a set of protein sequences. Alignments are constructed for each node in the tree. These alignments predict the structurally conserved elements of the sequences in a subtree and are therefore of different lengths, and represent different amino acid preferences, at different nodes. Hidden Markov Models (HMMs) are also generated for each node and are used to determine branching order, to align sequences and to predict structurally alignable regions. In experiments on the BAliBASE benchmark alignment database, SATCHMO is shown to perform comparably to ClustalW and the UCSC SAM HMM software. Results using SATCHMO to identify protein domains are demonstrated on potassium channels, with implications for the mechanism by which tumor necrosis factor alpha affects potassium current.
Related Concept Videos
 02:54
Evolutionary Relationships through Genome Comparisons
 02:51
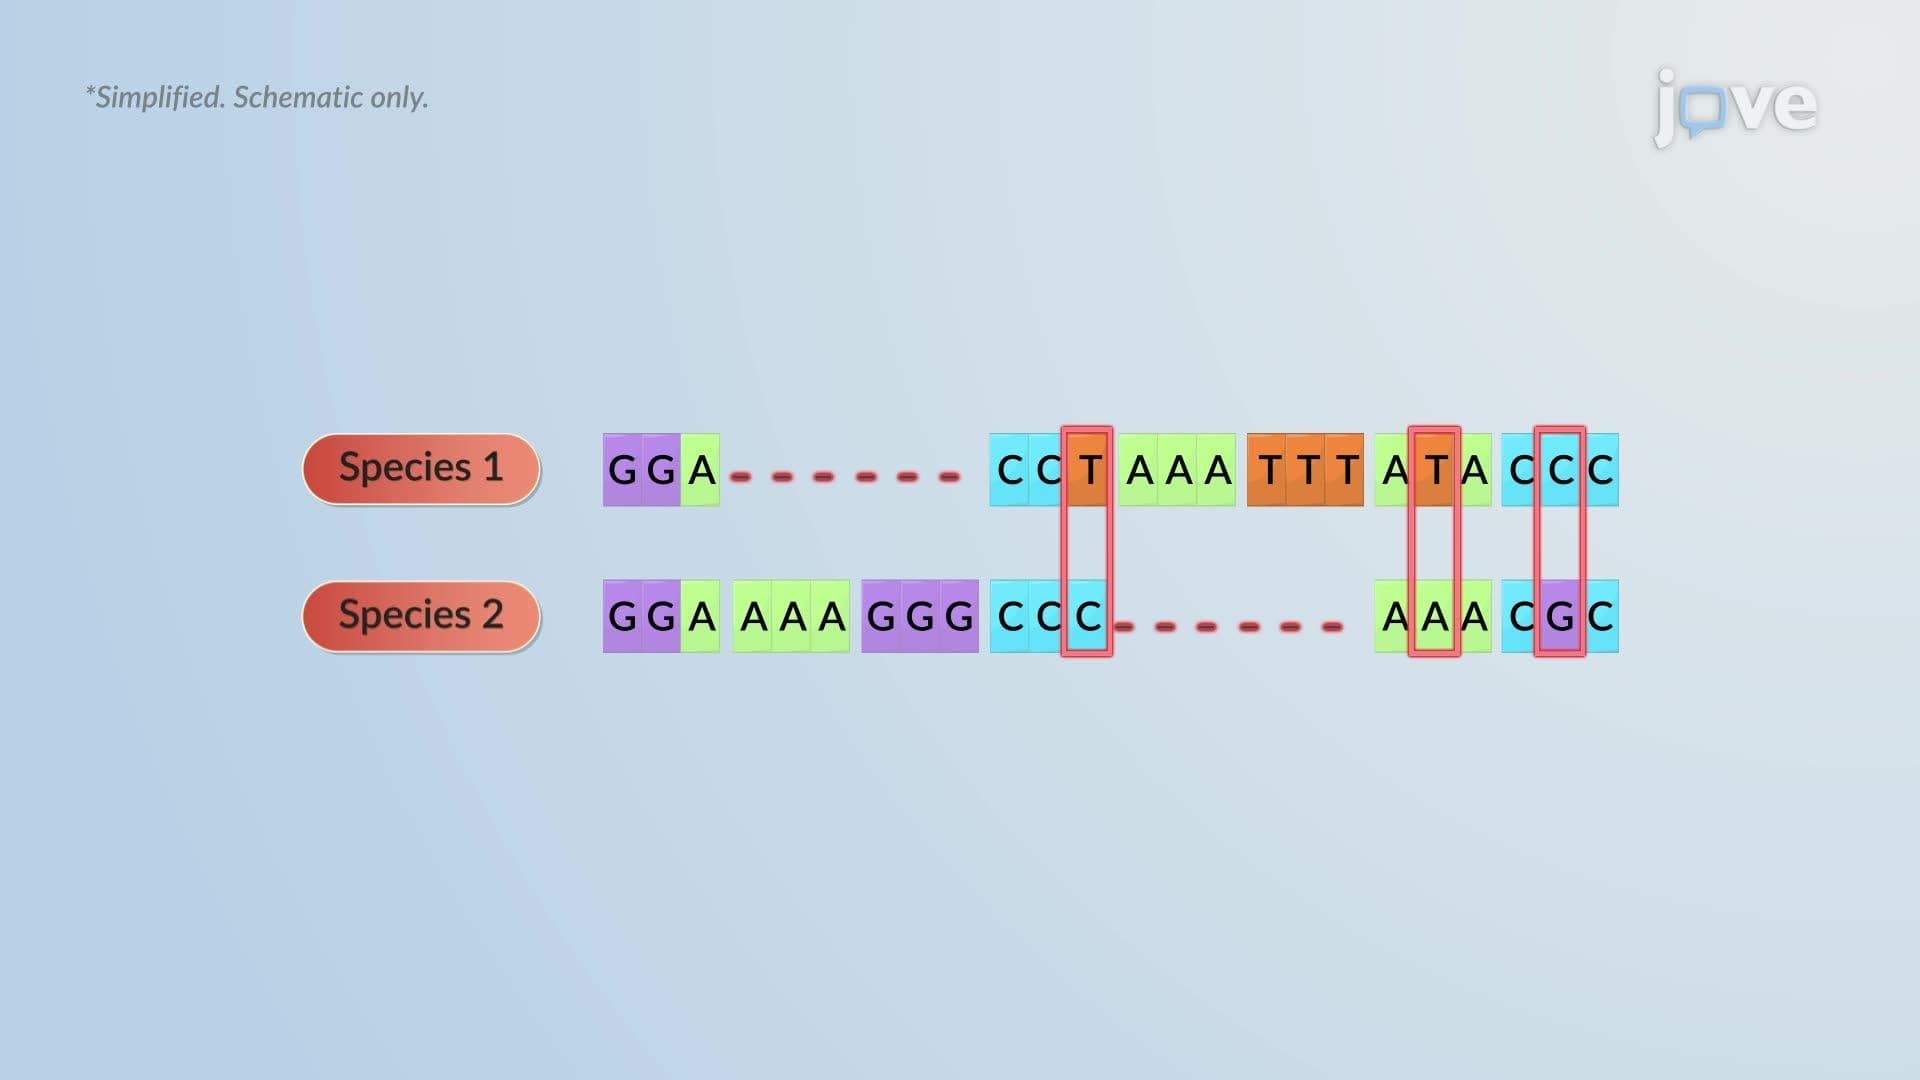
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved DNA...
 01:28
Microbial Phylogeny
 03:21
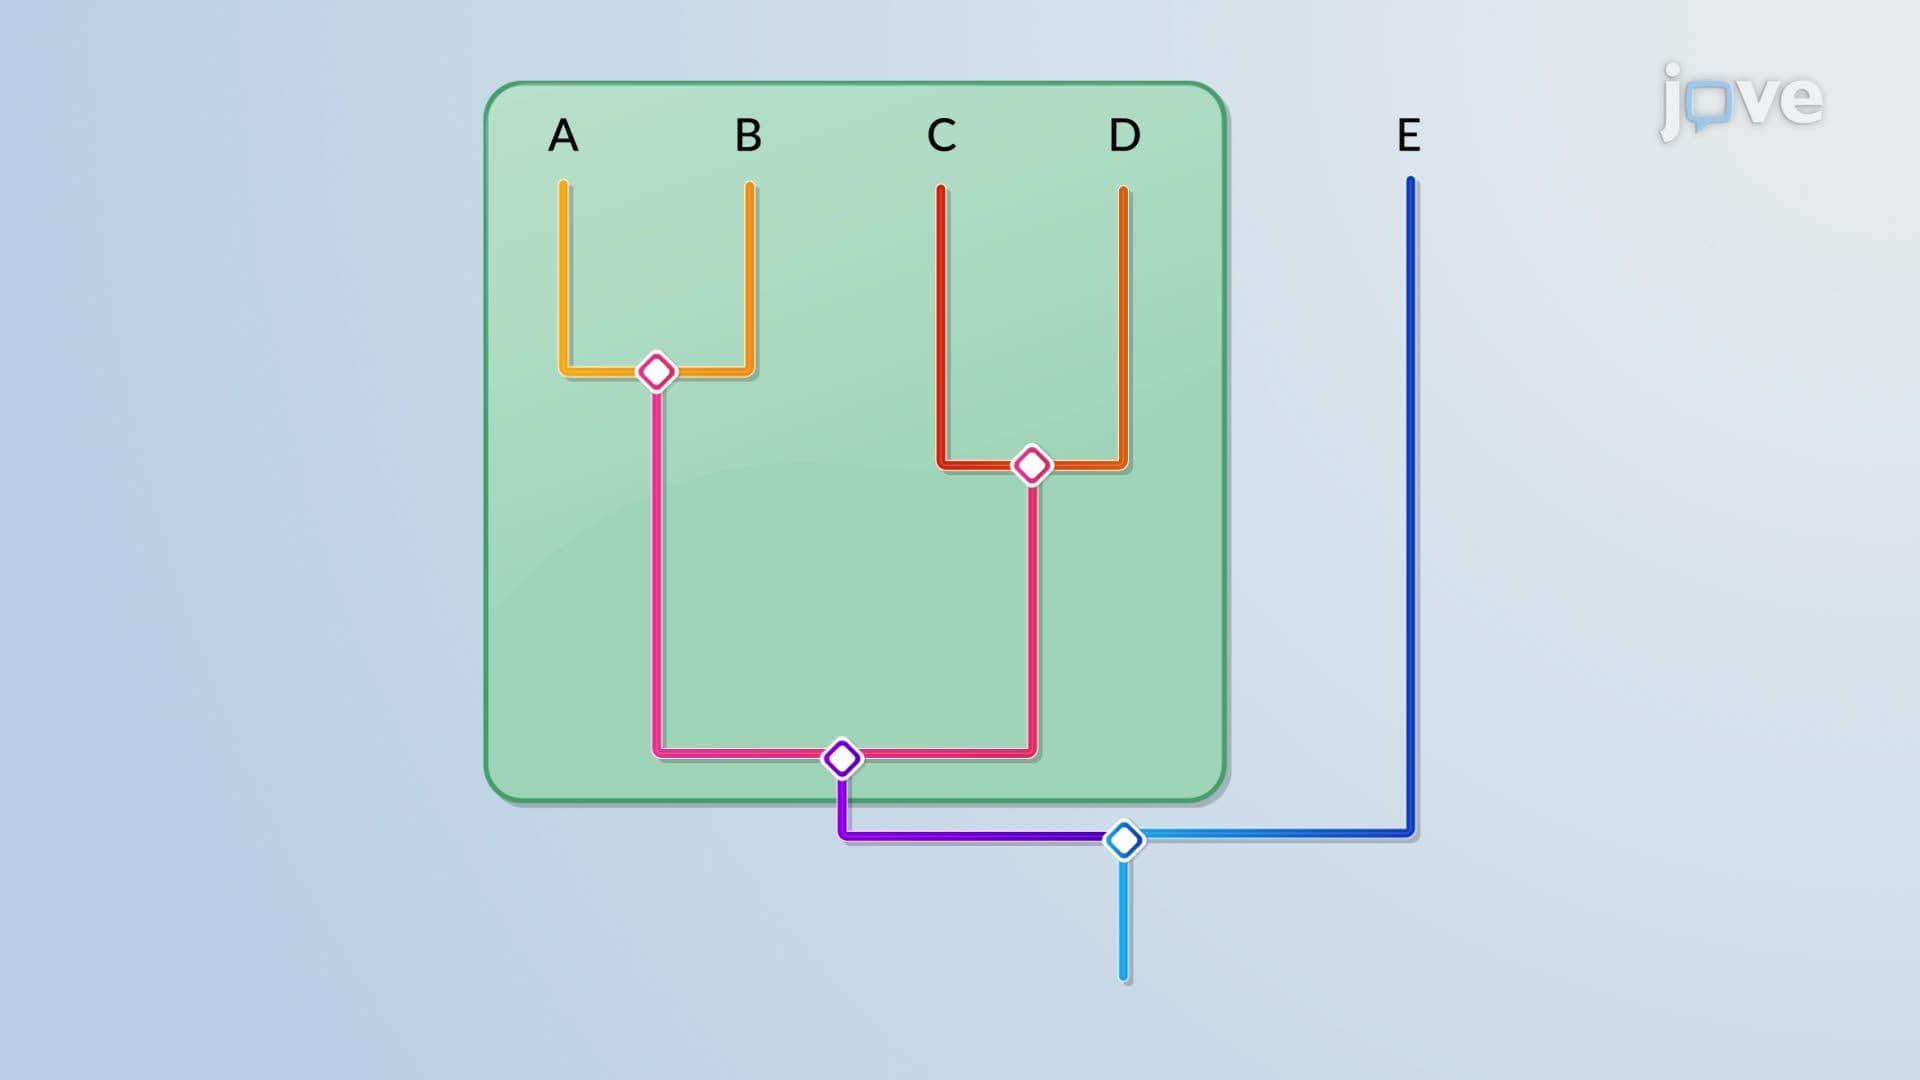
Phylogenetic Trees
03:21Phylogenetic Trees
 01:29
Modern Molecular Taxonomy