Related Experiment Video
Updated: Sep 26, 2026

Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
Pulmonary alveolar proteinosis mimicking idiopathic pulmonary fibrosis
Zoya K Arbiser1, David M Guidot, Jeffrey R Pine
1Department of Pathology and Laboratory Medicine, Emory University Hospital, Atlanta, GA 30322, USA.

Abstract:
Pulmonary alveolar proteinosis (PAP) is a rare disorder of unknown etiology. While most biopsies from patients with PAP disclose a prominent positive periodic acid-Schiff, diastase-resistant intra-alveolar exudate, and minimal interstitial thickening, interstitial fibrosis and honeycombing can be the predominant histologic findings. We recently identified two patients with PAP who initially were thought to have idiopathic pulmonary fibrosis, but whose biopsies ultimately disclosed PAP with considerable interstitial fibrosis. Because the clinical and radiographic presentation of PAP can overlap with other interstitial lung disorders, it should be considered in the differential diagnosis of unexplained idiopathic interstitial pneumonias in young patients.
Related Concept Videos
 01:23
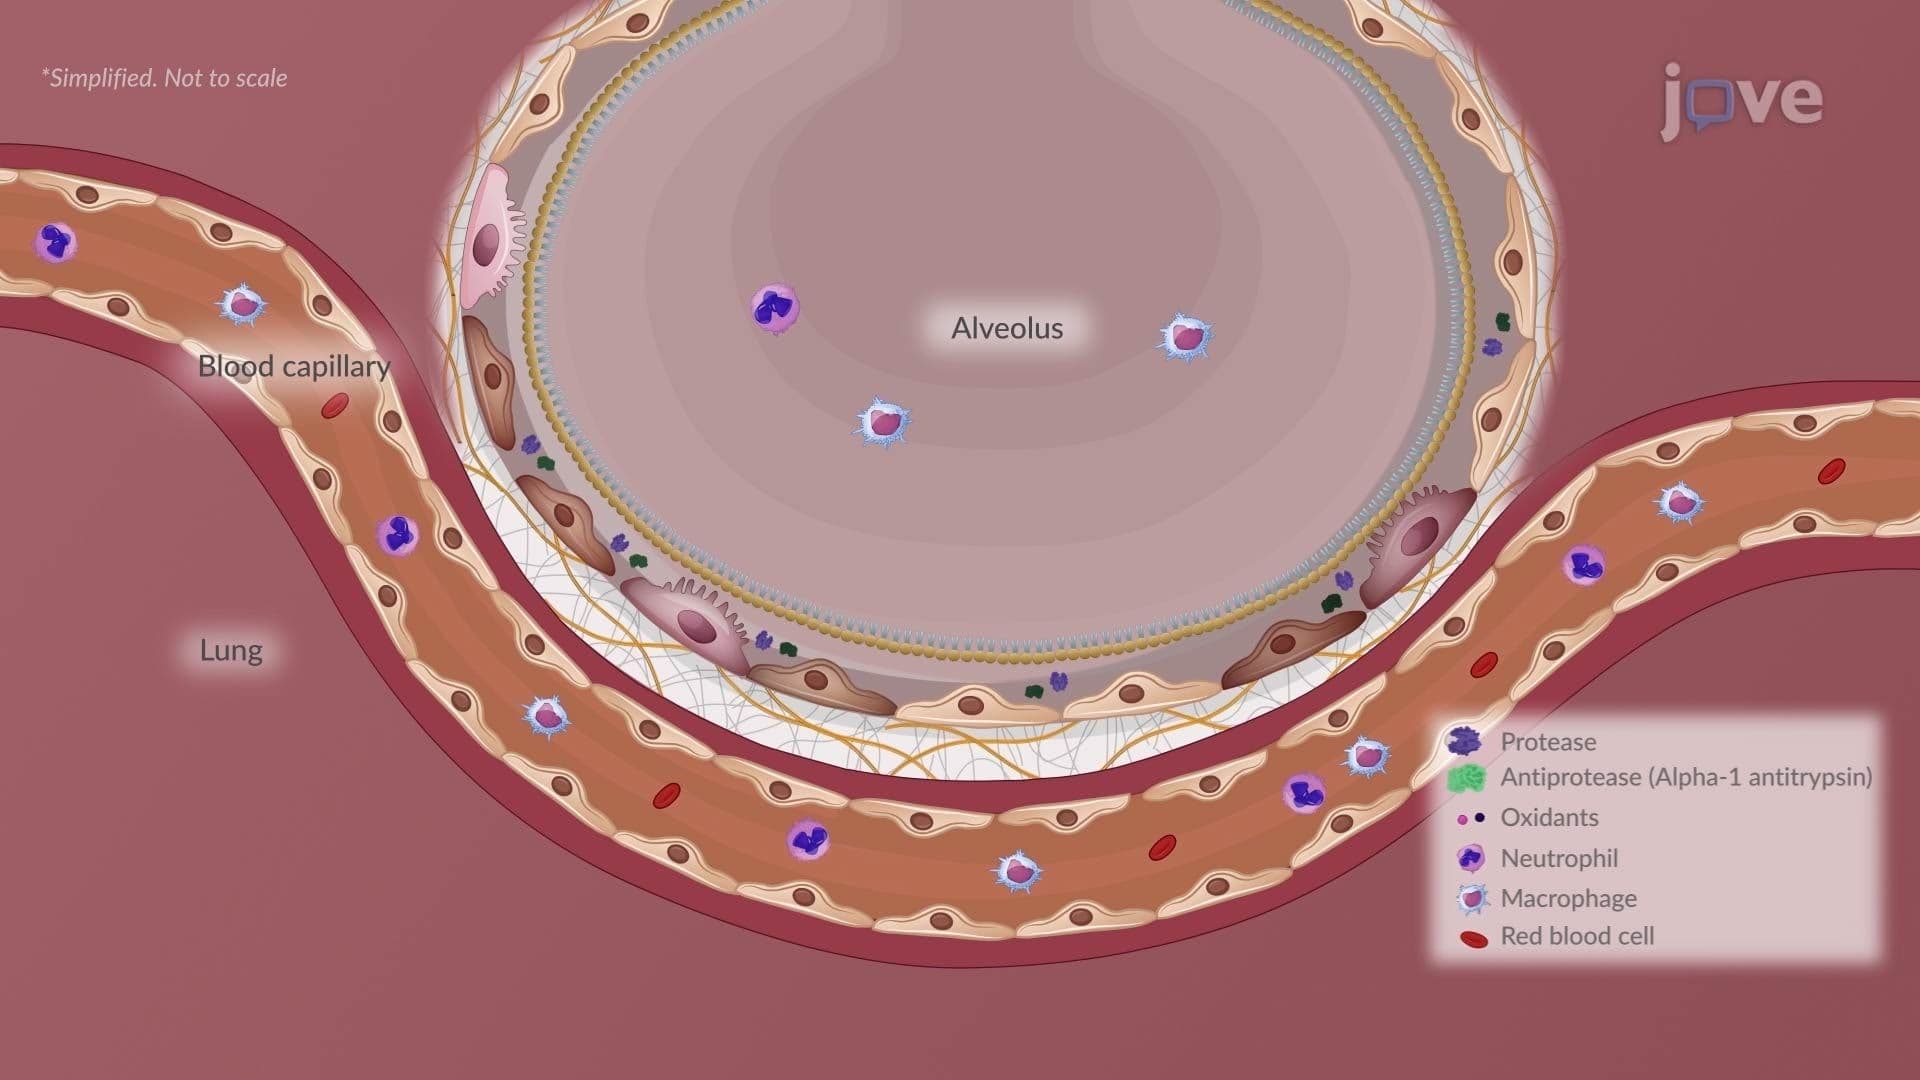
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:18
Pulmonary Edema II: Pathophysiology
 01:10
Atelectasis II: Pathophysiology
 01:21
Acute Respiratory Failure-II
The underlying physiological abnormalities that contribute to hypoxemic respiratory failure include:
 01:14
Atypical Pneumonia
 01:29
Pneumonia II: Pathophysiology