Arachidonyl dopamine as a ligand for the vanilloid receptor VR1 of the rat
Related Experiment Video
Updated: Sep 26, 2026
Establishing a Mouse Model of a Pure Small Fiber Neuropathy with the Ultrapotent Agonist of Transient Receptor Potential Vanilloid Type 1
Published on: February 13, 2018
Attila Tóth1, Noémi Kedei, Yun Wang
1Molecular Mechanisms of Tumor Promotion Section, Laboratory of Cellular Carcinogenesis and Tumor Promotion, National Cancer Institute, National Institutes of Health, Bldg. 37, Rm. 4048, 37 Convent Dr., MSC 4255, Bethesda, MD 20892-4255, USA.

Abstract:
The vanilloid receptor VR1 is a nonspecific Ca(2+) channel, expressed in sensory neurons in the peripheral nervous system and in various brain regions, which is believed to be an important molecular integrator of several chemical (acid, vanilloids) and physical stimuli (heat) that cause pain. Recently, several endogenous ligands for VR1 have been identified such as arachidonyl ethanolamide (anandamide) and the more potent arachidonyl dopamine (AA-DO). Here, we further characterize AA-DO as a ligand for rat VR1, heterologously expressed in CHO and HEK293 cells. AA-DO inhibited the binding of [3H]RTX to VR1 with a K(d) value of 5.49 +/- 0.68 microM and with positive cooperativity (p = 1.89 +/- 0.27), indicating that AA-DO was about 5-fold more potent than anandamide in this system. The K(d) (9.7 +/- 3.3 microM), and p values (1.54 +/- 0.04) for the binding of AA-DO to spinal cord membranes are in good correlation with the CHO-VR1 data. AA-DO stimulated 45Ca(2+) uptake on CHO-VR1 and HEK-VR1 cells with EC(50) values of 4.76 +/- 1.43 and 7.17 +/- 1.64 microM and Hill coefficients of 1.28 +/- 0.11 and 1.13 +/- 0.13, respectively, consistent with the binding measurements. In contrast to anandamide, AA-DO induced virtually the same level of 45Ca(2+) uptake as did capsaicin (90 +/- 6.6% in the CHO cells expressing VR1 and 89.3 +/- 9.4% in HEK293 cells expressing VR1). In a time dependent fashion following activation, AA-DO partially desensitized VR1 both in 45Ca(2+) uptake assays (IC(50) = 3.24 +/- 0.84 microM, inhibition is 68.5 +/- 6.85%) as well as in Ca(2+) imaging experiments (35.8 +/- 5.1% inhibition) using the CHO-VR1 system. The extent of desensitization was similar to that caused by capsaicin itself. We conclude that AA-DO is a full agonist for VR1 with a potency in the low micromolar range and is able to significantly desensitize the cells in a time and dose dependent manner.
Related Concept Videos
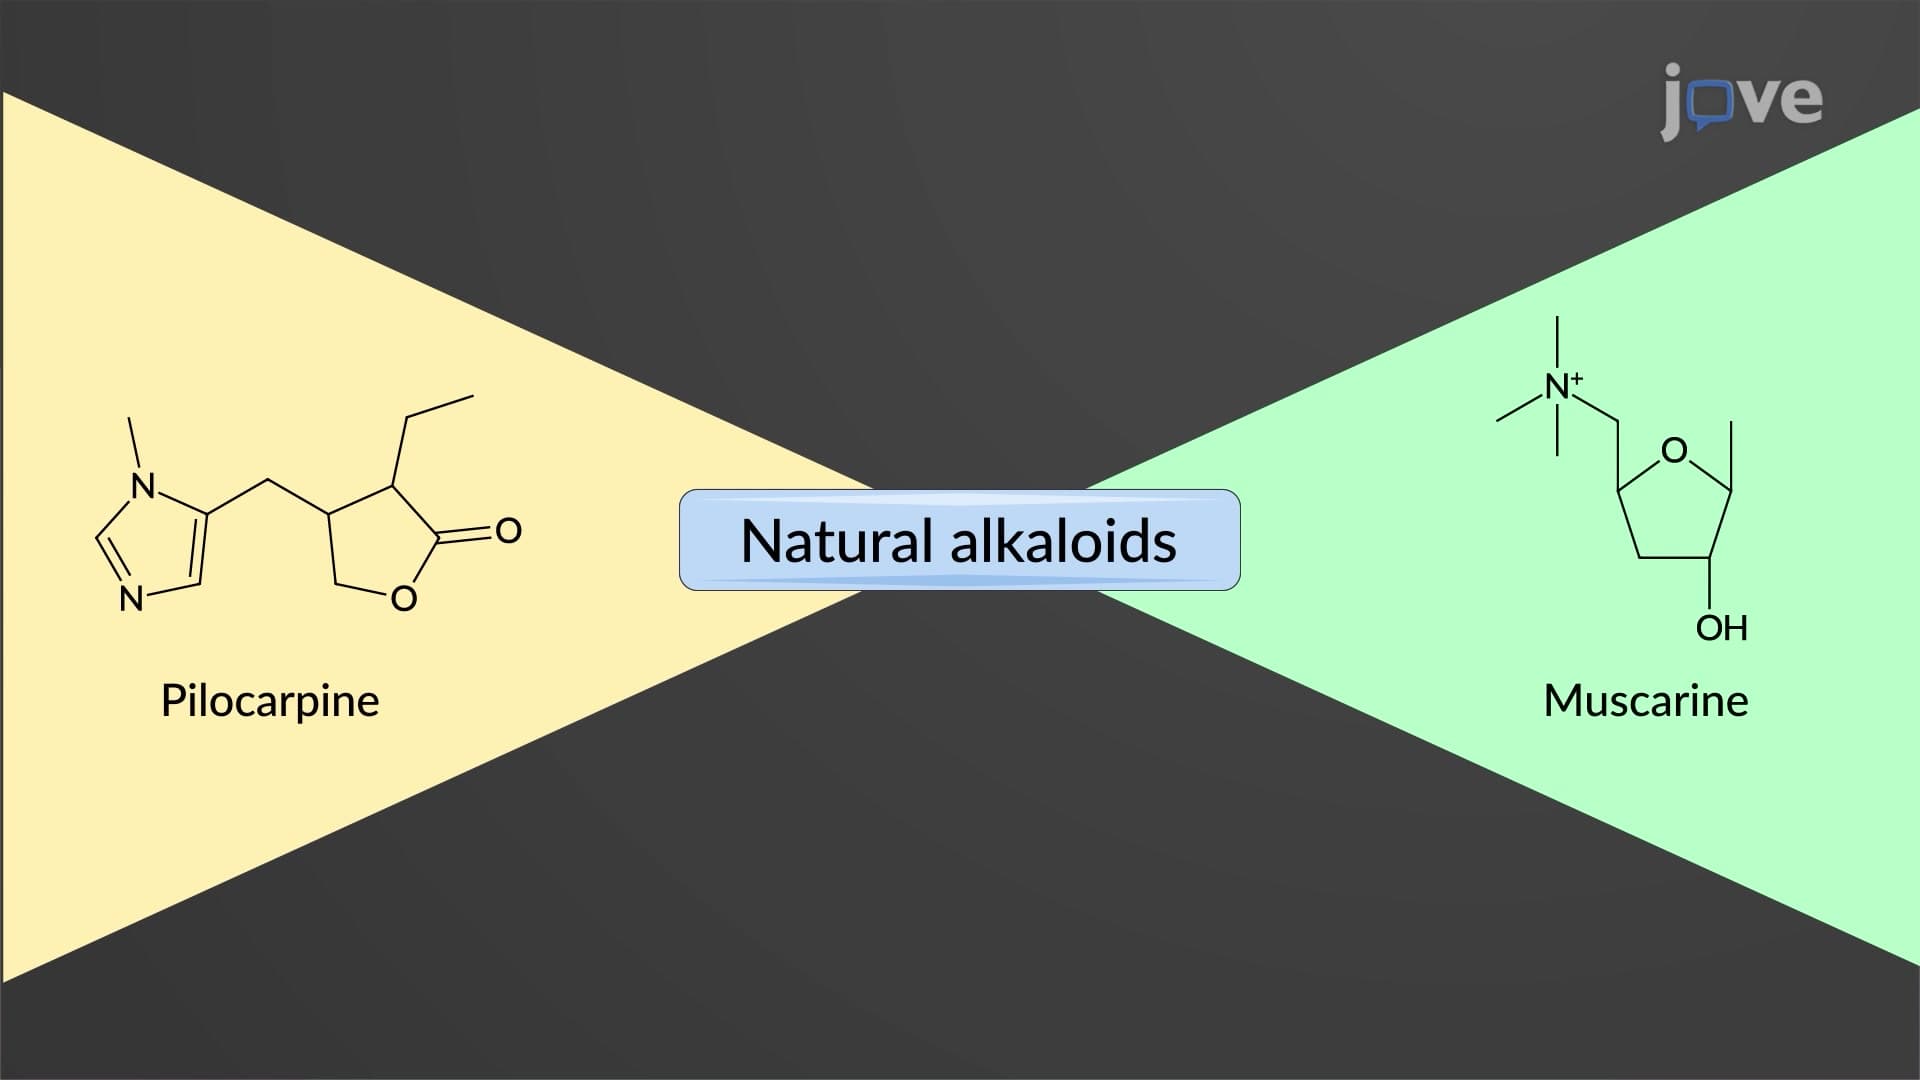 01:22
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...
 01:25
Cholinergic Receptors: Muscarinic
The subtypes M1, M3, and M5 couple with the Gq subunit and activate the phospholipase C (PLC) activity, mobilizing intracellular Ca2+. Activation...
 01:30
Ligand-Gated Ion Channel Receptor: Gating Mechanism