Related Experiment Video
Updated: Sep 15, 2026
A Bioluminescent and Fluorescent Orthotopic Syngeneic Murine Model of Androgen-dependent and Castration-resistant Prostate Cancer
Published on: March 6, 2018
Pharmacological profile of MEN 11066, a novel potent and selective aromatase inhibitor
M Muratori1, A Lippi, R Mancina
1Department of Clinical Physiopathology, University of Florence, V. le Pieraccini 6, I-50139 Firenze, Italy.

Abstract:
MEN 11066 is a new non-steroidal compound which potently inhibits human placenta (K(i)=0.5 nM) and rat ovarian (K(i)=0.2 nM) aromatase in vitro. In vivo, a single oral dose of 0.3 mgkg(-1) significantly decreased uterus weight in immature rats after stimulation of uterus growth by androstenedione. MEN 11066 reduced in a dose-dependent manner plasma estradiol levels in adult female rats treated with pregnant mare serum gonadotropin (PMSG). After 2 weeks of repeated daily treatment in adult rats, a significant decrease in uterine weight was observed together with a 65% decrease in plasma estradiol, whereas plasma levels of testosterone, progesterone, aldosterone, corticosterone, cholesterol, LH and FSH were not affected. The lack of any effect by MEN 11066 on adrenal steroids was confirmed by the unchanged plasma corticosterone and aldosterone levels in immature rats and also in adult rats when the repeated treatment with MEN 11066 (15 days) was followed by the administration of a synthetic ACTH analogue. No change in 11beta-hydroxylase or 21-hydroxylase activities was produced in vitro by the addition of 10 microM MEN 11066. Fifteen-day treatment with MEN 11066 did not produce changes in several rat hepatic enzymatic activities involved in the metabolism of xenobiotics. These results demonstrated that MEN 11066 is a potent inhibitor of aromatase which does not interfere with the cytochrome P450 involved in the synthesis of other steroids or in the metabolism of xenobiotics.
Related Concept Videos
 01:31
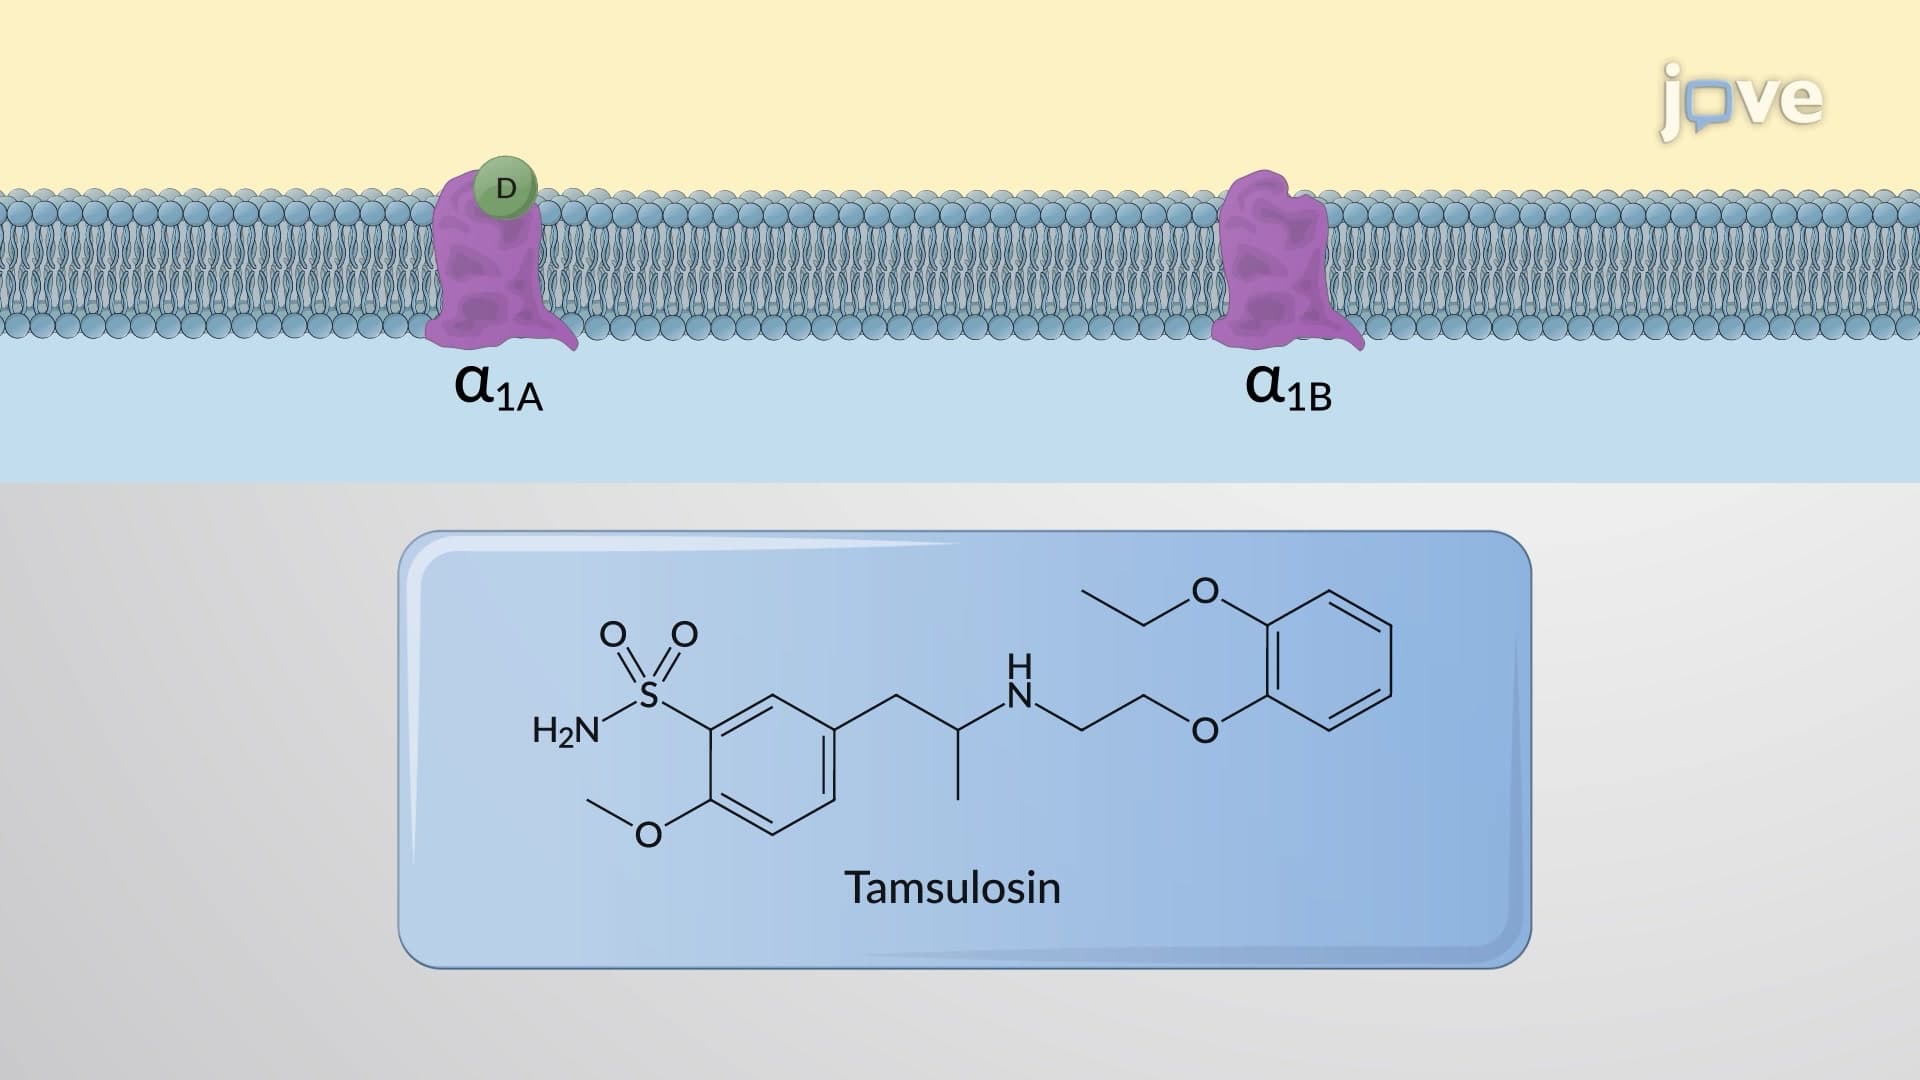
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
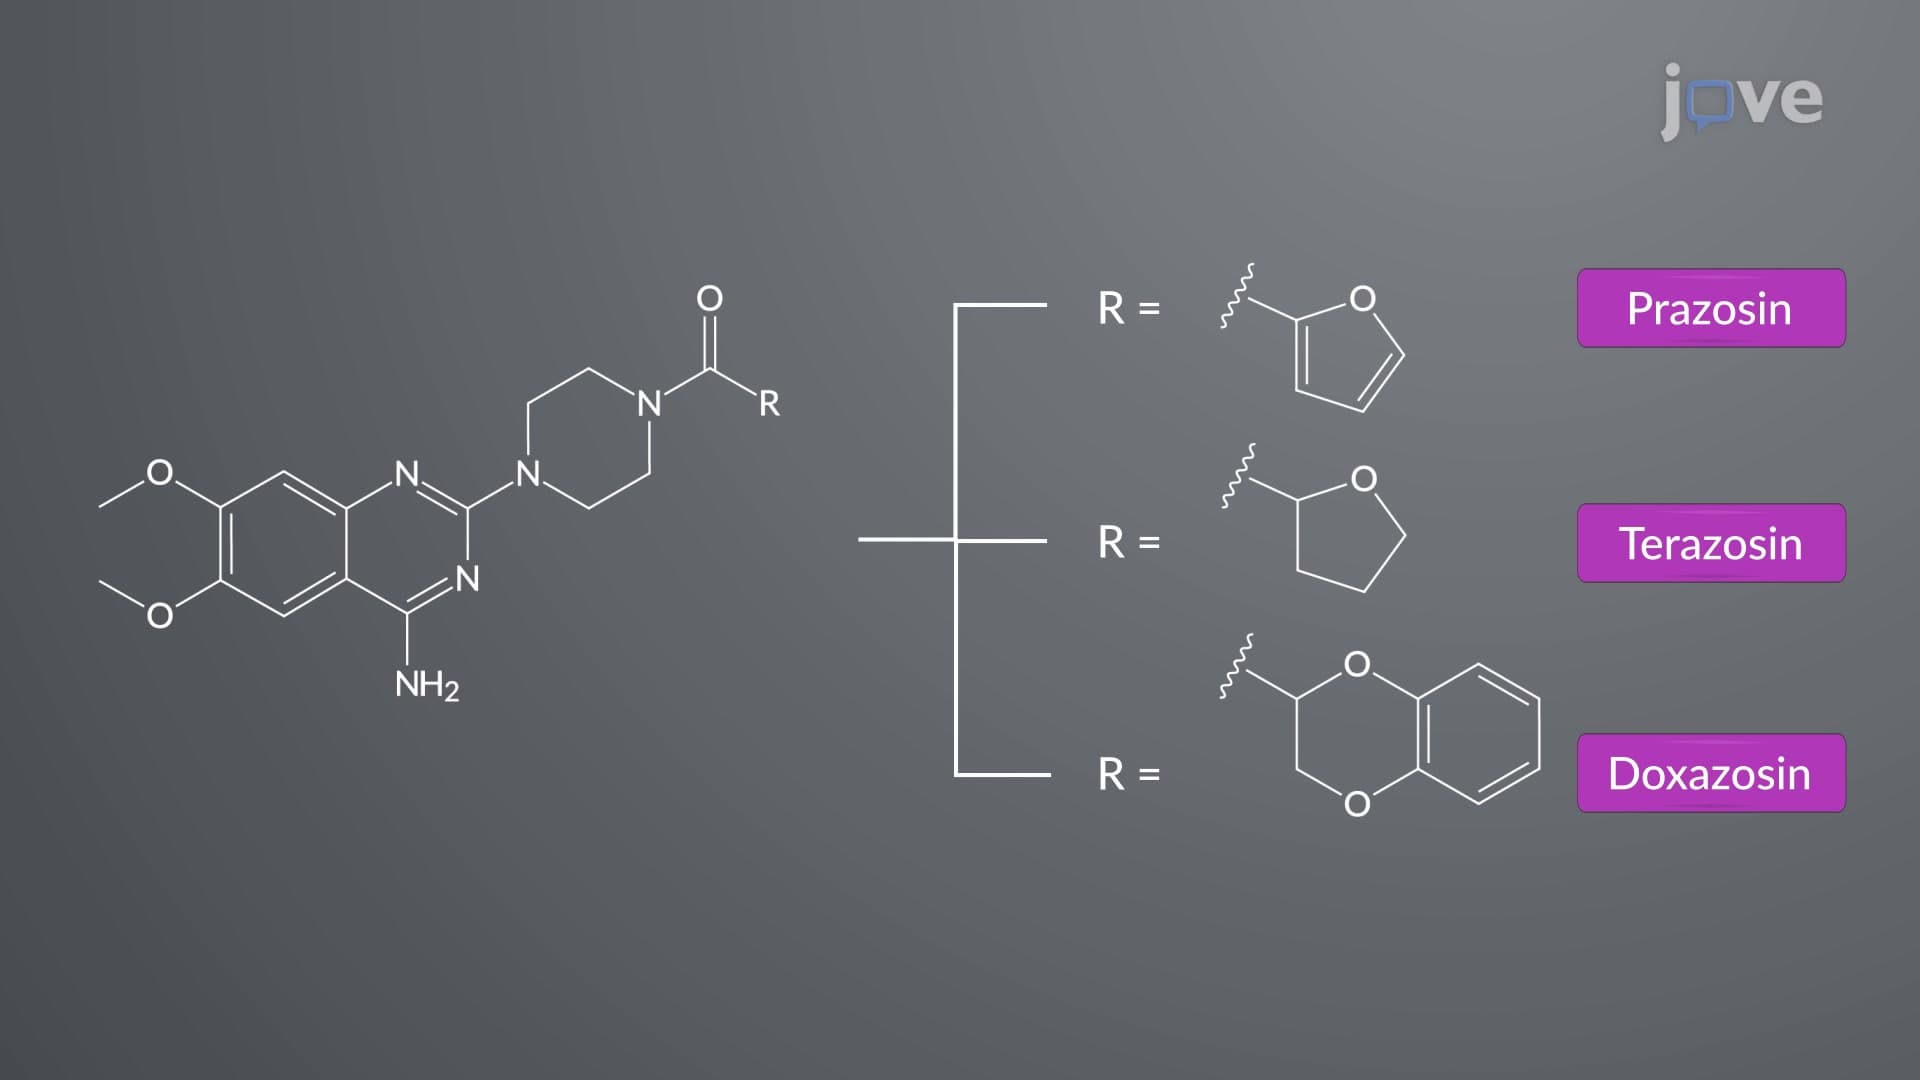
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:22
Adrenergic Antagonists: Pharmacological Actions of ɑ-Receptor Blockers
α1-blockers: These drugs inhibit α1-adrenoceptors on smooth muscle cells, resulting in vasodilation. This vasodilation lowers blood pressure, making α1-blockers valuable in treating hypertension. Additionally, α1-blockers effectively address urinary obstruction...
 01:28
Treatment for Pulmonary Arterial Hypertension: Phosphodiesterase Inhibitors
Among the PDE5 inhibitors, sildenafil (Revatio) stands out as a competitive and selective inhibitor. It operates by elevating cellular levels of cGMP and augmenting signaling through the cGMP-PKG pathway, promoting vasodilation. Upon oral...
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase