Related Experiment Video
Updated: Sep 25, 2026

Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Testing hypotheses about determinants of protein structure with high-precision, high-throughput stability
Fang Yi1, Dorothy A Sims, Gary J Pielak
1Departments of Biochemistry and Biophysics, University of North Carolina, Chapel Hill, North Carolina 27599, USA.

Abstract:
Statistical modeling provides the mathematics to use data from large numbers of mutant proteins to generate information about hypotheses concerning protein structure not easily obtained from anecdotal studies on small numbers of mutants. Here we use the unfolding free energies of 303 unique eglin c mutant proteins obtained from high-precision, high-throughput chemical denaturation measurements to assess models concerning helix stability. A model with helix propensity as the sole determinant of stability accounts for 83% of the mutant-to-mutant variation in stability for 99% of the mutant proteins (three outliers). When position effects and side chain-side chain interactions are added to the model, the fraction of variation explained increases to 92%. The propensity parameters in this model are identical to helix propensity values derived from other approaches. Measurement error accounts for another 1% of the mutant-to-mutant variation in stability. While the data support terms for several of the expected stabilizing/destabilizing effects, it does not support terms for several others, including i, i + 3 effects in the center of the helix and helix-dipole effects. In addition, the model does better with terms for several stabilizing/destabilizing effects for which we cannot identify the physical basis. The precision of our unfolding stability measurements (+/-0.087 kcal/mol) allows us to conclude that the 7% of variation in stabilities of the mutant proteins not accounted for by the model or by measurement variation is both real and large with respect to the nonpropensity terms in the model. The analysis also shows that the common practice of using C(m)m(av) instead of C(m)m(mut) to calculate DeltaG(HOH,N-D) values for each mutant protein results in a loss of information. We see no correlation between the residuals derived from the full model and m(mut) - m(wt), and hence it is unlikely our m(mut) values reflect mutant-to-mutant differences in the denatured state.
Related Concept Videos
 01:24
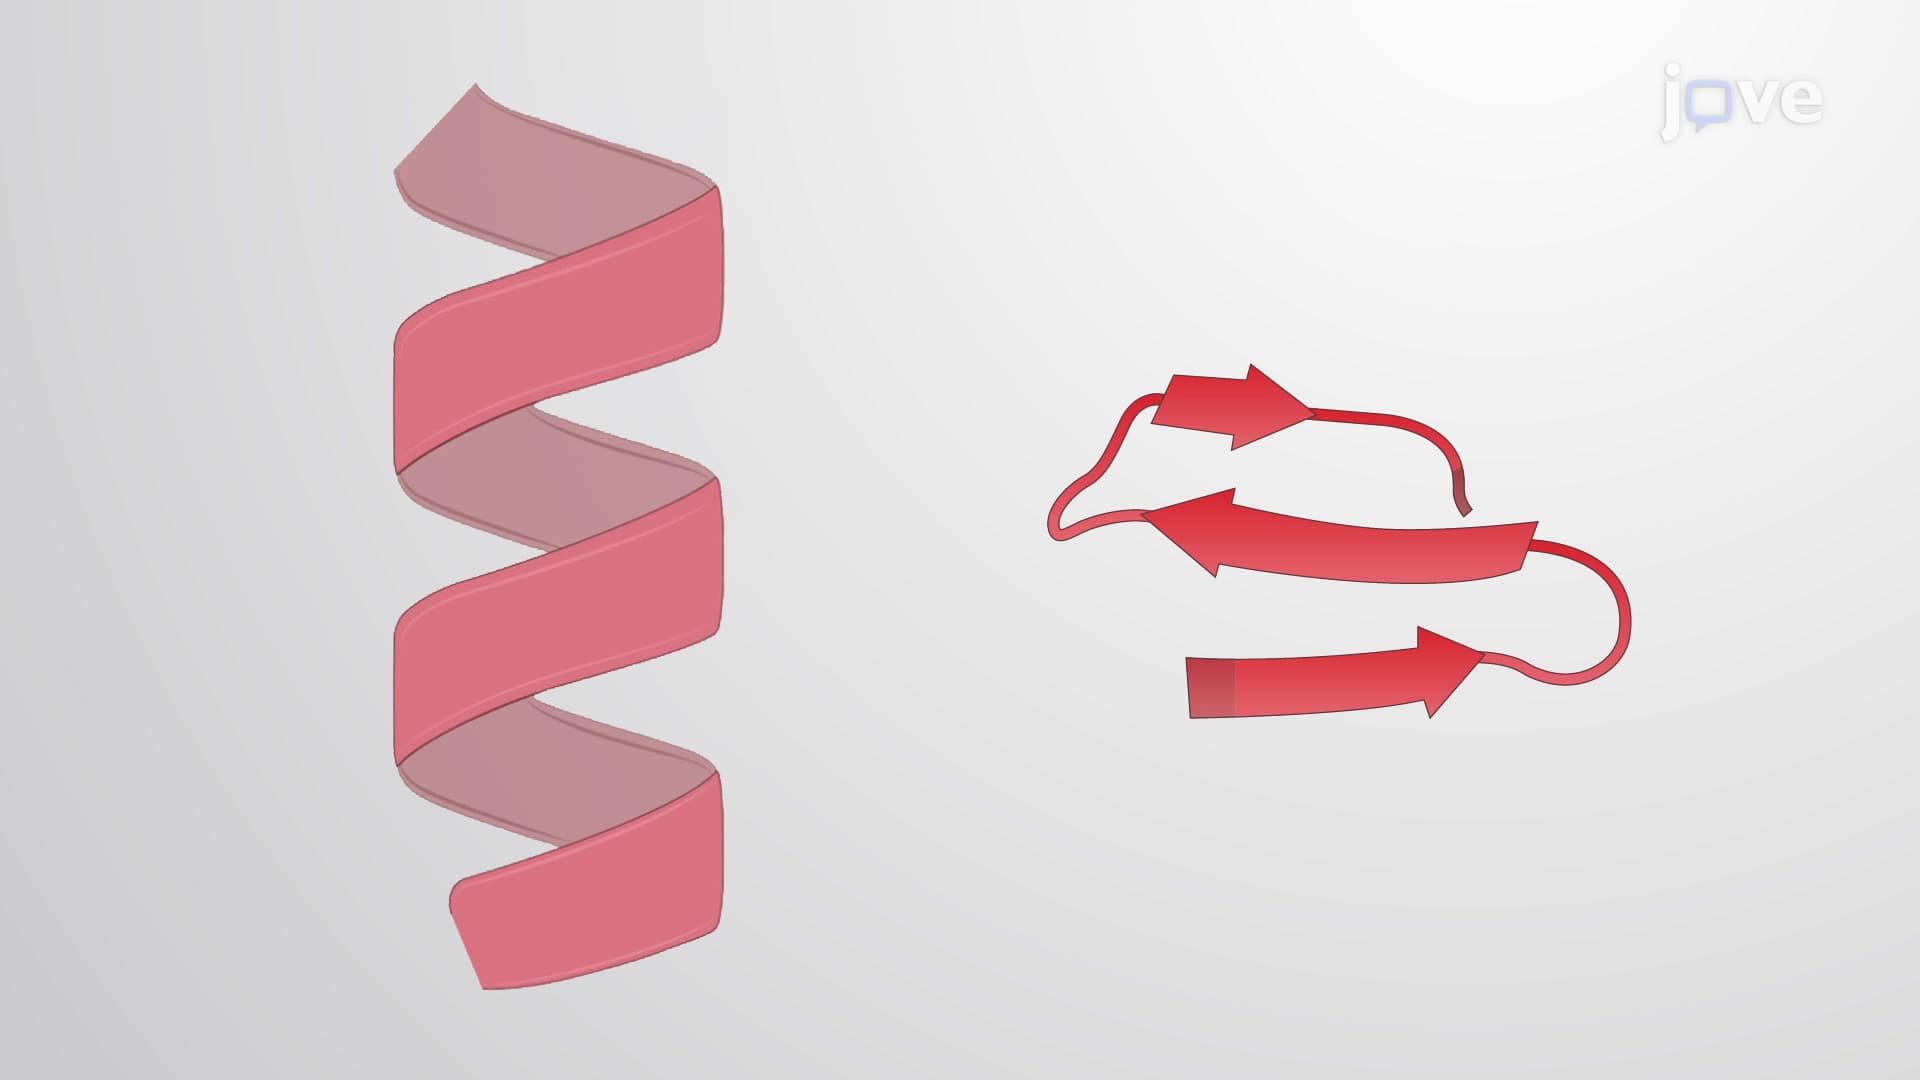
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:49
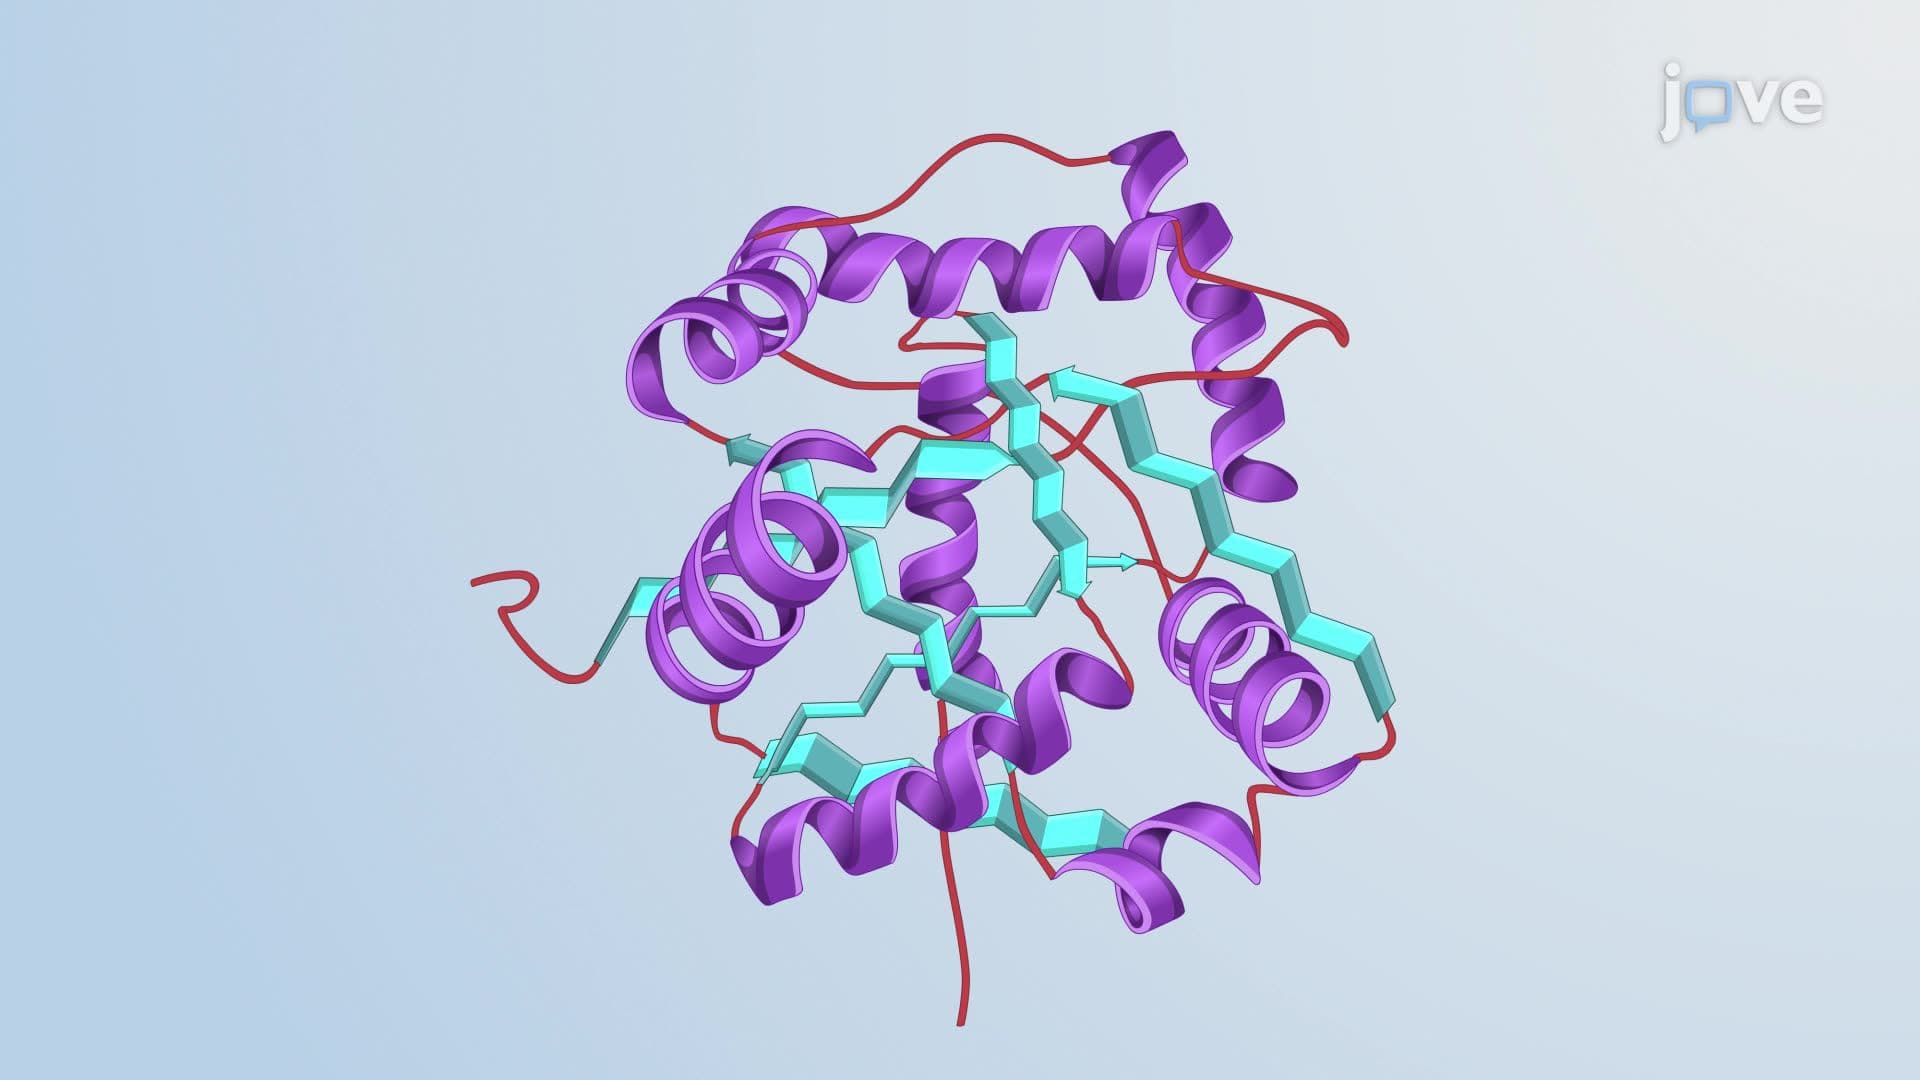
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:39
Protein and Protein Structure