Related Experiment Video
Updated: Jul 13, 2026

Functional Reconstitution and Channel Activity Measurements of Purified Wildtype and Mutant CFTR Protein
Published on: March 9, 2015
Abnormal passive chloride absorption in cystic fibrosis jejunum functionally opposes the classic chloride secretory
Michael A Russo1, Christoph Hogenauer, Stephen W Coates
1Baylor University Medical Center, Dallas, Texas 75246, USA.

Abstract:
Due to genetic defects in apical membrane chloride channels, the cystic fibrosis (CF) intestine does not secrete chloride normally. Depressed chloride secretion leaves CF intestinal absorptive processes unopposed, which results in net fluid hyperabsorption, dehydration of intestinal contents, and a propensity to inspissated intestinal obstruction. This theory is based primarily on in vitro studies of jejunal mucosa. To determine if CF patients actually hyperabsorb fluid in vivo, we measured electrolyte and water absorption during steady-state perfusion of the jejunum. As expected, chloride secretion was abnormally low in CF, but surprisingly, there was no net hyperabsorption of sodium or water during perfusion of a balanced electrolyte solution. This suggested that fluid absorption processes are reduced in CF jejunum, and further studies revealed that this was due to a marked depression of passive chloride absorption. Although Na+-glucose cotransport was normal in the CF jejunum, absence of passive chloride absorption completely blocked glucose-stimulated net sodium absorption and reduced glucose-stimulated water absorption 66%. This chloride absorptive abnormality acts in physiological opposition to the classic chloride secretory defect in the CF intestine. By increasing the fluidity of intraluminal contents, absence of passive chloride absorption may reduce the incidence and severity of intestinal disease in patients with CF.
Related Concept Videos
 01:23
Transcellular Transport of Solutes
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic sinusitis...
 01:28
Reabsorption and Secretion in the PCT
Transport mechanisms involving sodium ions (Na+) contribute significantly to solute reabsorption. These mechanisms include symport and antiport processes.
A key example is the...
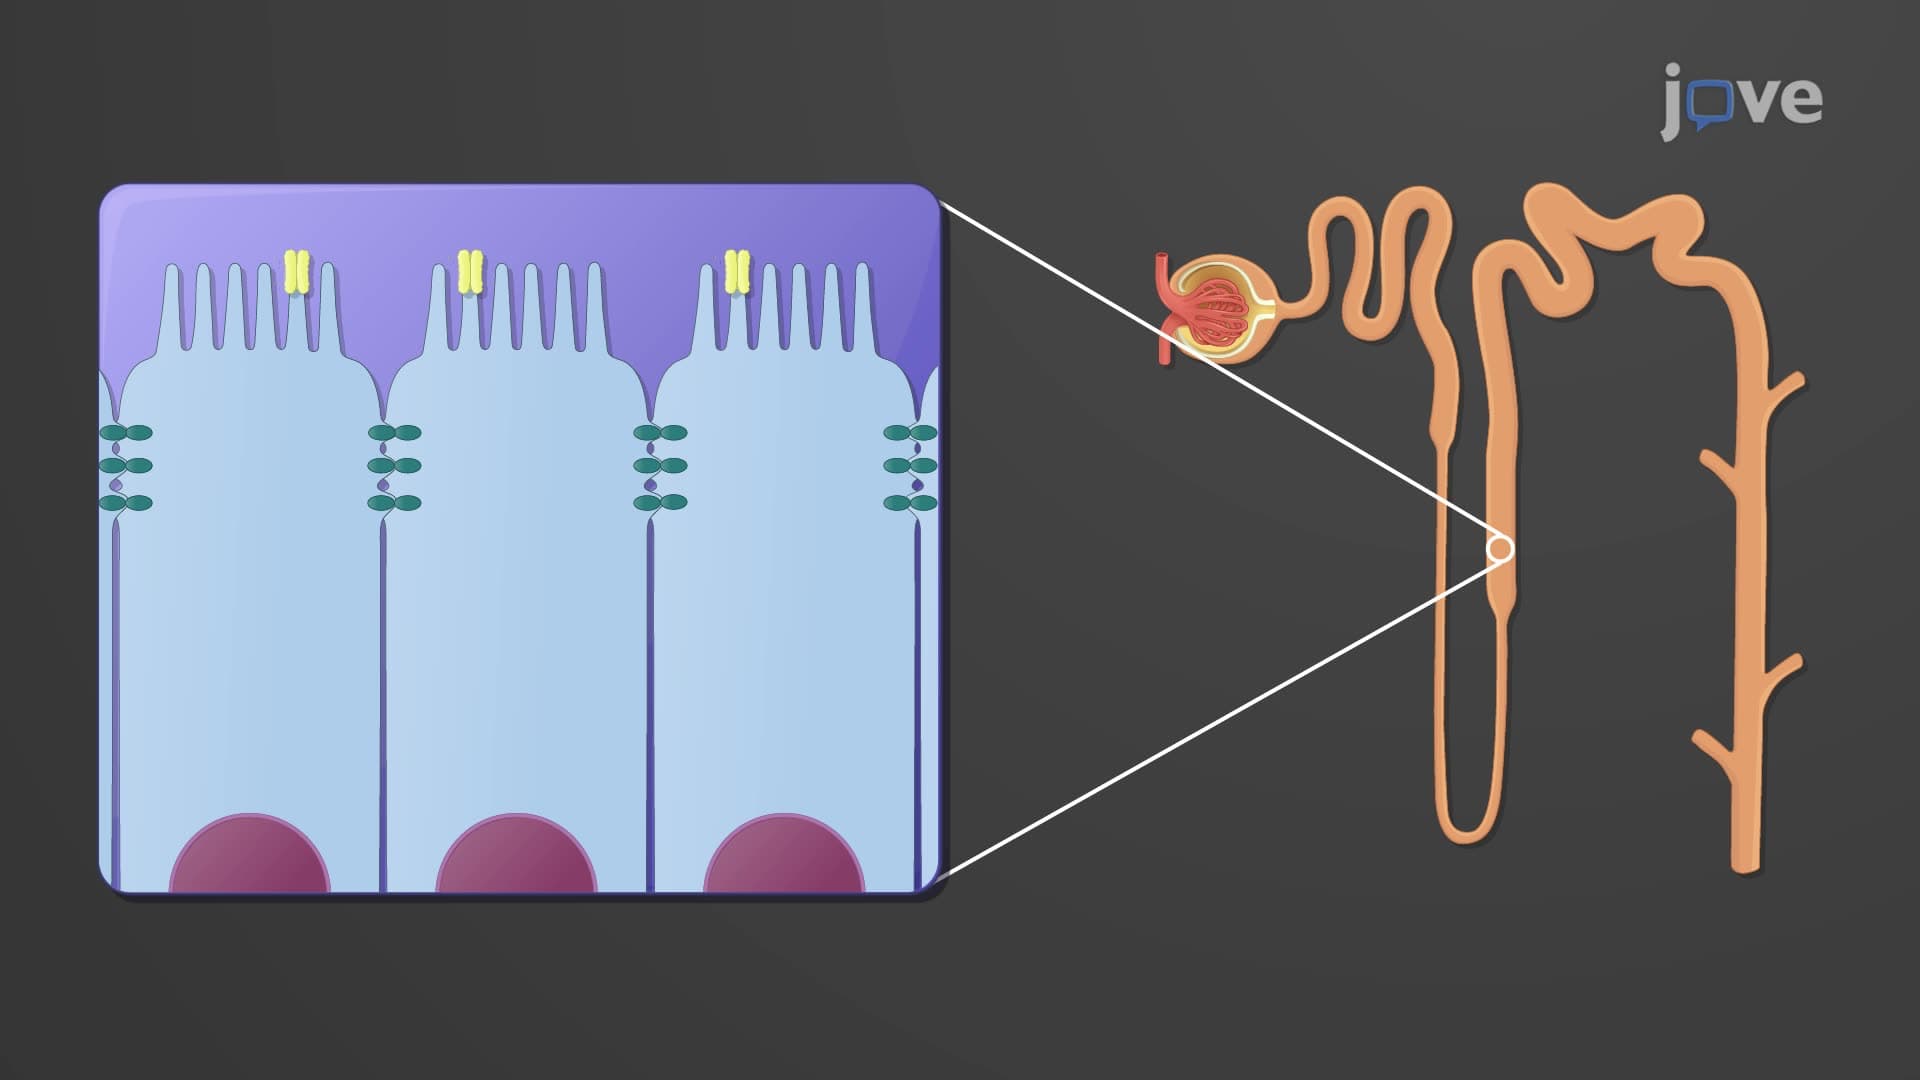 01:17
Reabsorption and Secretion in the Loop of Henle
 01:26
Reabsorption and Secretion in the DCT and Collecting Duct
The distal part of the DCT, along with the...