endo-Fullerene and doped diamond nanocrystallite-based models of qubits for solid-state quantum computers
Related Experiment Video
Updated: Sep 20, 2026
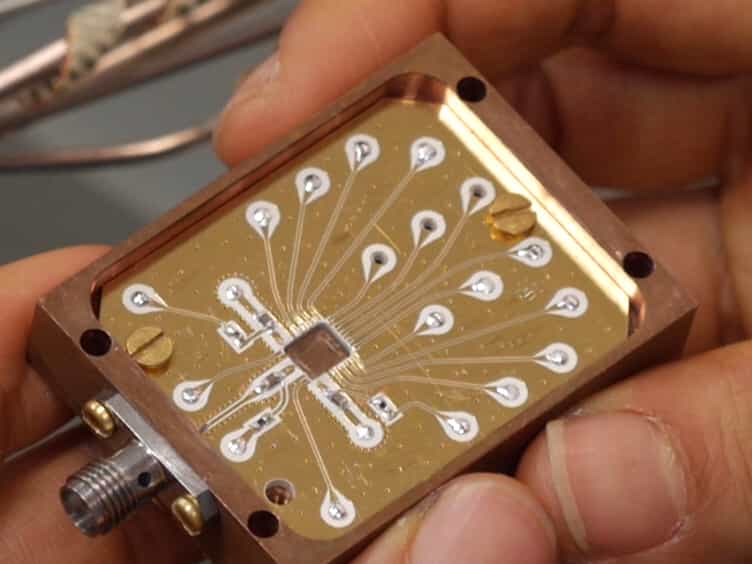
Silicon Metal-oxide-semiconductor Quantum Dots for Single-electron Pumping
Published on: June 3, 2015
1Department of Chemical Engineering, Stanford University, Stanford, California 94305-5025, USA.

Abstract:
Models of encapsulated nuclear spin 1/2 1H and 31P atoms in fullerene and diamond nanocrystallite, respectively, are proposed and examined with an ab initio local density functional method for possible applications as single quantum bits (qubits) in solid-state quantum computers. A 1H atom encapsulated in a fully deuterated fullerene, C20D20, forms the first model system and ab initio calculation shows that the 1H atom is stable in its atomic state at the center of the fullerene with a barrier of about 1 eV to escape. A 31P atom positioned at the center of a diamond nanocrystallite is the second model system, and 31P atom is found to be stable at the substitutional site relative to interstitial sites by 15 eV. Vacancy formation energy is 6 eV in diamond, so the substitutional 31P atom will be stable against diffusion during the formation mechanisms within the nanocrystallite. The coupling between the nuclear spin and the weakly bound (valance) donor electron in both systems is found to be suitable for single qubit applications, whereas the spatial distributions of (valance) donor electron wave functions are found to be preferentially spread along certain lattice directions, facilitating two or more qubit applications. The feasibility of the fabrication pathways for both model solid-state qubit systems within practical quantum computers is discussed within the context of our proposed solid-state qubits.
Related Concept Videos
 02:45
The Quantum-Mechanical Model of an Atom
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:42
Valence Bond Theory
 02:54
Molecular and Ionic Solids
Molecular Solids
Molecular crystalline solids, such as ice, sucrose (table sugar), and iodine, are solids that are composed of neutral molecules as their constituent units. These molecules are held together by weak intermolecular forces such as London dispersion forces, dipole-dipole interactions, or hydrogen bonds, which...