Regional self-interaction correction of density functional theory
Related Experiment Video
Updated: Sep 20, 2026
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Takao Tsuneda1, Muneaki Kamiya, Kimihiko Hirao
1Graduate School of Engineering, The University of Tokyo, Tokyo 113-8656, Japan. tune@qcl.t.u-tokyo.ac.jp

Abstract:
We propose a new simple scheme for self-interaction correction (SIC) of exchange functionals in the density functional theory. In the new scheme, exchange energies are corrected by substituting exchange self-interactions for exchange functionals in regions of self-interaction. To classify the regions of self-interaction, we take advantage of the property of the total kinetic energy density approaching the Weizsäcker density in the case of electrons in isolated orbitals. The scheme differs from conventional SIC methods in that it produces optimized molecular structures. Applying the scheme to the calculation of reaction energy barriers showed that it provides a clear improvement in cases where the barriers are underestimated by conventional "pure" functionals. In particular, we found that this scheme even reproduces a transition state that is not given by pure functionals.
More Related Videos

Related Concept Videos
 01:24
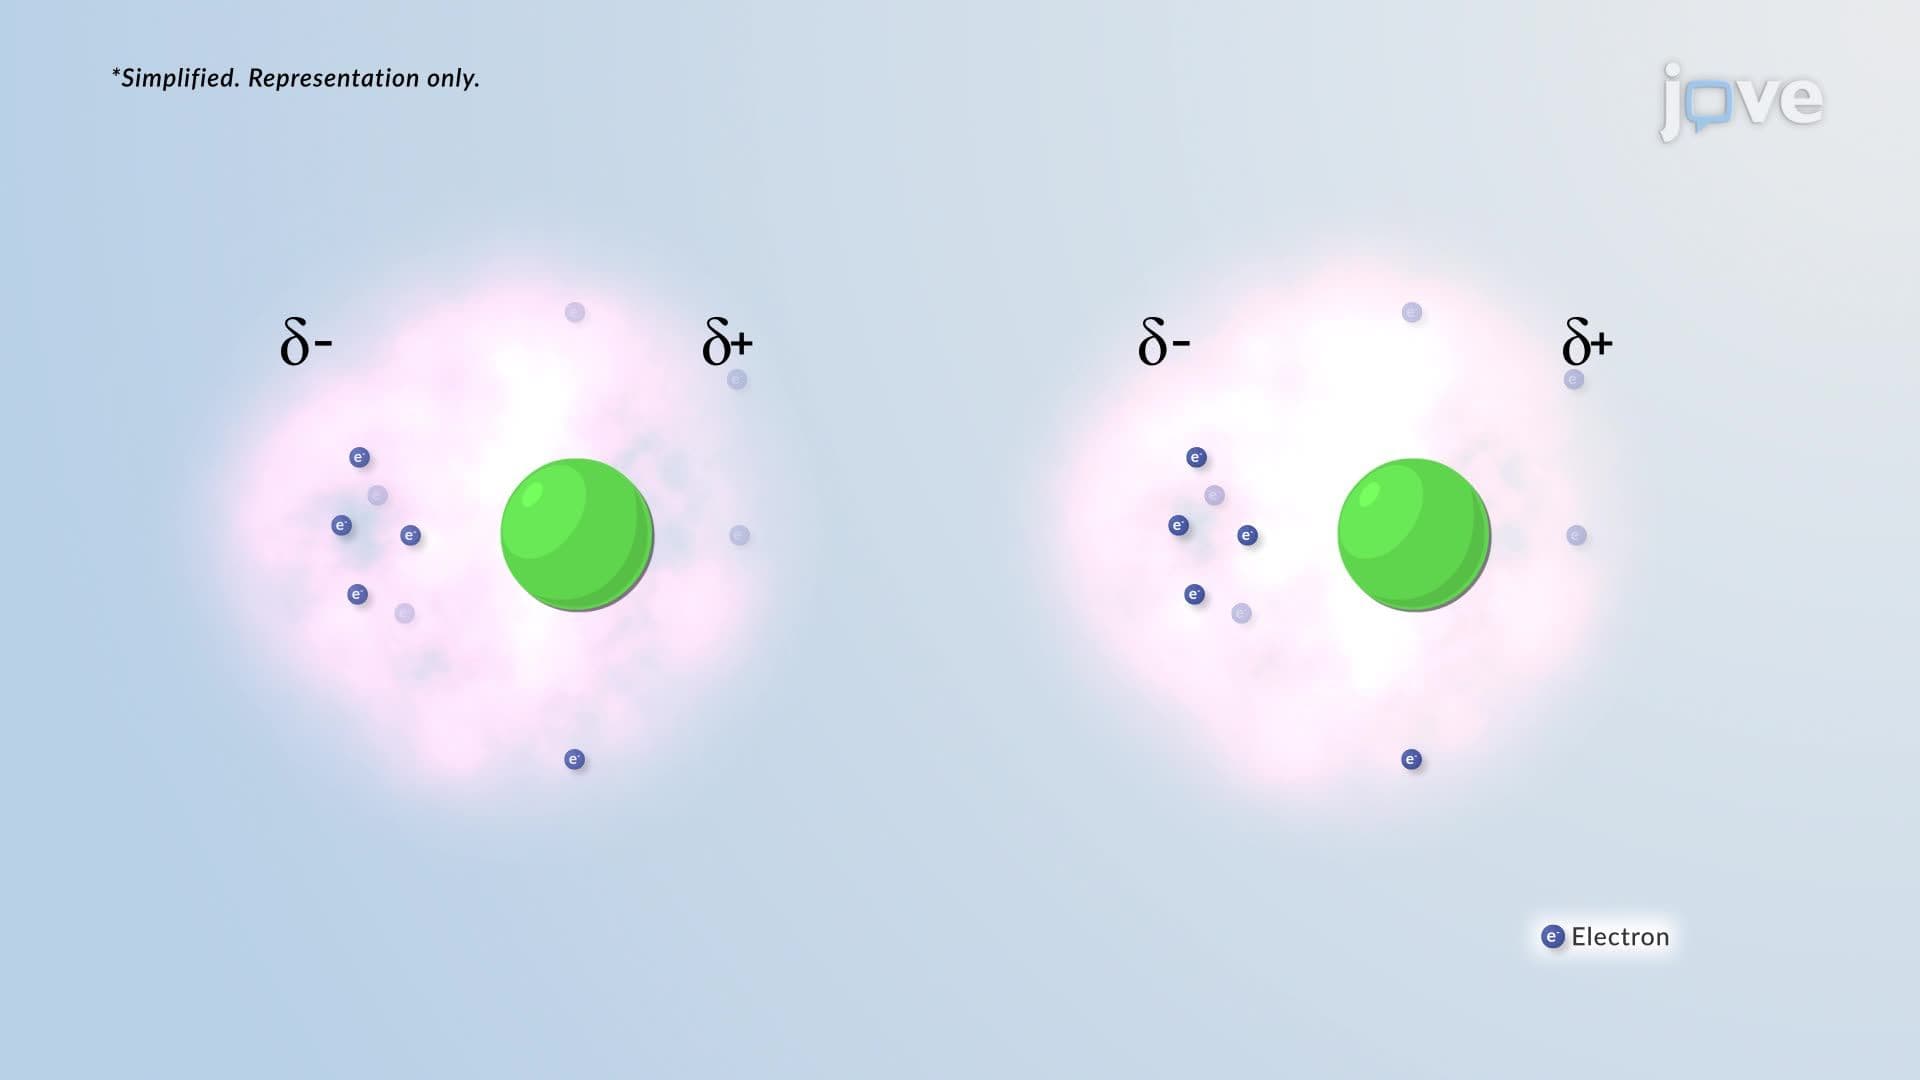
Van der Waals Interactions
 01:10
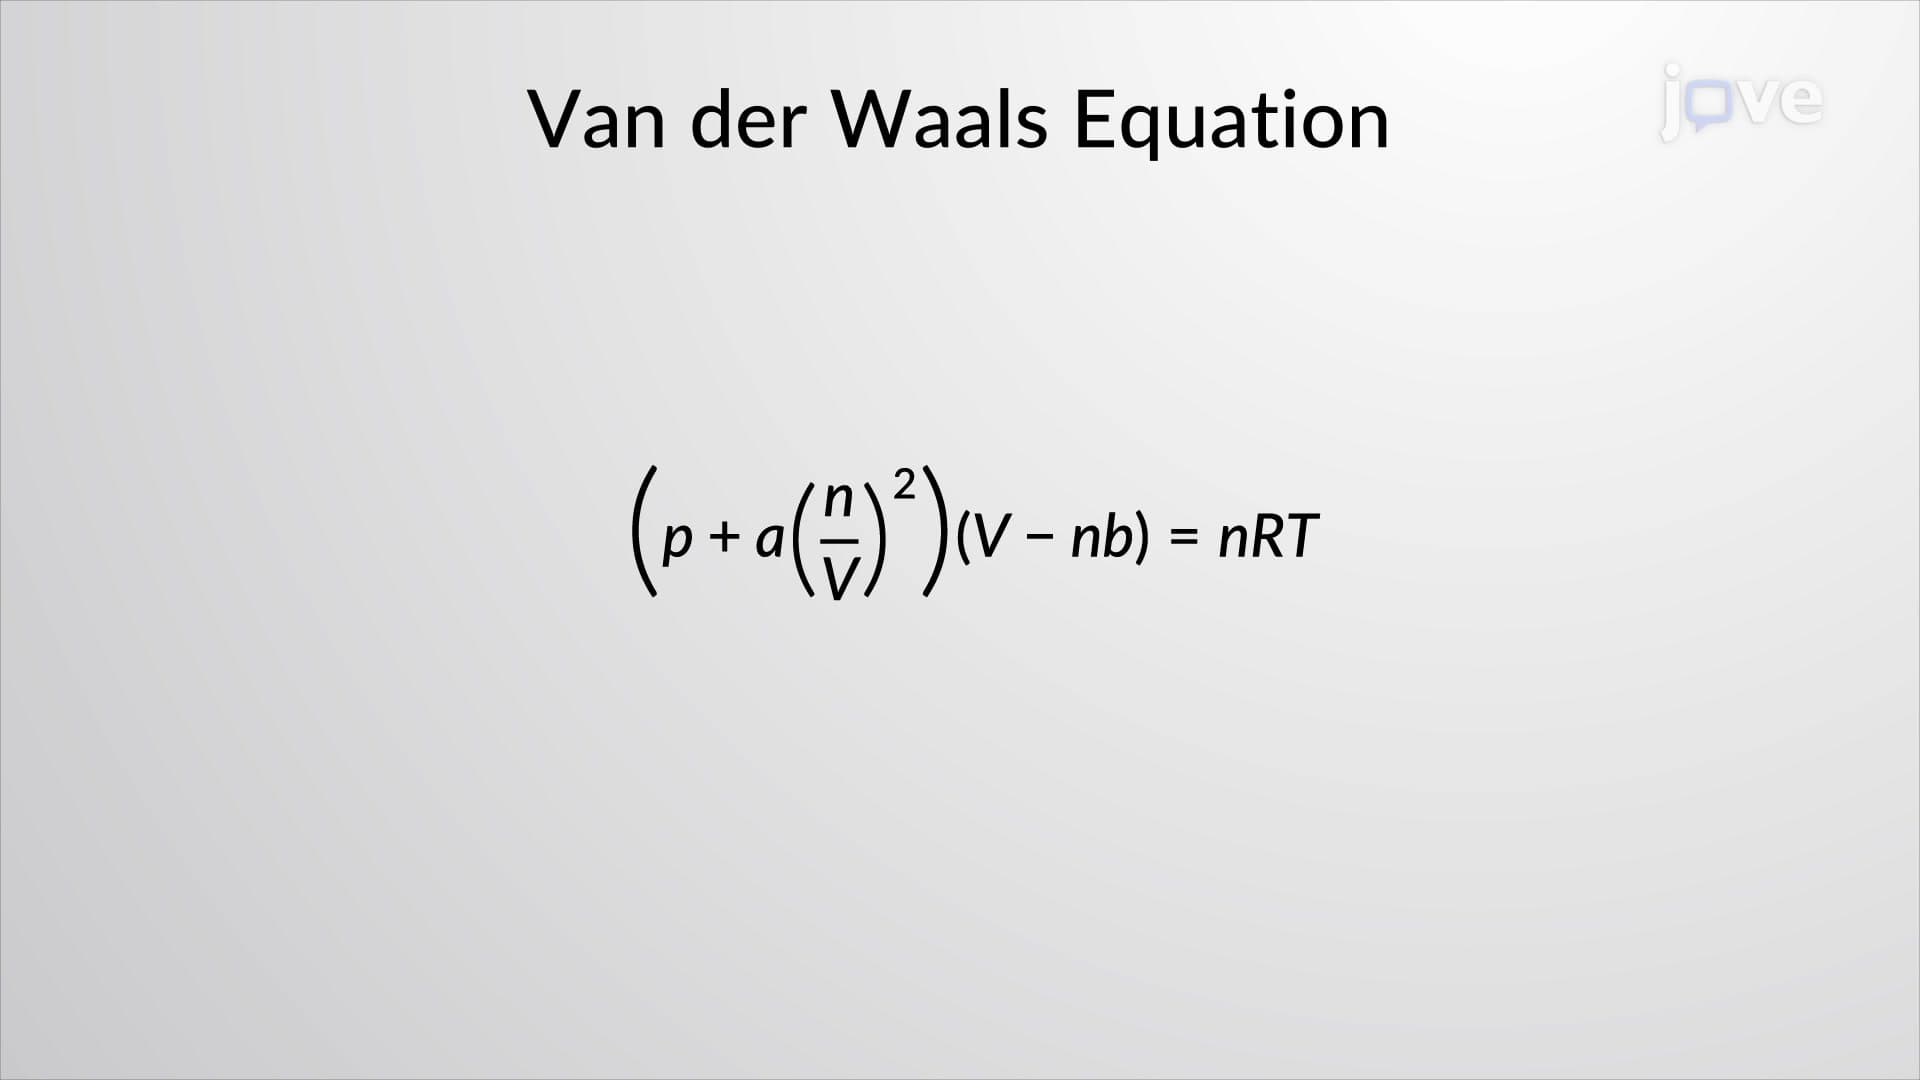
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...
 03:51
Molecular Orbital Theory II
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 02:35
Molecular Orbital Theory I
 01:28
Debye–Huckel–Onsager Conductance Equation