Creutzfeldt-Jakob disease--identifying prions and carriers
Related Experiment Video
Updated: Sep 20, 2026
09:50
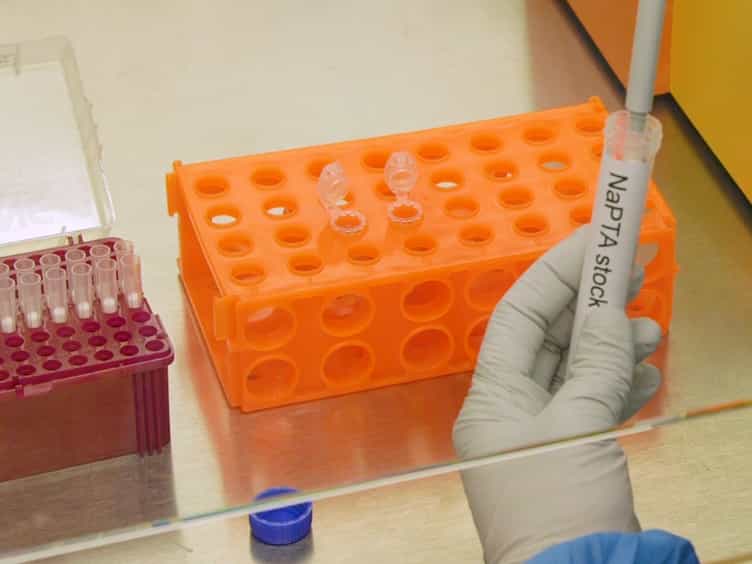
Real-time Quaking-induced Conversion Assay for Detection of CWD Prions in Fecal Material
Published on: September 29, 2017
AORN Journal
|August 28, 2003

Abstract:
Two major obstacles to be overcome in minimizing the possibility of transmitting Creutzfeldt-Jakob disease or variant Creutzfeldt-Jakob disease are associated with identifying the prion on disinfected surgical instruments and identifying carriers of the protease-resistant form of the prion. New developments indicate that the means for doing both soon may be available.
Related Concept Videos
 03:03
Amyloid Fibrils
Amyloid fibrils are aggregates of misfolded proteins. Under most circumstances, misfolded proteins are either refolded by chaperone proteins or degraded by the proteasome. However, in the case of a mutation or a disease, these proteins can accumulate to form large clusters and often further assemble to form elongated fibers, called fibrils.
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:29
Subviral Agents
Subviral agents are infectious entities that resemble viruses but lack one or more viral components, such as a capsid or essential replication machinery. These agents include viroids, prions, and satellites, each possessing distinct structural and functional characteristics that influence their mode of infection and replication.Viroids are the simplest subviral agents, consisting of circular, single-stranded RNA molecules without a protein coat. They exclusively infect plants, relying entirely...
 01:21
Huntington Disease l: Introduction
Huntington disease or HD is a progressive, fatal neurodegenerative disorder inherited in an autosomal dominant pattern.PathophysiologyIt is caused by expansion of the CAG trinucleotide repeat in the HTT gene on chromosome 4 (4p16.3), producing an abnormal huntingtin protein with an expanded polyglutamine tract. This misfolded protein disrupts cellular function, leading to neuronal death. Normal alleles have ≤26 repeats, 27–35 are intermediate (risk of expansion), 36–39 show reduced penetrance,...