Related Experiment Video
Updated: Sep 13, 2026

Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
Surfactant protein A and B genetic variants predispose to idiopathic pulmonary fibrosis
Moises Selman1, Hung-Mo Lin, Martha Montaño
1Instituto Nacional de Enfermedades Respiratorias, México DF, México.

Abstract:
Derangement in pulmonary surfactant or its components and alveolar collapse are common findings in idiopathic pulmonary fibrosis (IPF). Surfactant proteins play important roles in innate host defense and normal function of the lung. We examined associations between IPF and genetic polymorphic variants of surfactant proteins, SP-A1, SP-A2, SP-B, SP-C, and SP-D. One SP-A1 (6A(4)) allele and single nucleotide polymorphisms (SNPs) that characterize the 6A(4) allele, and one SP-B (B1580_C) were found with higher frequency ( P
Related Concept Videos
 01:05
Breathing
 01:23
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase
 01:16
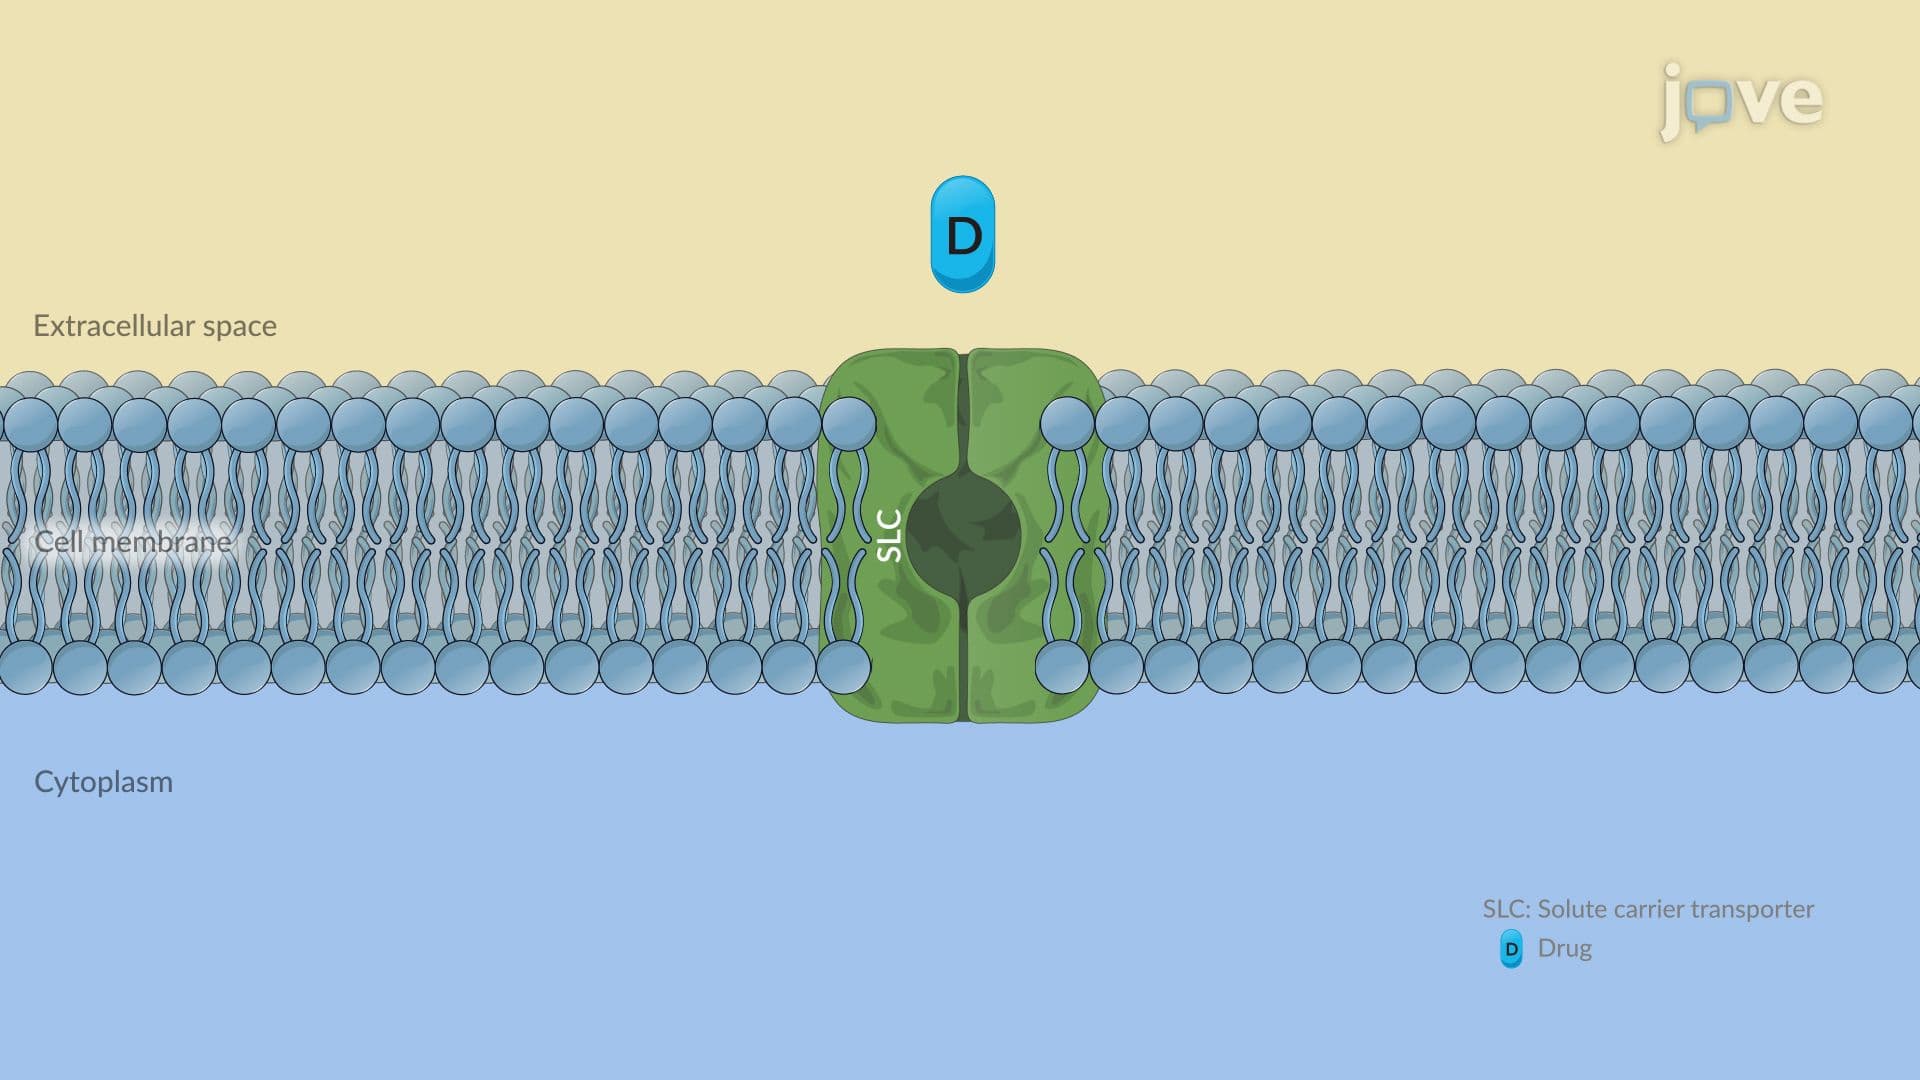
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:29
Respiratory Syncytial Virus Disease