Related Experiment Video
Updated: Aug 14, 2026
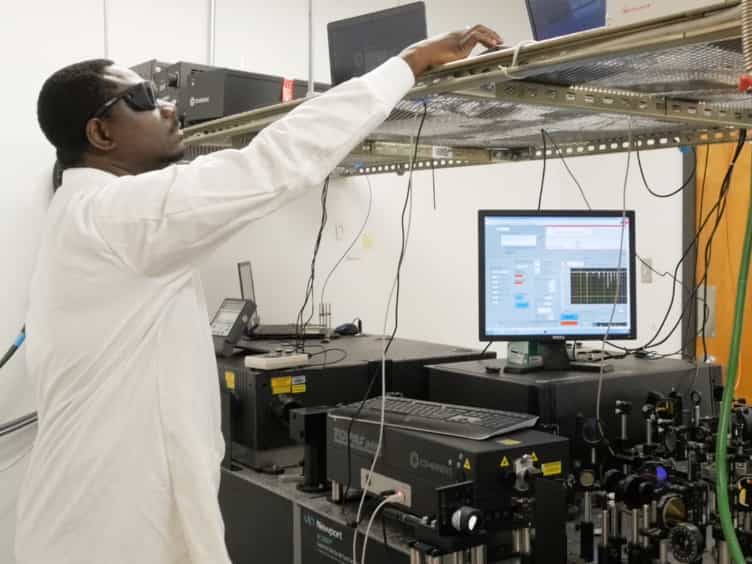
Measurement of Ultrafast Vibrational Coherences in Polyatomic Radical Cations with Strong-Field Adiabatic Ionization
Published on: August 6, 2018
Car-Parrinello molecular dynamics study of the rearrangement of the valeramide radical cation
Marija Semialjac1, Detlef Schröder, Helmut Schwarz
1Institut für Chemie, Technischen Universität Berlin, Strasse des 17. Juni 135, 10623 Berlin, Germany. Marija.Semialjac@www.chem.tu-berlin.de

Abstract:
Car-Parrinello molecular dynamics (CPMD) studies of neutral (1) and ionized (1 (+.)) valeramide are performed with the aim of providing a rationalization for the unusual temperature effect on the dissociation pattern of 1(+.) observed in mass spectrometric experiments. According to CPMD simulations of neutral valeramide 1 performed at approximately 500 K, the conformation with the fully relaxed carbon backbone predominates (96 %). Conformational changes involving folding of the carbon backbone into conformers that would allow intramolecular H transfers are predicted not to take place spontaneously at this temperature because of the barrier heights associated with these transitions (3.5 and 6.9 kcal mol(-1)), which cannot be overcome by thermal motion alone. For 1(+.), CPMD simulations performed at approximately 300 K reveal a substantial stability of a conformation in which the carbon backbone is fully relaxed; no reaction is observed even after 7 ps. However, when conformers with already folded carbon-backbones are used as initial geometries in the CPMD simulations, the gamma-hydrogen migration (McLafferty rearrangement resulting in C(3)H(6)) is already completed within 2 ps. For this important process, the free activation energy associated with both a required conformational change and the subsequent H transfer equals 4.5 kcal mol(-1), while for the formally related delta-H shift (which eventually gives rise to the elimination of C(2)H(4)/C(2)H(5.)) it amounts to 7.0 kcal mol(-1). Since the barriers associated with conformational changes are energetically more demanding than those of the corresponding hydrogen transfers, 1(+.) is essentially trapped by conformational barriers and long-lived at approximately 300 K. At elevated temperatures (500 K), the preferred reaction (within 7.3 ps) in the CPMD simulation corresponds to the McLafferty rearrangement. The estimated free activation energy associated with this process amounts to 2.5 kcal mol(-1), while the free activation energy for the delta-H transfer equals 4.4 kcal mol(-1). This relatively small free activation energy for the McLafferty rearrangement might cause dissociation of a substantial fraction of 1(+.) prior to the time-delayed mass selection, which would reduce the C3/C2 ratio in the experiments conducted with metastable ions that have a lifetime in the order of some micros at a source temperature of 500 K.
Related Concept Videos
 01:21
[3,3] Sigmatropic Rearrangement of 1,5-Dienes: Cope Rearrangement
 01:07
Radicals: Electronic Structure and Geometry
Accordingly, the structure of a trivalent radical lies between the geometries of carbocations and carbanions. An sp2-hybridized carbocation is trigonal planar, while an sp3-hybridized carbanion is trigonal pyramidal. Here, the difference in geometry is...
 01:11
Radical Reactivity: Overview
 01:10
Radical Reactivity: Steric Effects
Along with electronic factors, steric factors also account...
 01:02
Radical Reactivity: Electrophilic Radicals
 01:33
Radical Reactivity: Intramolecular vs Intermolecular