Multipoles and interaction potentials in ionic materials from planewave-DFT calculations

Andrés Aguado1, Leonardo Bernasconi, Sandro Jahn
1Physical and Theoretical Chemistry Laboratory, University of Oxford, South Parks Road, Oxford, UK OX1 3QZ.

Abstract:
Oxide potentials which transfer well between different materials have to account explicitly for many-body contributions to the interaction potentials between the ions. These include dipole and quadrupole polarization effects and the compression and deformation of an oxide ion by its immediate coordination environment. Such complex potentials necessarily involve many parameters. We examine how the results of ab initio electronic structure calculations, based upon planewave DFT methods, on general configurations of ions derived from simulations at finite temperature, may be used to parameterize an "aspherical ion method" (AIM) potential (A. J. Rowley, P. Jemmer, M. Wilson and P. A. Madden, J Chem. Phys., 1998, 108, 10209). Dipoles and quadrupoles on the individual ions are obtained via a transformation of the Kohn Sham orbitals to localized orbitals on each ion, which enables a distorted charge density for each ion to be obtained. The dipoles and quadrupoles appearing in polarization parts of the AIM potential are fit to those obtained from the ab initio ionic charge densities obtained in this way. The remaining parts of the potential, describing short-range repulsive interactions between ions with compressed and deformed charge densities, are fit to the ab initio forces and the stress tensor. By using a sufficiently large and varied set of configurations on which to carry out this optimization, an excellent transferable potential is obtained.
More Related Videos

Related Concept Videos
 03:13
Intermolecular Forces
03:13Intermolecular Forces
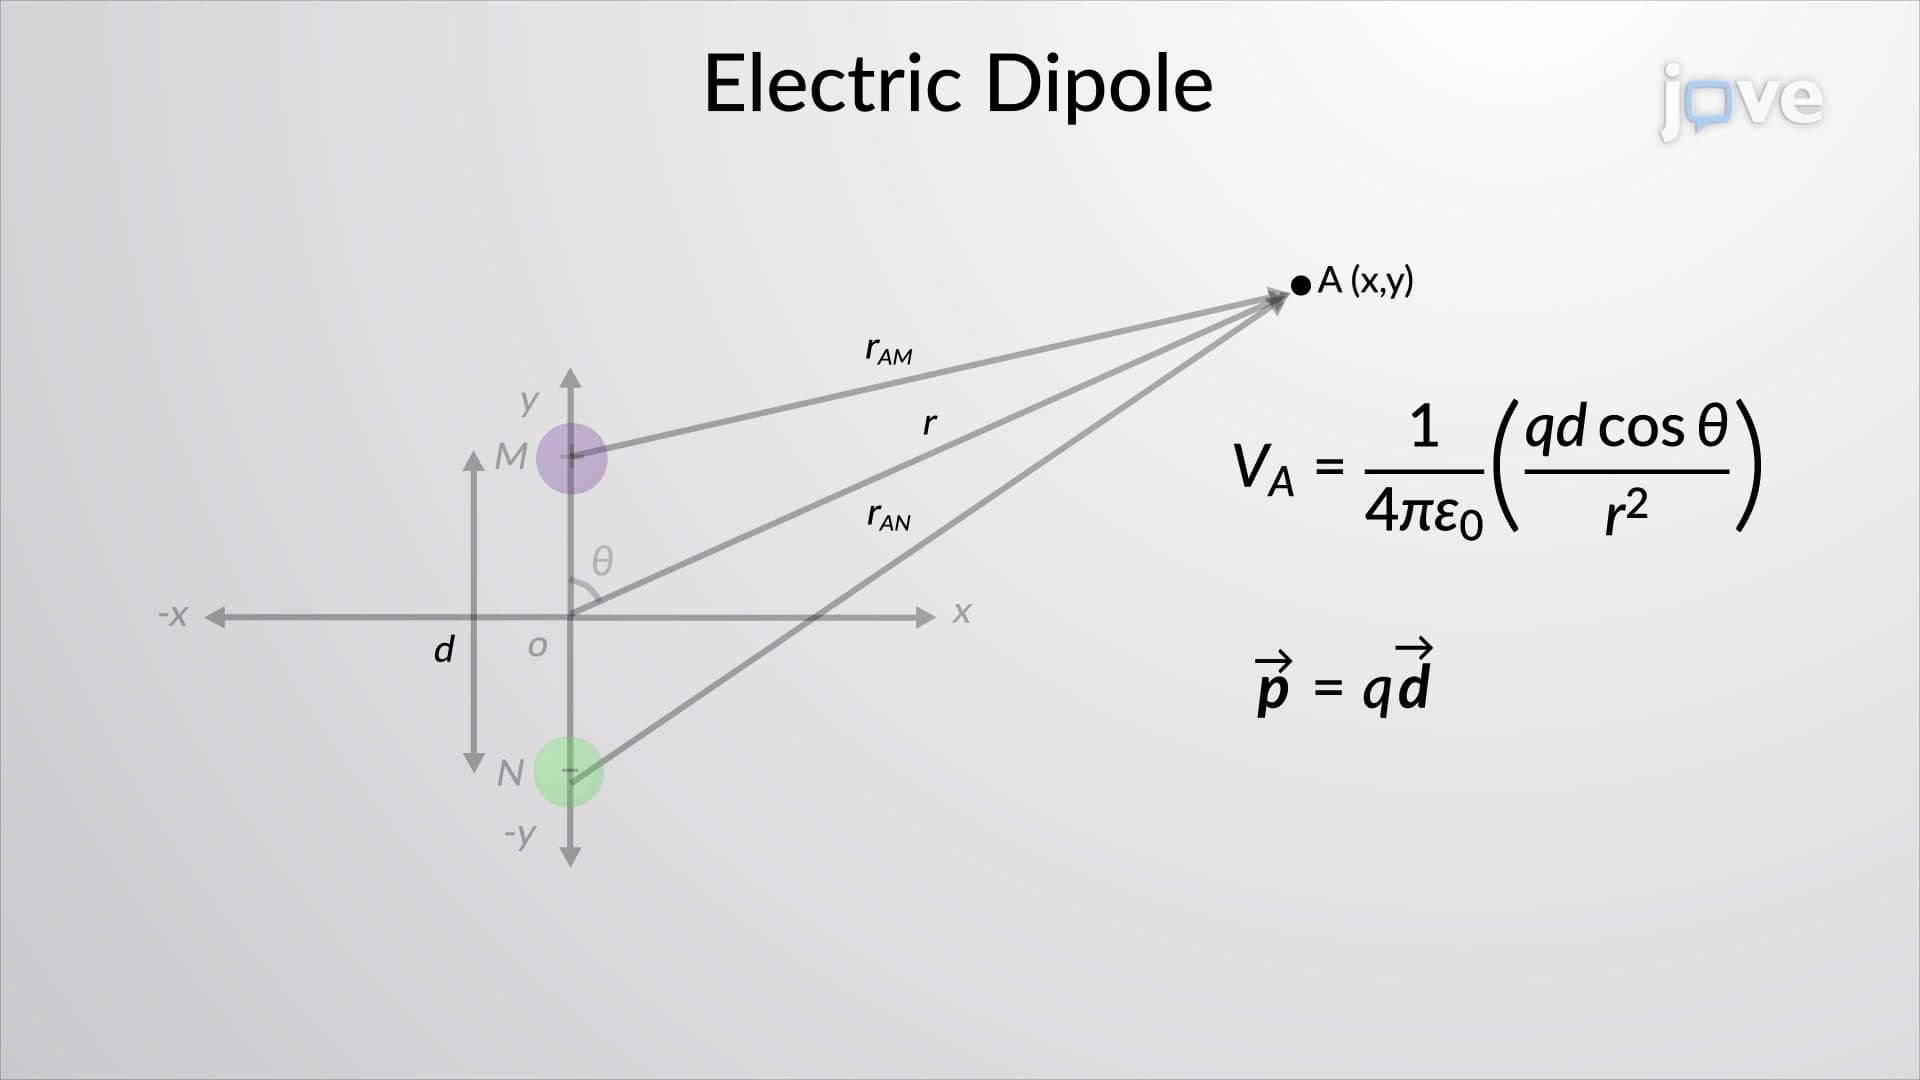 01:27
Calculations of Electric Potential II
Consider a...
 02:42
Valence Bond Theory
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
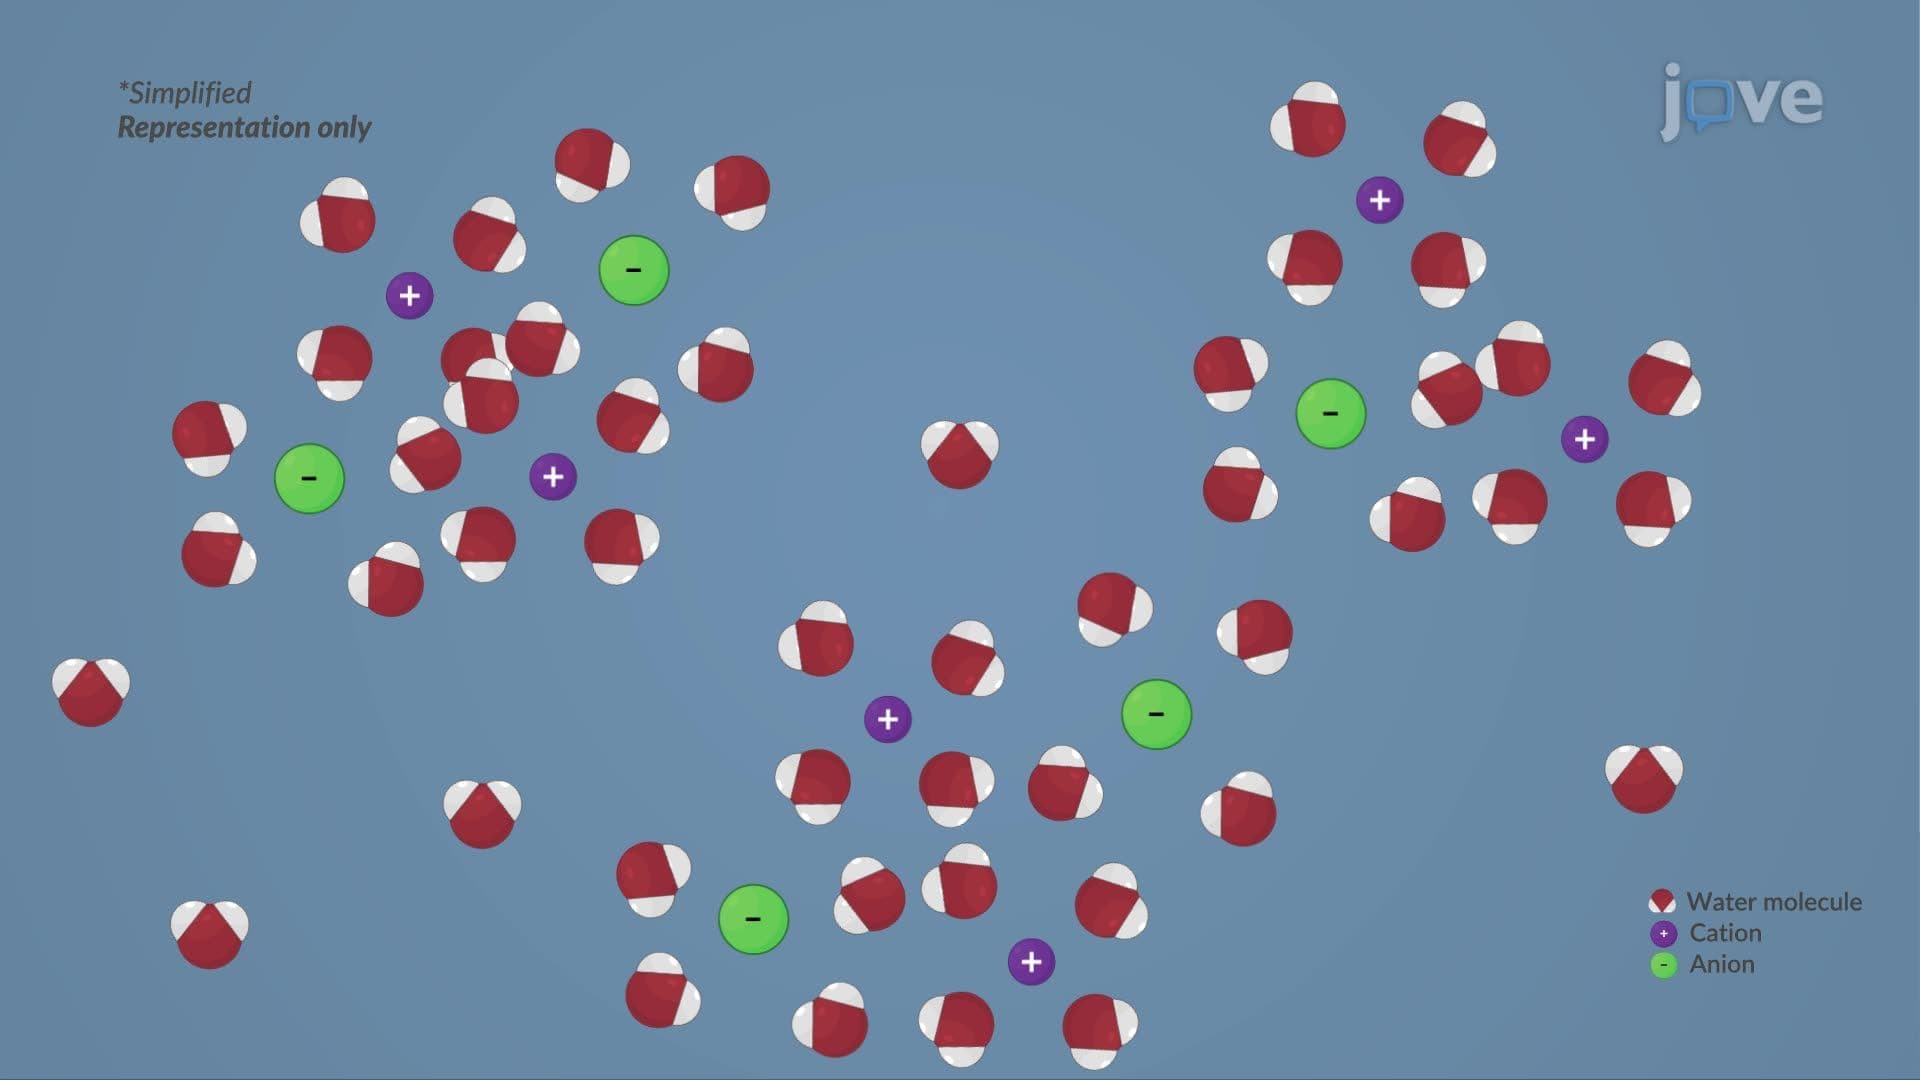 01:23
Theory of Strong Electrolytes