Molecular dynamics simulations on HIV-1 Tat
Related Experiment Video
Updated: Aug 30, 2026
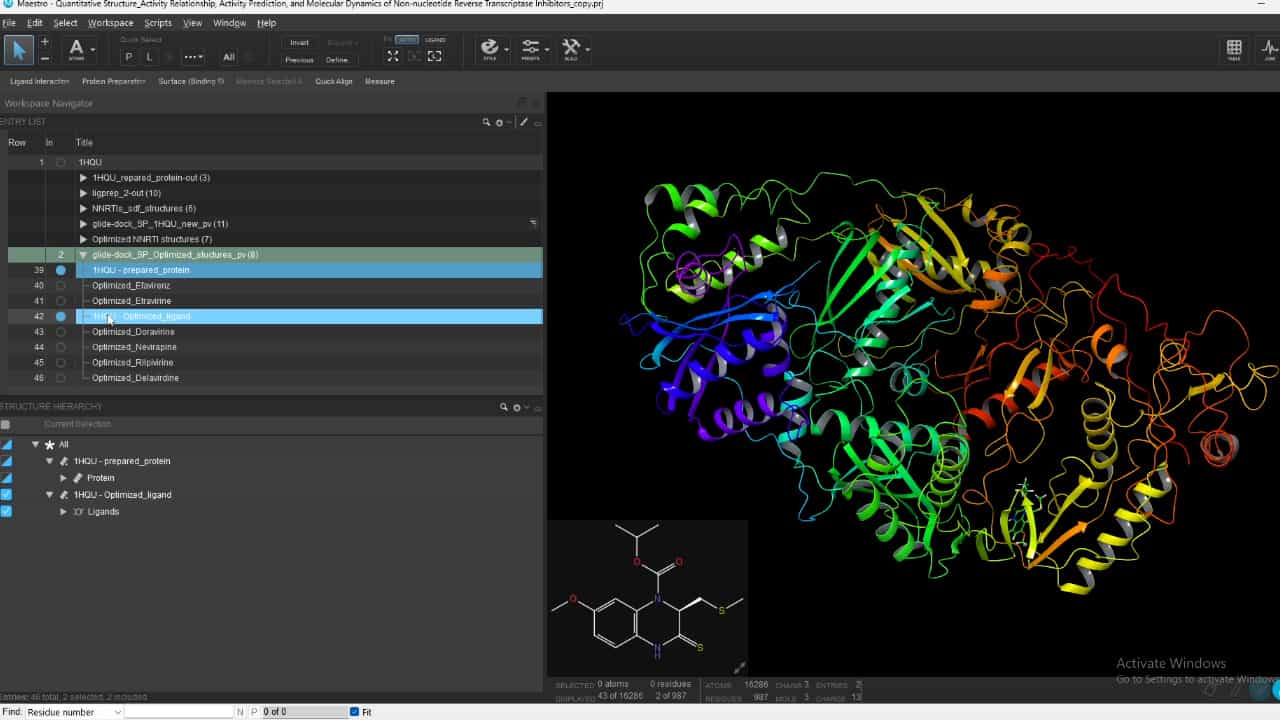
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Sergio Pantano1, Mudit Tyagi, Mauro Giacca
1International School for Advanced Studies (ISAS) and INFM, DEMOCRITOS Modeling Center for Research in Atomistic Simulation, Via Beirut 2-4, 34014, Trieste, Italy.

Abstract:
Molecular dynamics simulations are used to investigate dynamics and intramolecular interactions of the HIV-1 transactivator (Tat) in aqueous solution. The calculations are based on the AMBER force field with particle mesh Ewald treatment for long-range electrostatics. The Tat structure exhibits a large flexibility, consistent with its absence of secondary structure elements. From an analysis of the correlation matrix and of electrostatic interactions we suggest that segments expressed by the two exons (amino acids 1-72 and 73-86, respectively) exhibit rather separated dynamic and energetic properties. We also identify intramolecular interactions of importance for structure stabilization. In particular, significant electrostatic interactions are recognized between the N-terminus and the basic domain of the protein, consistent with site-directed mutagenesis performed in this work.
More Related Videos
 07:29
Determination of Molecular Structures of HIV Envelope Glycoproteins using Cryo-Electron Tomography and Automated Sub-tomogram Averaging
Published on: December 1, 2011
 11:19
Pairwise Growth Competition Assay for Determining the Replication Fitness of Human Immunodeficiency Viruses
Published on: May 4, 2015