Related Experiment Video
Updated: Aug 30, 2026
A High-throughput Calcium-flux Assay to Study NMDA-receptors with Sensitivity to Glycine/D-serine and Glutamate
Published on: July 10, 2018
Optically active mexiletine analogues as stereoselective blockers of voltage-gated Na(+) channels
Carlo Franchini1, Alessia Carocci, Alessia Catalano
1Dipartimento Farmaco-Chimico and Farmaco-Biologico, Università degli Studi di Bari - Via E Orabona 4, 70125 Bari, Italy. cfranc@farmchim.uniba.it

Abstract:
Optically active mexiletine analogues were synthesized and evaluated in vitro as use-dependent blockers of skeletal muscle sodium channels. The mexiletine analogues were obtained by replacing either the methyl group on the stereogenic center of mexiletine [1-(2,6-dimethylphenoxy)propan-2-amine] with a phenyl group or modifying the phenoxy moiety (by removal of one or both of the methyl groups, or introducing a chlorine atom), or both. The voltage clamp recordings showed that, regardless of the substitution pattern of the aryloxy moiety, all the compounds bearing a phenyl group on the stereogenic center (3a-f) were more active than mexiletine both in tonic and phasic block. This observation was in contrast with what was observed for mexiletine, where the removal of both methyls from the aryloxy moiety caused a dramatic reduction of potency. The most potent congener, (R)-2-(2-methylphenoxy)-1-phenylethanamine [(R)-3b], was 27-fold more potent than (R)-mexiletine in producing a tonic block, i.e., the reduction of peak sodium current in resting conditions after application of the compound. (R)-3b maintained a use-dependent behavior, being 23-fold more potent in condition of high frequency of stimulation (phasic block). Despite what was observed with mexiletine, the stereoselectivity held in phasic block conditions. Stereoselectivity indexes were generally low, ranging from 1 to 4, but except for that of the 2,6-xylyloxy congener 3c, they were higher for the congeners bearing a phenyl ring on the stereogenic center than for mexiletine and its strictly related analogue 1-methyl-2-phenoxyethanamine (1). This finding was in agreement with Pfeiffer's rule. The introduction of a chlorine atom in the 4-position of the aryloxy moiety caused a reduction of potency and a reversal of stereoselectivity as well. On the basis of the model to date accepted for the sodium channel local anesthetic-like molecule receptor, some possible explanations of our observations will be proposed.
More Related Videos
Related Concept Videos
 01:22
Antiarrhythmic Drugs: Class I Agents as Sodium Channel Blockers
Class 1A Antiarrhythmic Drugs: These drugs work by moderately blocking sodium channels,...
 01:28
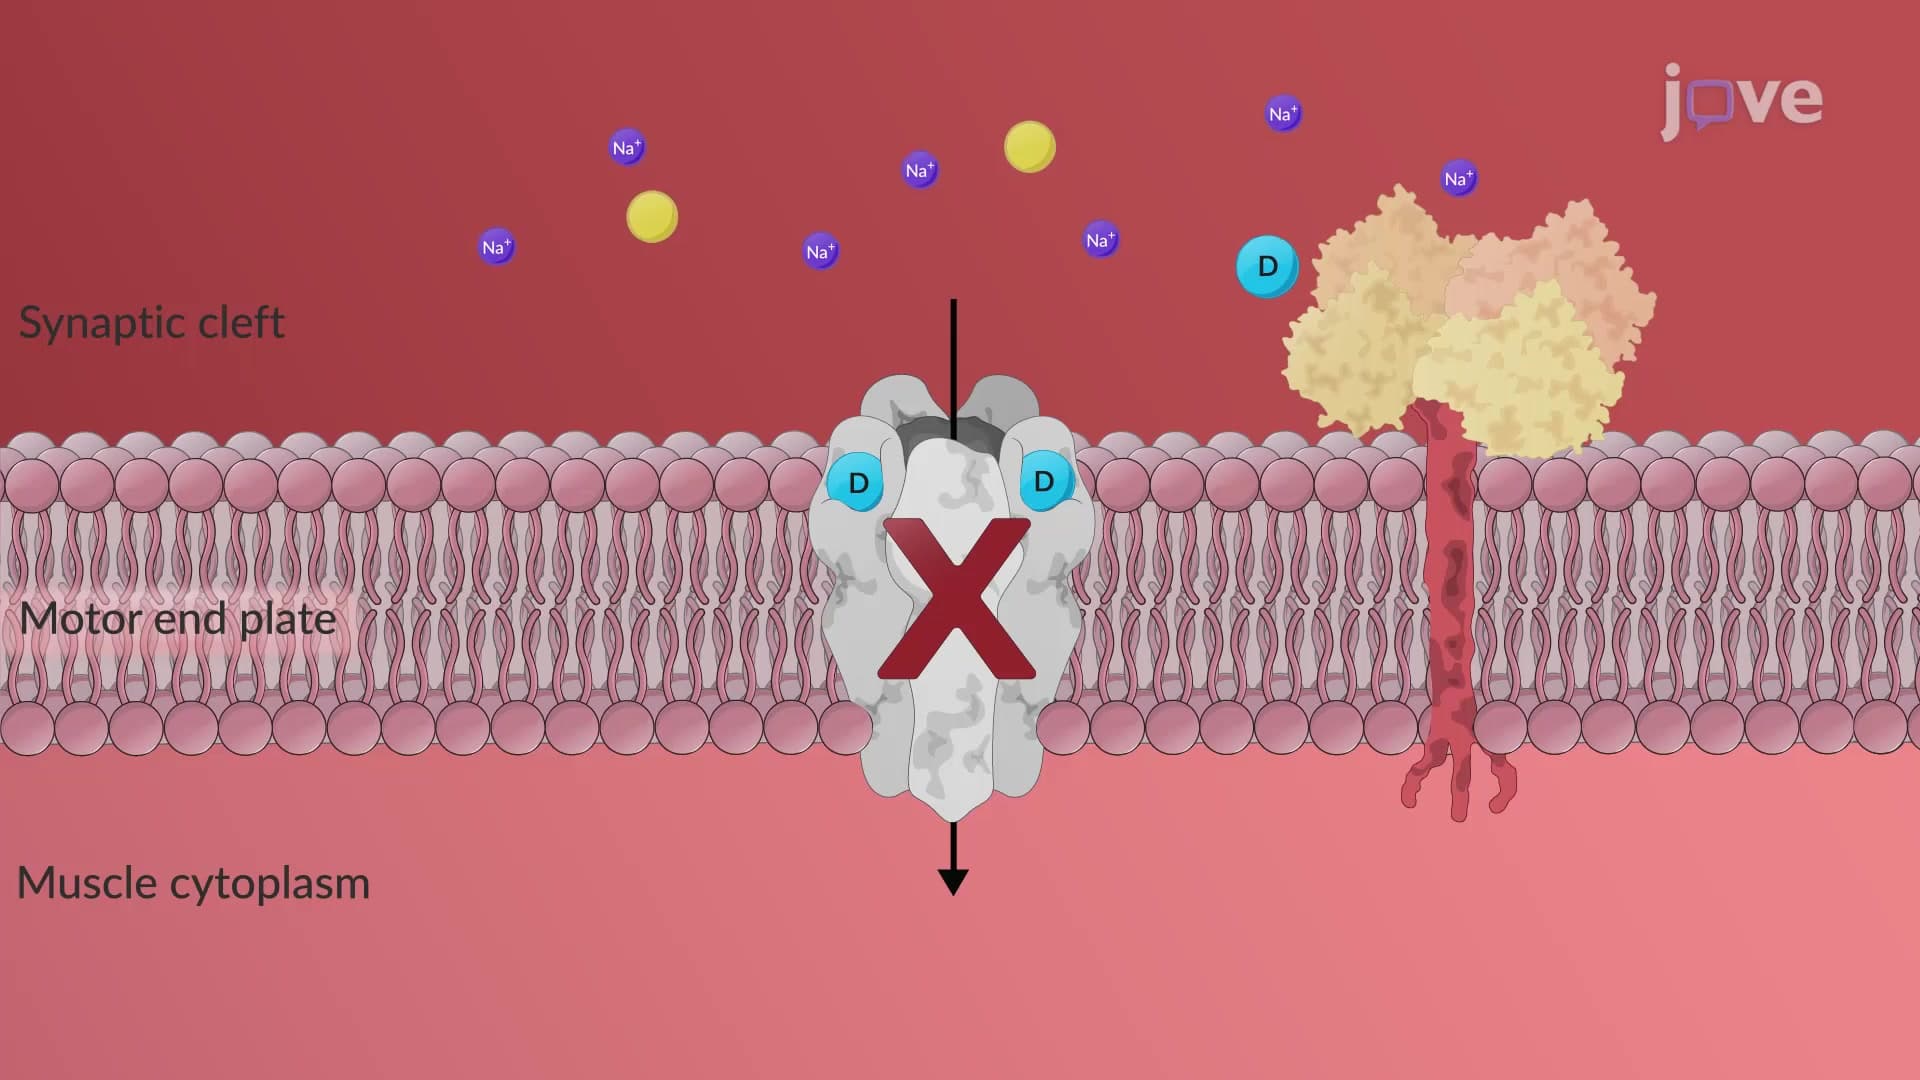
Depolarizing Blockers: Mechanism of Action
Succinylcholine is the most commonly used depolarizing blocker. Chemically, it constitutes two molecules of acetylcholine joined together by an acetate methyl group. They act on the receptors in the same way as acetylcholine. Because succinylcholine...
 01:17
Nondepolarizing (Competitive) Neuromuscular Blockers: Mechanism of Action
Competitive antagonists prevent acetylcholine from binding to its receptor, inhibiting membrane depolarization. Without conformational changes or intrinsic...
 01:12
Antiarrhythmic Drugs: Class III Agents as Potassium Channel Blockers
 01:20
Antiarrhythmic Drugs: Class IV Agents as Calcium Channel Blockers
Verapamil, a calcium channel blocker, inhibits calcium movement across myocardial cell membranes and vascular smooth muscle. This results in the dilation of coronary and...
 01:08
Antiepileptic Drugs: Sodium Channel Blockers
Sodium channel blockers modulate ion channels, particularly voltage-gated sodium channels. They block only sodium ion movement.
Among the most commonly prescribed antiepileptic drugs are...