Related Experiment Video
Updated: Aug 30, 2026


In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
New GAA mutations in Japanese patients with GSDII (Pompe disease)
Judy R Pipo1, Jian-Hua Feng, Toshiyuki Yamamoto
1Division of Child Neurology, Institute of Neurological Sciences, Faculty of Medicine, Tottori University, Yonago, Japan.

Abstract:
Glycogen storage disease type II (Pompe disease) is inherited by autosomal recessive transmission and caused by a deficiency of acid alpha-glucosidase (GAA), resulting in impaired degradation and lysosomal accumulation of glycogen. The GAA gene, responsible for this disease, has been mapped to chromosome 17q25.2-25.3. To date, more than 70 disease-causing mutations have been identified. In this study, we present four mutations found in three Japanese patients with the juvenile form of glycogen storage disease type II; three of these mutations were new (R224W, S619R, and R660H). The pathogenicity of these new mutations was verified by the loss of function of the mutant enzymes expressed in COS cells.
More Related Videos
Related Concept Videos
 02:27
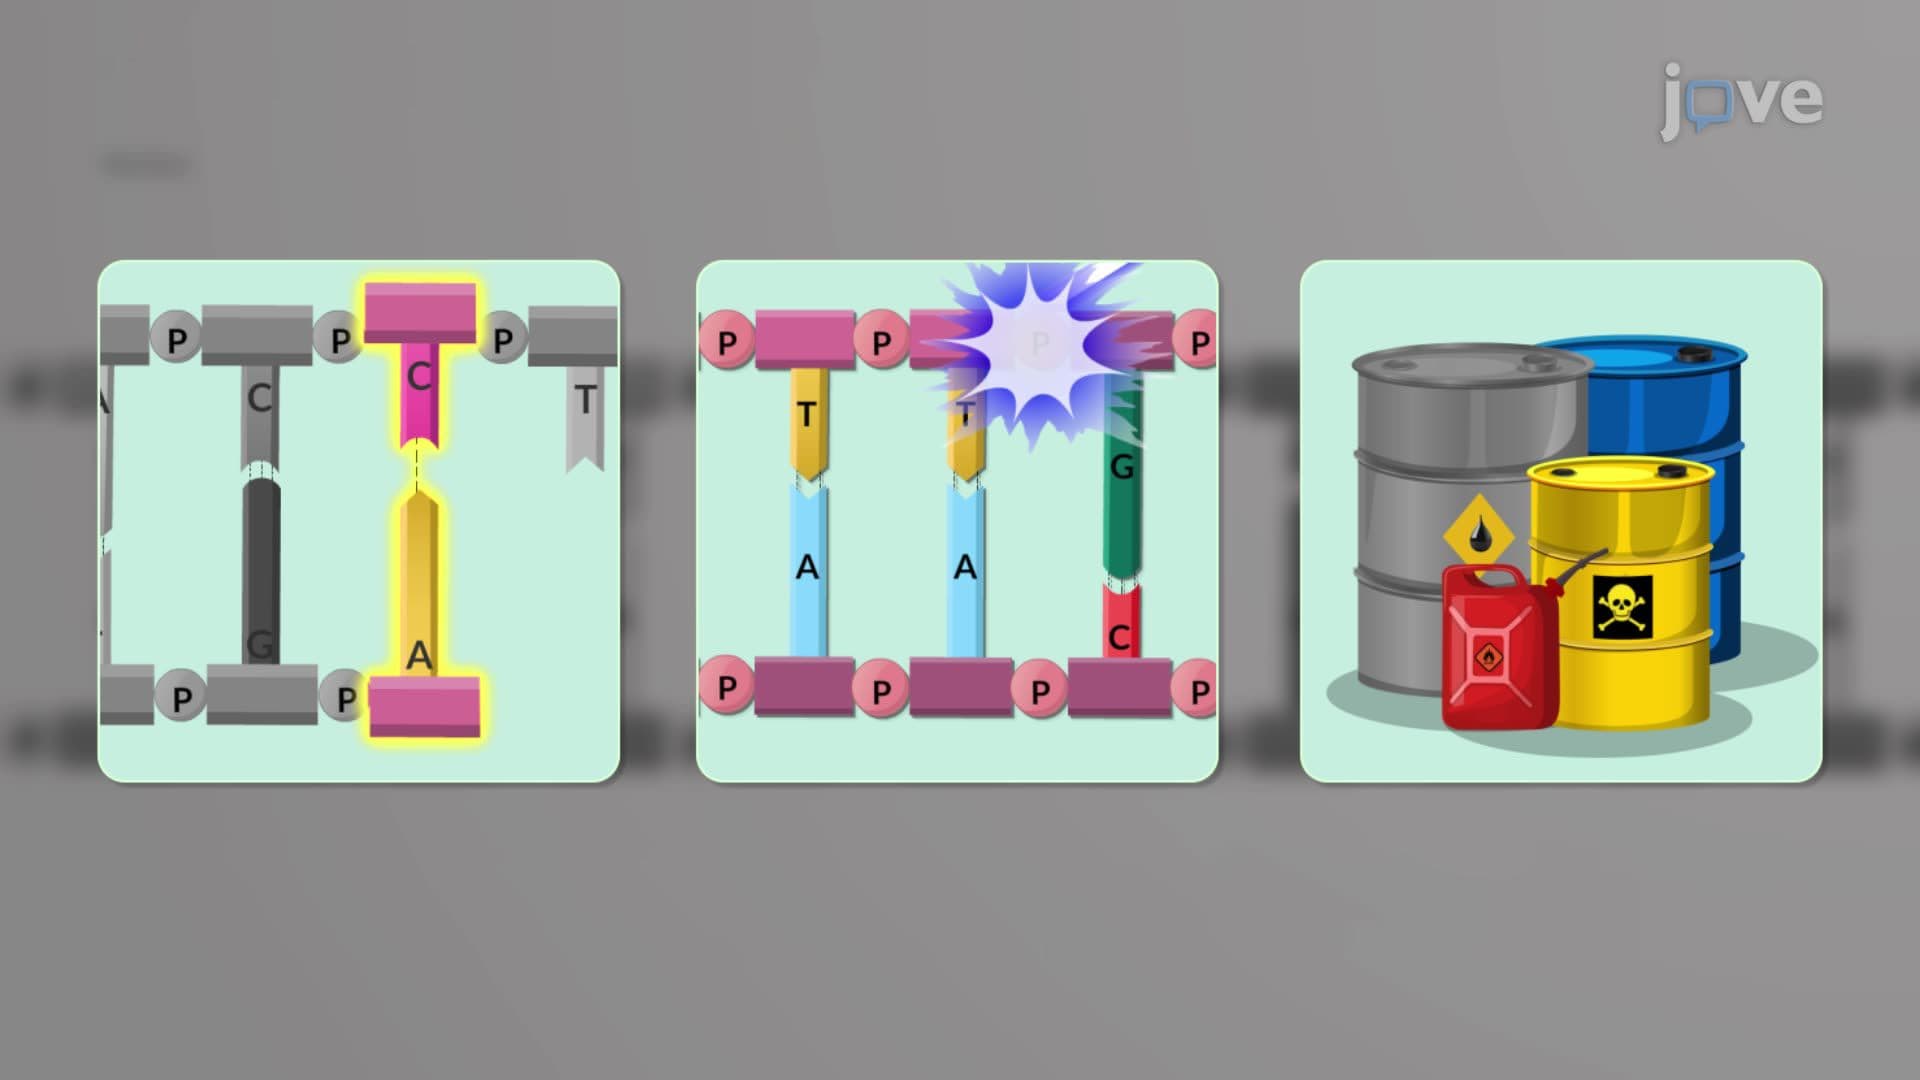
Mutations
 01:37
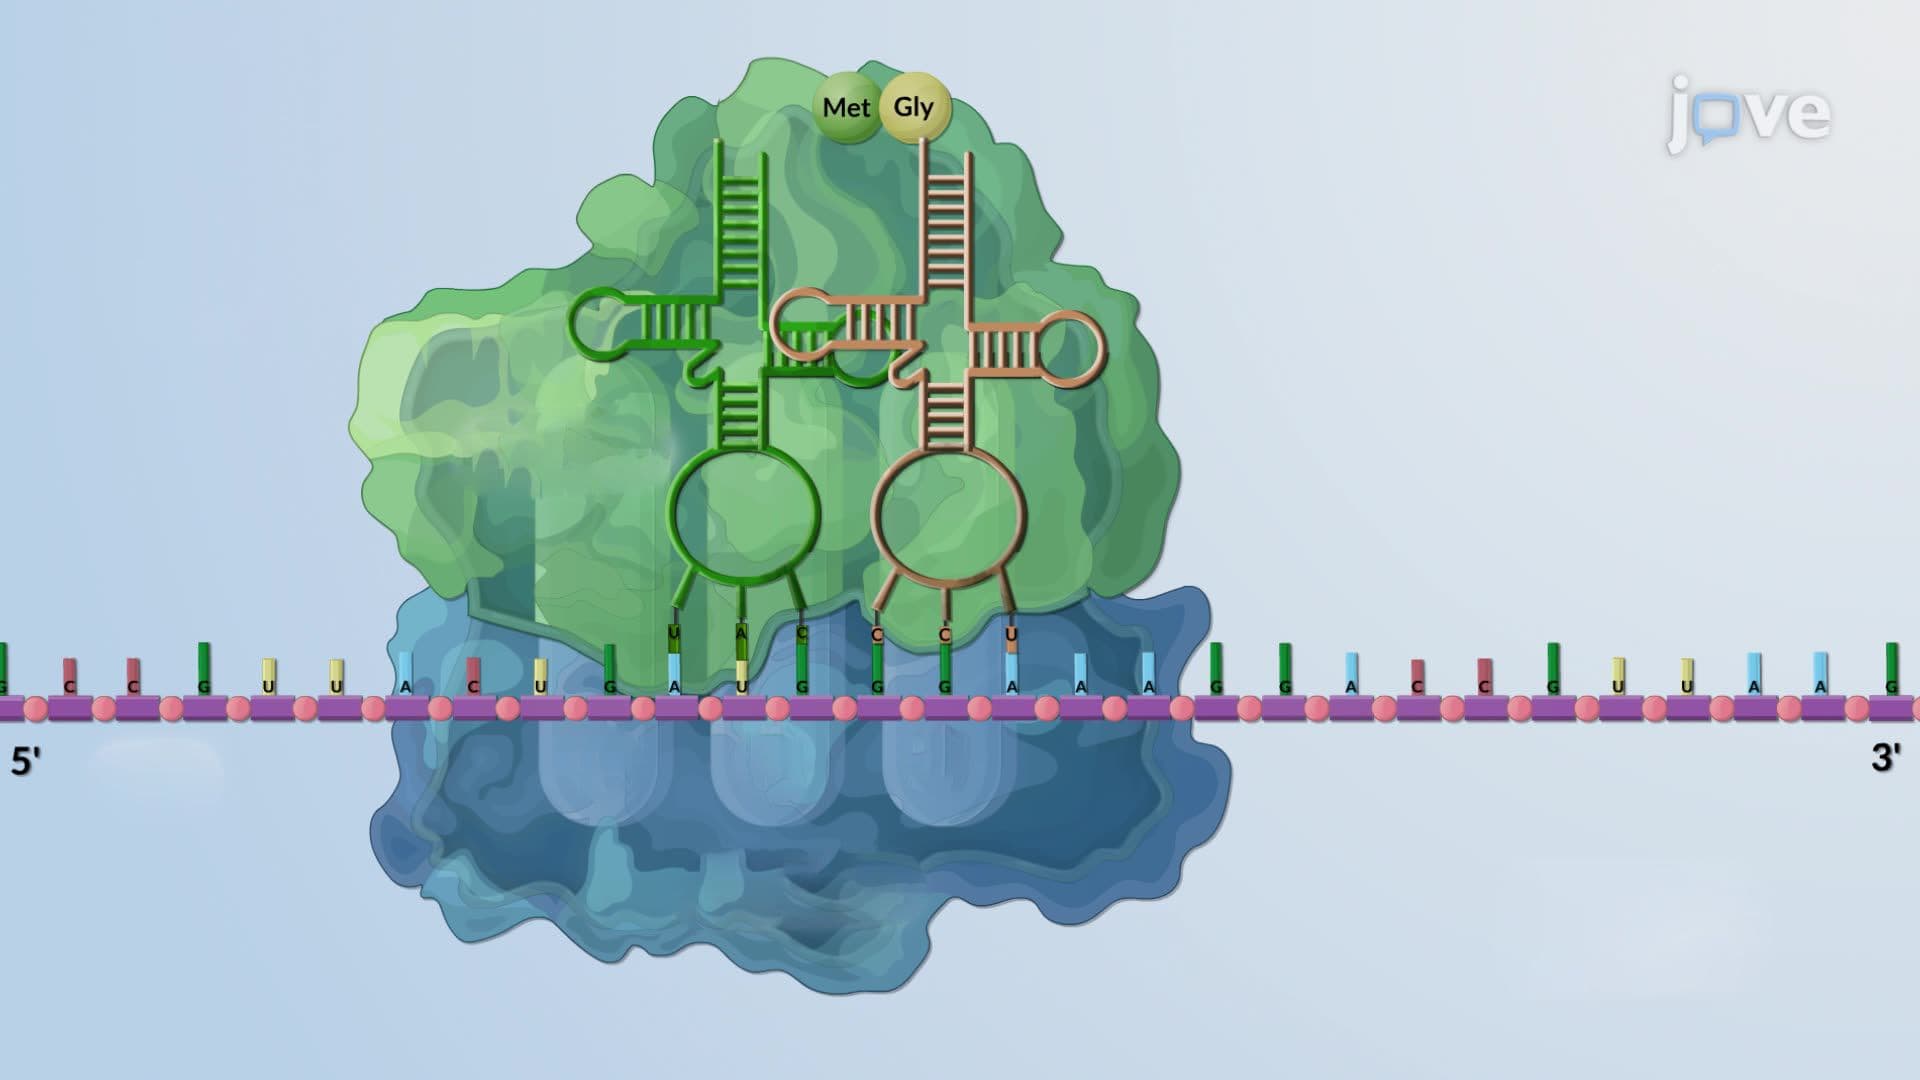
Translation
 01:05
Single Nucleotide Polymorphisms-SNPs
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 02:14
GTPases and their Regulation
Large G-proteins, also known...