Related Experiment Video
Updated: Aug 29, 2026

Determination of Lipid Raft Partitioning of Fluorescently-tagged Probes in Living Cells by Fluorescence Correlation Spectroscopy (FCS)
Published on: April 6, 2012
[Lipid rafts, flotillin-1 and Alzheimer disease]
Nadège Girardot1, Bernadette Allinquant, Charles Duyckaerts
1Laboratoire de Neuropathologie Raymond Escourolle et Inserm U106, Hôpital de La Salpêtrière, Paris.

Abstract:
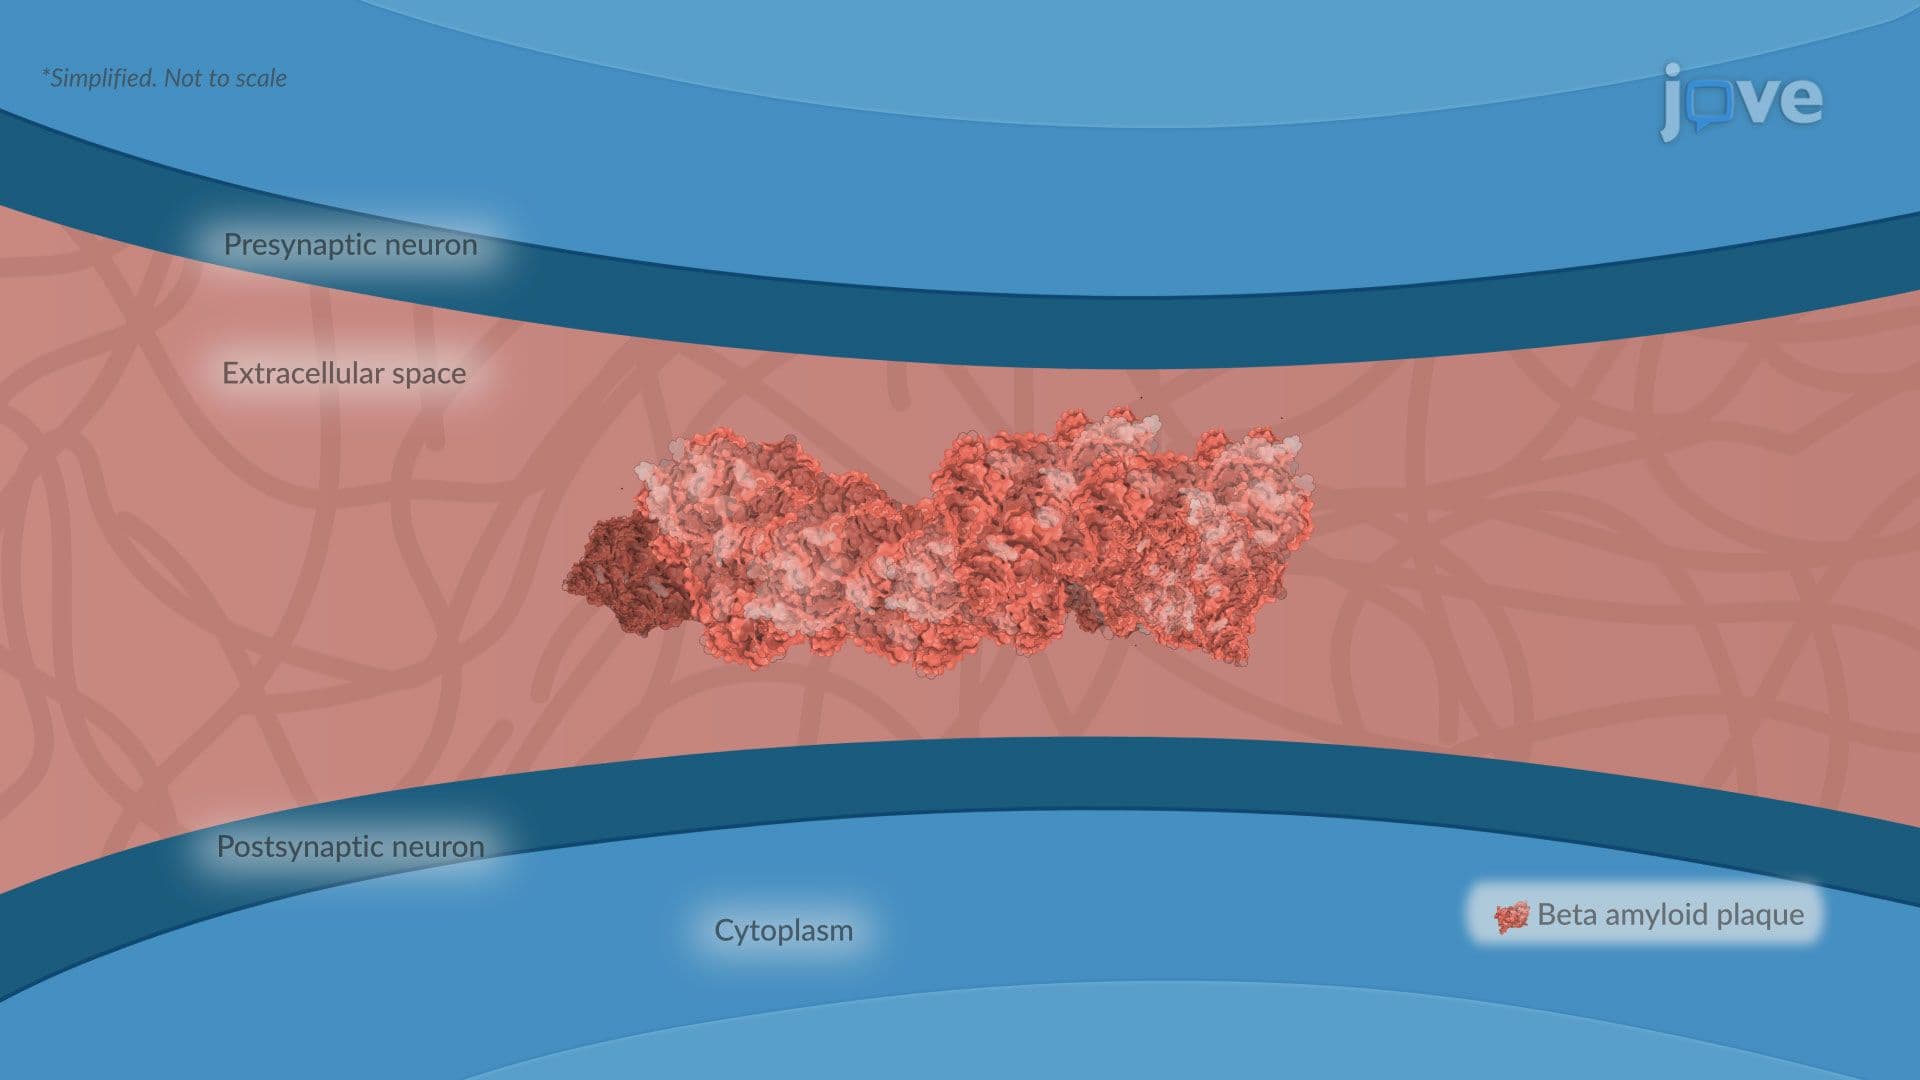
A beta peptide accumulates in the extracellular space during Alzheimer's disease. It is the cleavage product of APP (Amyloid Precursor Protein), a large transmembrane protein. After ultracentrifugation, APP is found in a low-density fraction, enriched in cholesterol. These properties are characteristic of lipid rafts, which are microdomains that "float" like rafts on the plasma membrane. We have confirmed the presence of cholesterol in the core of the senile plaque, using the fluorescent probe filipin. In addition, we have shown that flotillin-1, a marker of rafts, accumulated in lysosomes of neurons in Alzheimer's disease. In most cases (76% of the flotillin-1 positive neurons), the accumulation was associated with the presence of neurofibrillary tangles. Our data suggest that the A beta peptide, which is poorly soluble in water, is actually linked with cholesterol, possibly from cellular membranes, in the extracellular space.
Related Concept Videos
 01:23
Alzheimer Disease ll: Pathophysiology
 01:29
Alzheimer Disease l: Introduction
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
03:03Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:22
Dementia l: Introduction
 01:19
Membrane Asymmetry Regulating Transporters
Flippase
Eukaryotic flippases are type-IV P-type ATPases or P4-ATPases belonging to P-type ATPase family proteins that are membrane-bound pumps involved in the ATP-mediated transport of ions and molecules across the membrane. Flippases flip specific phospholipids from the outer to the inner leaflet of a membrane. All P4-ATPases have one...