Related Experiment Video
Updated: Aug 29, 2026
Implementation of Non-invasive Point of Care Transient Elastography for Evaluation of Liver Disease in Pediatric Populations with Cystic Fibrosis
Published on: August 29, 2025
[Cystic fibrosis from the exocrine pancreatic point of view]
1Clalit Health Services, Sharon-Shomron District, Hadera Child Health Center, Israel.

Abstract:
Cystic fibrosis (CF) is the most common life-limiting autosomal recessive genetic disorder in Caucasians and is characterized by a wide variability of clinical expression. The vast majority of patients with CF have pancreatic insufficiency (PI) requiring exogenous pancreatic enzyme replacement therapy with meals. It is caused by mutations in the cystic fibrosis transmembrane conductance regulator (CFTR) gene, which encodes a protein that functions as a chloride channel and is regulated by cAMP. CFTR gene mutations may be classified as severe or mild with respect to pancreatic function status. Patients homozygous for two severe mutations experience severe clinical presentation including PI. Patients carrying at least one mild mutation are considered to be pancreatic sufficient (PS) and carry an overall prognosis that is vastly superior to CF patients with PI. Thus mild mutations appear to be dominant, providing enough functional CFTR to avoid PI. Five general mechanisms have been proposed which describe how CFTR gene mutations influence CFTR-mediated chloride secretion. Classes 1, 2, and 3. that confer little or no chloride channel function, confer the PI phenotype. In contrast, classes 4 and 5 allow CFTR residual function, that are expected to confer the less severe PS phenotype. Recent advances in mutation detection technology and the demonstration of characteristic abnormalities in trans-epithelial potential difference measurements, have expanded the spectrum of diseases associated with the CFTR mutant genes, and paradoxically, in some ways complicated rather than simplified CF diagnosis. Until new diagnostic criteria is conclusively determined, CF diagnosis should be made on clinical rather than laboratory grounds.
More Related Videos
 15:12
Purification of the Cystic Fibrosis Transmembrane Conductance Regulator Protein Expressed in Saccharomyces cerevisiae
Published on: May 10, 2014
 08:00
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:25
Chronic Pancreatitis I: Introduction
 01:24
Chronic Pancreatitis I: Introduction
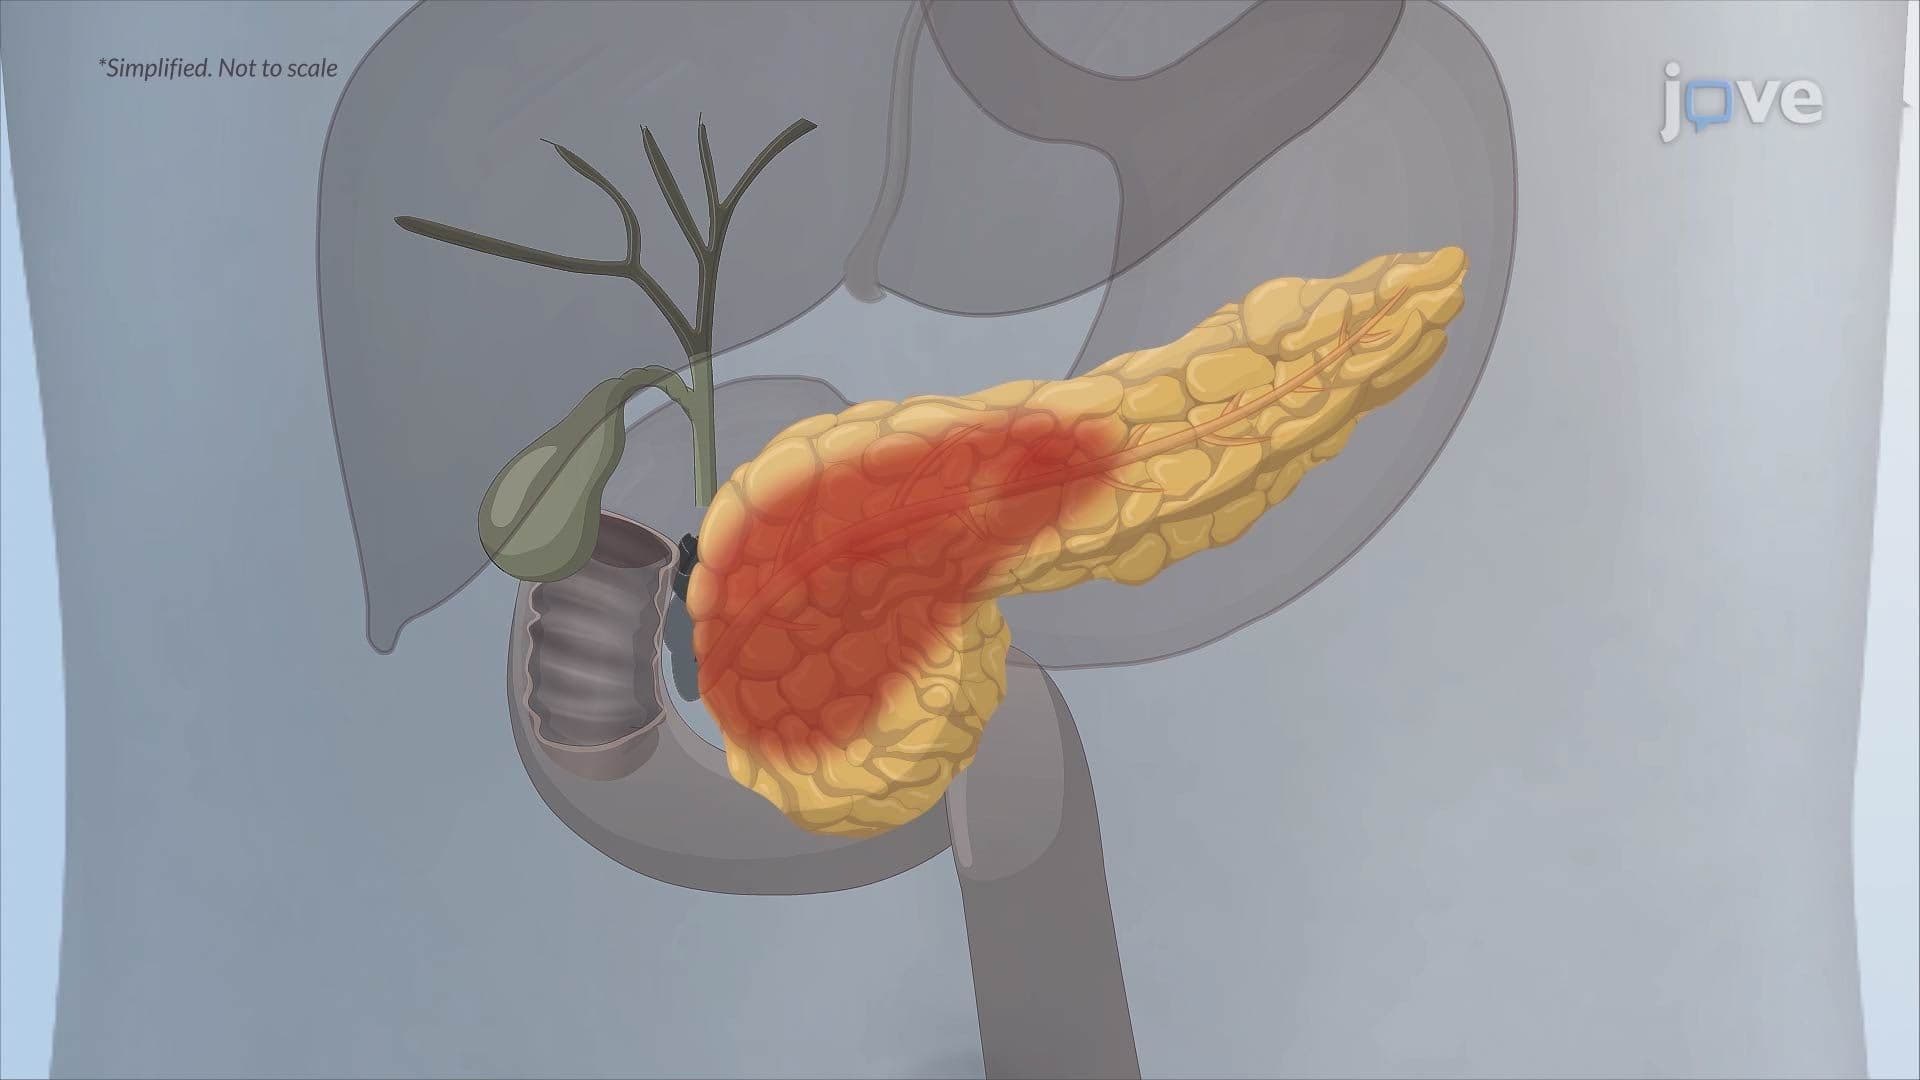
Pancreatitis is the inflammation of the pancreas, which occurs when the immune system becomes active and causes swelling, pain, and disruptions in organ function. Pancreatitis can manifest as either an acute or chronic condition.
Acute pancreatitis arises suddenly and lasts for a brief duration, while chronic pancreatitis is a long-term affliction...
 01:21
Chronic Pancreatitis II: Pathophysiology
 01:19
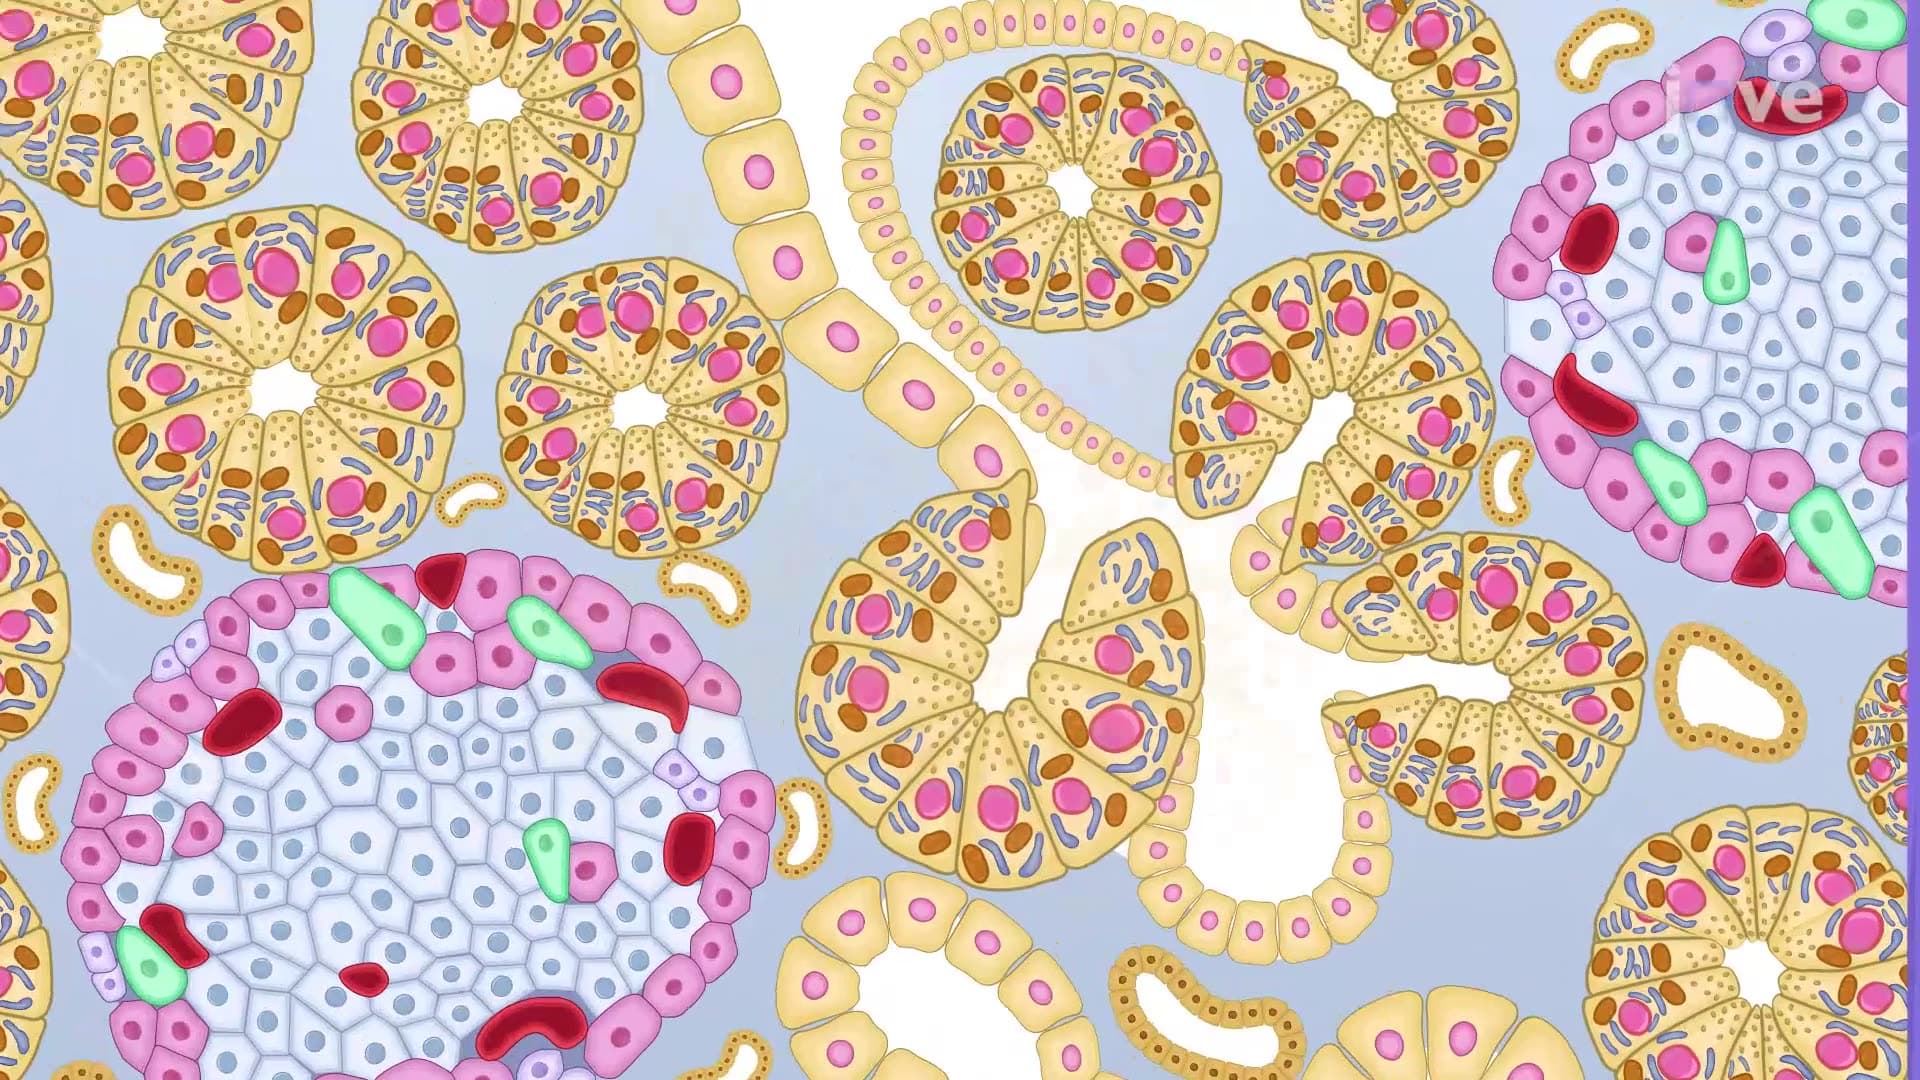
Pancreas
The broad head of the pancreas lies within the loop formed by the duodenum, while its slender body reaches towards the spleen. The tail of the pancreas is short and...
 01:16
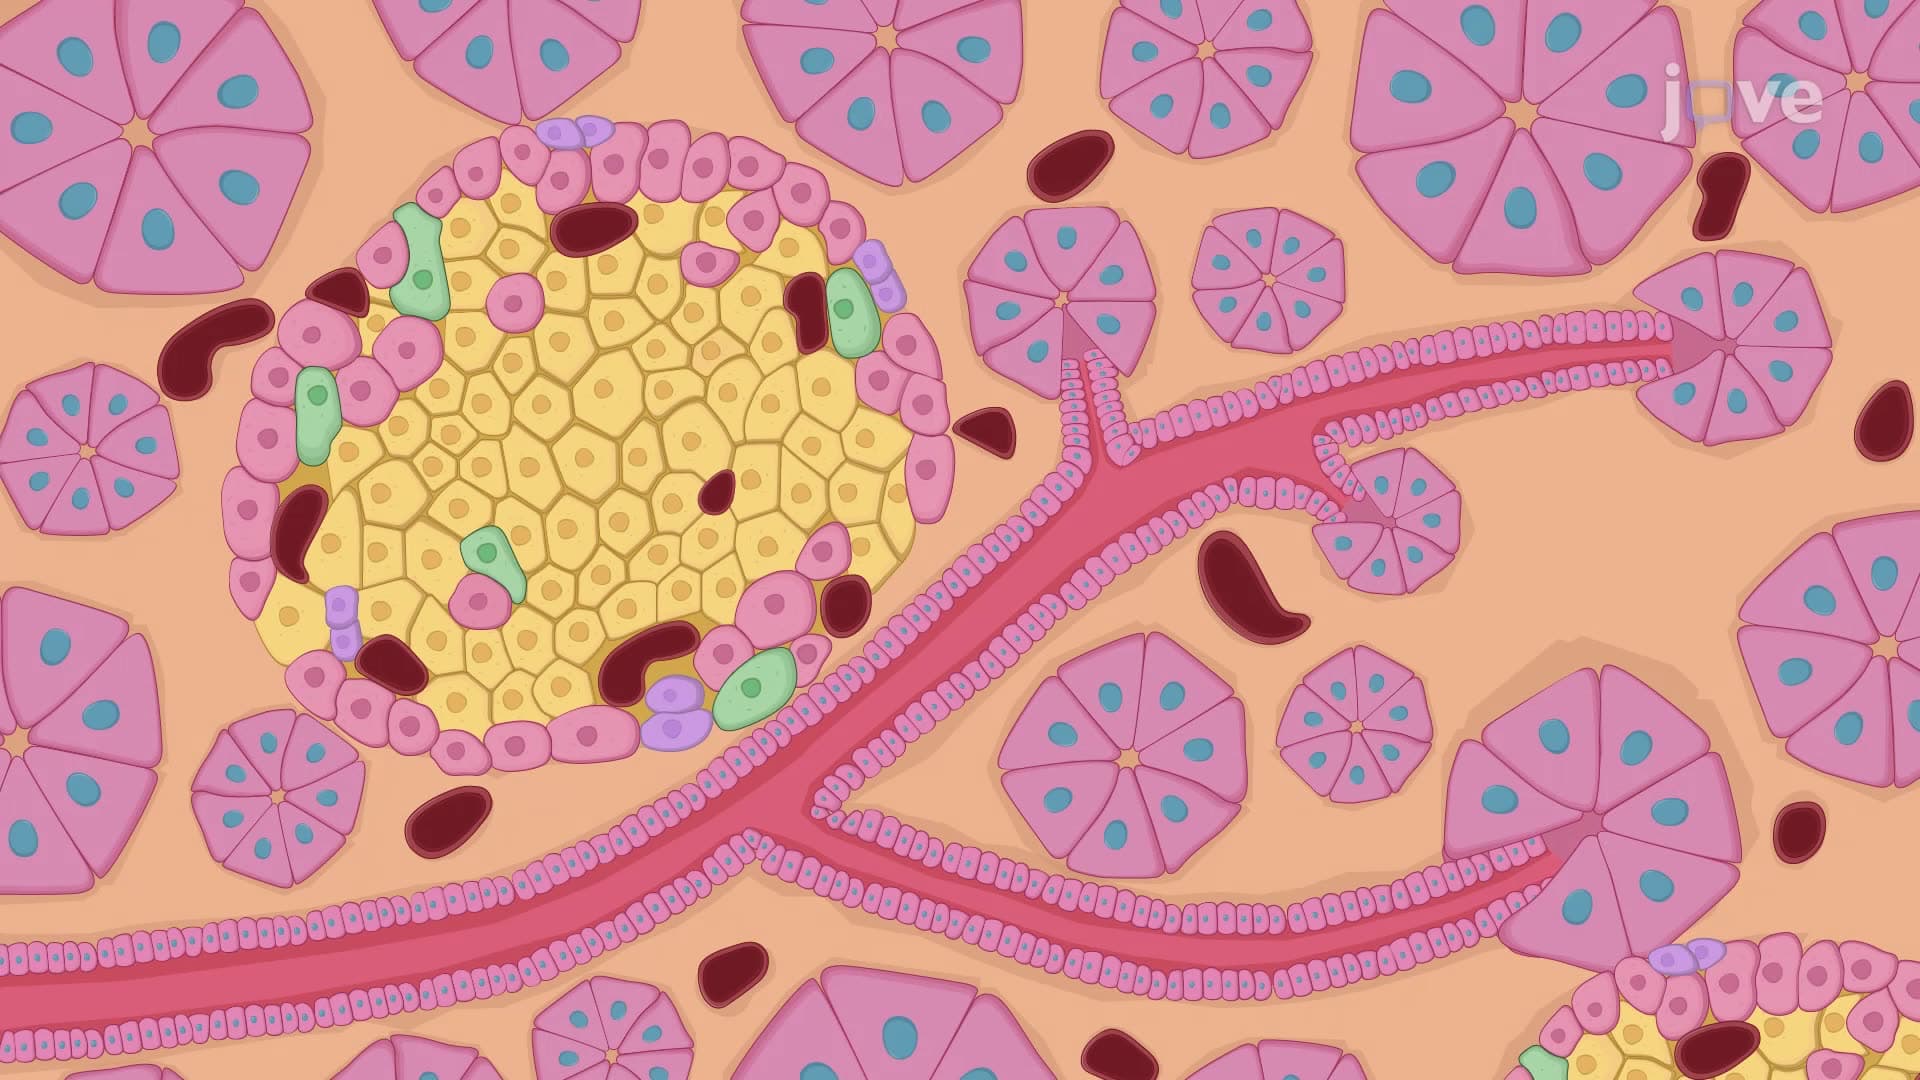
Cells and Secretions of the Pancreas
Exocrine function is carried out by acinar cells, organized into clusters known as acini. These cells contribute to digestion by releasing substantial quantities of enzyme-rich, alkaline digestive juices.
Concurrently, the dispersed clusters of endocrine cells throughout the...