Related Experiment Video
Updated: Jul 9, 2026

Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Rapid protein-ligand docking using soft modes from molecular dynamics simulations to account for protein
1Computational Biology, School of Engineering and Science, International University Bremen, Bremen, Germany. m.zacharias@iu-bremen.de

Abstract:
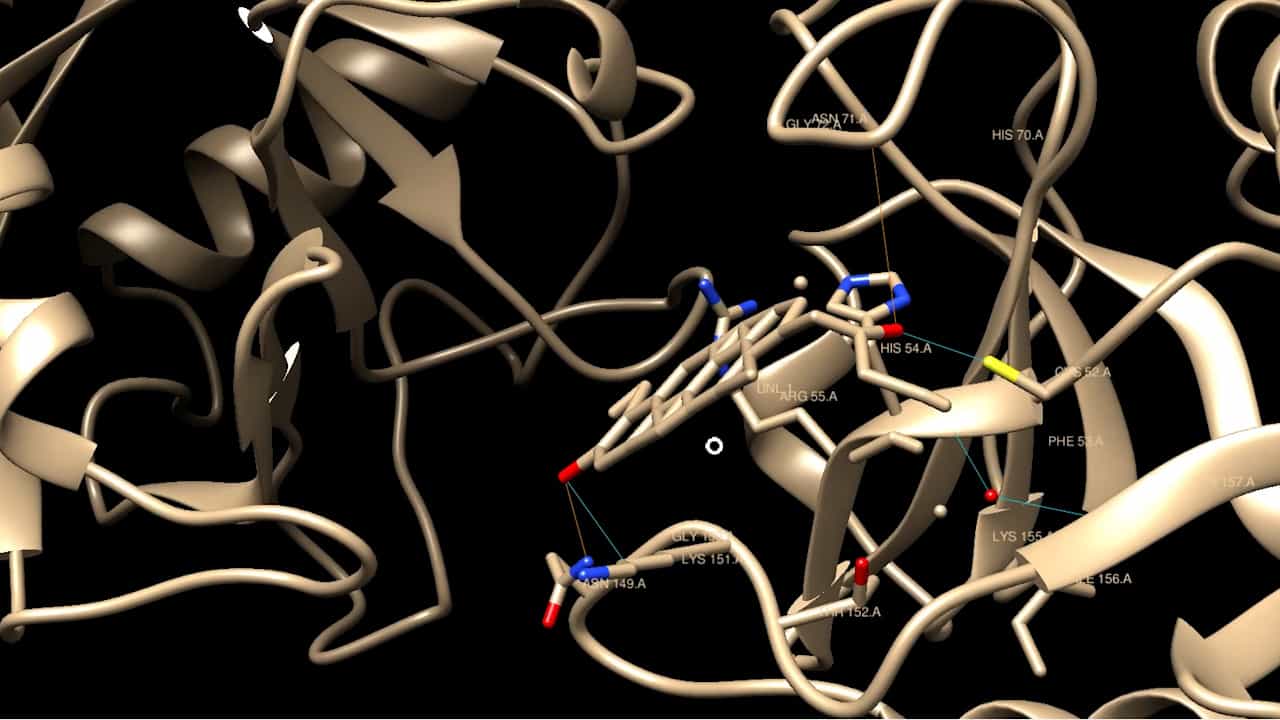
Most current docking methods to identify possible ligands and putative binding sites on a receptor molecule assume a rigid receptor structure to allow virtual screening of large ligand databases. However, binding of a ligand can lead to changes in the receptor protein conformation that are sterically necessary to accommodate a bound ligand. An approach is presented that allows relaxation of the protein conformation in precalculated soft flexible degrees of freedom during ligand-receptor docking. For the immunosuppressant FK506-binding protein FKBP, the soft flexible modes are extracted as principal components of motion from a molecular dynamics simulation. A simple penalty function for deformations in the soft flexible mode is used to limit receptor protein deformations during docking that avoids a costly recalculation of the receptor energy by summing over all receptor atom pairs at each step. Rigid docking of the FK506 ligand binding to an unbound FKBP conformation failed to identify a geometry close to experiment as favorable binding site. In contrast, inclusion of the flexible soft modes during systematic docking runs selected a binding geometry close to experiment as lowest energy conformation. This has been achieved at a modest increase of computational cost compared to rigid docking. The approach could provide a computationally efficient way to approximately account for receptor flexibility during docking of large numbers of putative ligands and putative docking geometries.
More Related Videos
 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Related Concept Videos
 02:40
Ligand Binding Sites
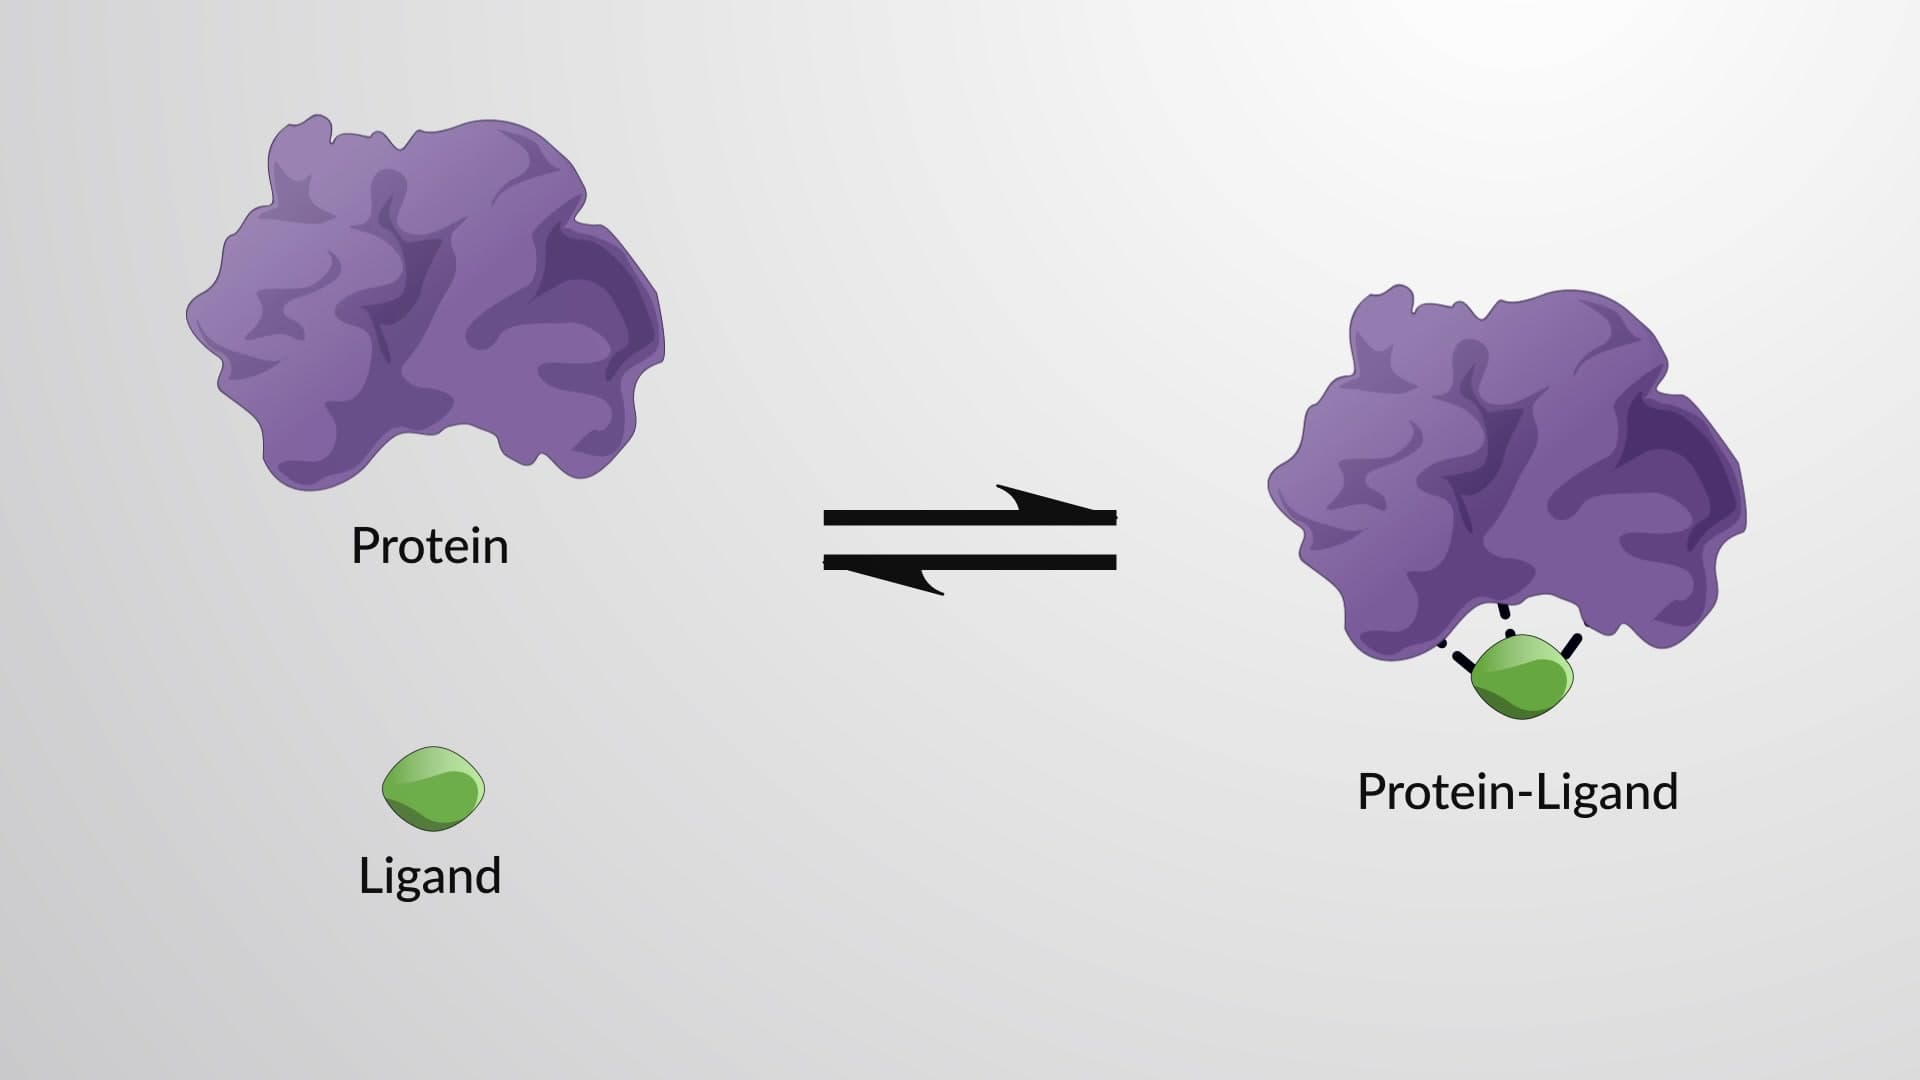
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:04
Protein-protein Interfaces
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
 01:22
Protein-Drug Binding: Determination Methods
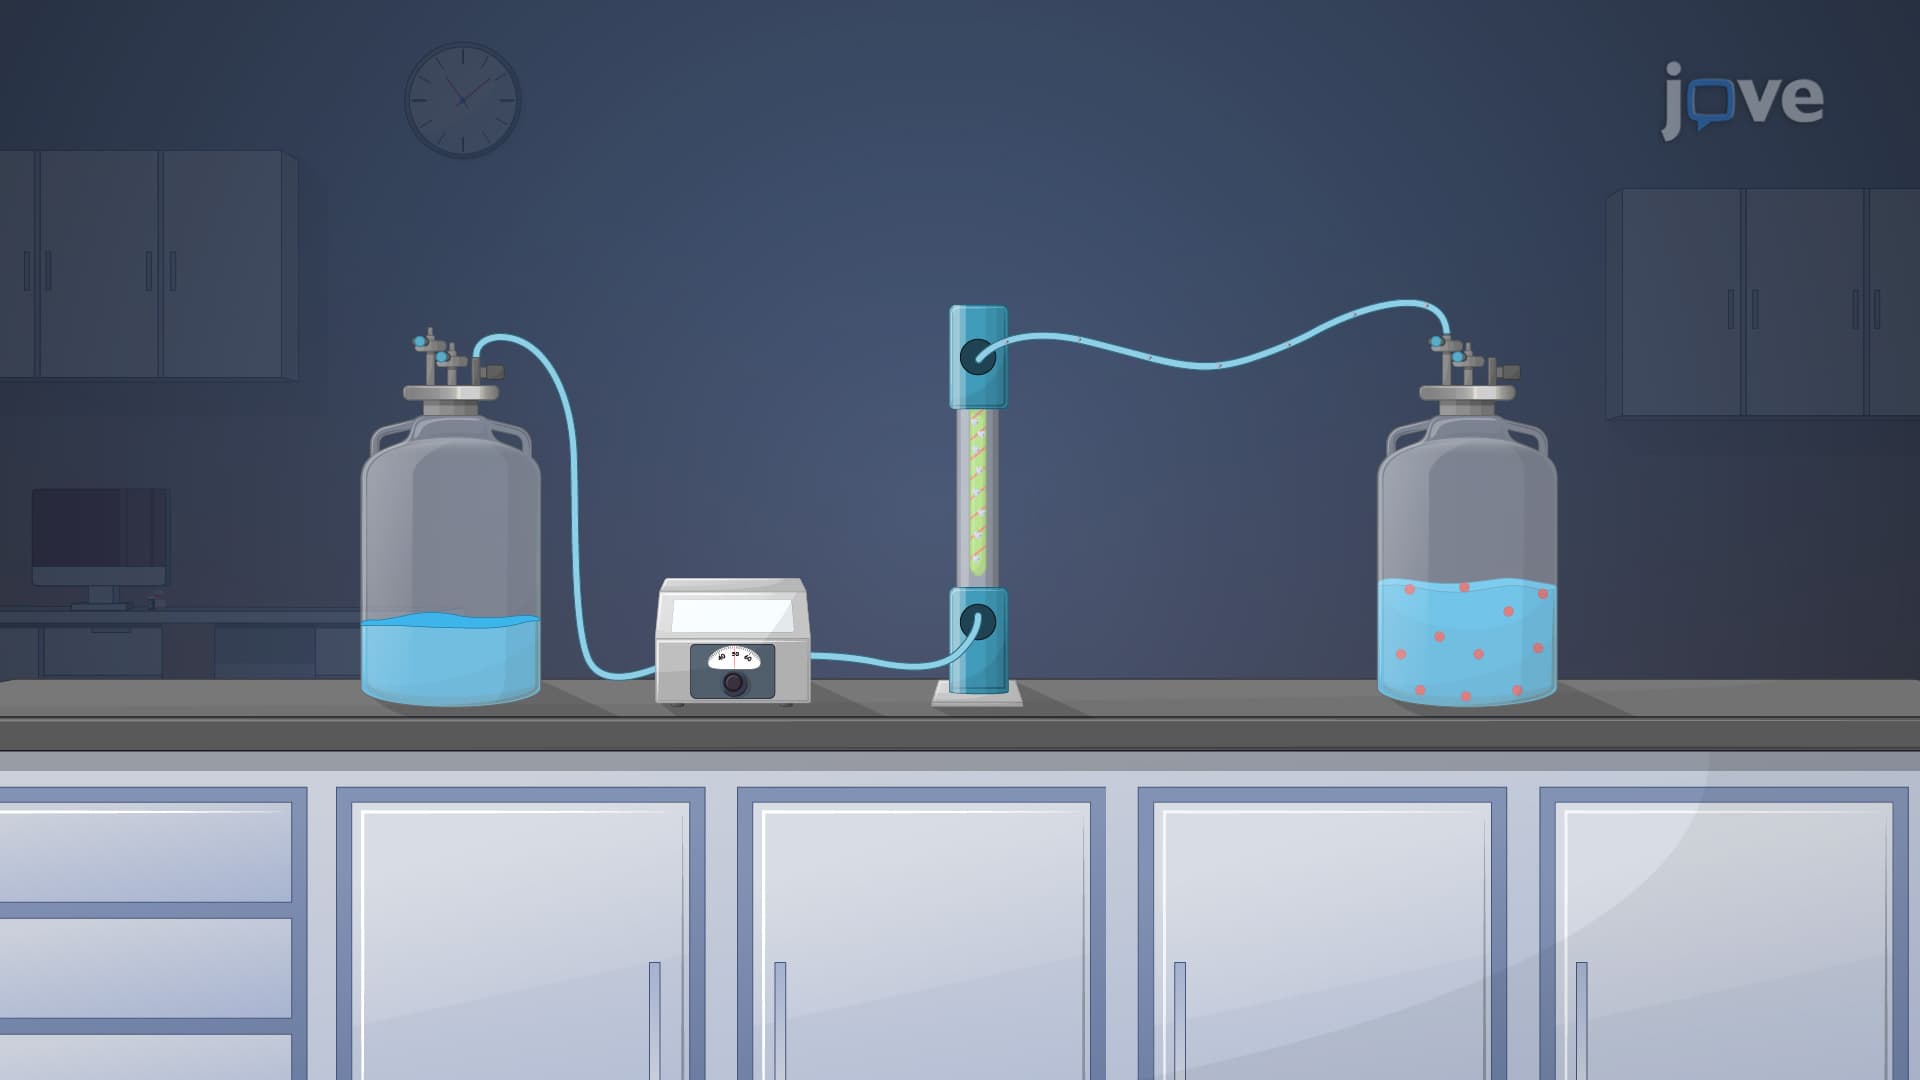
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...