Related Experiment Video
Updated: Aug 26, 2026
Assay to Measure Nucleocytoplasmic Transport in Real Time within Motor Neuron-like NSC-34 Cells
Published on: May 16, 2017
The intracellular location and function of proteins of neuronal ceroid lipofuscinoses
1Department of Biochemistry, Juntendo University School of Medicine, 2-1-1 Hongo, Bunkyo-ku, Tokyo 11 3-8421, Japan.

Abstract:
Neuronal ceroid lipofuscinoses are a group of diseases characterized by accumulation of hydrophobic proteins in lysosomes of neurons and other types of cells. NCLs are caused by at least 8 mutant genes (CLN1-CLN8), though CLN4 and CLN7 have not yet been identified. Except for Cln1p, the protein encoded by CLN1, the defective proteins are associated with lysosomal accumulation of mitochondrial ATP synthase subunit c. Cln1p and Cln2p are soluble lysosomal enzymes, targeted to lysosomes in a mannose 6-phosphate dependent manner. Mutations in the lysosomal protease cathepsin D cause another NCL. Cln3p, Cln5p, Cln6p and Cln8p are thought to be transmembrane proteins. Cln3p and Cln5p are localized in the endosome-lysosomal compartment. Deficiency of endosomal membrane protein CLC-3, a member of the chloride channel family, causes NCL-like phenotype and lysosomal storage of subunit c. Herein, we review the features of NCL and NCL-related proteins and discuss the involvement of the proteins in lysosomal degradation of subunit c.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 01:45
ER Retrieval Pathway
The ER uses many checkpoints to prevent the entry of incorrectly folded or a resident protein as cargo onto a transport vesicle. These mechanisms...
 01:45
Regulation of Nuclear Protein Sorting
 01:28
The Early Endosome: Endocytosis of Transferrin
 01:43
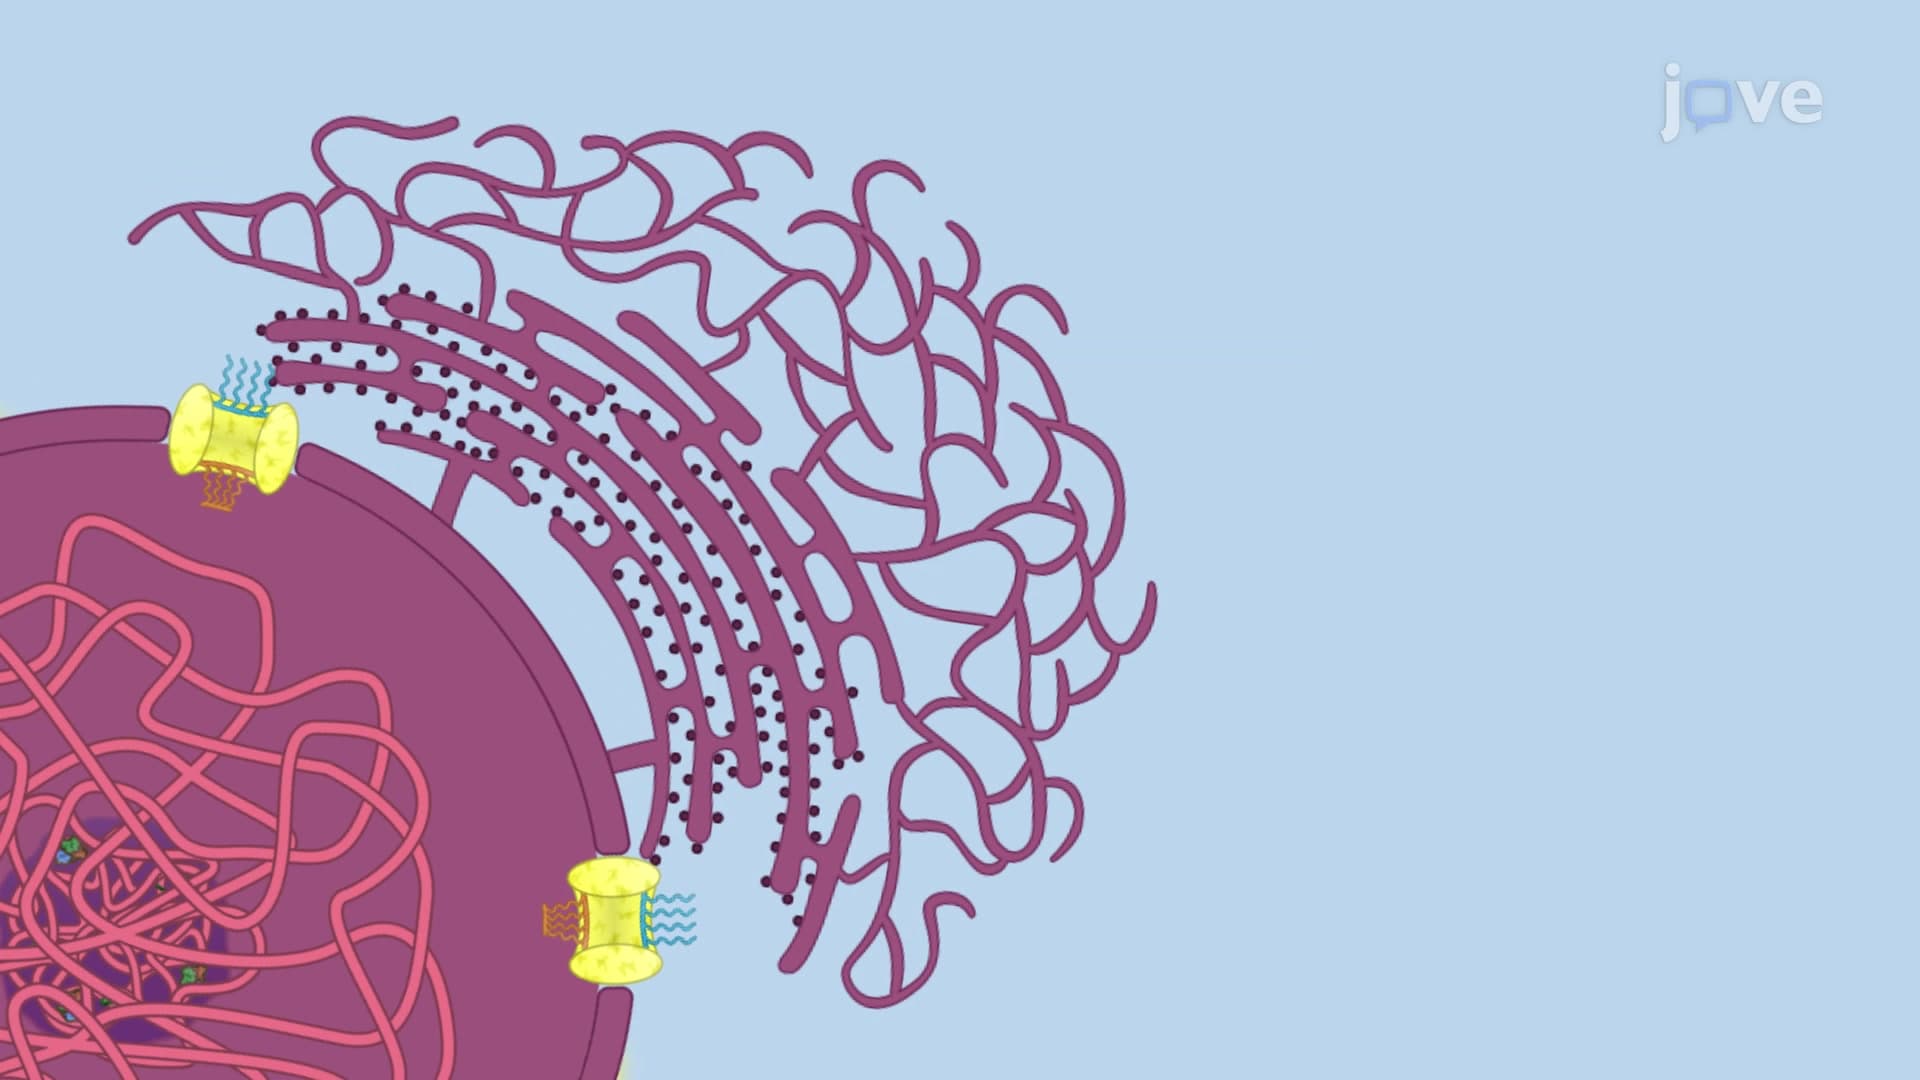
The Endoplasmic Reticulum
 01:49
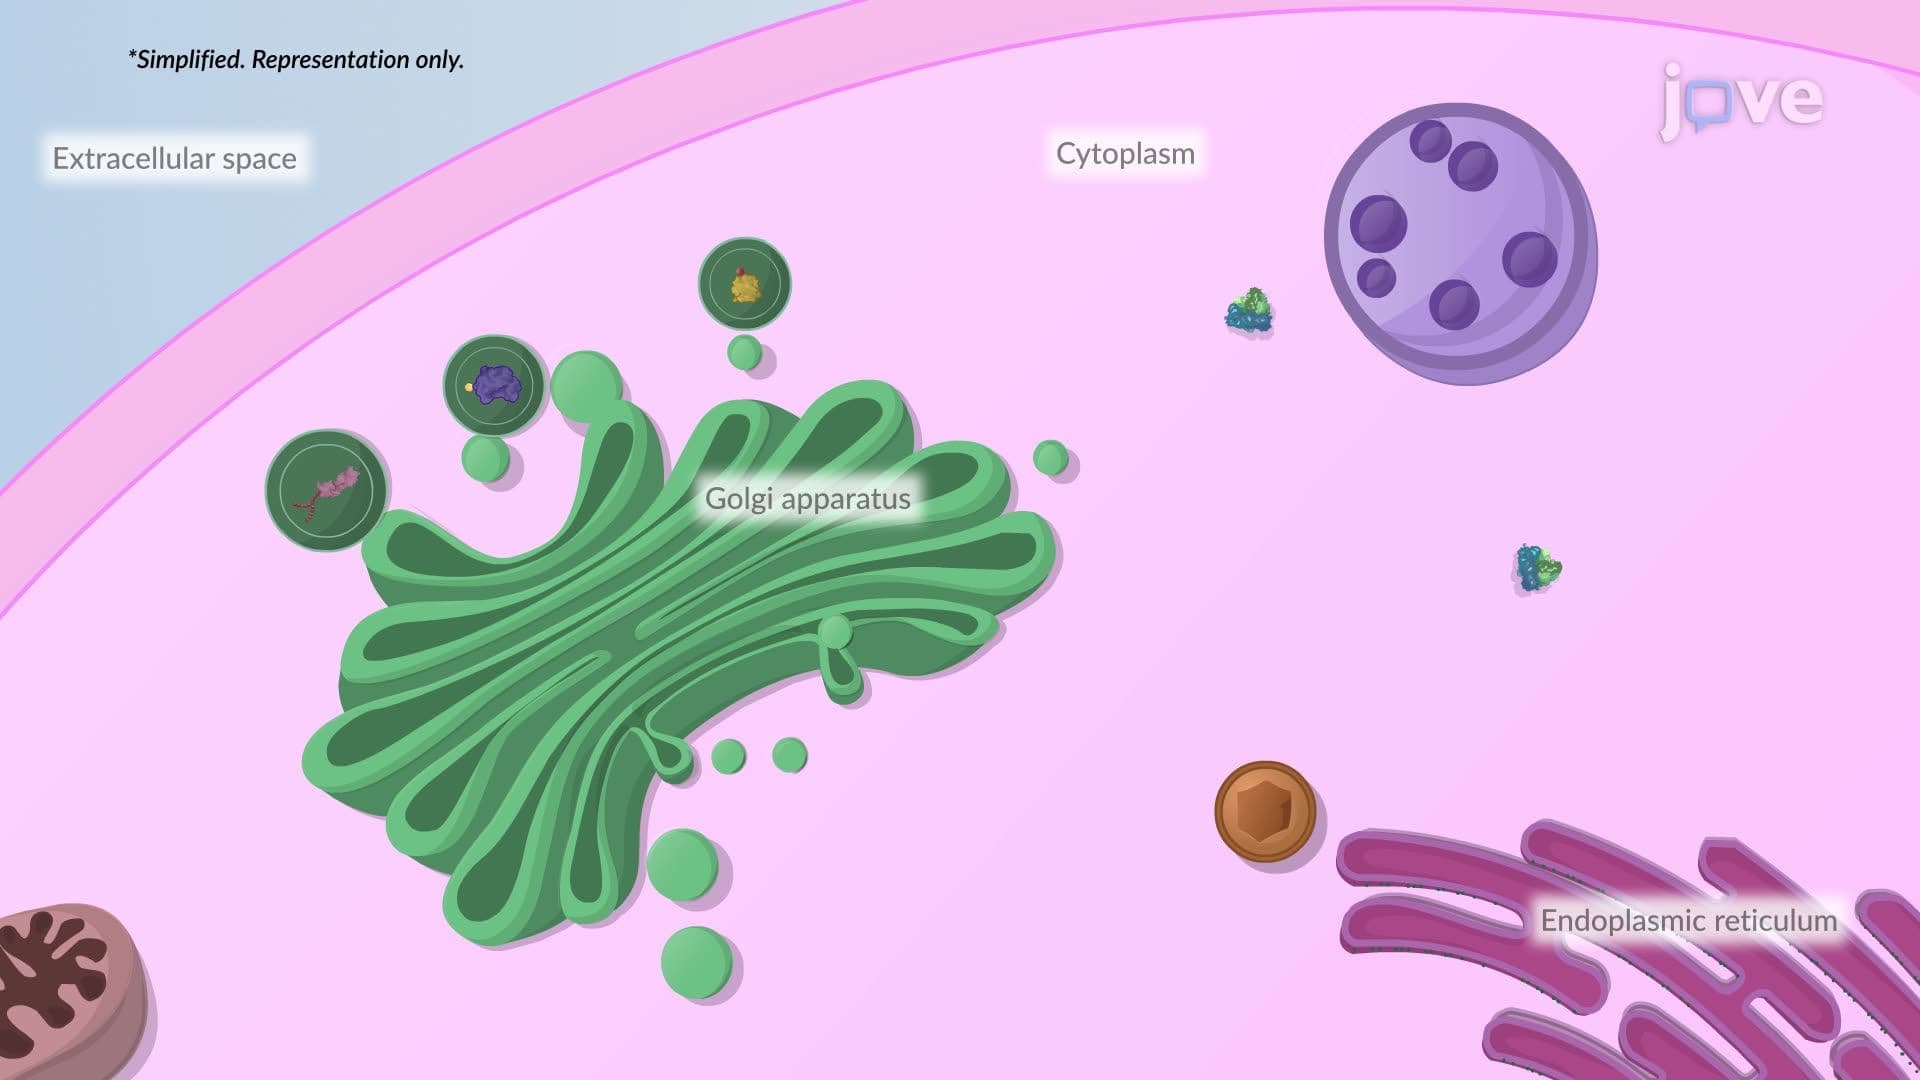
Golgi Apparatus