

Mitochondrial disease: mutations and mechanisms
Matthew McKenzie1, Danae Liolitsa, Michael G Hanna
1Department of Physiology, University College London, Gower Street, London, United Kingdom WC1E 6BT. m.mckenzie@ucl.ac.uk

Abstract:
The mitochondrial diseases encompass a diverse group of disorders that can exhibit various combinations of clinical features. Defects in mitochondrial DNA (mtDNA) have been associated with these diseases, and studies have been able to assign biochemical defects. Deficiencies in mitochondrial oxidative phosphorylation appear to be the main pathogenic factors, although recent studies suggest that other mechanisms are involved. Reactive oxygen species (ROS) generation has been implicated in a wide variety of neurodegenerative diseases, and mitochondrial ROS generation may be an important factor in mitochondrial disease pathogenesis. Altered apoptotic signaling as a consequence of defective mitochondrial function has also been observed in both in vitro and in vivo disease models. Our current understanding of the contribution of these various mechanisms to mitochondrial disease pathophysiology will be discussed.
Insights
Mitochondrial diseases stem from mitochondrial DNA (mtDNA) defects, often involving oxidative phosphorylation deficiencies. Emerging research highlights the roles of reactive oxygen species (ROS) and altered apoptosis in disease development.
Area of Science:
- Biochemistry
- Genetics
- Cell Biology
Background:
- Mitochondrial diseases are a complex group of disorders with varied clinical presentations.
- Defects in mitochondrial DNA (mtDNA) are linked to these diseases, with biochemical defects often identifiable.
- While deficiencies in mitochondrial oxidative phosphorylation are primary pathogenic factors, other mechanisms are increasingly recognized.
Purpose of the Study:
- To discuss the current understanding of mitochondrial disease pathophysiology.
- To explore the roles of various contributing mechanisms beyond oxidative phosphorylation defects.
Main Methods:
- Review of existing literature on mitochondrial diseases.
- Analysis of studies investigating biochemical defects and genetic associations.
- Examination of research on reactive oxygen species (ROS) and apoptosis in disease models.
Main Results:
- Mitochondrial DNA (mtDNA) defects and oxidative phosphorylation deficiencies are key factors.
- Mitochondrial reactive oxygen species (ROS) generation is implicated in pathogenesis.
- Altered apoptotic signaling is observed in mitochondrial dysfunction.
Conclusions:
- Mitochondrial diseases involve complex pathophysiology with multiple contributing factors.
- Reactive oxygen species (ROS) and apoptosis play significant roles alongside primary energy metabolism defects.
- Further research is needed to fully elucidate the interplay of these mechanisms.
Related Concept Videos
 01:48
ATP Synthase: Mechanism
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...
 02:59
Animal Mitochondrial Genetics
 01:19
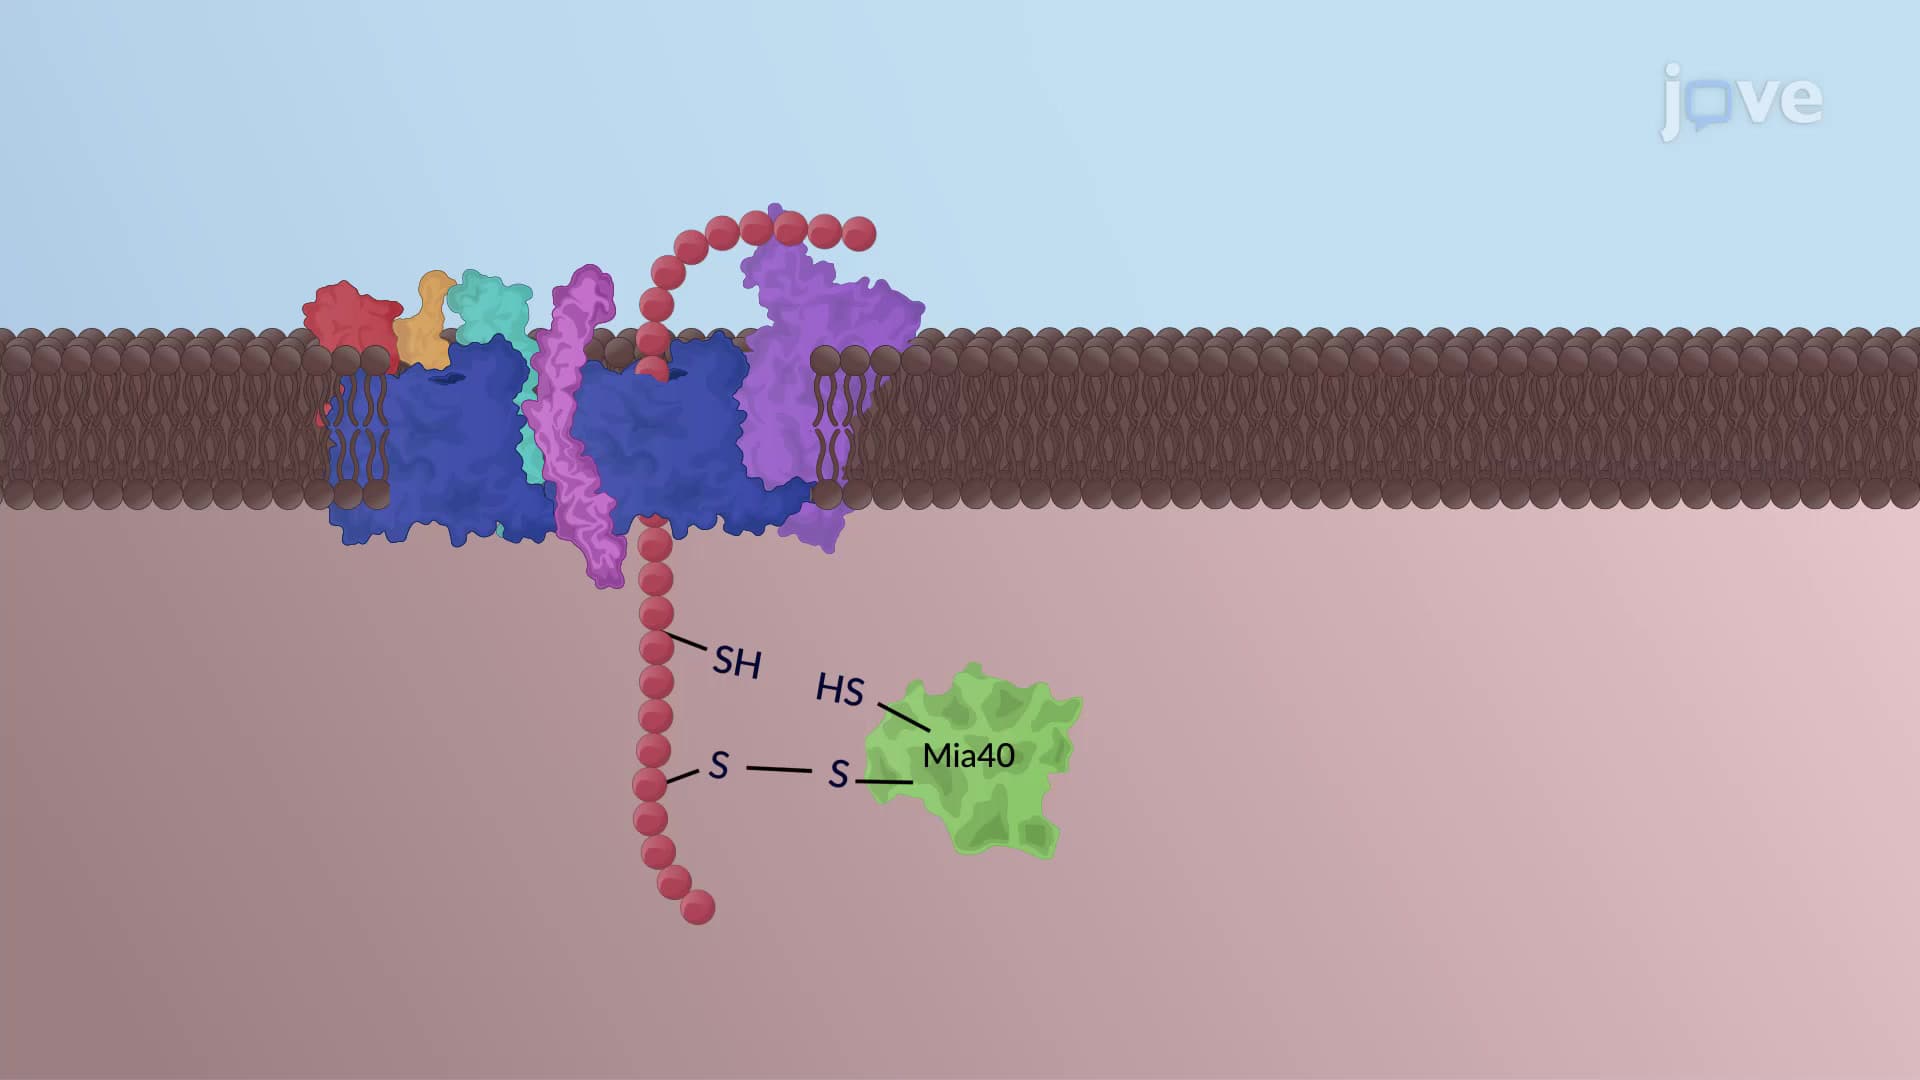
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:39
Mutations
 01:35
Mutations
Chromosomal Alterations Are Large-Scale Mutations
While point mutations are changes in a single nucleotide in...