A hierarchical approach to all-atom protein loop prediction
Related Experiment Video
Updated: Aug 25, 2026
Protein WISDOM: A Workbench for In silico De novo Design of BioMolecules
Published on: July 25, 2013
Matthew P Jacobson1, David L Pincus, Chaya S Rapp
1Department of Pharmaceutical Chemistry, University of California, San Francisco 94143-2240, USA. matt@cgl.ucsf.edu

Abstract:
The application of all-atom force fields (and explicit or implicit solvent models) to protein homology-modeling tasks such as side-chain and loop prediction remains challenging both because of the expense of the individual energy calculations and because of the difficulty of sampling the rugged all-atom energy surface. Here we address this challenge for the problem of loop prediction through the development of numerous new algorithms, with an emphasis on multiscale and hierarchical techniques. As a first step in evaluating the performance of our loop prediction algorithm, we have applied it to the problem of reconstructing loops in native structures; we also explicitly include crystal packing to provide a fair comparison with crystal structures. In brief, large numbers of loops are generated by using a dihedral angle-based buildup procedure followed by iterative cycles of clustering, side-chain optimization, and complete energy minimization of selected loop structures. We evaluate this method by using the largest test set yet used for validation of a loop prediction method, with a total of 833 loops ranging from 4 to 12 residues in length. Average/median backbone root-mean-square deviations (RMSDs) to the native structures (superimposing the body of the protein, not the loop itself) are 0.42/0.24 A for 5 residue loops, 1.00/0.44 A for 8 residue loops, and 2.47/1.83 A for 11 residue loops. Median RMSDs are substantially lower than the averages because of a small number of outliers; the causes of these failures are examined in some detail, and many can be attributed to errors in assignment of protonation states of titratable residues, omission of ligands from the simulation, and, in a few cases, probable errors in the experimentally determined structures. When these obvious problems in the data sets are filtered out, average RMSDs to the native structures improve to 0.43 A for 5 residue loops, 0.84 A for 8 residue loops, and 1.63 A for 11 residue loops. In the vast majority of cases, the method locates energy minima that are lower than or equal to that of the minimized native loop, thus indicating that sampling rarely limits prediction accuracy. The overall results are, to our knowledge, the best reported to date, and we attribute this success to the combination of an accurate all-atom energy function, efficient methods for loop buildup and side-chain optimization, and, especially for the longer loops, the hierarchical refinement protocol.
More Related Videos
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
 06:50
Computational Prediction of Amino Acid Preferences of Potentially Multispecific Peptide-Binding Domains Involved in Protein-Protein Interactions
Published on: January 26, 2024
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:13
Protein Organization
 01:24
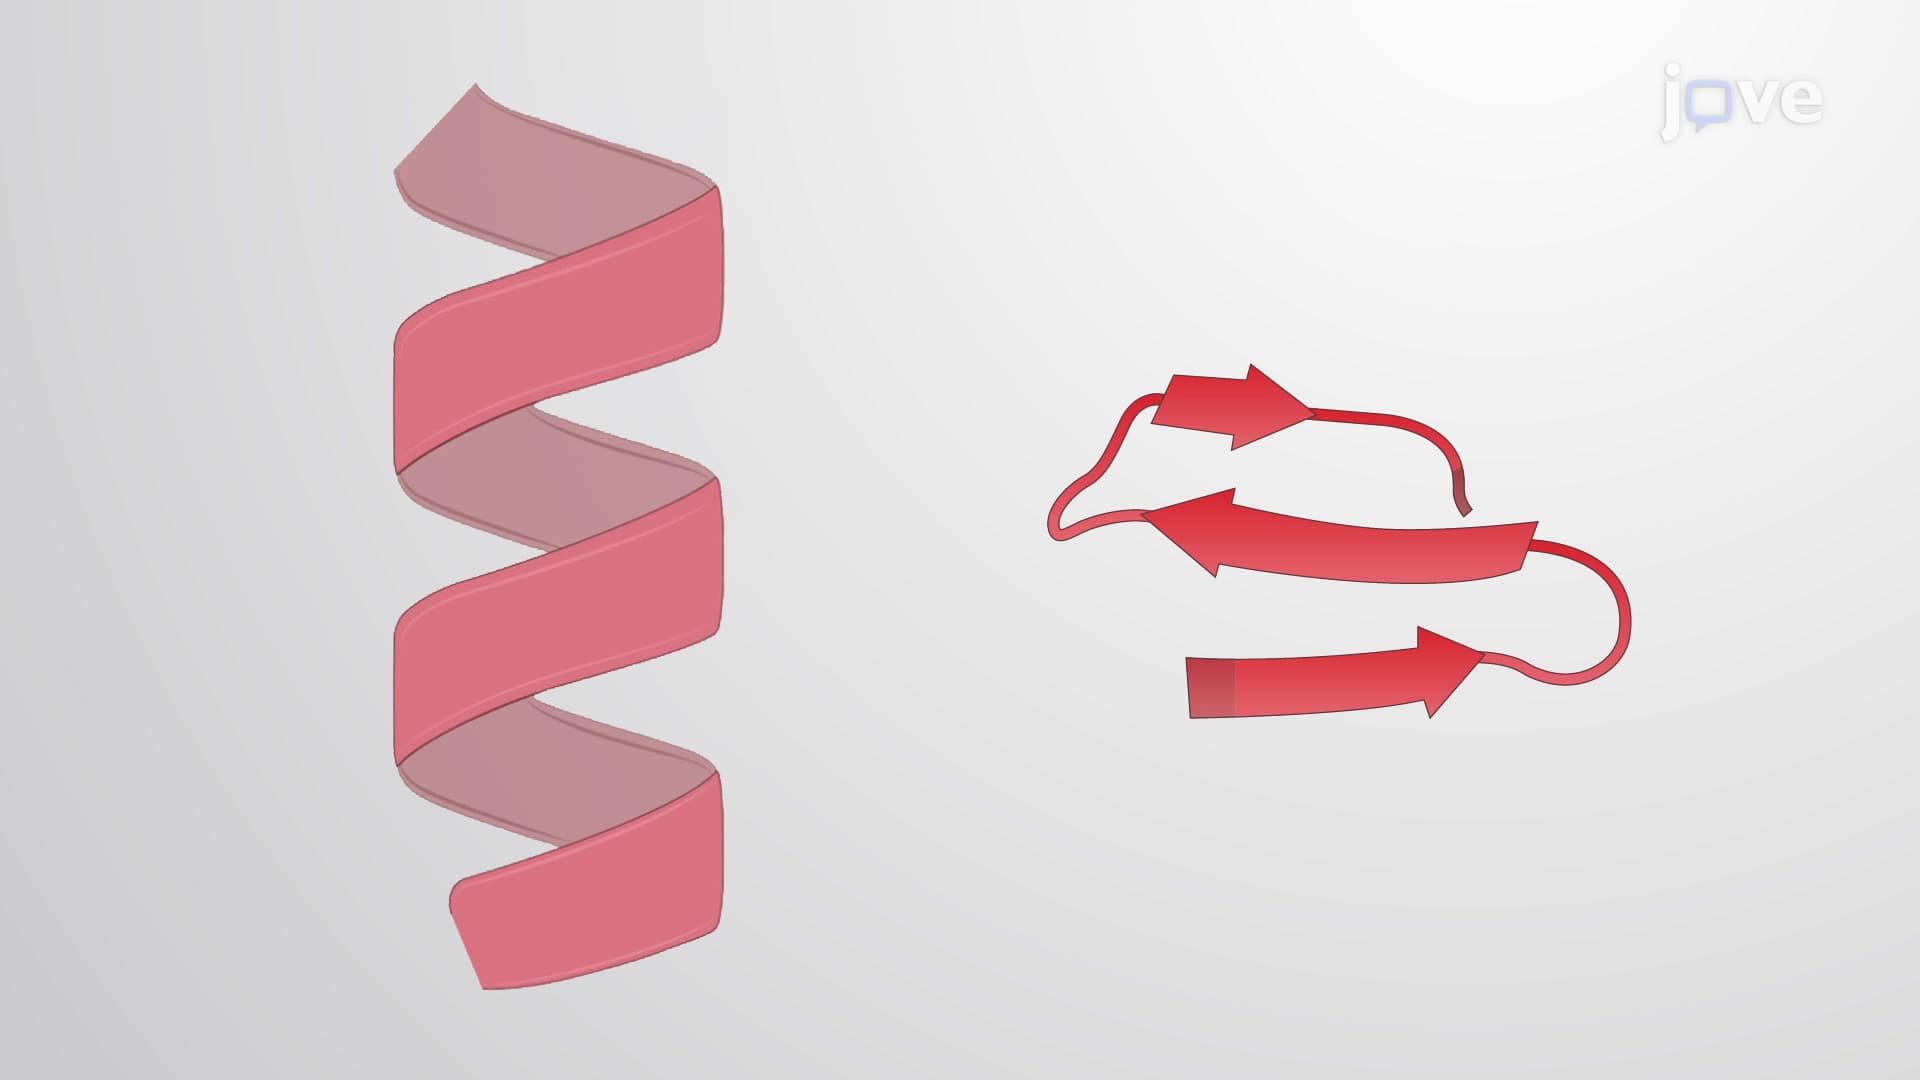
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
01:22Protein Folding