Related Experiment Video
Updated: Aug 24, 2026
Determining the Serum Stability of Human Adenosine Deaminase 1 Enzyme
Published on: September 27, 2024
Exploring distal regions of the A3 adenosine receptor binding site: sterically constrained
Susanna Tchilibon1, Soo-Kyung Kim, Zhan-Guo Gao
1Molecular Recognition Section, Laboratory of Bioorganic Chemistry, National Institute of Diabetes & Digestive & Kidney Diseases, National Institutes of Health, Department of Health, and Human Services, Bethesda, MD 20892-0810, USA.

Abstract:
We synthesized phenyl ring-substituted analogues of N(6)-(1S,2R)-(2-phenyl-1-cyclopropyl)adenosine, which is highly potent in binding to the human A(3)AR with a Ki value of 0.63 nM. The effects of these structural changes on affinity at human and rat adenosine receptors and on intrinsic efficacy at the hA(3)AR were measured. A 3-nitrophenyl analogue was resolved chromatographically into pure diastereomers, which displayed 10-fold stereoselectivity in A(3)AR binding in favor of the 1S,2R isomer. A molecular model defined a hydrophobic region (Phe168) in the putative A(3)AR binding site around the phenyl moiety. A heteroaromatic group (3-thienyl) could substitute for the phenyl moiety with retention of high affinity of A(3)AR binding. Other related N(6)-substituted adenosine derivatives were included for comparison. Although the N(6)-(2-phenyl-1-cyclopropyl) derivatives were full A(3)AR agonists, several other derivatives had greatly reduced efficacy. N(6)-Cyclopropyladenosine was an A(3)AR antagonist, and adding either one or two phenyl rings at the 2-position of the cyclopropyl moiety restored efficacy. N(6)-(2,2-Diphenylethyl)adenosine was an A(3)AR antagonist, and either adding a bond between the two phenyl rings (N(6)-9-fluorenylmethyl) or shortening the ethyl moiety (N(6)-diphenylmethyl) restored efficacy. A QSAR study of the N(6) region provided a model that was complementary to the putative A(3)AR binding site in a rhodopsin-based homology model. Thus, a new series of high-affinity A(3)AR agonists and related nucleoside antagonists was explored through both empirical and theoretical approaches.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:17
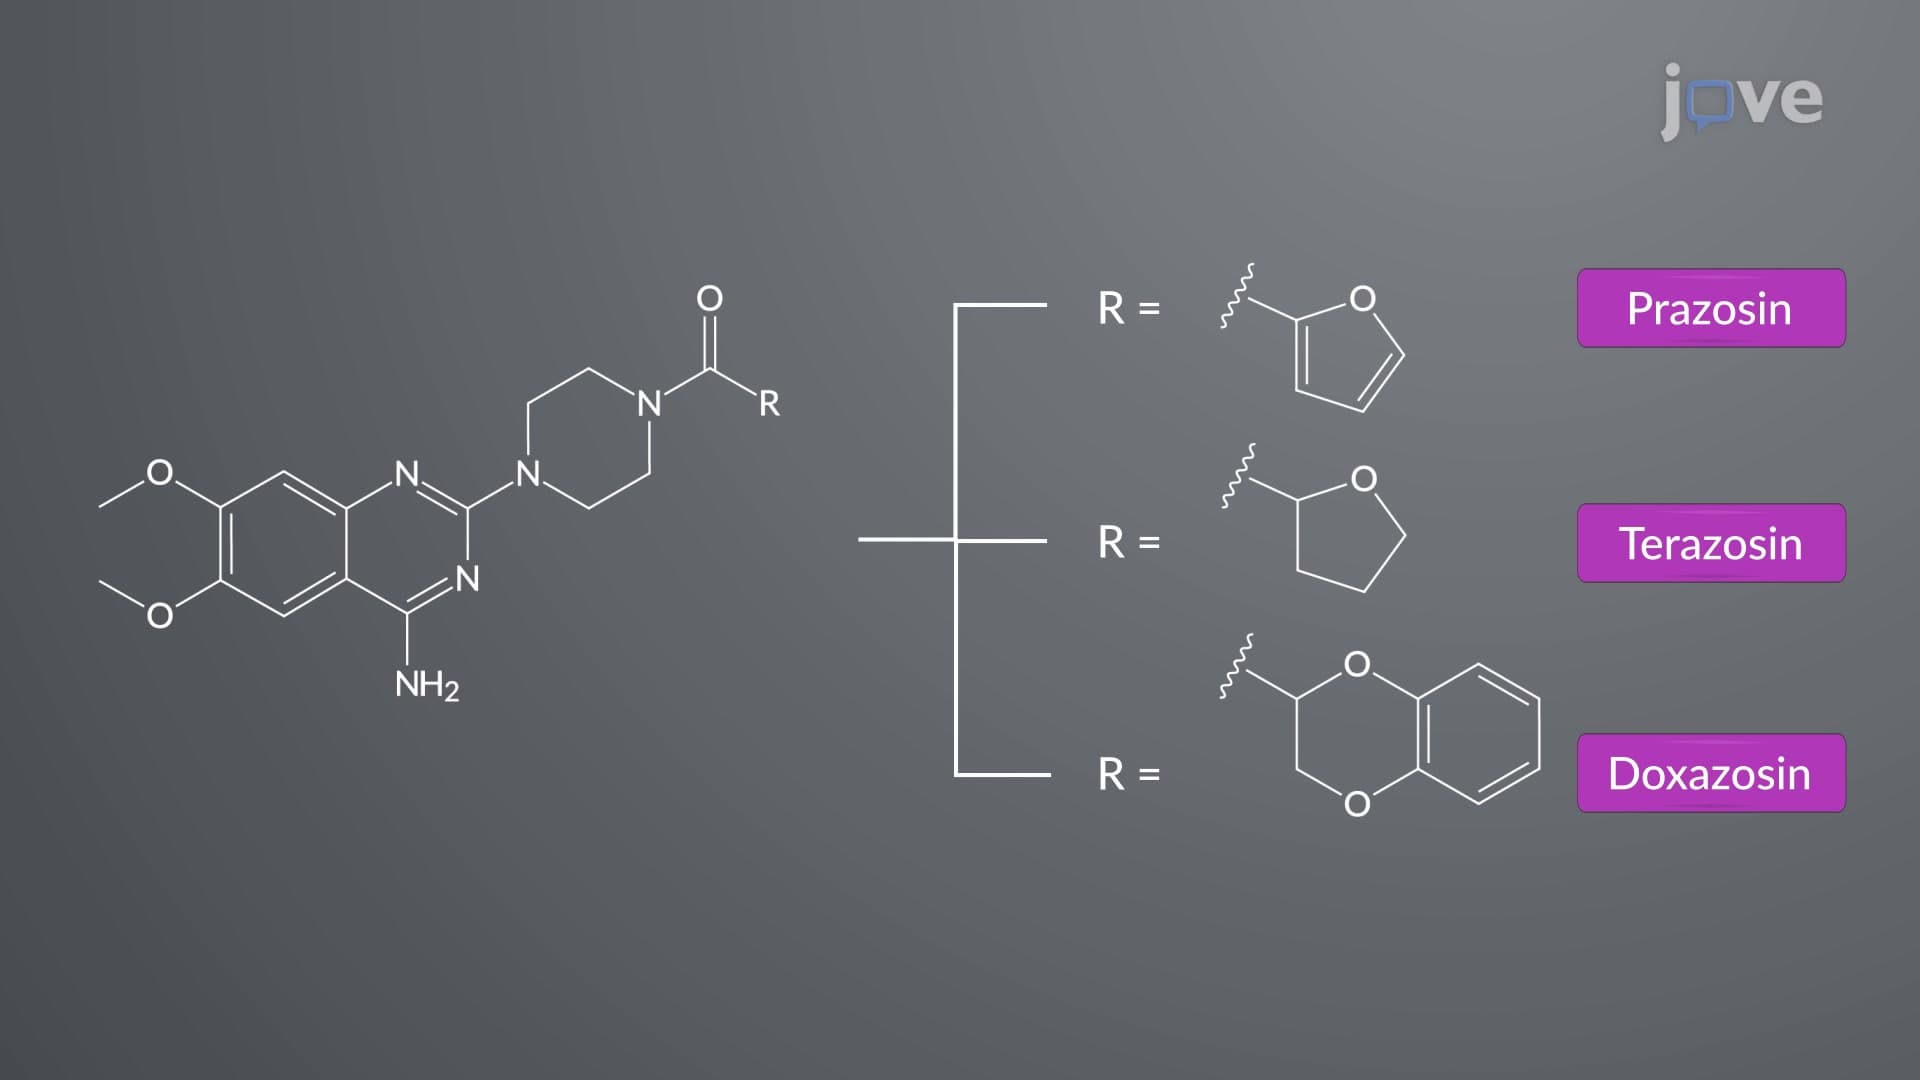
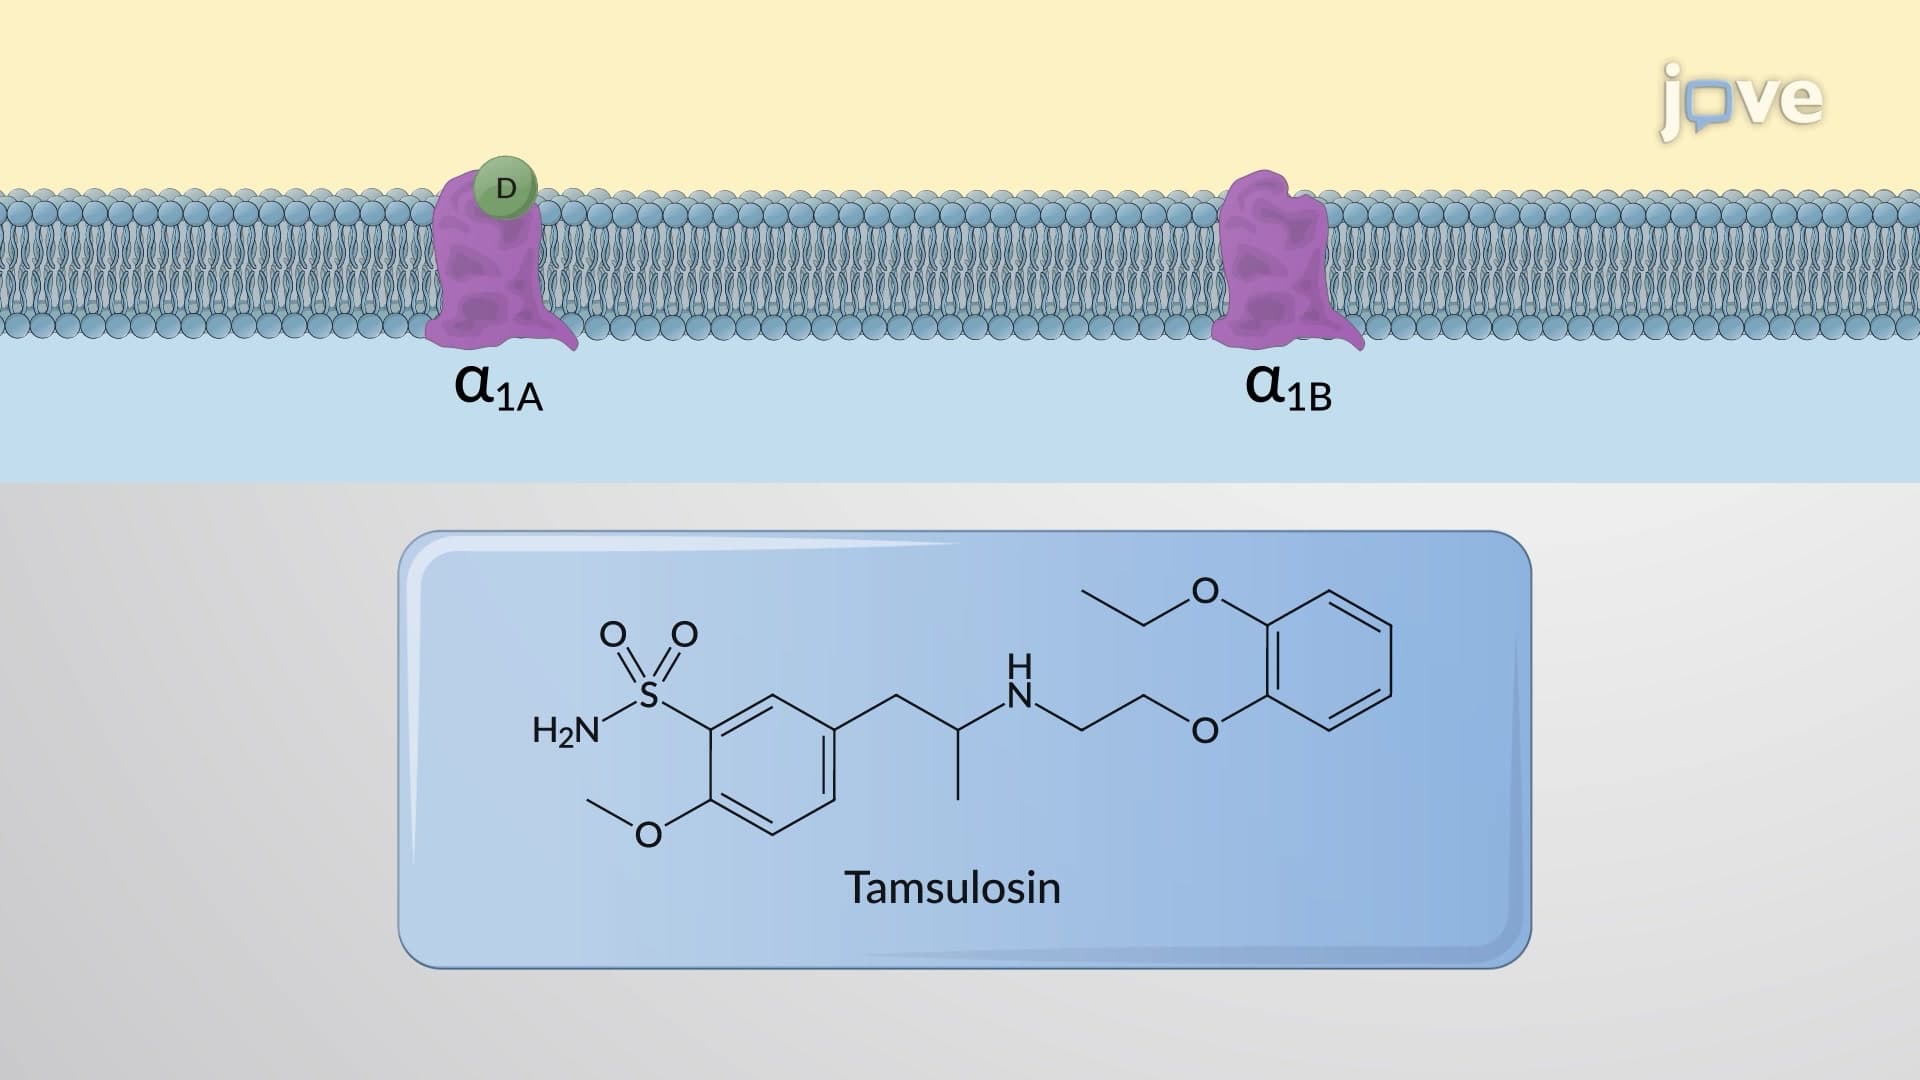
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:26
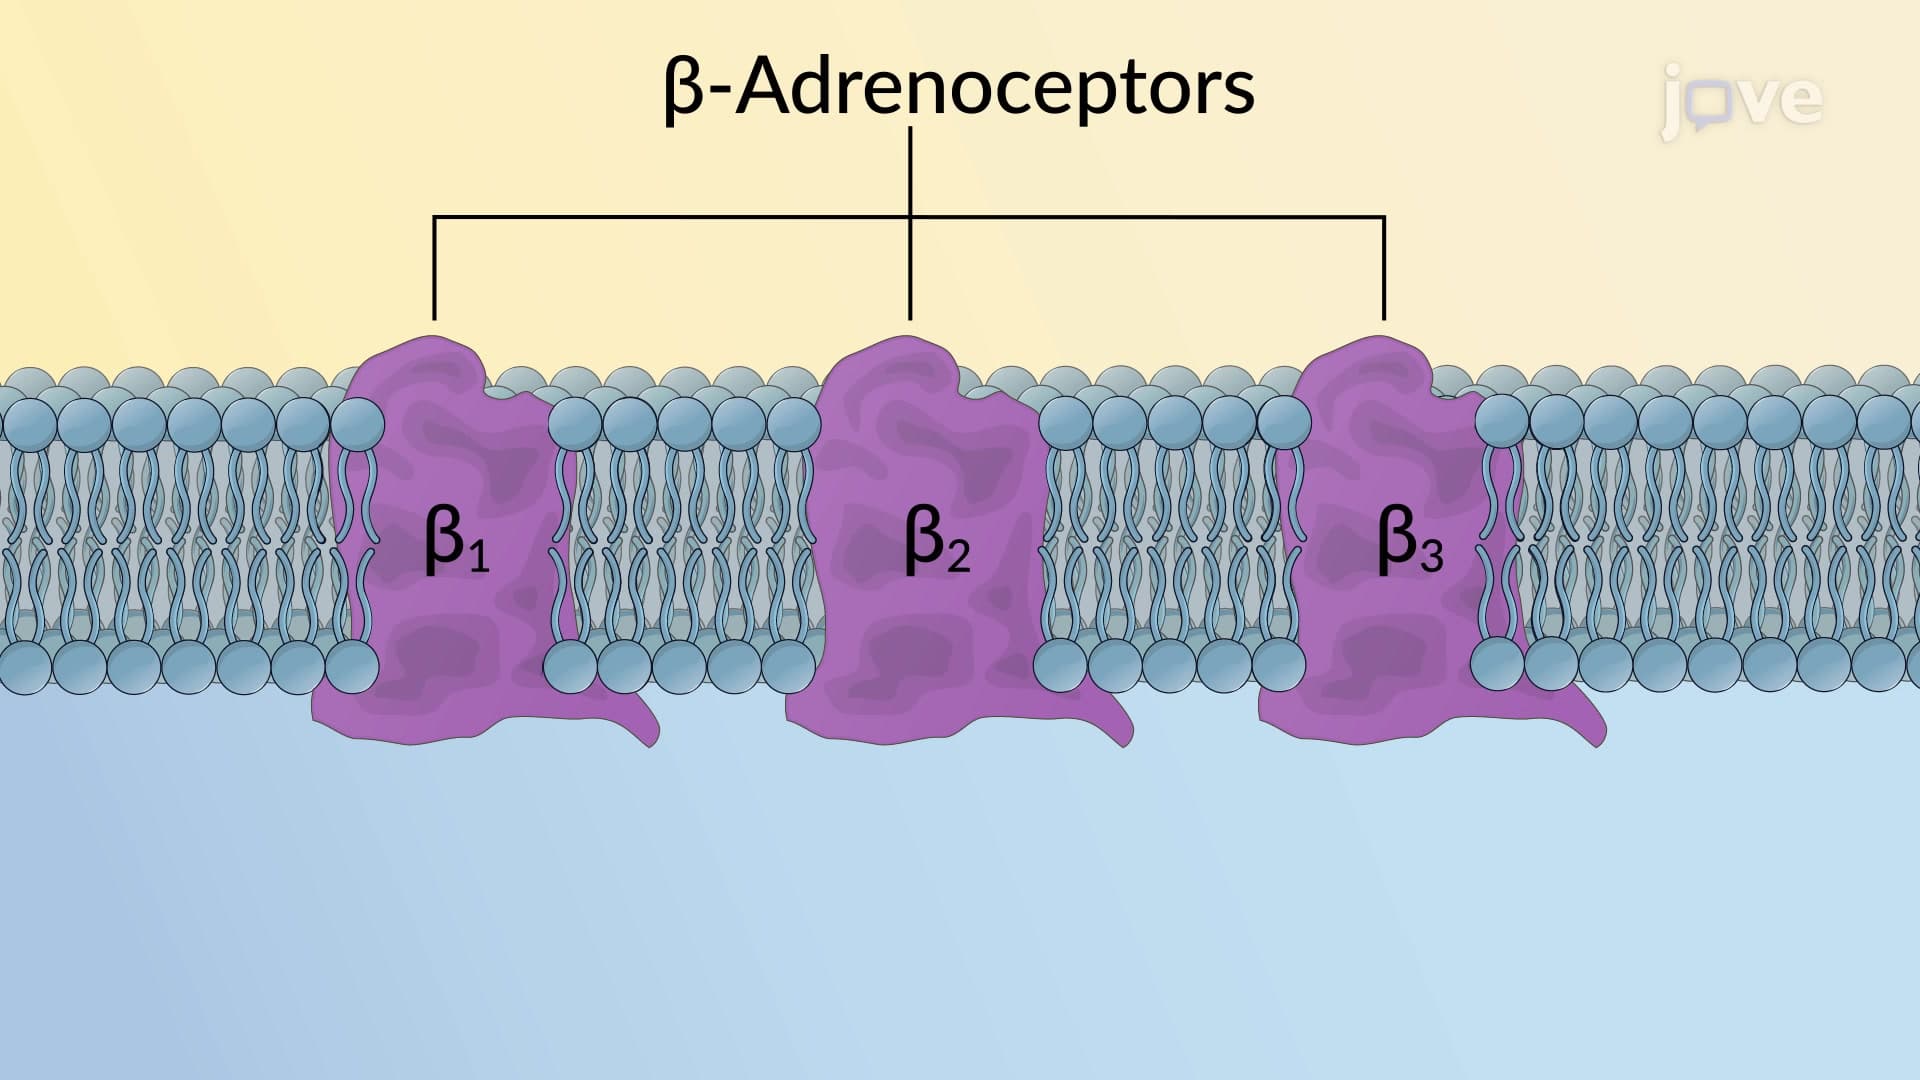
Adrenergic Receptors: β Subtype
Isoprenaline > Adrenaline > Noradrenaline
Neurotransmitter binding to these receptors causes activation of adenylyl cyclase resulting in increased concentrations of cAMP and modulation of calcium ion channels within the cell. They are further classified into β1, β2, and β3 subtypes.
β1-adrenoceptors: β1-adrenoceptors have equal affinities for...
 01:30
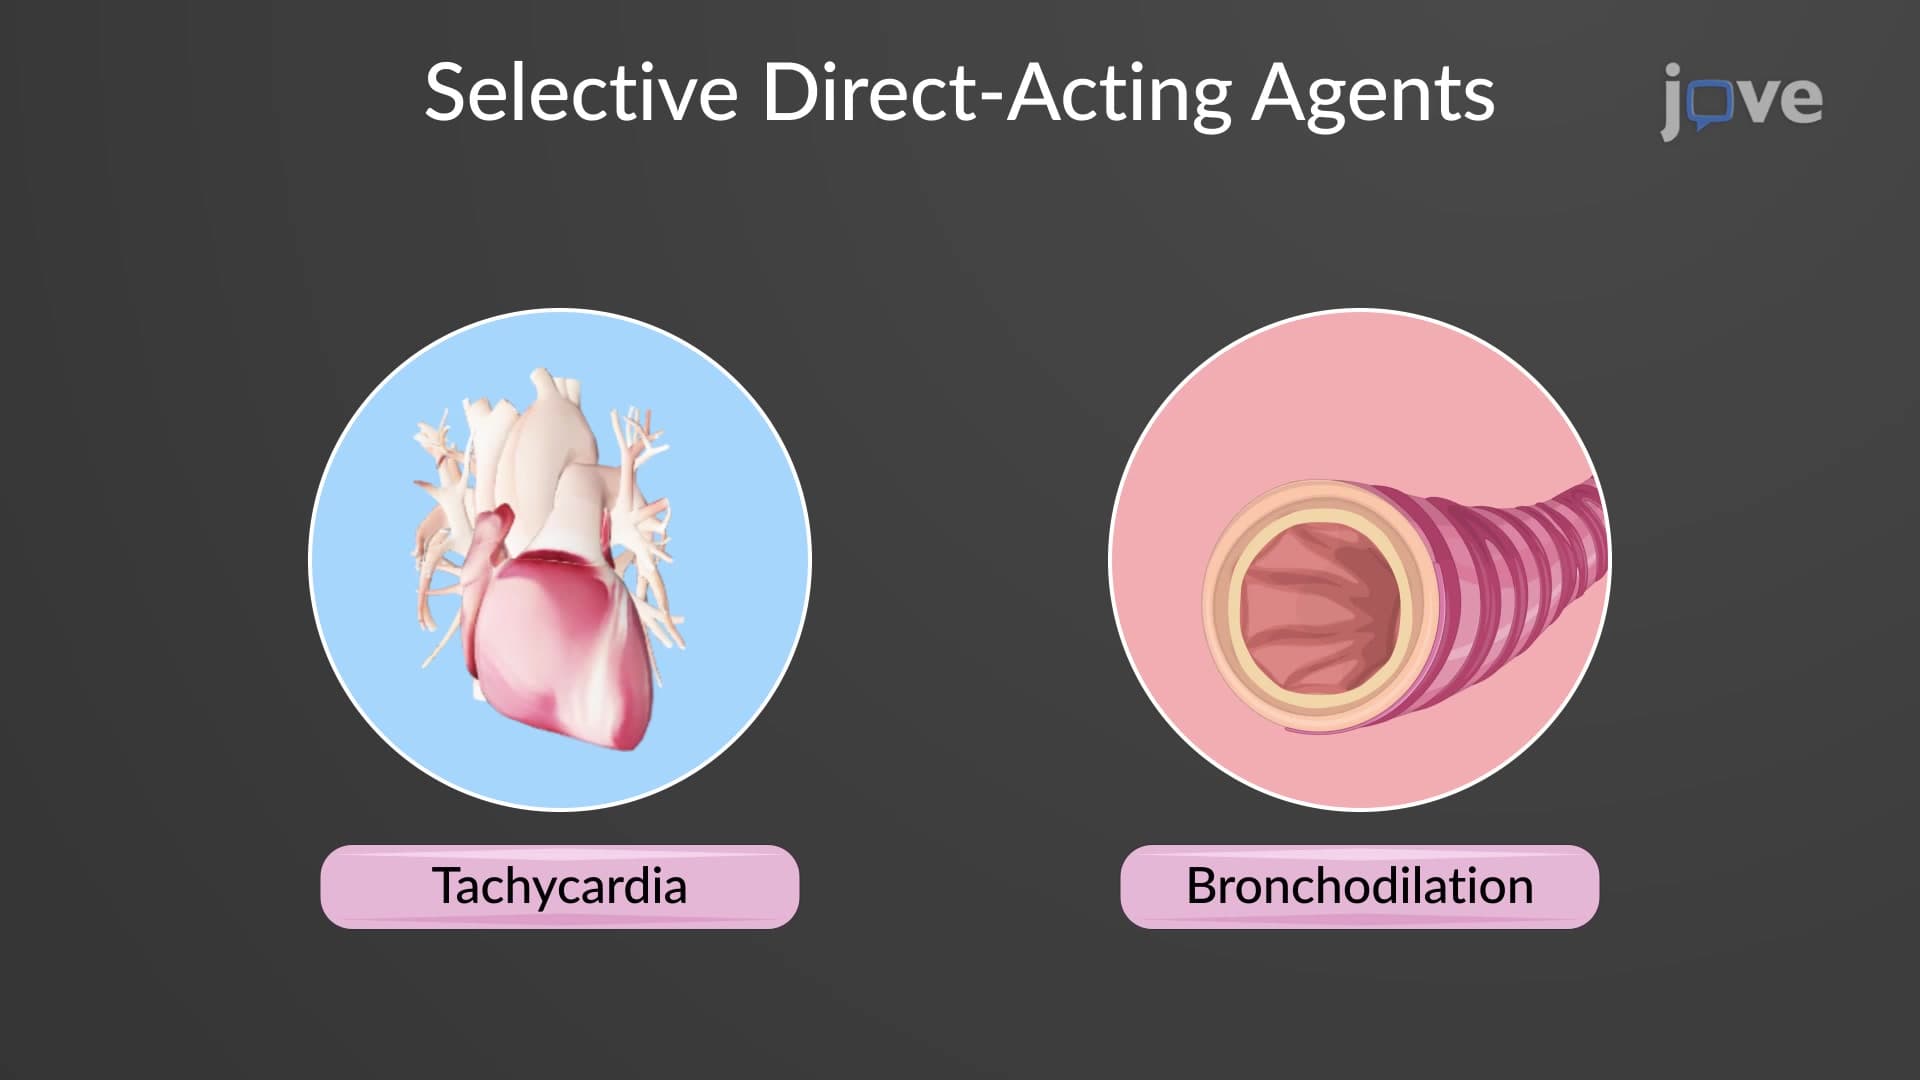
Adrenergic Agonists: Direct-Acting Agents
These agents can be classified...
 01:22
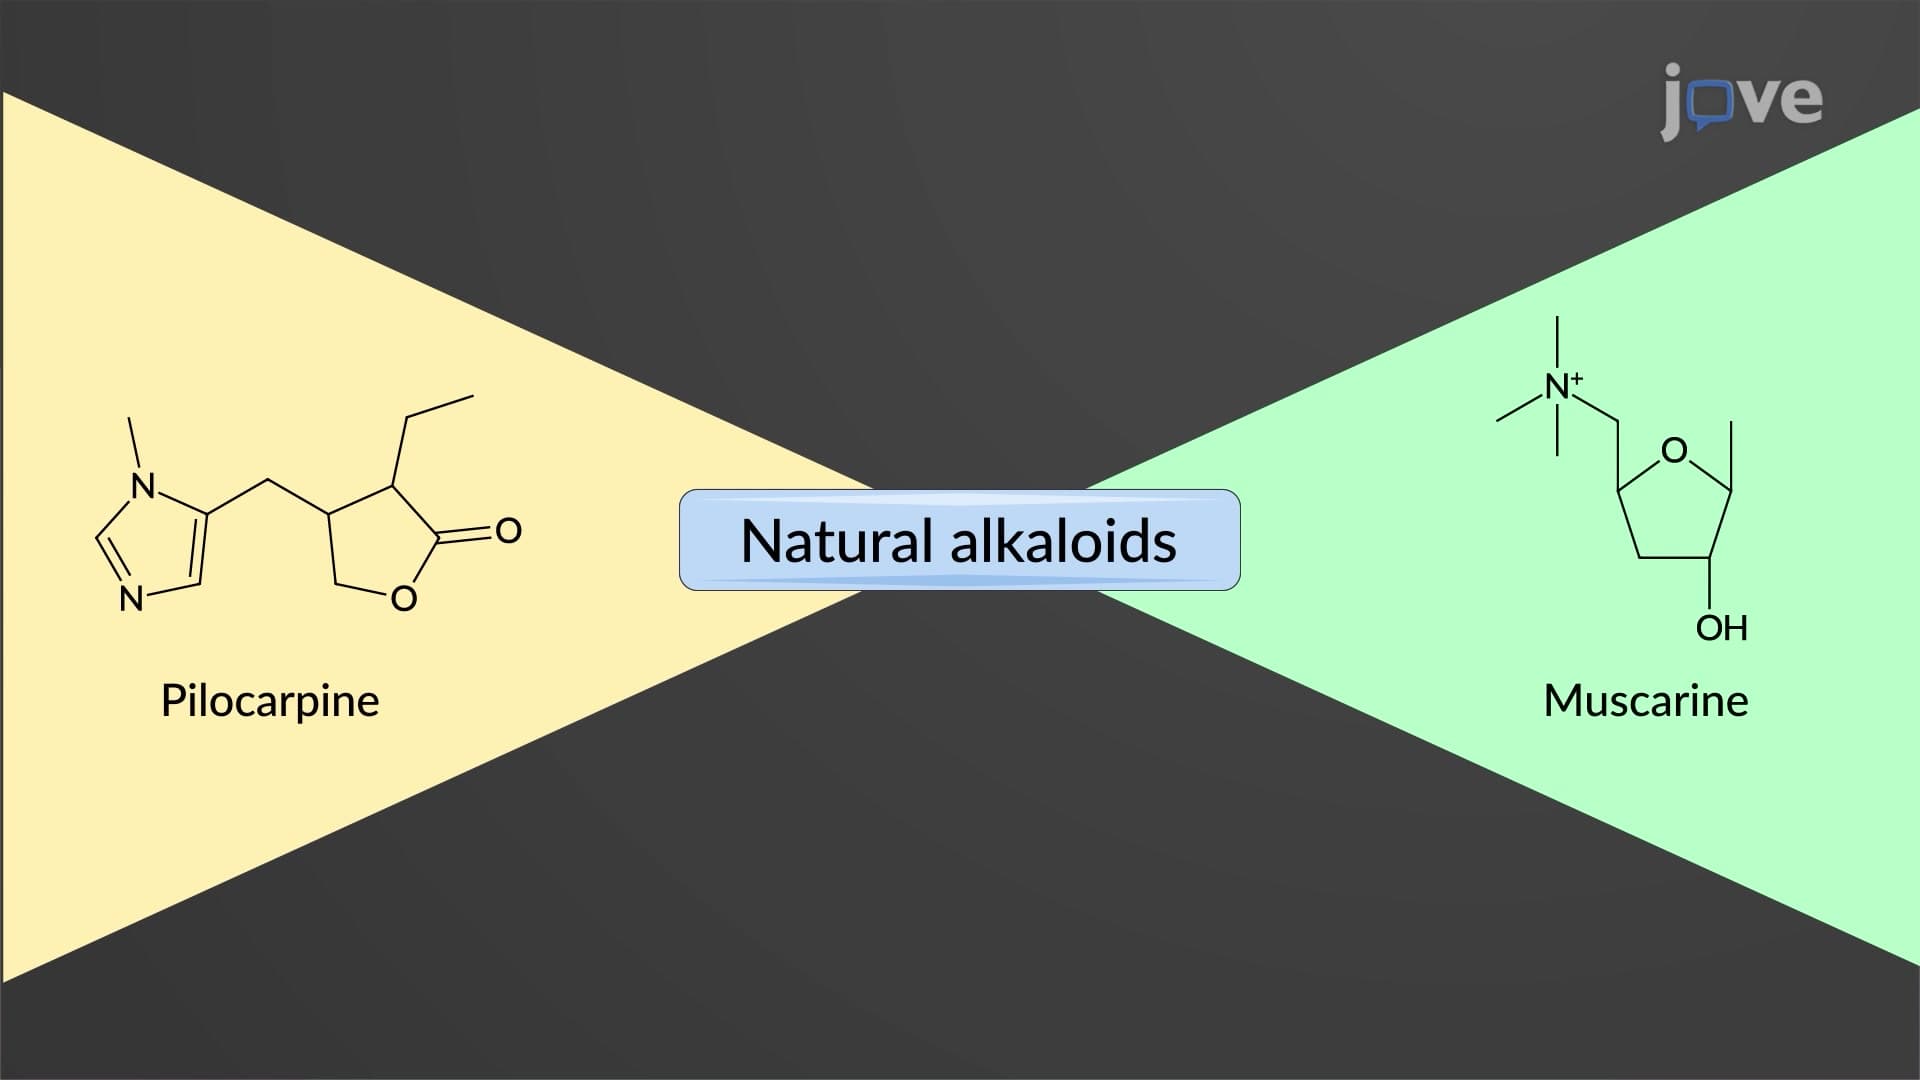
Direct-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
The direct-acting...