Related Experiment Video
Updated: Aug 24, 2026

Determining the Serum Stability of Human Adenosine Deaminase 1 Enzyme
Published on: September 27, 2024
Structural determinants of efficacy at A3 adenosine receptors: modification of the ribose moiety
Zhan-Guo Gao1, Lak Shin Jeong, Hyung Ryong Moon
1Laboratory of Bioorganic Chemistry, Molecular Recognition Section, National Institute of Diabetes and Digestive and Kidney Diseases, National Institutes of Health, Bethesda, MD 20892, USA.

Abstract:
We have found previously that structural features of adenosine derivatives, particularly at the N6- and 2-positions of adenine, determine the intrinsic efficacy as A3 adenosine receptor (AR) agonists. Here, we have probed this phenomenon with respect to the ribose moiety using a series of ribose-modified adenosine derivatives, examining binding affinity and activation of the human A3 AR expressed in CHO cells. Both 2'- and 3'-hydroxyl groups in the ribose moiety contribute to A3 AR binding and activation, with 2'-OH being more essential. Thus, the 2'-fluoro substitution eliminated both binding and activation, while a 3'-fluoro substitution led to only a partial reduction of potency and efficacy at the A3 AR. A 5'-uronamide group, known to restore full efficacy in other derivatives, failed to fully overcome the diminished efficacy of 3'-fluoro derivatives. The 4'-thio substitution, which generally enhanced A3 AR potency and selectivity, resulted in 5'-CH2OH analogues (10 and 12) which were partial agonists of the A3 AR. Interestingly, the shifting of the N6-(3-iodobenzyl)adenine moiety from the 1'- to 4'-position had a minor influence on A3 AR selectivity, but transformed 15 into a potent antagonist (16) (Ki = 4.3 nM). Compound 16 antagonized human A3 AR agonist-induced inhibition of cyclic AMP with a K(B) value of 3.0 nM. A novel apio analogue (20) of neplanocin A, was a full A3 AR agonist. The affinities of selected, novel analogues at rat ARs were examined, revealing species differences. In summary, critical structural determinants for human A3 AR activation have been identified, which should prove useful for further understanding the mechanism of receptor activation and development of more potent and selective full agonists, partial agonists and antagonists for A3 ARs.
More Related Videos
 11:29
HPLC-based Assay to Monitor Extracellular Nucleotide/Nucleoside Metabolism in Human Chronic Lymphocytic Leukemia Cells
Published on: July 20, 2016
 10:24
NMR-Based Activity Assays for Determining Compound Inhibition, IC50 Values, Artifactual Activity, and Whole-Cell Activity of Nucleoside Ribohydrolases
Published on: June 30, 2019
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:17
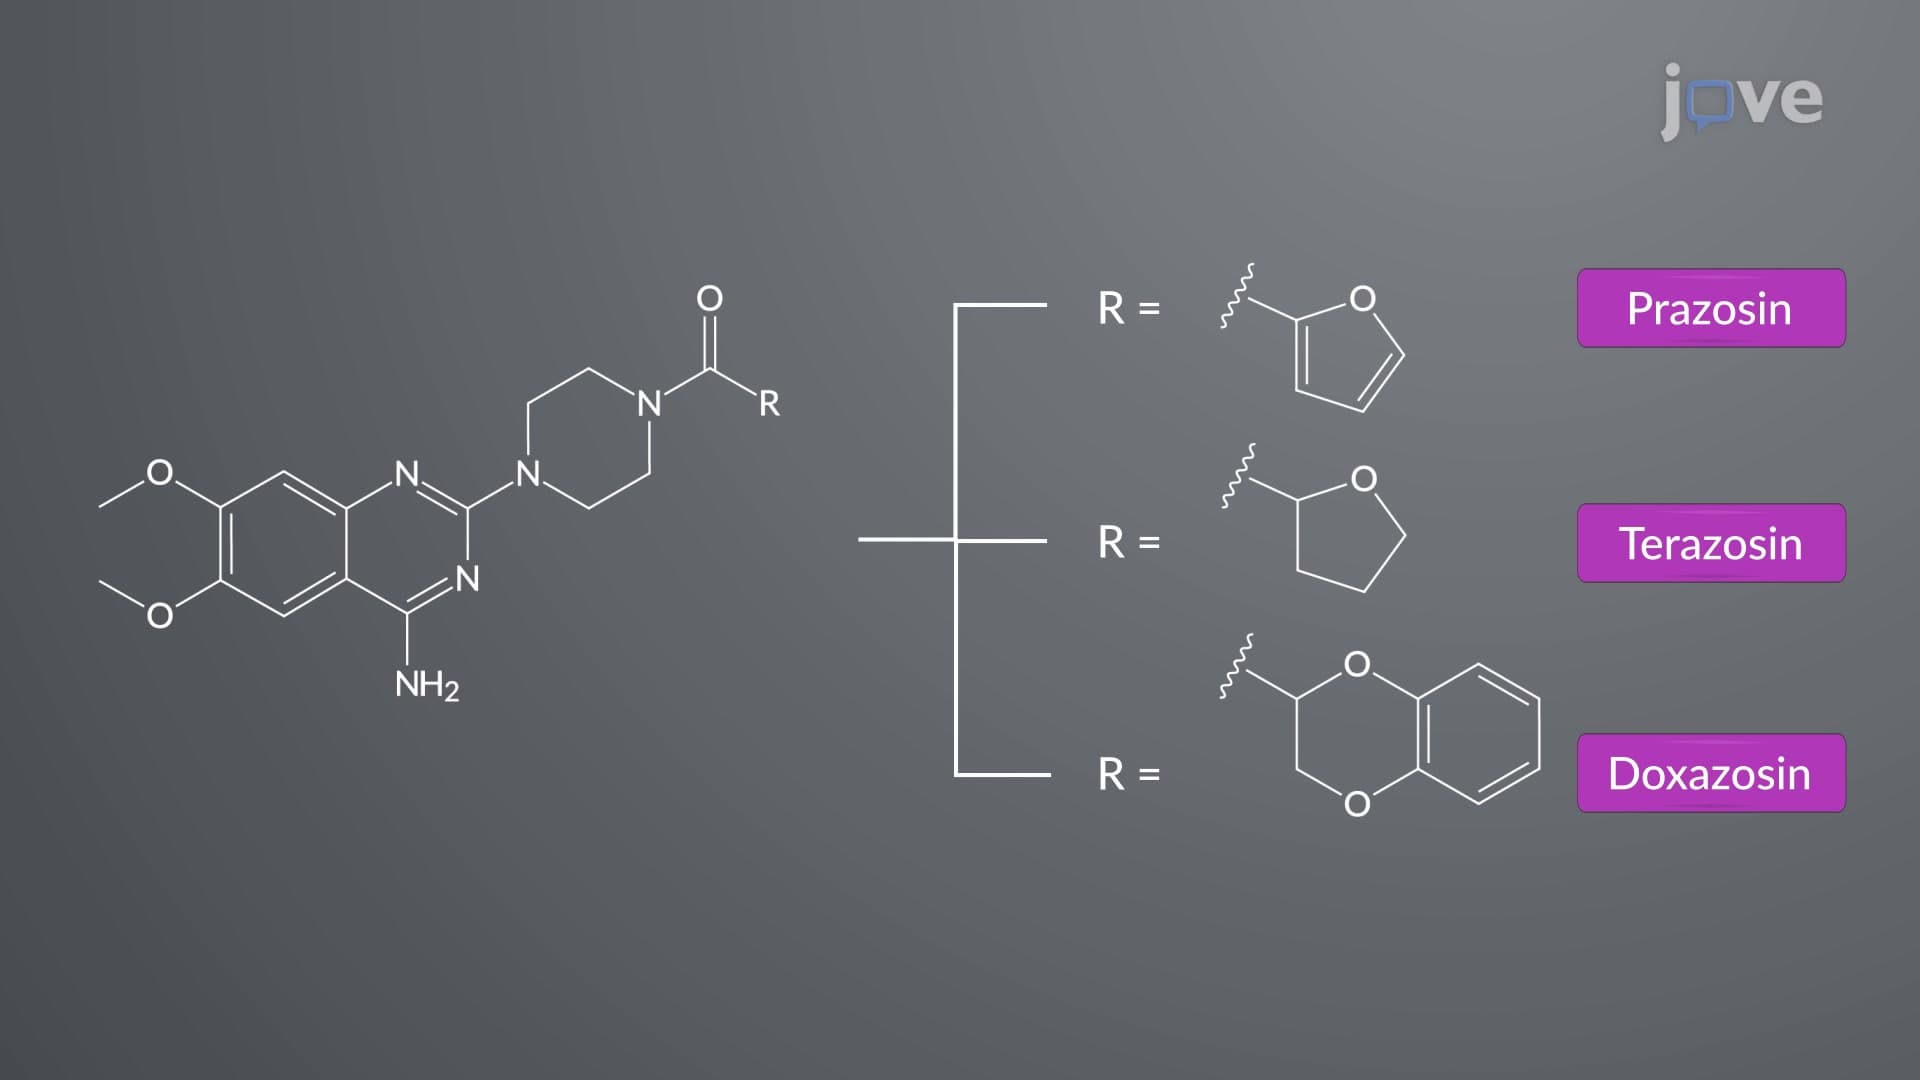
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:26
Adrenergic Receptors: β Subtype
Isoprenaline > Adrenaline > Noradrenaline
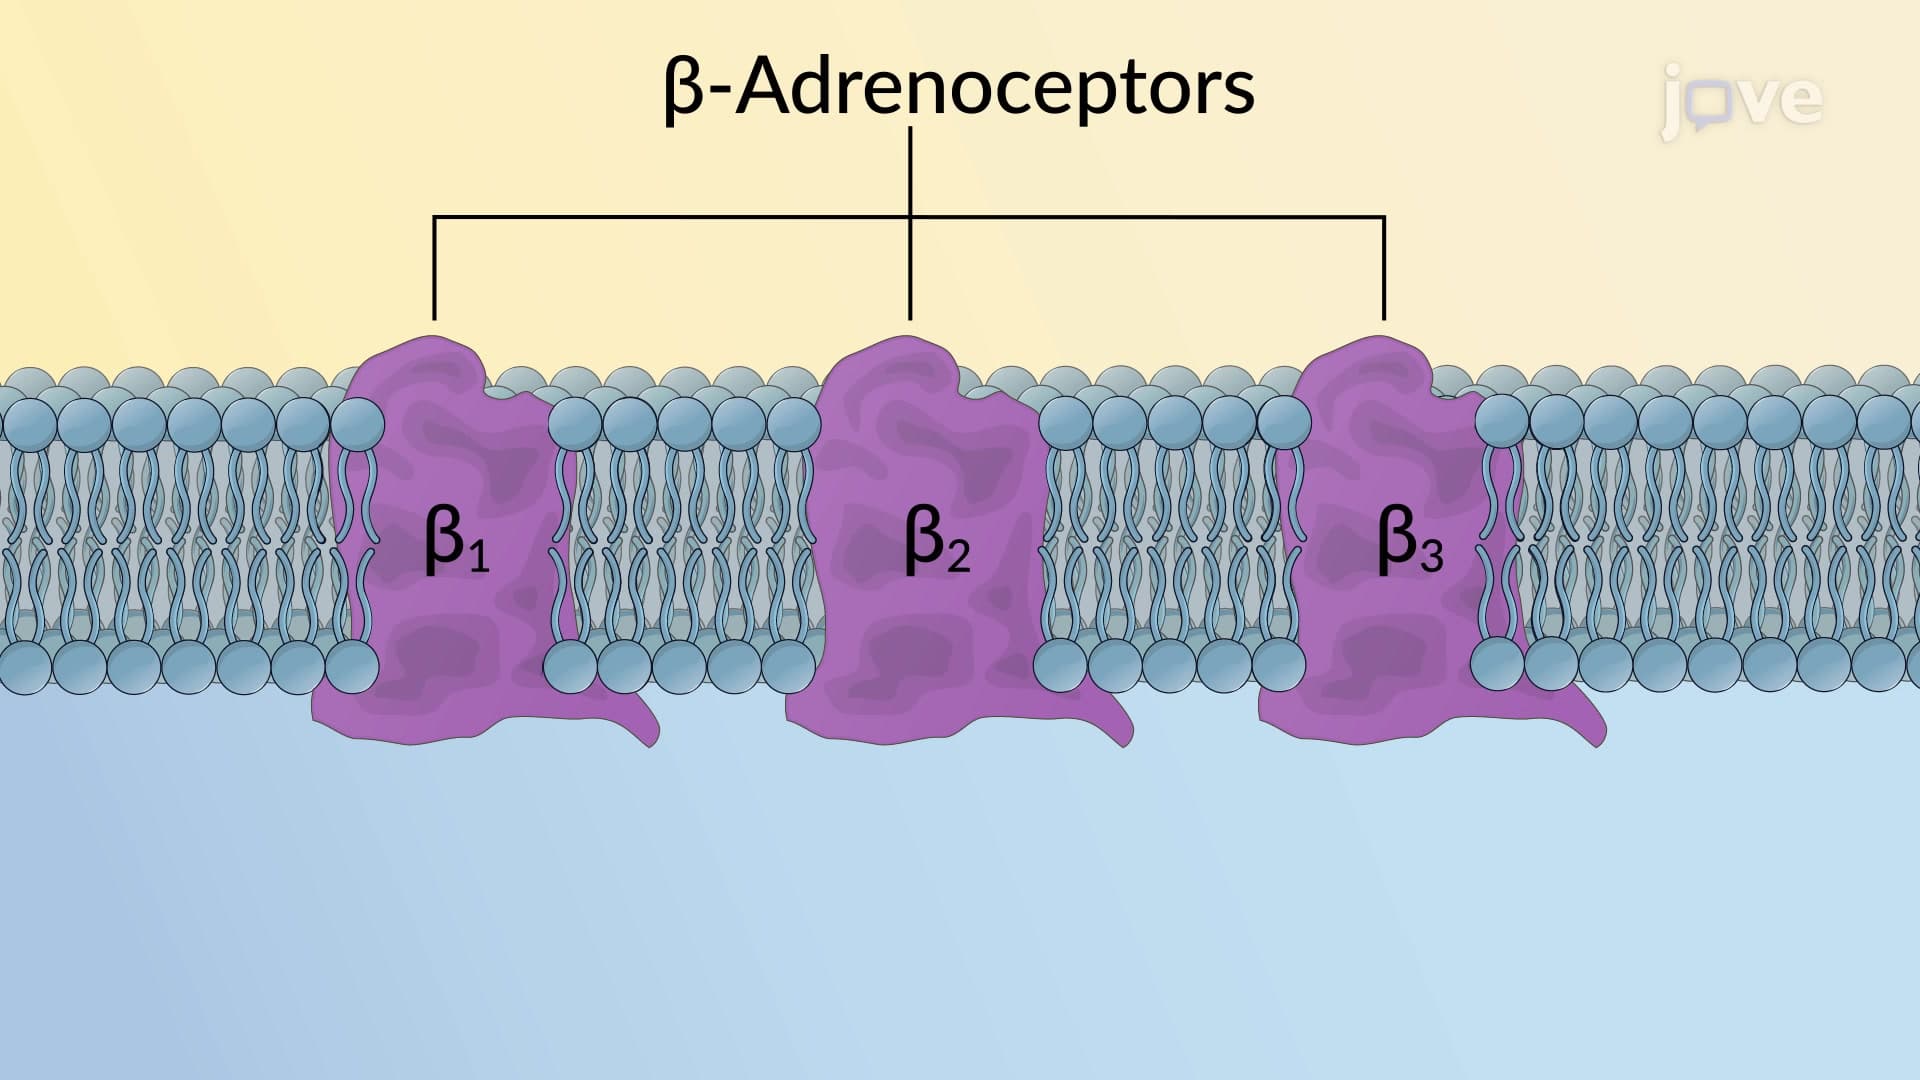
Neurotransmitter binding to these receptors causes activation of adenylyl cyclase resulting in increased concentrations of cAMP and modulation of calcium ion channels within the cell. They are further classified into β1, β2, and β3 subtypes.
β1-adrenoceptors: β1-adrenoceptors have equal affinities for...
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...