Computational identification of transcription factor binding sites by functional analysis of sets of genes sharing
Davide Corà1, Ferdinando Di Cunto, Paolo Provero
1Dipartimento di Fisica Teorica, Università di Torino, and INFN, sezione di Torino, Italy. cora@to.infn.it

Background:
Transcriptional regulation is a key mechanism in the functioning of the cell, and is mostly effected through transcription factors binding to specific recognition motifs located upstream of the coding region of the regulated gene. The computational identification of such motifs is made easier by the fact that they often appear several times in the upstream region of the regulated genes, so that the number of occurrences of relevant motifs is often significantly larger than expected by pure chance.
Results:
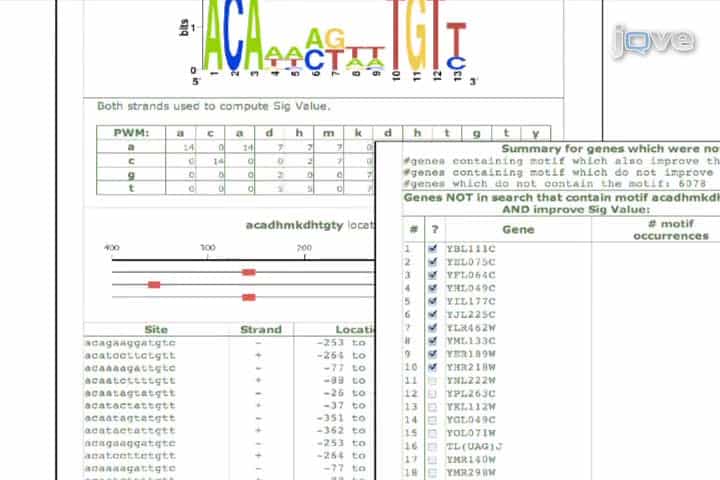
To exploit this fact, we construct sets of genes characterized by the statistical overrepresentation of a certain motif in their upstream regions. Then we study the functional characterization of these sets by analyzing their annotation to Gene Ontology terms. For the sets showing a statistically significant specific functional characterization, we conjecture that the upstream motif characterizing the set is a binding site for a transcription factor involved in the regulation of the genes in the set.
Conclusions:
The method we propose is able to identify many known binding sites in S. cerevisiae and new candidate targets of regulation by known transcription factors. Its application to less well studied organisms is likely to be valuable in the exploration of their regulatory interaction network.
Related Concept Videos
 02:16
Transcription Factors
02:16Transcription Factors
 02:13
Cooperative Binding of Transcription Regulators
02:13Cooperative Binding of Transcription Regulators
 02:33
Combinatorial Gene Control
The expression of more than 30,000 genes is controlled by approximately 2000-3000 transcription factors. This is possible because a single transcription factor can recognize more than one regulatory sequence. The specificity in gene...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...