Mitochondrial disorders: prevalence, myths and advances
1The Murdoch Children's Research Institute and Genetic Health Services Victoria, Royal Children's Hospital, and Department of Paediatrics, University of Melbourne, Melbourne, Australia. david.thorburn@mcri.edu.au

Abstract:
Disorders of mitochondrial oxidative phosphorylation (OXPHOS) are renowned for their variability in clinical features and genetic causes. This makes it difficult to determine their true prevalence, but recent studies have documented a minimum birth prevalence of 13.1/100000 or 1/7634 for oxidative phosphorylation disorders with onset at any age. This clearly remains an underestimate but it indicates that oxidative phosphorylation disorders can be regarded as the most common group of inborn errors of metabolism. Pathogenic mutations causing human oxidative phosphorylation disorders have now been identified in more than 30 of the 37 mitochondrial DNA genes and in more than 30 nuclear genes. Most of the nuclear gene defects cause autosomal recessive diseases, but autosomal dominant and X-linked disorders also occur. It is likely that at least another 30, and perhaps over 100, nuclear-encoded oxidative phosphorylation disorders await identification. Oxidative phosphorylation genetics are complex and there appear to be a number of common misconceptions about mitochondrial DNA mutations that may impede optimal investigation and management of patients. In our experience, mitochondrial DNA mutations are not a negligible cause of OXPHOS disorders in children but account for 20-25% of cases. Similarly, a family history suggesting maternal inheritance is the exception rather than the norm for children with mitochondrial DNA mutations, many of whom have de novo mutations. Only some mitochondrial DNA mutations disappear from cultured cells, so deficient enzyme activity in fibroblasts does not imply the presence of a nuclear defect. Finally, it is still widely thought that there are very few reproductive options that can be offered to women at risk of transmitting a mitochondrial DNA mutation. While a cautious approach is needed, there is now a consensus that prenatal diagnosis should be offered to some women, particularly those at lower recurrence risk. Preimplantation genetic diagnosis can also be an option.
Insights
Oxidative phosphorylation (OXPHOS) disorders are common inborn errors of metabolism, with genetic causes involving both mitochondrial and nuclear DNA. Understanding their complex genetics is crucial for diagnosis and management.
Area of Science:
- Biochemistry
- Genetics
- Metabolic Disorders
Background:
- Mitochondrial oxidative phosphorylation (OXPHOS) disorders exhibit significant clinical and genetic heterogeneity, complicating prevalence estimation.
- Recent data suggest a minimum birth prevalence of 13.1/100,000, positioning OXPHOS disorders as the most common inborn errors of metabolism.
Purpose of the Study:
- To clarify the genetic basis and prevalence of OXPHOS disorders.
- To address common misconceptions regarding mitochondrial DNA mutations and their inheritance patterns.
- To review diagnostic approaches and reproductive options for affected families.
Main Methods:
- Analysis of genetic causes, including mutations in mitochondrial and nuclear DNA genes.
- Review of clinical data and diagnostic findings in patients with OXPHOS disorders.
- Evaluation of current understanding of inheritance patterns and diagnostic interpretations.
Main Results:
- Pathogenic mutations identified in over 30 mitochondrial DNA genes and more than 30 nuclear genes; numerous additional nuclear defects are anticipated.
- Mitochondrial DNA mutations account for 20-25% of pediatric OXPHOS cases, often arising from de novo events rather than maternal inheritance.
- Fibroblast enzyme activity does not definitively indicate a nuclear defect due to the behavior of some mitochondrial DNA mutations.
Conclusions:
- OXPHOS disorders are a significant group of metabolic diseases with complex genetic underpinnings.
- Misconceptions about mitochondrial DNA mutation inheritance and diagnostic interpretation require clarification.
- Reproductive options, including prenatal and preimplantation genetic diagnosis, are increasingly available for at-risk women.
Related Concept Videos
 01:37
Mitochondria
 01:45
Mitochondrial Membranes
 02:59
Animal Mitochondrial Genetics
 01:39
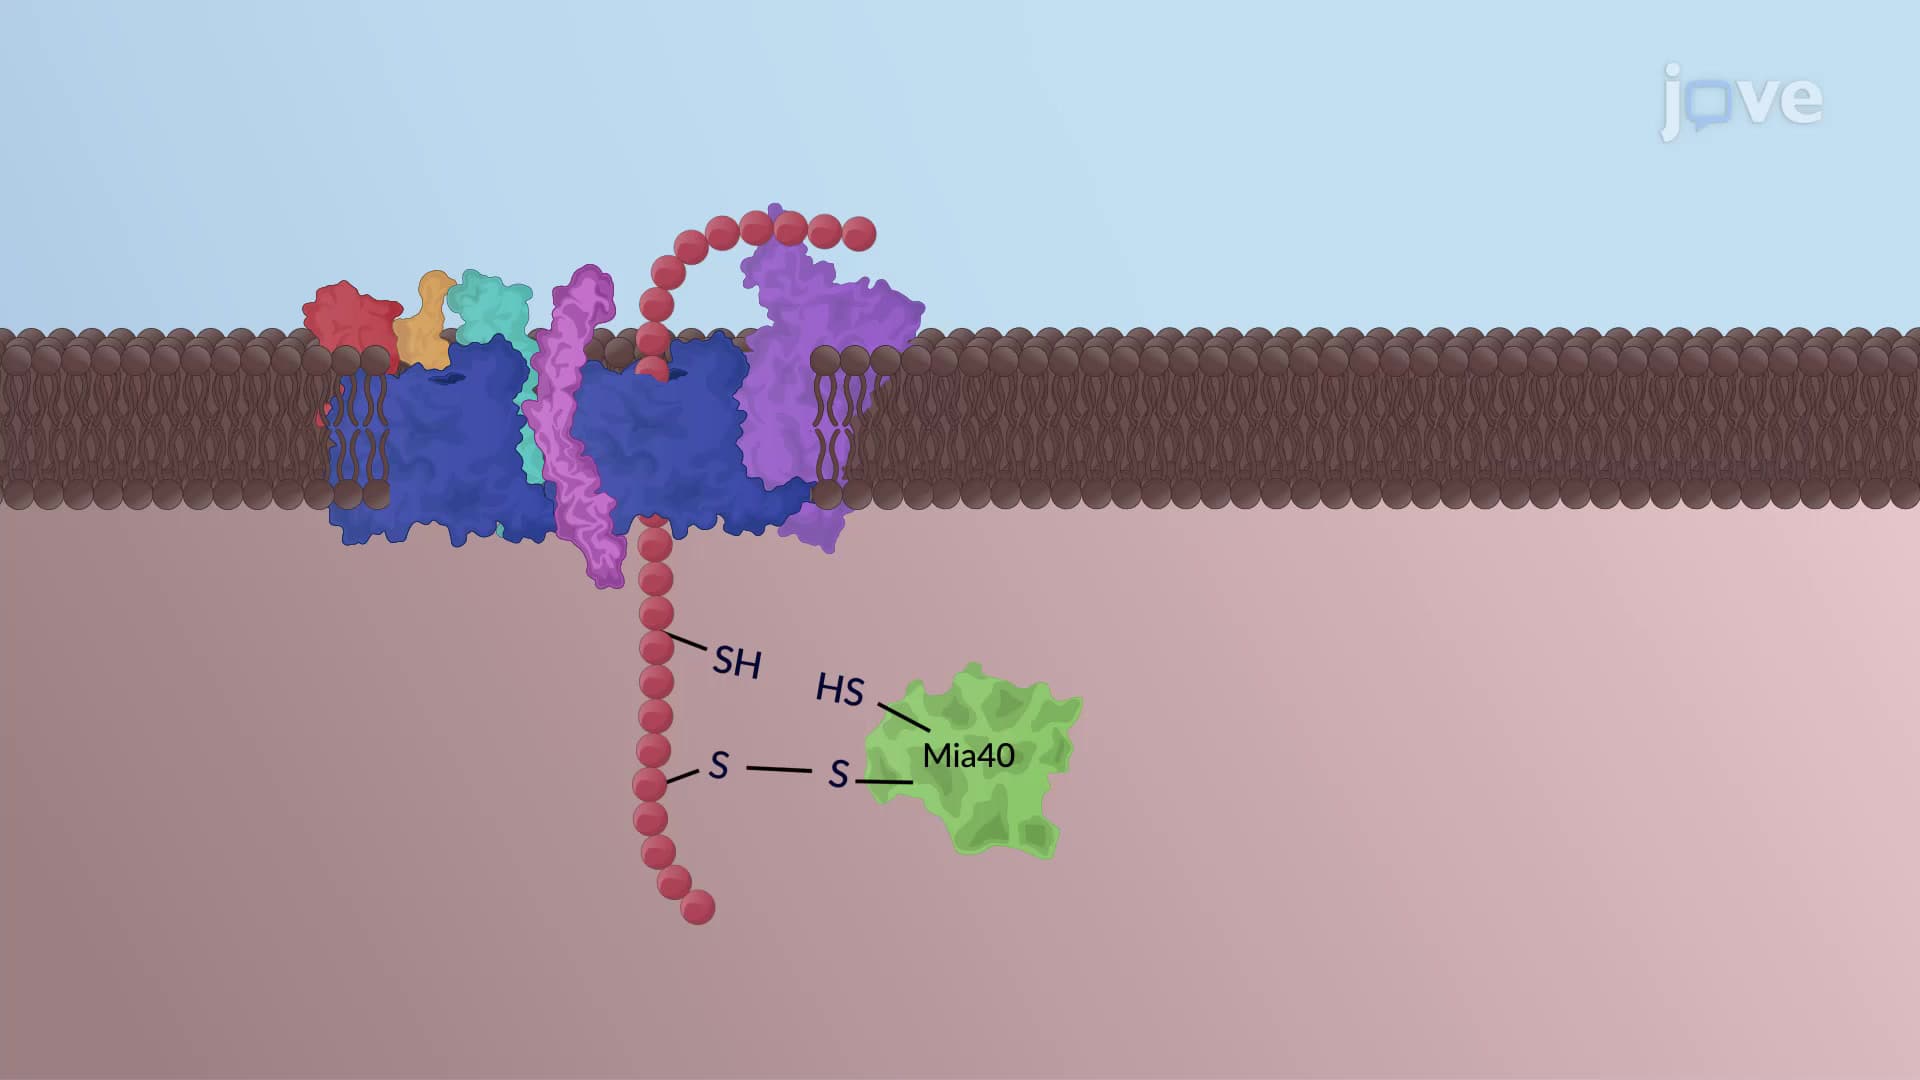
Mitochondrial Protein Sorting
Most of these mitochondrial proteins are encoded by the nucleus and imported to the mitochondria as unfolded or loosely folded precursors. Mitochondrial precursors...
 01:19
Translocation of Proteins into the Mitochondria
Sorting of outer membrane proteins:
Mitochondrial outer membrane proteins are of two types: the transmembrane, beta-barrel porins, and the membrane-anchored, alpha-helical proteins. Beta-barrel porin precursors are translocated by the TOM complex and inserted into the outer mitochondrial membrane by the SAM complex. In contrast,...
 01:46
Electron Transport Chain: Complex I and II
ROS generation is regulated and maintained at moderate levels necessary...