Related Experiment Video
Updated: Aug 23, 2026

Demonstration of the Sequence Alignment to Predict Across Species Susceptibility Tool for Rapid Assessment of Protein Conservation
Published on: February 10, 2023
Influence of conservation on calculations of amino acid covariance in multiple sequence alignments
Anthony A Fodor1, Richard W Aldrich
1Department of Molecular and Cellular Physiology, and Howard Hughes Medical Institute, Stanford University School of Medicine, Stanford, California 94305-5345, USA.

Abstract:
It has long been argued that algorithms that find correlated mutations in multiple sequence alignments can be used to find structurally or functionally important residues in proteins. We examined the properties of four different methods for detecting these correlated mutations. On both simple, artificial alignments and real alignments from the Pfam database, we found a surprising lack of agreement between the four correlated mutation methods. We argue that these differences are caused in part by differing sensitivities to background conservation. Correlated mutation algorithms can be envisioned as "filters" of background conservation with each algorithm searching for correlated mutations that occur at a different background conservation frequency.
More Related Videos
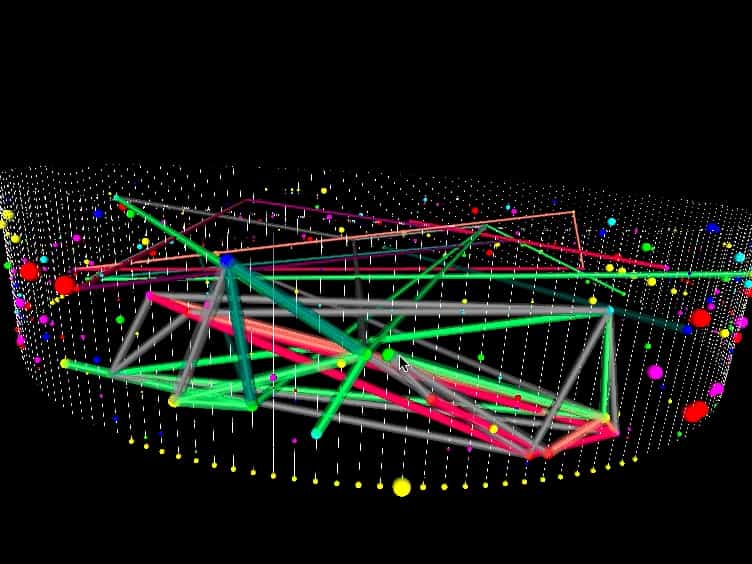 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
 07:49
Creating and Applying a Reference to Facilitate the Discussion and Classification of Proteins in a Diverse Group
Published on: August 16, 2017
Related Concept Videos
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
02:26Conservation of Protein Domains
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 02:51
Multi-species Conserved Sequences
Although the genome of each species varies greatly from each other, a few sequences are highly conserved. Such conserved DNA...
 02:54
Evolutionary Relationships through Genome Comparisons