Related Experiment Video
Updated: Aug 23, 2026

Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Comparison of kinetic Monte Carlo and molecular dynamics simulations of diffusion in a model glass former
Thomas F Middleton1, David J Wales
1University Chemical Laboratories, Lensfield Road, Cambridge CB2 1EW, United Kingdom.

Abstract:
We present results from kinetic Monte Carlo (KMC) simulations of diffusion in a model glass former. We find that the diffusion constants obtained from KMC simulations have Arrhenius temperature dependence, while the correct behavior, obtained from molecular dynamics simulations, can be super-Arrhenius. We conclude that the discrepancy is due to undersampling of higher-lying local minima in the KMC runs. We suggest that the relevant connectivity of minima on the potential energy surface is proportional to the energy density of the local minima, which determines the "inherent structure entropy." The changing connectivity with potential energy may produce a correlation between dynamics and thermodynamics.
More Related Videos
Related Concept Videos
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion
 01:24
The Kinetic Model of Gases
 01:27
Distribution of Molecular Speeds
 01:34
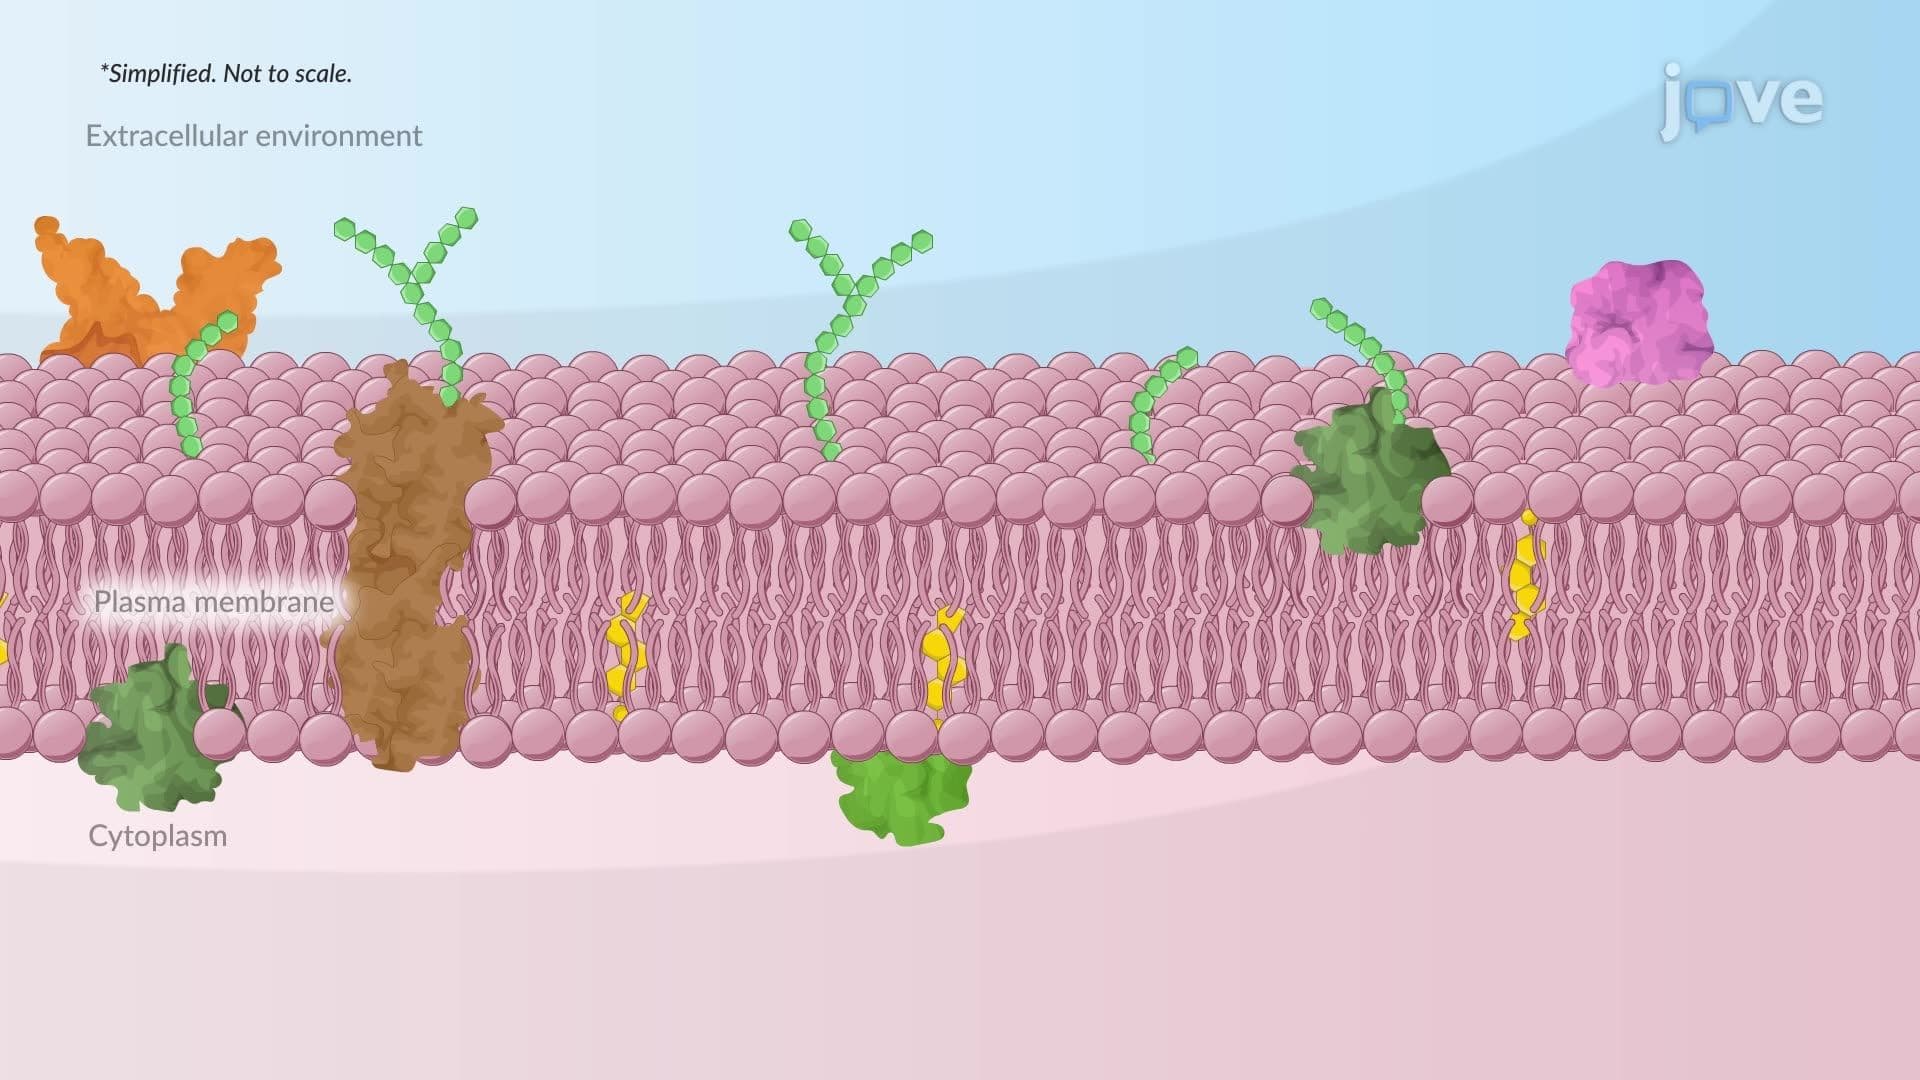
Fluid Mosaic Model
 01:12
Kinetic Theory of an Ideal Gas
The number of molecules in one mole is called Avogadro's number...
 02:26
Molecular Comparison of Gases, Liquids, and Solids