Related Experiment Video
Updated: Aug 23, 2026

Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
Adsorption and vibrational spectroscopy of ammonia at mordenite: ab initio study
1Institut fur Materialphysik and Center for Computational Material Science, Universitat Wien, Sensengasse, A-1090 Vienna, Austria.

Abstract:
The adsorption of ammonia at various active centers at the outer and inner surfaces of mordenite, involving Brønsted acid (BA) sites, terminal silanol groups, and Lewis sites has been investigated using periodic ab initio density-functional theory. It is shown that ammonia forms an ammonium ion when adsorbed at strong BA sites. The calculated adsorption energies for different BA sites vary in the interval from 111.5 to 174.7 kJ/mol depending on the local environment of the adduct. The lowest adsorption energy is found for a monodentate complex in the main channel, the highest for a tetradentate configuration in the side pocket. At weak BA sites such as terminal silanol groups or a defect with a BA site in a two-membered ring ammonia is H bonded via the N atom. Additional weak H bonds are formed between H atoms of ammonia and O atoms of neighboring terminal silanol groups. The calculated adsorption energies for such adducts range between 61.7 and 70.9 kJ/mol. The interaction of ammonia with different Lewis sites is shown to range between weak (DeltaE(ads)=17.8 kJ/mol) and very strong (DeltaE(ads)=161.7 kJ/mol), the strongest Lewis site being a tricoordinated Al atom at the outer surface. Our results are in very good agreement with the distribution of desorption energies estimated from temperature-programmed desorption (TPD) and microcalorimetry experiments, the multipeaked structure of the TPD spectra is shown to arise from strong and weak Brønsted and Lewis sites. The vibrational properties of the adsorption complexes are investigated using a force-constant approach. The stretching and bending modes of NH(4) (+) adsorbed to the zeolite are strongly influenced by the local environment. The strongest redshift is calculated for the asymmetric stretching mode involving the NH group hydrogen bonded to the bridging O atom of the BA site, the shift is largest for a monodentate and smallest for a tetradentate adsorption complex. The reduced symmetry of the adsorbate also leads to a substantial splitting of the stretching and bending modes. In agreement with experiment we show that the main vibrational feature which differentiates coordinatively bonded ammonia from a hydrogen-bonded ammonium ion is the absence of bending modes above 1630 cm(-1) and in the region between 1260 and 1600 cm(-1), and a low-frequency bending band in the range from 1130 to 1260 cm(-1). The calculated distribution of vibrational frequencies agrees very well with the measured infrared adsorption spectra. From the comparison of the adsorption data and the vibrational spectra we conclude that due to the complex adsorption geometry the redshift of the asymmetric stretching is a better measure of the acidity of an active sites than the adsorption energy.

Related Concept Videos
 01:19
NMR Spectroscopy Of Amines
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:15
Mass Spectrometry of Amines
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 00:55
Mass Spectrometry: Amine Fragmentation
In amines, the number of nitrogen atoms affects the mass of the molecular ion, which is described by the nitrogen rule of mass spectrometry. This rule states that a compound containing a single or...
 01:19
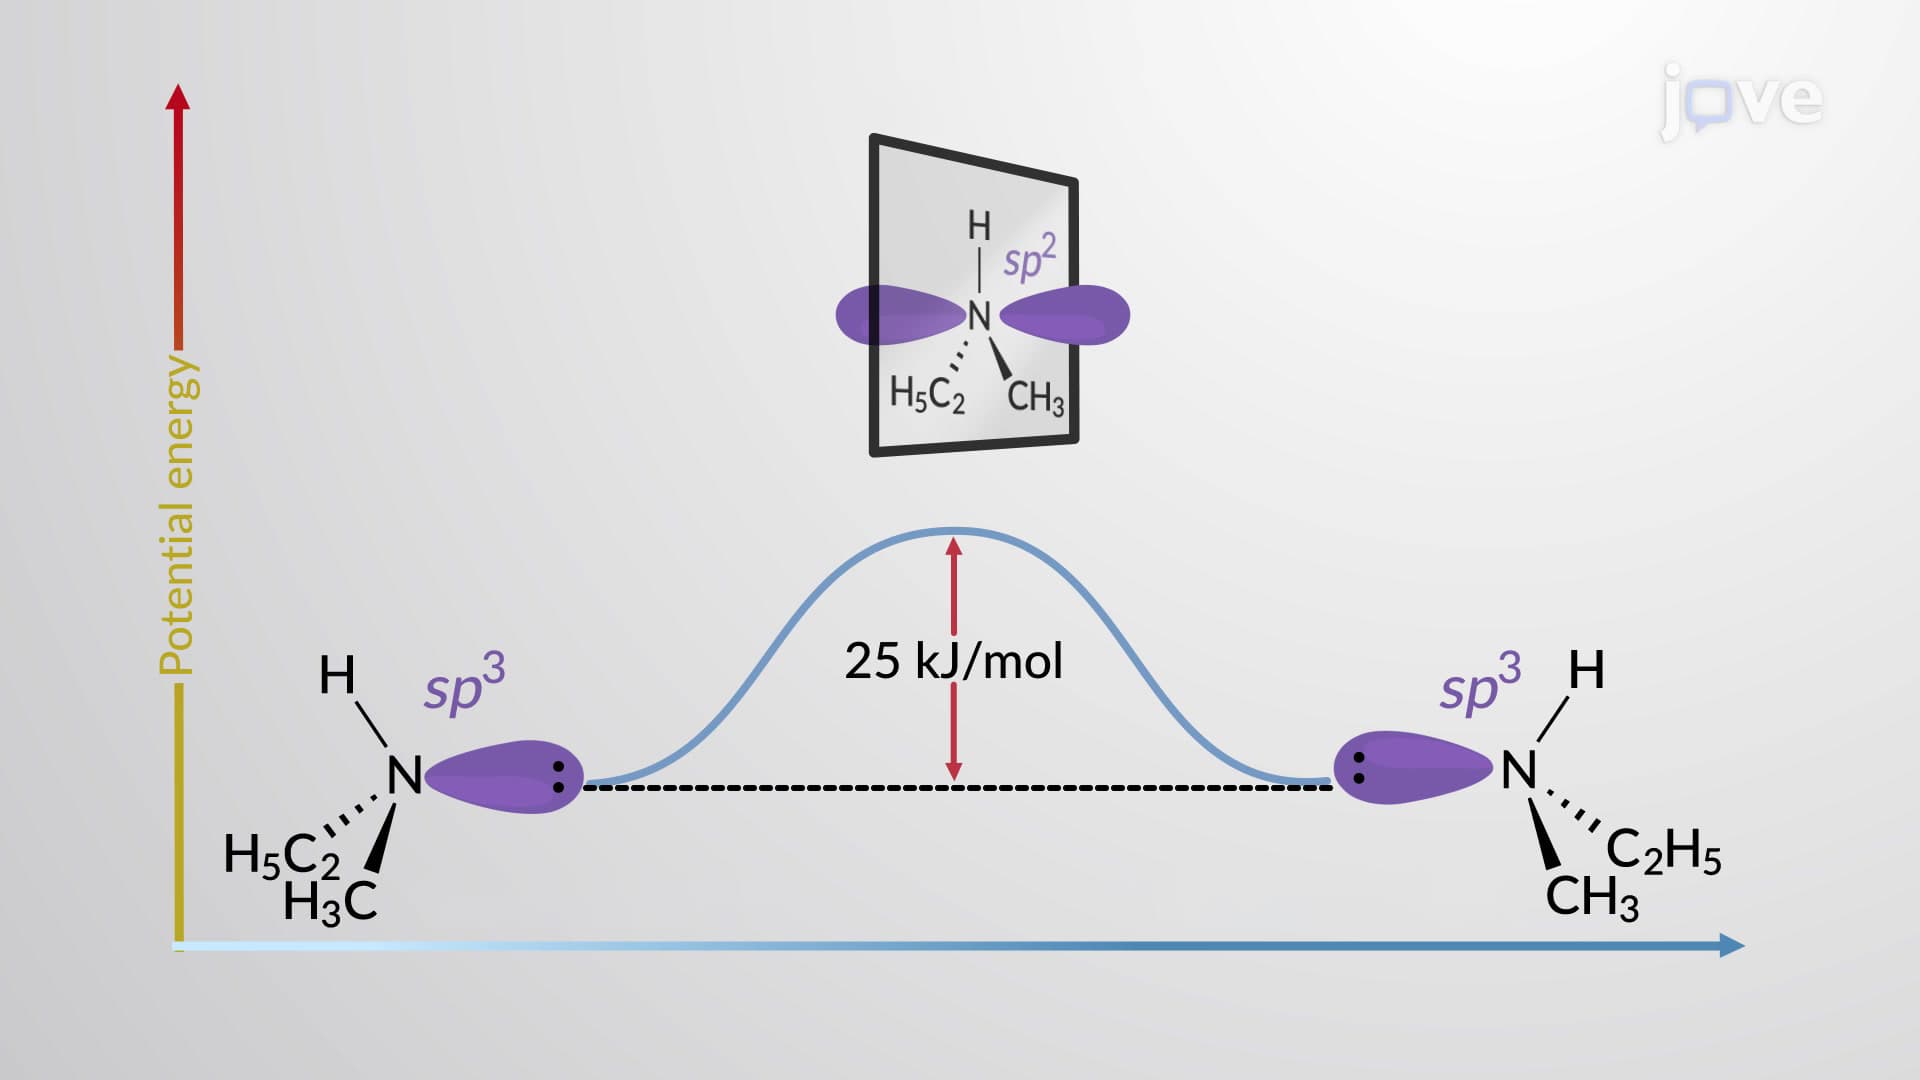
Structure of Amines