Related Experiment Video
Updated: Aug 23, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
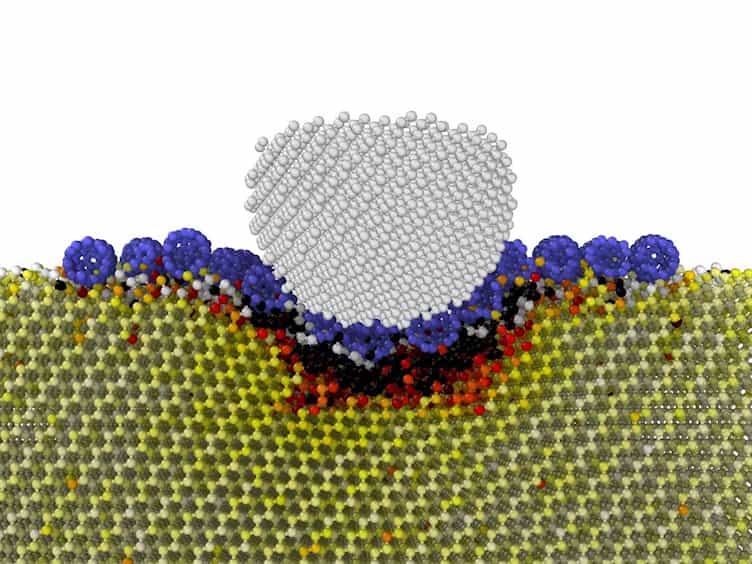
Molecular dynamics simulation of water in a contact with an iron pyrite FeS2 surface
Michael R Philpott1, Igor Yu Goliney, Ting Ting Lin
1Department of Materials Science, National University of Singapore, Kent Ridge, Singapore 119260. chmmp@nus.edu.sg

Abstract:
A strong adsorption of the water molecules to the pyrite surface is shown by a molecular dynamic simulation of the water-iron pyrite FeS2 interface. Water molecules closest to the pyrite surface are bound by an electrostatic interaction to the iron atoms in grooves running parallel to one of the crystal axes. The grooves are about two atoms wide and are directed along 010 for the (001) surface. The position of the water-surface potential minimum and the energy of adsorption were determined by optimization for a single water molecule at the interface. At room temperature and normal density there are altogether three distinguishable layers of water above the surface. One is associated with the groove: one with H bonding to the sulphur atoms comprising the ridges separating the grooves, and the third with the soft wall boundary between the absorbed water layers and bulk region of water. Simulations were also used to explore the effect of a temperature range significant for geophysical studies.