Related Experiment Video
Updated: Aug 23, 2026

Isolation and Kv Channel Recordings in Murine Atrial and Ventricular Cardiomyocytes
Published on: March 12, 2013
State-dependent trapping of flecainide in the cardiac sodium channel
Eugene Ramos1, Michael E O'leary
1Department of Pathology, Anatomy and Cell Biology, Jefferson Medical College, 1020 Locust Street, Philadelphia, PA 19107, USA.

Abstract:
Flecainide is a Class I antiarrhythmic drug and a potent inhibitor of the cardiac (Nav1.5) sodium channel. Although the flecainide inhibition of Nav1.5 is typically enhanced by depolarization, the contributions of the open and inactivated states to flecainide binding and inhibition remain controversial. We further investigated the state-dependent binding of flecainide by examining its inhibition of rapidly inactivating and non-inactivating mutants of Nav1.5 expressed in Xenopus oocytes. Applying flecainide while briefly depolarizing from a relatively negative holding potential resulted in a low-affinity inhibition of the channel (IC(50) = 345 microM). Increasing the frequency of stimulation potentiated the flecainide inhibition (IC(50) = 7.4 microM), which progressively increased over the range of voltages where Nav1.5 channels activated. This contrasts with sustained depolarizations that effectively stabilize the channels in inactivated states, which failed to promote significant flecainide inhibition. The voltage sensitivity and strong dependence of the flecainide inhibition on repetitive depolarization suggests that flecainide binding is facilitated by channel opening and that the drug does not directly bind to closed or inactivated channels. The binding of flecainide to open channels was further investigated in a non-inactivating mutant of Nav1.5. Flecainide produced a time-dependent decay in the current of the non-inactivating mutant that displayed kinetics consistent with a simple pore blocking mechanism (K(D) = 11 microM). At hyperpolarized voltages, flecainide slowed the recovery of both the rapidly inactivating (tau = 81 +/- 3 s) and non-inactivating (tau = 42 +/- 3 s) channels. Mutation of a conserved isoleucine of the D4S6 segment (I1756C) creates an alternative pathway that permits the rapid diffusion of anaesthetics out of the Nav1.5 channel. The I1756C mutation accelerated the recovery of both the rapidly inactivating (tau = 12.6 +/- 0.4 s) and non-inactivating (tau = 7.4 +/- 0.1 s) channels, suggesting that flecainide is trapped and not tightly bound within the pore when the channels are closed or inactivated. The data indicate that flecainide rapidly gains access to its binding site when the channel is open and inhibits Na(+) current by a pore blocking mechanism. Closing of either the activation or the inactivation gate traps flecainide within the pore resulting in the slow recovery of the drug-modified channels at hyperpolarized voltages.
Related Concept Videos
 01:22
Antiarrhythmic Drugs: Class I Agents as Sodium Channel Blockers
Class 1A Antiarrhythmic Drugs: These drugs work by moderately blocking sodium channels,...
 01:26
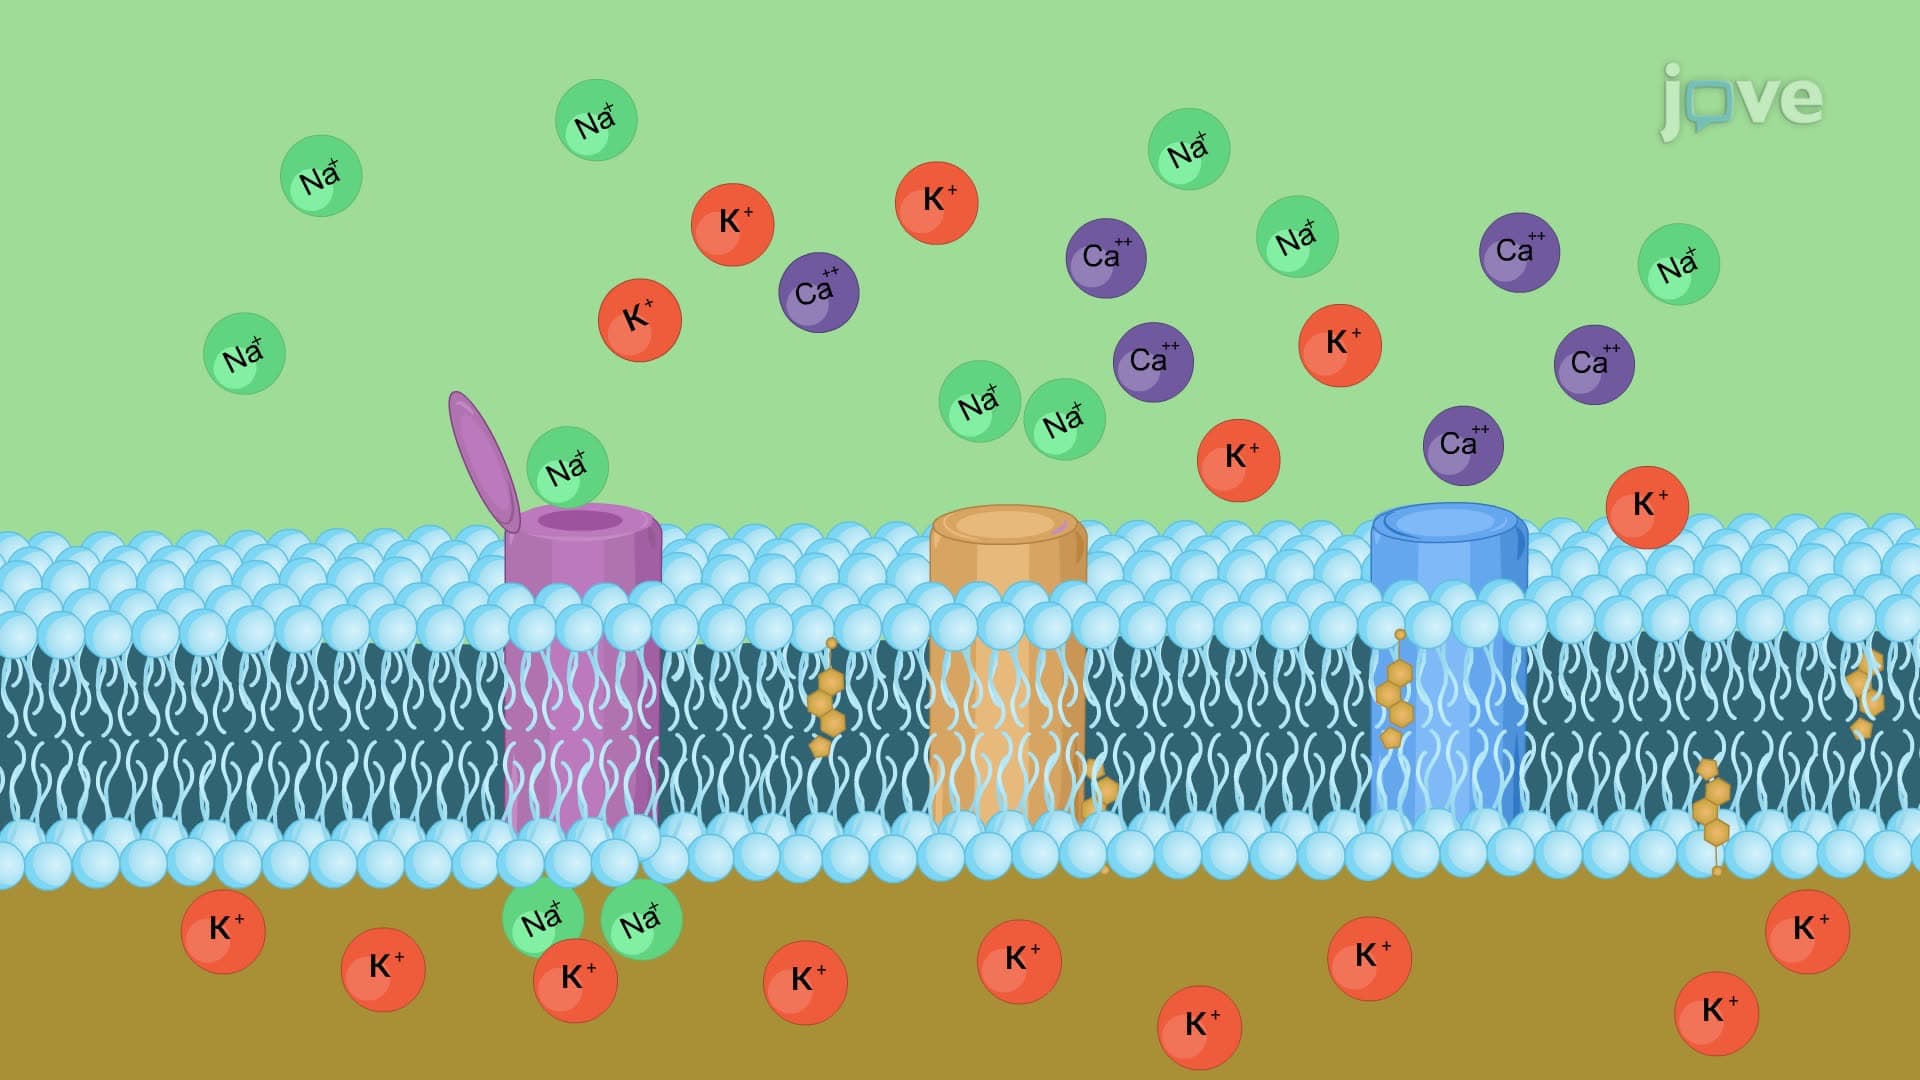
Voltage-gated Ion Channels
Generally, all voltage-gated ion channels have a 'voltage-sensing domain' that spans the lipid bilayer. The charged residues in the sensor move in response to the membrane potential changes that open the channel allowing ions movement. There are several types of...
 01:30
Cardiac Action Potential
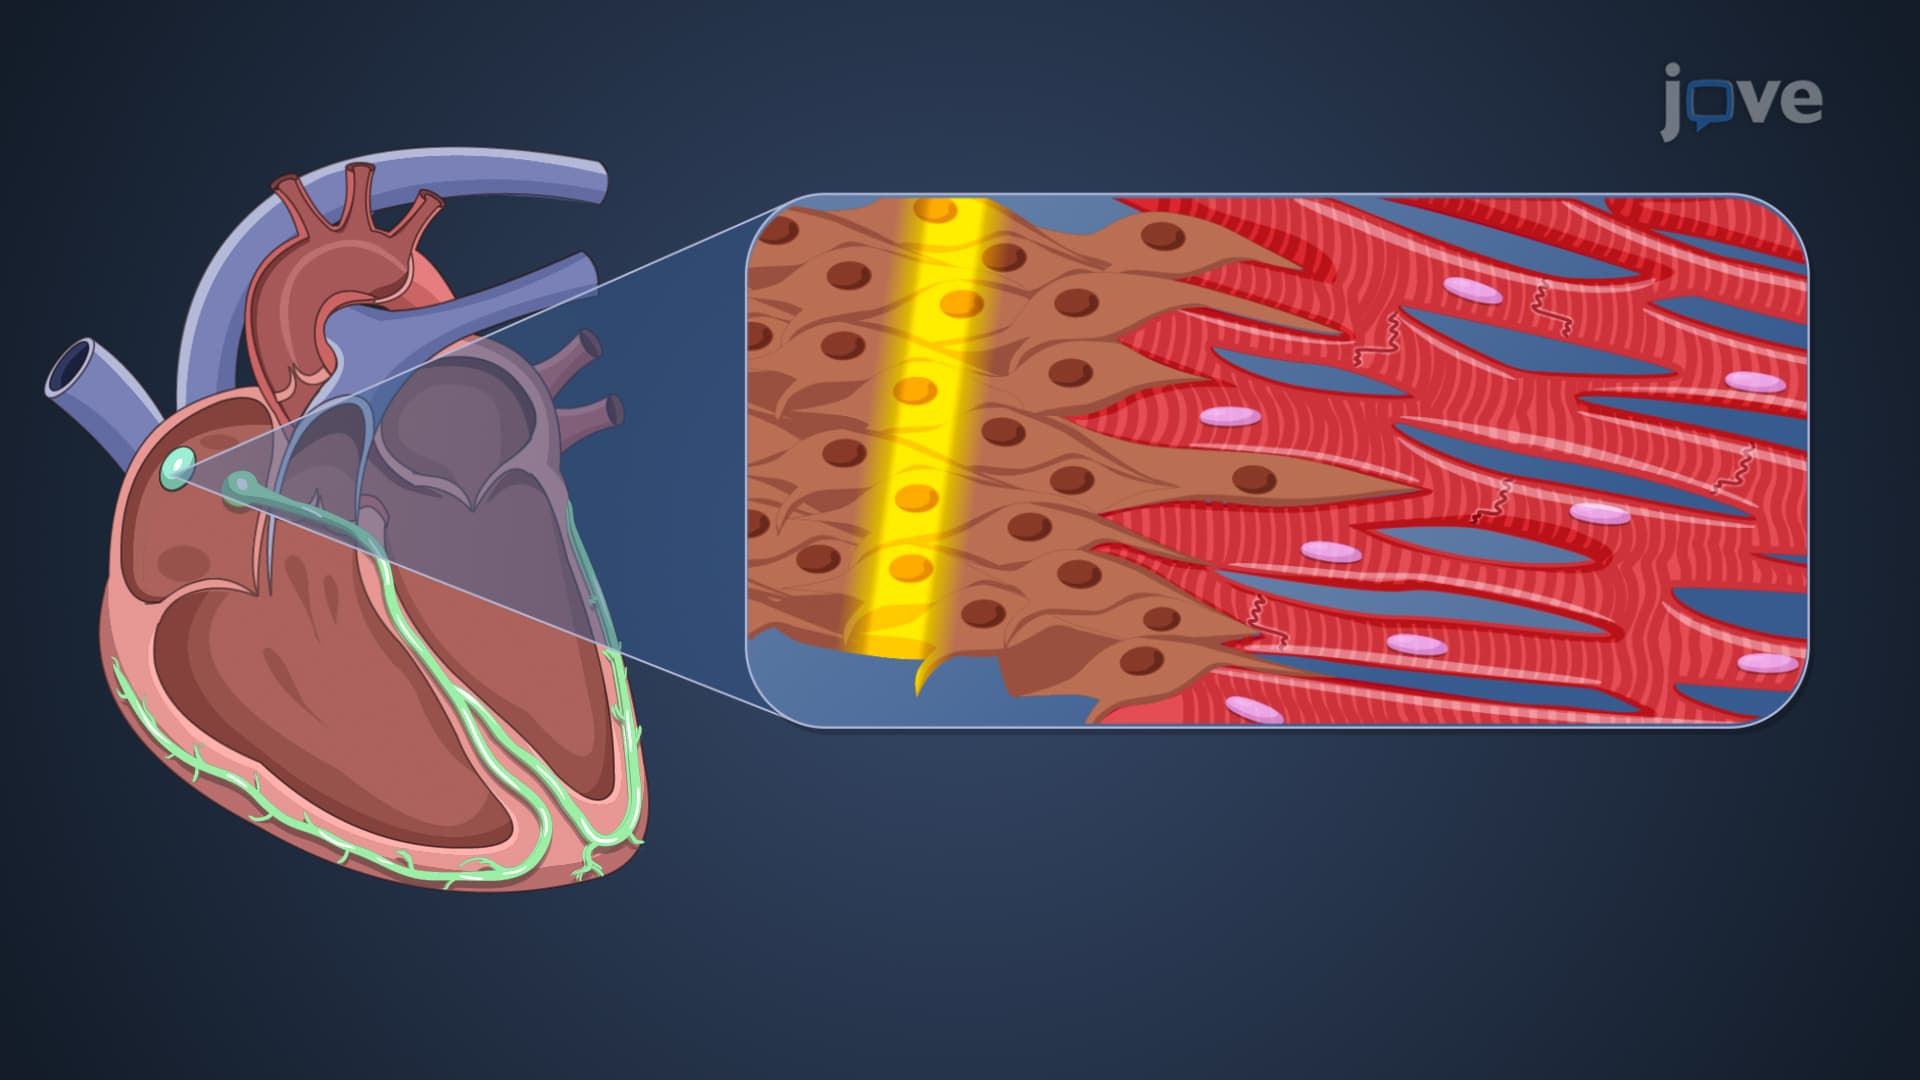
The cardiac action potential process involves a series of phases characterized by the movement of ions across the cardiac cell membranes, leading to the depolarization and repolarization of the cardiac myocytes.
Ionic Basis of Cardiac Action Potentials
 01:19
Electrophysiology of Normal Cardiac Rhythm
 01:25
ECG Interpretation of Arrhythmias II: Atrial, Junctional and Ventricular Arrhythmias
 01:21
The Resting Membrane Potential