Related Experiment Video
Updated: Aug 23, 2026

Facile Preparation of 4-Substituted Quinazoline Derivatives
Published on: February 15, 2016
Synthesis and multidrug resistance reversal activity of 1,2-disubstituted tetrahydroisoquinoline derivatives
Attila Mihályi1, Róbert Gáspár, Zita Zalán
1Institute of Pharmaceutical Chemistry, University of Szeged, H-6701 Szeged, POB 121, Hungary.

Background:
Cancer treatment often fails due to multidrug resistance (MDR) of the tumor cells. One of the major causes is overexpression of P-glycoprotein (P-gp).
Materials And Methods:
By N-substitution reactions of diamine, amino acid and amino alcohol derivatives with 1-substituted tetrahydroisoquinoline skeleton, structurally diverse 1,2-disubstituted 1,2,3,4-tetrahydroisoquinolines were synthesized. The compounds were assayed as P-gp inhibitors using a standard functional assay with rhodamine (6G) on MCF-7/Adr cells. Cytotoxicity was investigated on HeLa cells using an antiproliferative assay.
Results:
Five of the 24 compounds showed greater P-gp inhibition than the control compound verapamil with AC50 values (concentration of the compound eliciting 50% of the maximal rhodamine 6G accumulation) significantly lower than that of verapamil.
Conclusion:
Novel compounds were synthesized that showed MDR-reversal effect. One of them, (1'R*,2R*)-2-[2'-[2''-hydroxy-3''-(alpha-naphthyloxy)propyl]-6',7'-dimethoxy-1',2',3',4'-tetrahydro-1'-isoquinolyl]propan-1-ol hydrochloride, showed two times higher efficacy than verapamil at 10 times lower concentrations. The outcome makes this molecule an attractive subject for further investigation and development.
Insights
Researchers synthesized novel compounds to combat cancer multidrug resistance (MDR). One compound demonstrated significantly higher efficacy than verapamil in inhibiting P-glycoprotein (P-gp), offering a promising avenue for overcoming treatment failure.
Area of Science:
- Medicinal Chemistry
- Pharmacology
- Cancer Biology
Background:
- Multidrug resistance (MDR) in cancer is a major obstacle to effective treatment.
- Overexpression of P-glycoprotein (P-gp) is a primary mechanism driving MDR in tumor cells.
Purpose of the Study:
- To synthesize and evaluate novel 1,2-disubstituted 1,2,3,4-tetrahydroisoquinoline derivatives as P-gp inhibitors.
- To investigate the potential of these compounds to reverse multidrug resistance in cancer cells.
Main Methods:
- Synthesis of structurally diverse 1,2-disubstituted 1,2,3,4-tetrahydroisoquinolines via N-substitution reactions.
- Assay of P-gp inhibitory activity using rhodamine (6G) accumulation in MCF-7/Adr cells.
- Evaluation of cytotoxicity on HeLa cells using an antiproliferative assay.
Main Results:
- Five out of 24 synthesized compounds exhibited P-gp inhibition superior to the control, verapamil.
- These active compounds showed significantly lower AC50 values compared to verapamil.
- One specific compound demonstrated twice the efficacy of verapamil at ten times lower concentrations.
Conclusions:
- Novel tetrahydroisoquinoline derivatives effectively inhibit P-gp and show multidrug resistance-reversal effects.
- A lead compound exhibits significantly enhanced efficacy and potency compared to verapamil.
- This molecule represents a promising candidate for further preclinical investigation and development in cancer therapy.

Related Concept Videos
 01:28
Inhibitors of Bacterial DNA Synthesis
 01:15
Antihypertensive Drugs: Thiazide-Class Diuretics
 01:27
Combined Effects of Drugs: Synergism
Such synergistic combinations...
 01:22
Antiviral Nucleoside Inhibitors
 01:19
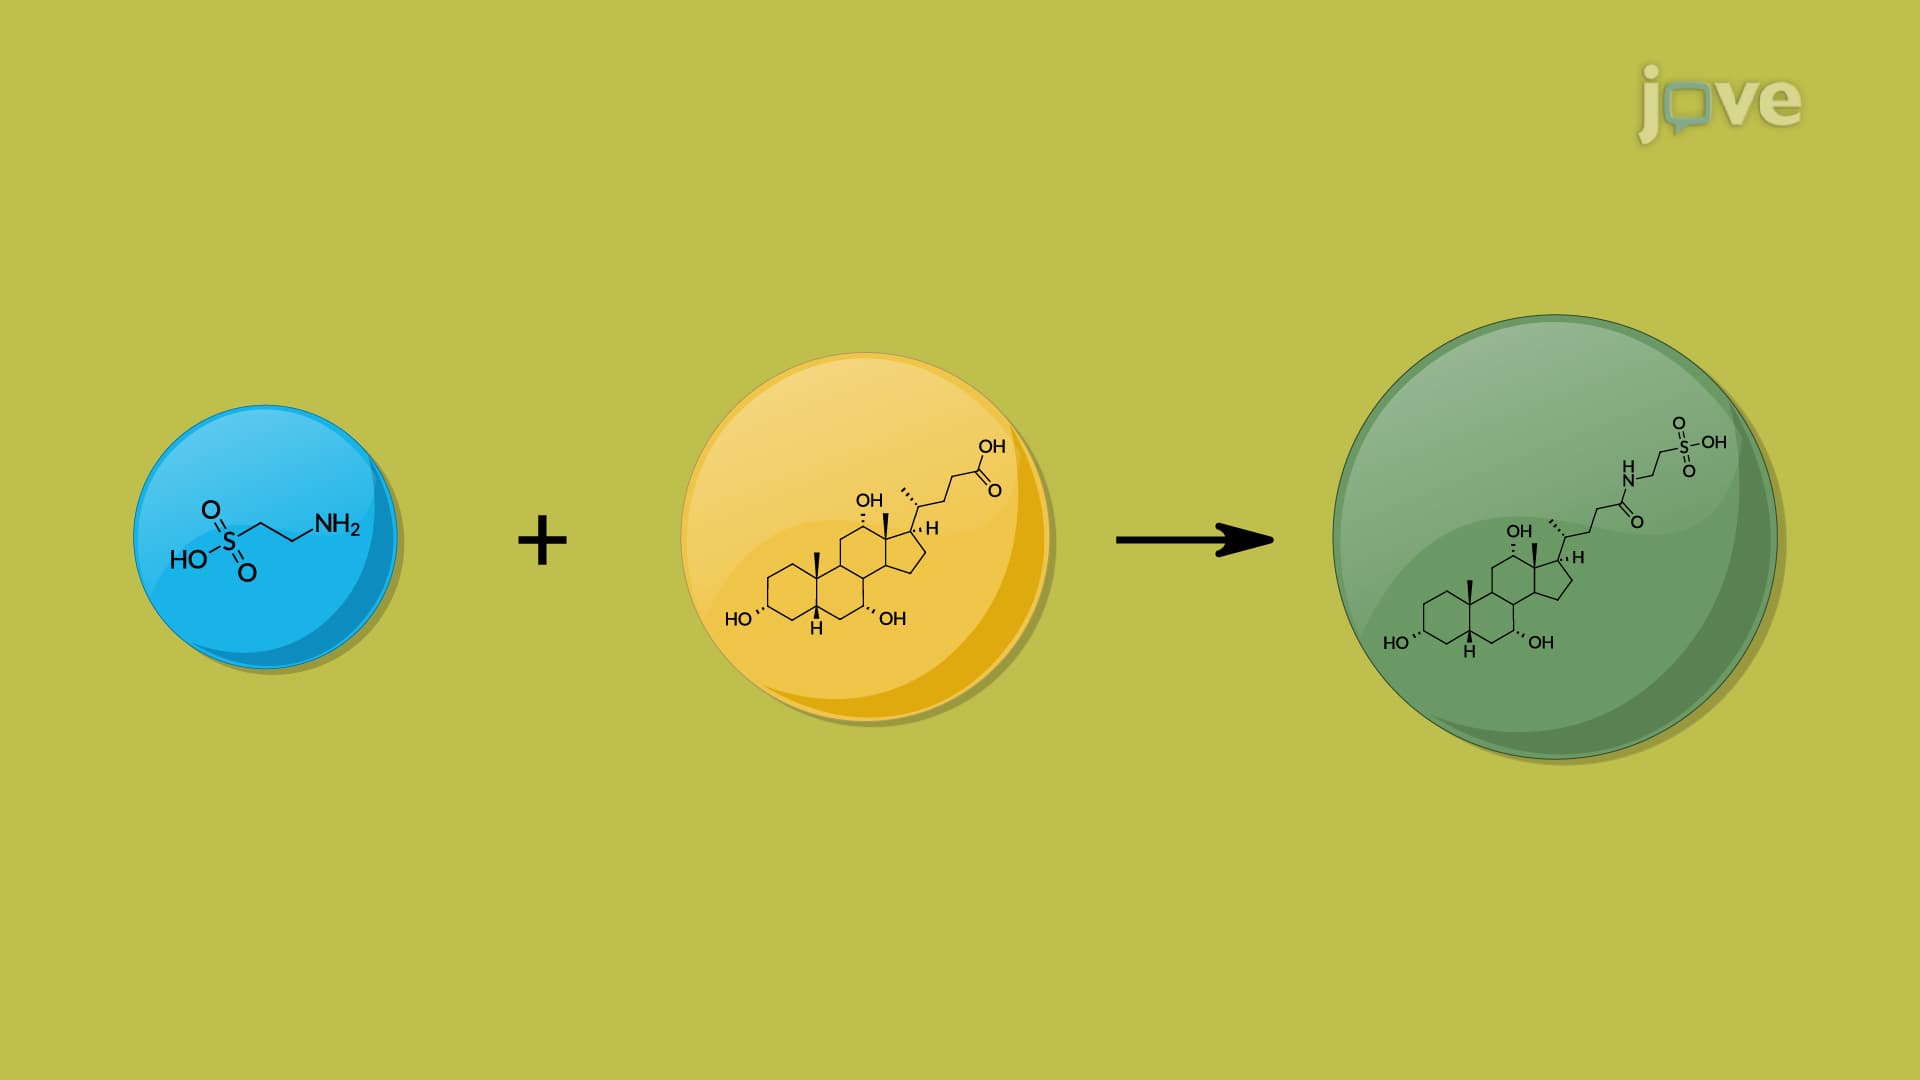
Phase II Reactions: Miscellaneous Conjugation Reactions
A key example involves the conjugation of cyanide ions, which impair cellular respiration and alter hemoglobin into non-oxygen-carrying cyanmethemoglobin. To neutralize this threat, a sulfur atom from thiosulphate is transferred to the cyanide ion, catalyzed by the enzyme rhodanese, resulting in an inactive compound called thiocyanate. The production of...
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase