2,6 di-tert-butylphenol, a nonanesthetic propofol analog, modulates alpha1beta glycine receptor function in a manner distinct from propofol
Related Experiment Video
Updated: Aug 23, 2026
Methods for the Discovery of Novel Compounds Modulating a Gamma-Aminobutyric Acid Receptor Type A Neurotransmission
Published on: August 16, 2018
Jörg Ahrens1, Gertrud Haeseler, Martin Leuwer
1Departments of *Anaesthesiology and ‡Neurology and Neurophysiology, Hannover Medical School, Hannover, Germany; and †University Department of Anaesthesia, The University of Liverpool, Liverpool, United Kingdom.

Abstract:
The anesthetic propofol (2,6 diisopropylphenol) mediates some of its effects by activating inhibitory chloride currents in the lower brainstem and spinal cord. The effects comprise direct activation of gamma-aminobutyric acid-A and glycine receptors in the absence of the natural agonist, as well as potentiation of the effect of submaximal agonist concentrations. Replacement of propofol's isopropyl groups by di-tert-butyl groups yields a compound without in vivo anesthetic effects. We have studied the effects of propofol and 2,6 di-tert-butylphenol on chloride inward currents via rat alpha1beta glycine receptors heterologously expressed in human embryonic kidney cells. Propofol, but not 2,6 di-tert-butylphenol, directly activated glycine receptors; half-maximal current activation was observed with propofol 114 +/- 27 microM. Both compounds potentiated the effect of a submaximal glycine concentration (10 microM) to a maximum value of 136% +/- 71% (propofol) and 279% +/- 109% (2,6 di-tert-butylphenol) of the response to glycine 10 microM. The 50% effective concentration for this effect was 12.5 +/- 6.4 microM and 9.4 +/- 10.2 microM for propofol and 2,6 di-tert-butylphenol, respectively. Propofol and its nonanesthetic structural analog do not differ in their ability to coactivate the glycine receptor but differ in their ability to directly activate the receptor in the absence of the natural agonist.
Related Concept Videos
 01:24
Parenteral Anesthetics: Overview
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...
 01:17
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:17
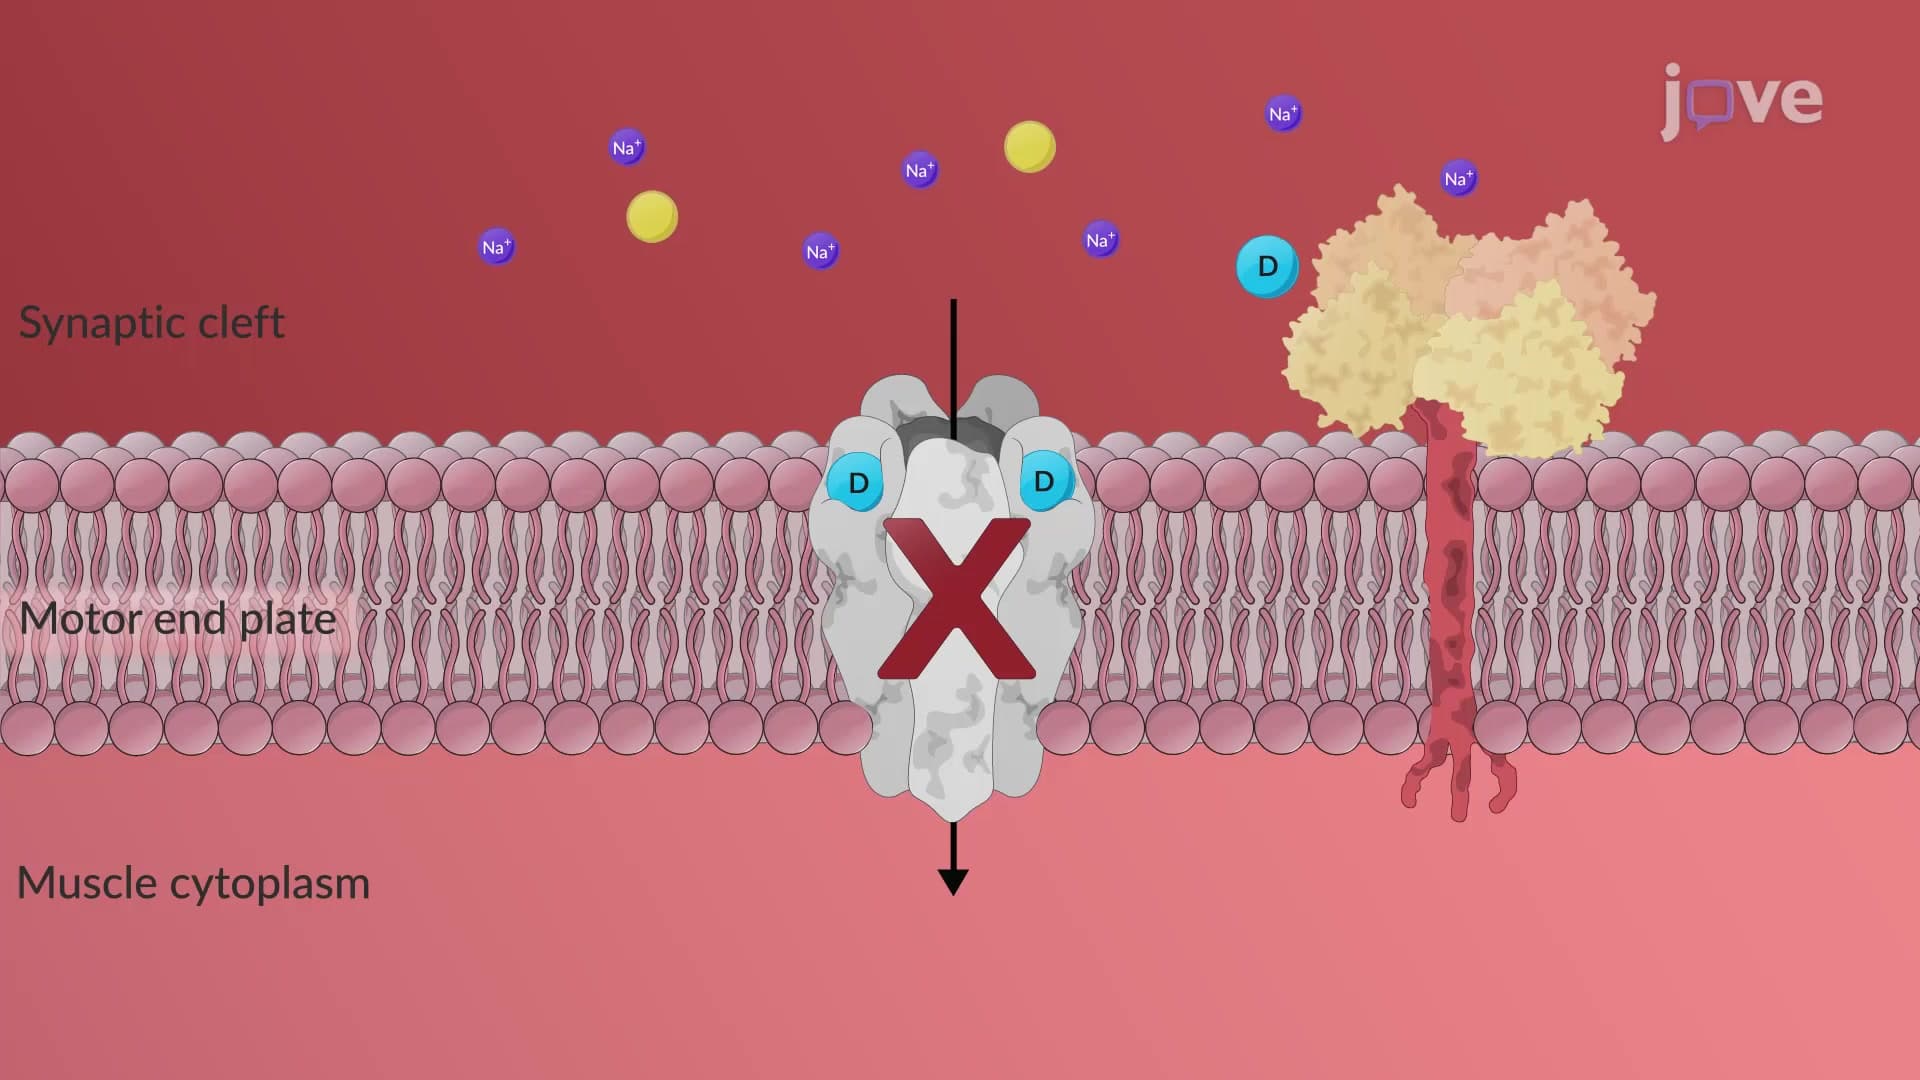
Nondepolarizing (Competitive) Neuromuscular Blockers: Mechanism of Action
Competitive antagonists prevent acetylcholine from binding to its receptor, inhibiting membrane depolarization. Without conformational changes or intrinsic...
 01:29
Indirect-Acting Cholinergic Agonists: Chemistry and Structure-Activity Relationship
Reversible inhibitors display short to medium durations of action. Short-acting agents include simple alcohols with...