Related Experiment Video
Updated: Aug 23, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Assessment of recently developed exchange-correlation functionals for the description of torsion potentials in
Juan Carlos Sancho-García1, Jérôme Cornil
1Laboratory for Chemistry of Novel Materials, Center for Research in Molecular Electronics and Photonics, University of Mons-Hainaut, Place du Parc 20, B-7000, Belgium. jc.sancho@averell.umh.ac.be

Abstract:
Newly developed exchange-correlation functionals in density functional theory (DFT) have been applied to describe conjugation effects in organic molecules. The performance of the various approaches is assessed through the calculation of torsion energy profiles and their critical comparison with available experimental data. Our results indicate that the OPTX-B95 exchange-correlation functional as well as its corresponding hybrid versions perform better than the well-established BLYP or B3LYP schemes when dealing with pi-conjugated molecules. In contrast, the recently introduced VSXC functional is not as reliable as other DFT methods for the systems examined here.
More Related Videos
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 02:40
MO Theory and Covalent Bonding
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:36
Molecular Geometry and Dipole Moments
 01:18
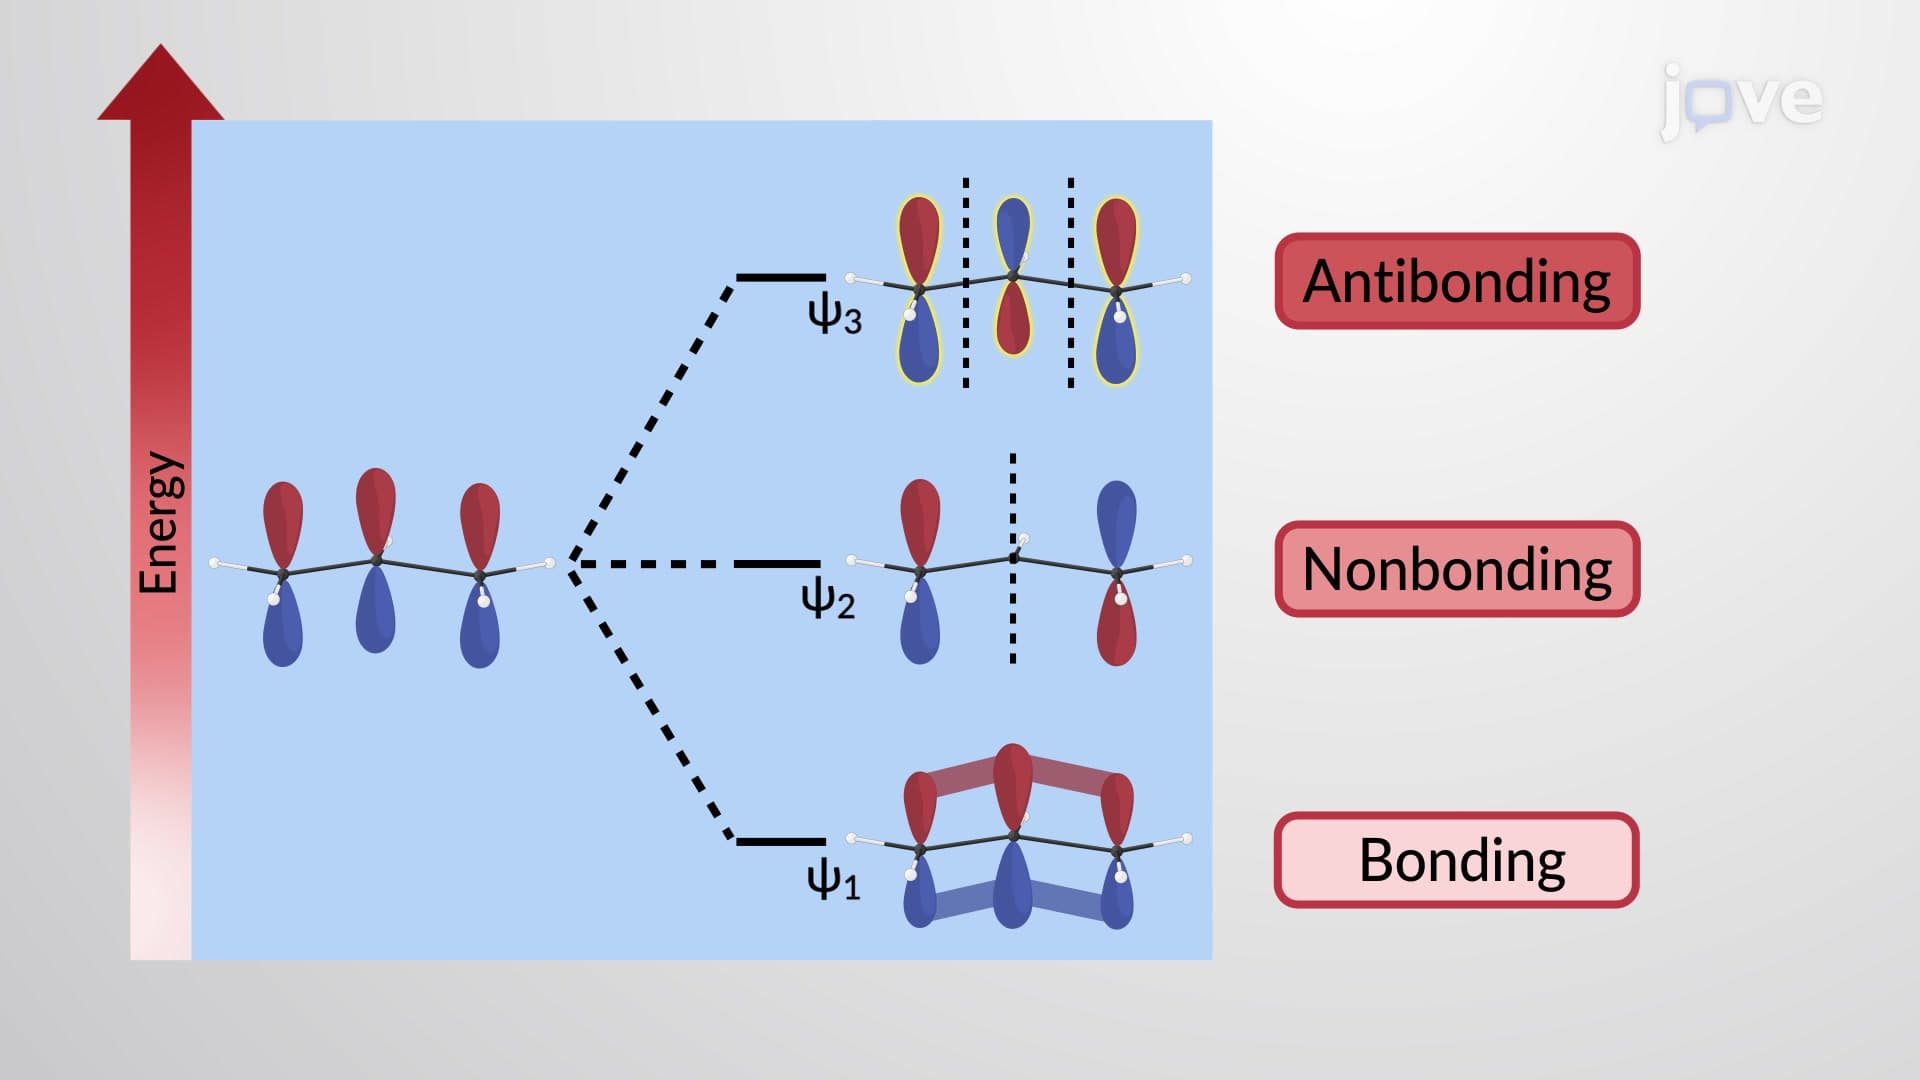
π Molecular Orbitals of the Allyl Cation and Anion