Related Experiment Video
Updated: Aug 22, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
The extended Perdew-Burke-Ernzerhof functional with improved accuracy for thermodynamic and electronic properties of
1State Key Laboratory for Physical Chemistry of Solid Surfaces, Center for Theoretical Chemistry, Department of Chemistry, Xiamen University, Xiamen 361005, China. xinxu@xmu.edu.cn

Abstract:
Density functional theory (DFT) has become the method of choice for many applications of quantum mechanics to the study of the electronic properties of molecules and solids. Despite the enormous progress in improving the functionals, the current generation is inadequate for many important applications. As part of the quest of finding better functionals, we consider in this paper the Perdew-Burke-Ernzerhof (PBE) functional, which we believe to have the best theoretical foundation, but which leads to unacceptable errors in predicting thermochemical data (heats of formation) of molecular systems [mean absolute deviation (MAD)=16.9 kcal/mol against the extended G2 data set of 148 molecules]. Much improved thermochemistry is obtained with hybrid DFT methods that include part of the Hartree-Fock exchange [thus B3LYP (Becke's three parameter scheme combining Hartree-Fock exchange, Becke gradient corrected exchange functional and Lee-Yang-Parr correlational functional) with MAD=3.1 kcal/mol and PBE0 (Perdew's hybrid scheme using PBE exchange and correlation functionals) with MAD=4.8 kcal/mol]. However we wish to continue the quest for a pure density-based DFT. Thus we optimized the four free parameters (micro, kappa, alpha, and beta) in PBE theory against experimental atomic data and the van der Waals interaction properties of Ne(2), leading to the xPBE extended functional, which significantly outperforms PBE for thermochemical properties MAD reduced to 8.0 kcal/mol while being competitive or better than PBE for predictions of geometric parameters, ionization potentials, electron affinities, and proton affinities and for the description of van der Waals and hydrogen bond interactions. Thus xPBE significantly enlarges the field of applications available for pure DFT. The functional forms thus obtained for the exchange and correlational functionals may be useful for discovering new improved functionals or formalisms.
More Related Videos


Related Concept Videos
 01:26
Thermodynamic Potentials
 01:19
Thermodynamic Properties of Ideal Solutions
 04:01
Real Gases: Effects of Intermolecular Forces and Molecular Volume Deriving Van der Waals Equation
 01:06
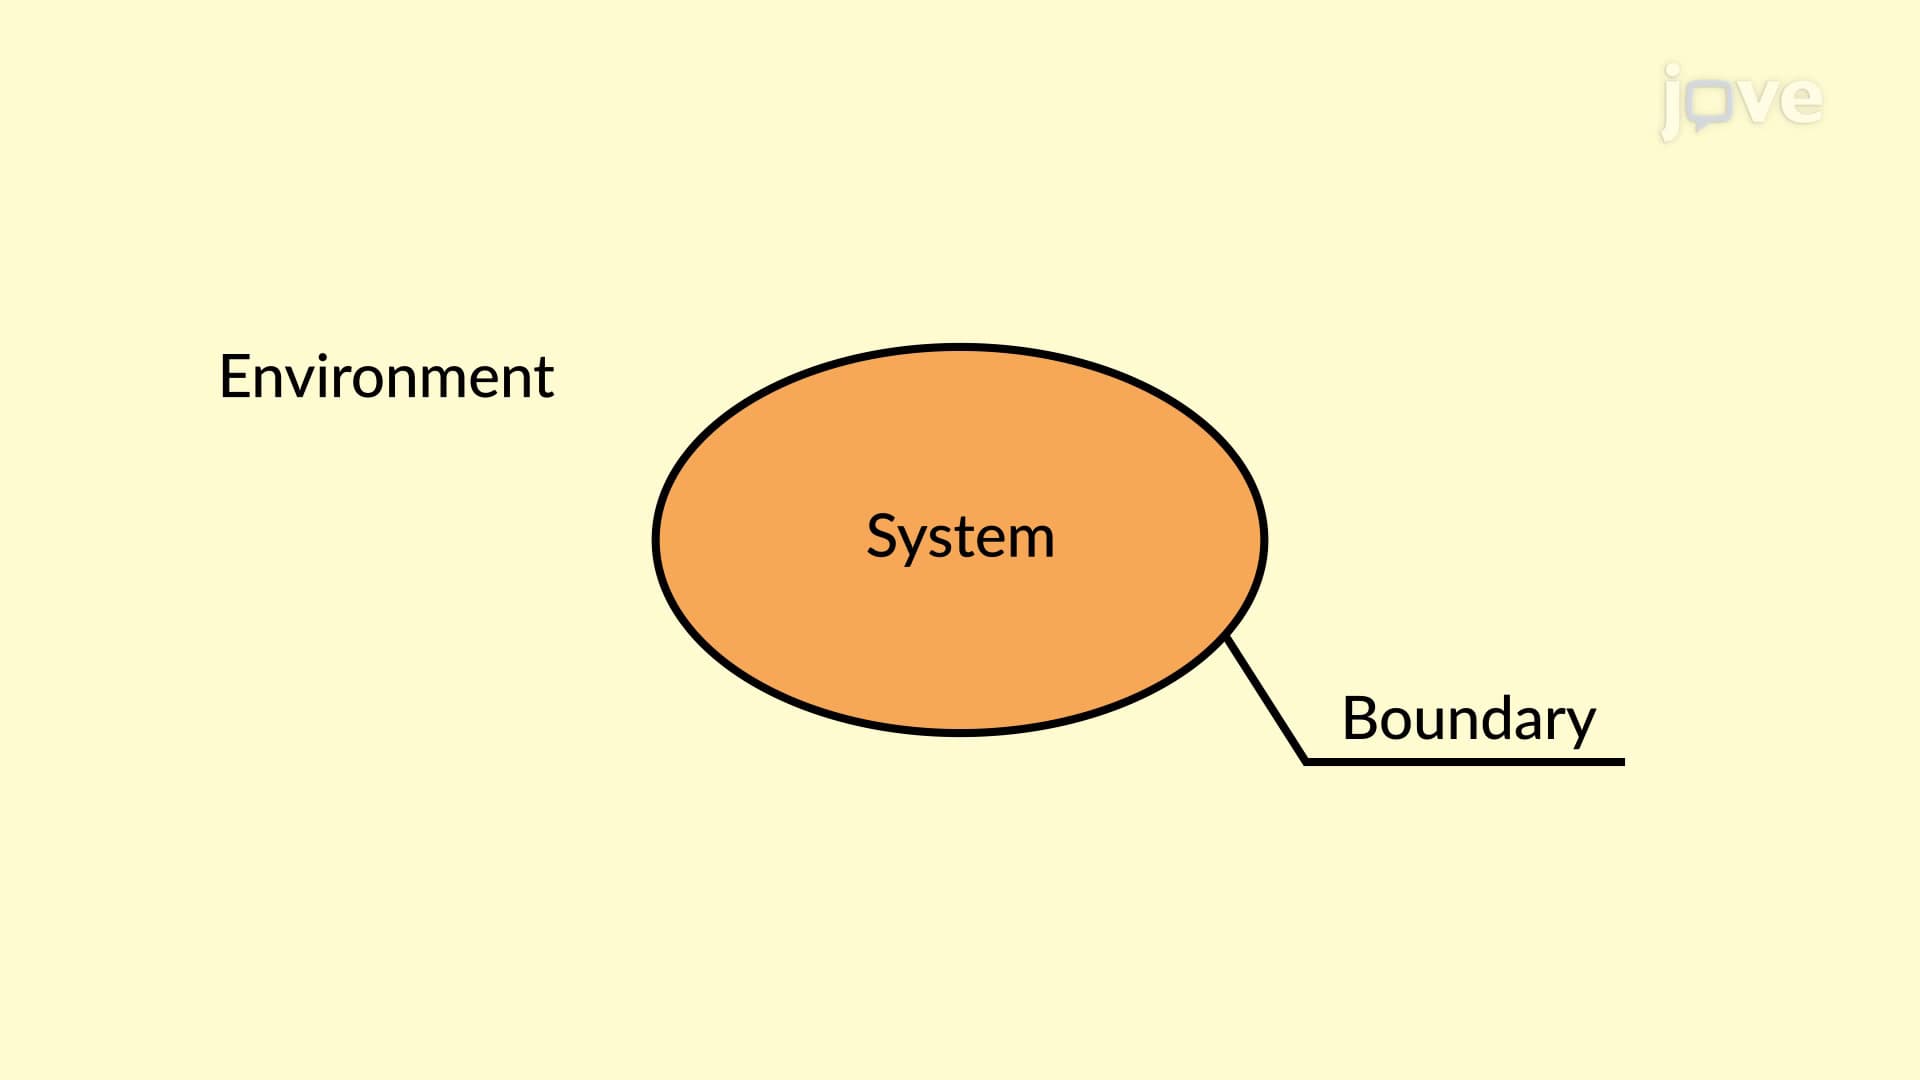
Thermodynamic Systems
Consider an example of tea boiling in a kettle. The tea and...
 01:10
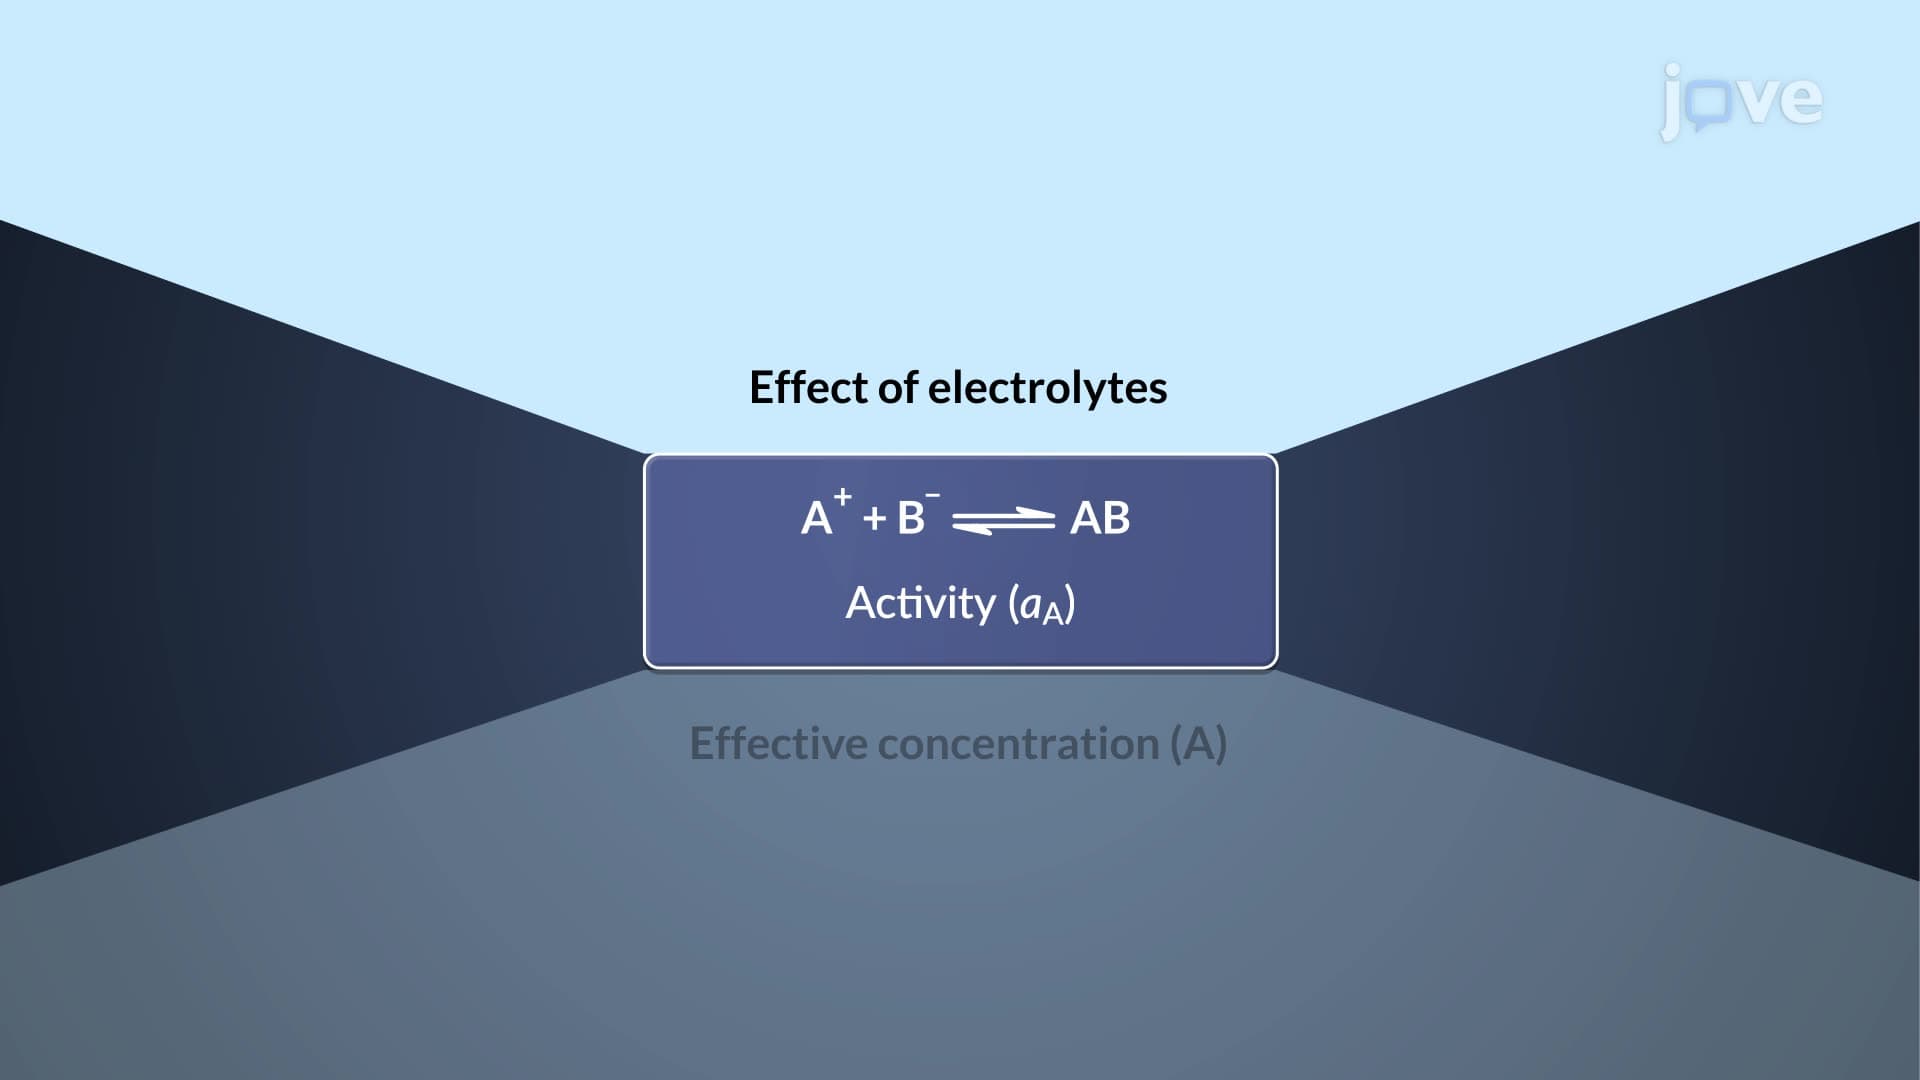
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 03:51
Molecular Orbital Theory II