Related Experiment Video
Updated: Jan 9, 2026
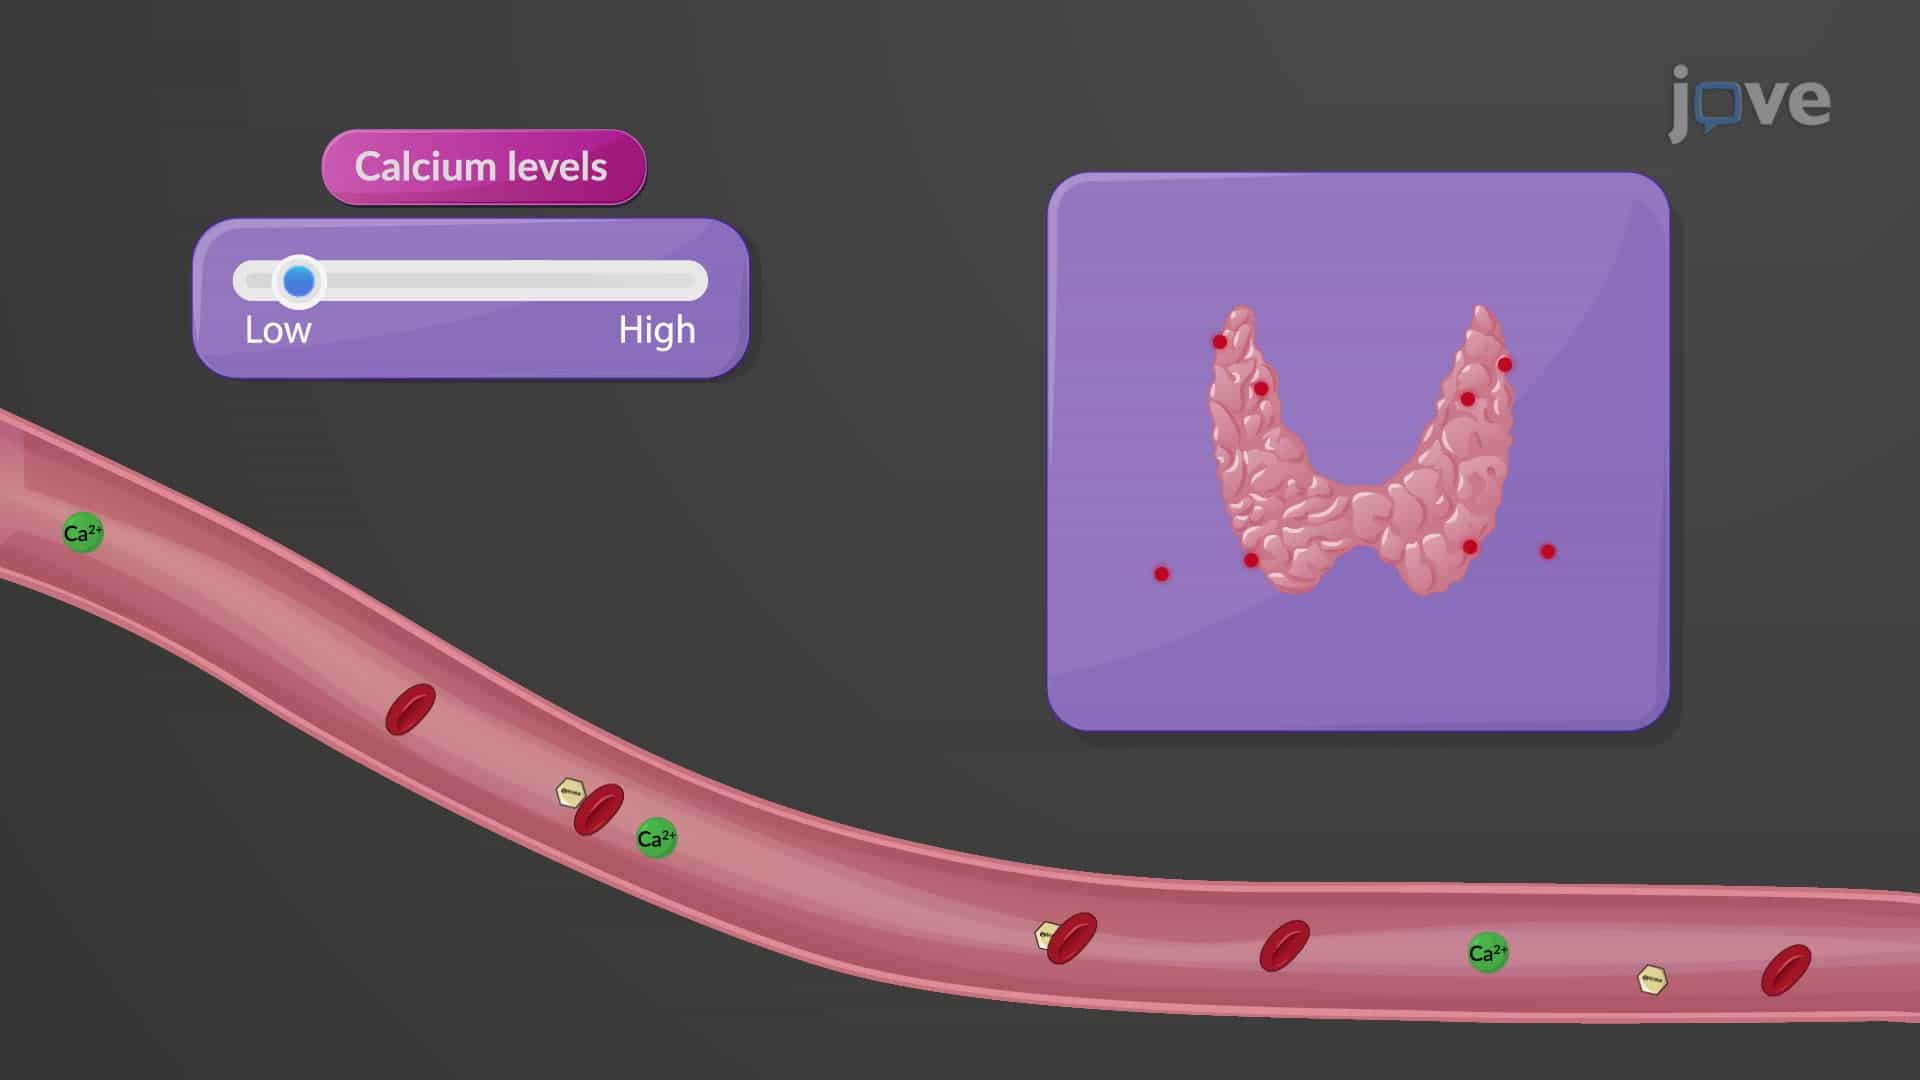

Regulation of Hormone Secretion
Hereditary and acquired angioedema: problems and progress: proceedings of the third C1 esterase inhibitor deficiency
Angelo Agostoni1, Emel Aygören-Pürsün, Karen E Binkley
1University of Milan, Italy.

Hereditary angioedema (HAE), a rare but life-threatening condition, manifests as acute attacks of facial, laryngeal, genital, or peripheral swelling or abdominal pain secondary to intra-abdominal edema. Resulting from mutations affecting C1 esterase inhibitor (C1-INH), inhibitor of the first complement system component, attacks are not histamine-mediated and do not respond to antihistamines or corticosteroids. Low awareness and resemblance to other disorders often delay diagnosis; despite availability of C1-INH replacement in some countries, no approved, safe acute attack therapy exists in the United States. The biennial C1 Esterase Inhibitor Deficiency Workshops resulted from a European initiative for better knowledge and treatment of HAE and related diseases. This supplement contains work presented at the third workshop and expanded content toward a definitive picture of angioedema in the absence of allergy. Most notably, it includes cumulative genetic investigations; multinational laboratory diagnosis recommendations; current pathogenesis hypotheses; suggested prophylaxis and acute attack treatment, including home treatment; future treatment options; and analysis of patient subpopulations, including pediatric patients and patients whose angioedema worsened during pregnancy or hormone administration. Causes and management of acquired angioedema and a new type of angioedema with normal C1-INH are also discussed. Collaborative patient and physician efforts, crucial in rare diseases, are emphasized. This supplement seeks to raise awareness and aid diagnosis of HAE, optimize treatment for all patients, and provide a platform for further research in this rare, partially understood disorder.
Hereditary angioedema (HAE), a rare but life-threatening condition, manifests as acute attacks of facial, laryngeal, genital, or peripheral swelling or abdominal pain secondary to intra-abdominal edema. Resulting from mutations affecting C1 esterase inhibitor (C1-INH), inhibitor of the first complement system component, attacks are not histamine-mediated and do not respond to antihistamines or corticosteroids. Low awareness and resemblance to other disorders often delay diagnosis; despite availability of C1-INH replacement in some countries, no approved, safe acute attack therapy exists in the United States. The biennial C1 Esterase Inhibitor Deficiency Workshops resulted from a European initiative for better knowledge and treatment of HAE and related diseases. This supplement contains work presented at the third workshop and expanded content toward a definitive picture of angioedema in the absence of allergy. Most notably, it includes cumulative genetic investigations; multinational laboratory diagnosis recommendations; current pathogenesis hypotheses; suggested prophylaxis and acute attack treatment, including home treatment; future treatment options; and analysis of patient subpopulations, including pediatric patients and patients whose angioedema worsened during pregnancy or hormone administration. Causes and management of acquired angioedema and a new type of angioedema with normal C1-INH are also discussed. Collaborative patient and physician efforts, crucial in rare diseases, are emphasized. This supplement seeks to raise awareness and aid diagnosis of HAE, optimize treatment for all patients, and provide a platform for further research in this rare, partially understood disorder.

Related Concept Videos
 01:19
Regulation of Hormone Secretion
Humoral...
 01:40
Hormonal Regulation
 01:33
Overview of Secretory Vesicles
Various proteins regulate the aggregation of molecules inside the secretory vesicles. Chromogranins...
 01:05
Insulin Secretory Vesicles
 01:27
Glucose Homeostasis: Pancreatic Islets and Insulin Secretion
Insulin and C-peptide are...
 00:46
Feedback Inhibition