A time-dependent wave packet study of the vibronic and spin-orbit interactions in the dynamics of Cl((2)P)+H(2)-->HCl(X (1)Sigma(g) (+))+H((2)S) reaction
Related Experiment Video
Updated: Aug 1, 2026
Proton Transfer and Protein Conformation Dynamics in Photosensitive Proteins by Time-resolved Step-scan Fourier-transform Infrared Spectroscopy
Published on: June 27, 2014
Subhas Ghosal1, Susanta Mahapatra
1School of Chemistry, University of Hyderabad, Hyderabad 500 046, India.

Abstract:
We investigate the vibronic and spin-orbit (SO) coupling effects in the state-selected dynamics of the title reaction with the aid of a time-dependent wave packet approach. The ab initio potential energy surfaces of Capecchi and Werner [Science 296, 715 (2002)] have been employed for this purpose. Collinear approach of the Cl((2)P) atom to the H(2) molecule splits the degeneracy of the (2)P state and gives rise to (2)Sigma and (2)Pi electronic states. These two surfaces form a conical intersection at this geometry. These states transform as 1 (2)A('), 1 (2)A("), and 2 (2)A('), respectively, at the nonlinear configurations of the nuclei. In addition, the SO interaction due to Cl atom further splits these states into (2)Sigma(1/2), (2)Pi(3/2), and (2)Pi(1/2) components at the linear geometry. The ground-state reagent Cl((2)P(3/2))+H(2) correlates with (2)Sigma(1/2) and (2)Pi(3/2), where as the SO excited reagent Cl(*)((2)P(1/2))+H(2) correlates with (2)Pi(1/2) at the linear geometry. In order to elucidate the impact of the vibronic and SO coupling effects on the initial state-selected reactivity of these electronic states we carry out quantum scattering calculations based on a flux operator formalism and a time-dependent wave packet approach. In this work, total reaction probabilities and the time dependence of electronic population of the system by initiating the reaction on each of the above electronic states are presented. The role of conical intersection alone on the reaction dynamics is investigated with a coupled two-state model and for the total angular momentum J=0 (neglecting the electronic orbital angular momentum) both in a diabatic as well as in the adiabatic electronic representation. The SO interaction is then included and the dynamics is studied with a coupled three-state model comprising six diabatic surfaces for the total angular momentum J=0.5 neglecting the Coriolis Coupling terms of the Hamiltonian. Companion calculations are carried out for the uncoupled adiabatic and diabatic surfaces in order to explicitly reveal the impact of two different surface coupling mechanisms in the dynamics of this prototypical reaction.
Related Concept Videos
 03:24
Hybridization of Atomic Orbitals I
 03:35
Hybridization of Atomic Orbitals II
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:14
SN2 Reaction: Kinetics
In a chemical reaction, a relationship exists between the concentration of reactants and the rate at which the reaction proceeds. The study to measure this relationship is known as the kinetics of a chemical reaction. Kinetic studies are used to deduce the rate law of a chemical reaction, which provides information about the species involved during the transition state of the rate-determining step. Thus, kinetic studies help to derive the mechanism of a reaction.
 01:08
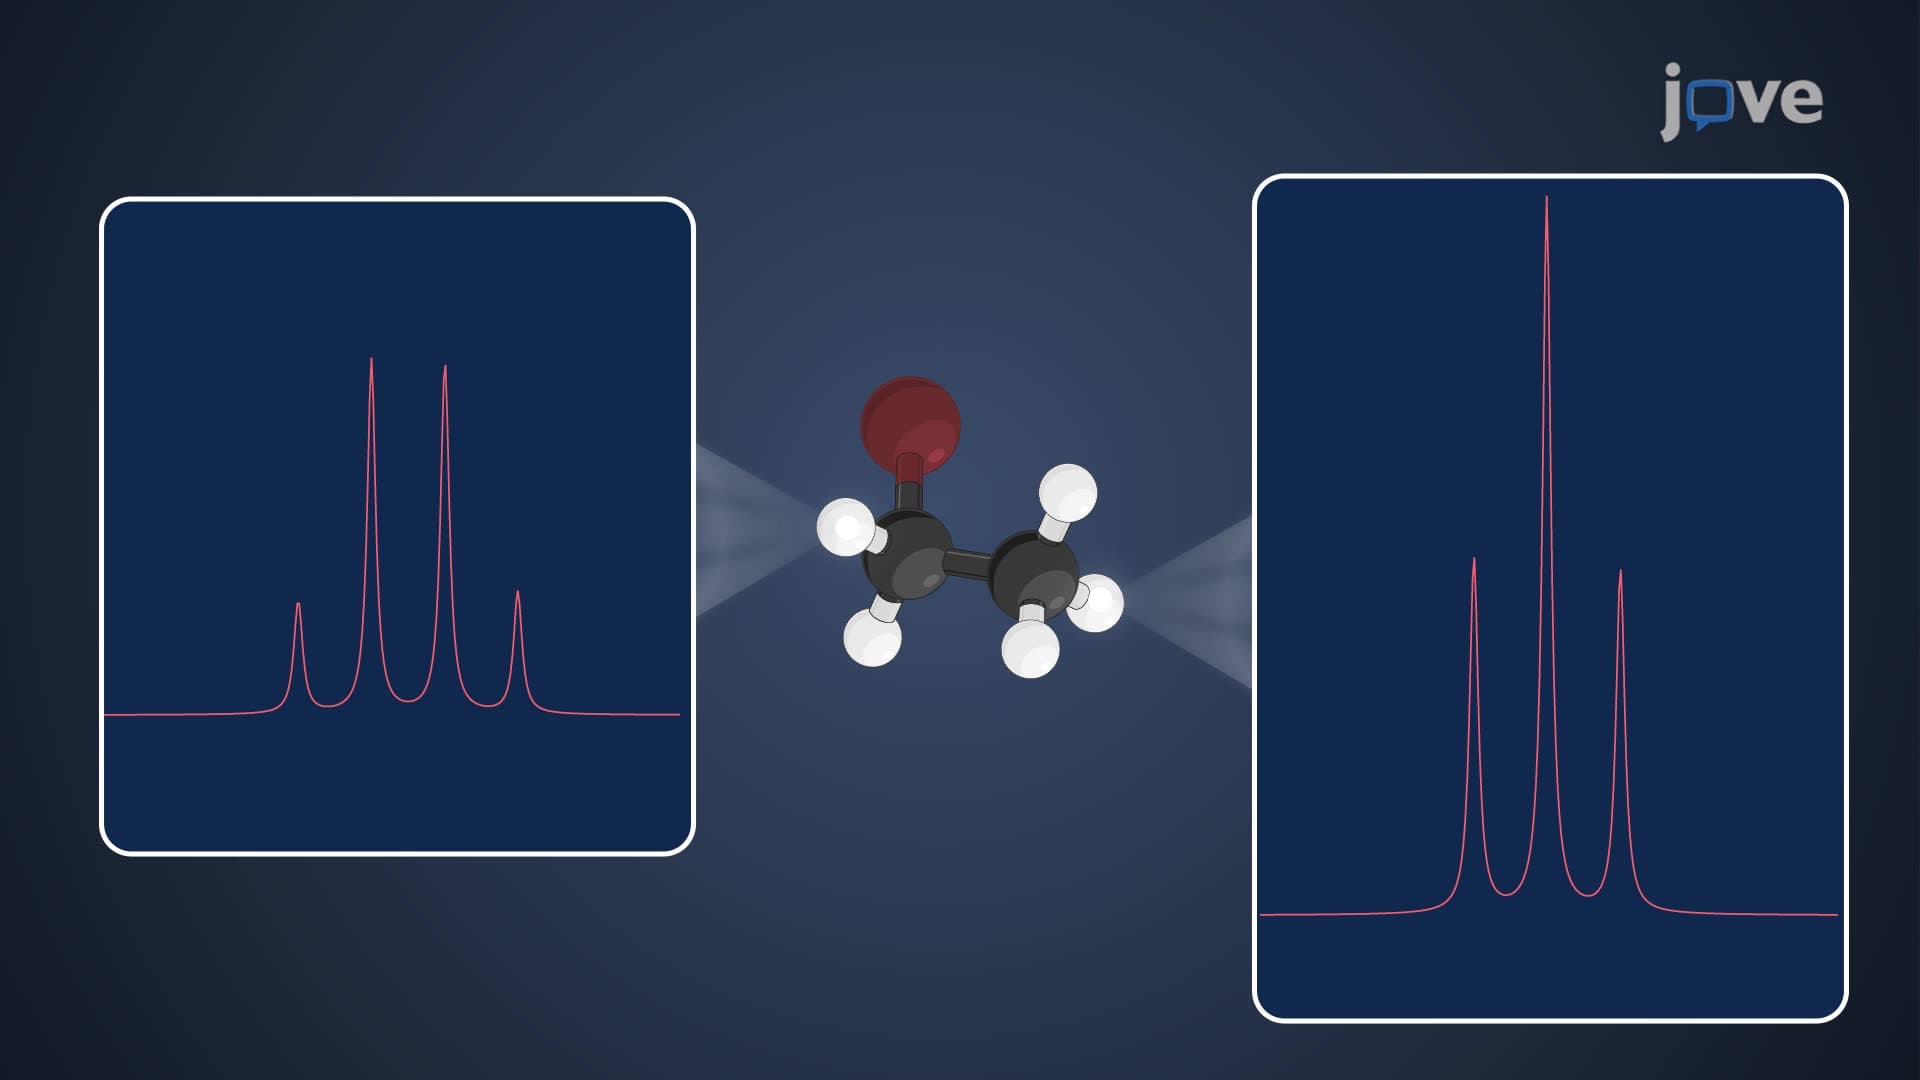
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must have a...