Related Experiment Video
Updated: Aug 22, 2026
Catalytic Reactions at Amine-Stabilized and Ligand-Free Platinum Nanoparticles Supported on Titania During Hydrogenation of Alkenes and Aldehydes
Published on: June 24, 2022
Torsional vibration and central bond length of N-benzylideneanilines
Jun Harada1, Mayuko Harakawa, Keiichiro Ogawa
1Department of Chemistry, Graduate School of Arts and Sciences, The University of Tokyo, Komaba, Meguro-ku, Tokyo 153-8902, Japan.

Abstract:
The crystal structures of N-benzylideneaniline (1), N-benzylidene-4-carboxyaniline (2), N-(4-methylbenzylidene)-4-nitroaniline (3), N-(4-nitrobenzylidene)-4-methoxyaniline (4), N-(4-nitrobenzylidene)-4-methylaniline (5), N-(4-methoxybenzylidene)aniline (6) and N-(4-methoxybenzylidene)-4-methylaniline (7) were determined by X-ray diffraction analyses at various temperatures. In the crystal structures of all the compounds, an apparent shortening of the central C=N bond was observed at room temperature. As the temperature was lowered, the observed bond lengths increased to approximately 1.28 A at 90 K, irrespective of substituents in the molecules. The shortening and the temperature dependence of the C=N bond length are interpreted in terms of an artifact caused by the torsional vibration of the C-Ph and N-Ph bonds in the crystals. In the crystal structures of (1) and (7), a static disorder around the C=N bond was observed, which is also responsible for the apparent shortening of the C=N bond.
More Related Videos

Related Concept Videos
 01:18
Structure of Benzene: Molecular Orbital Model
 01:37
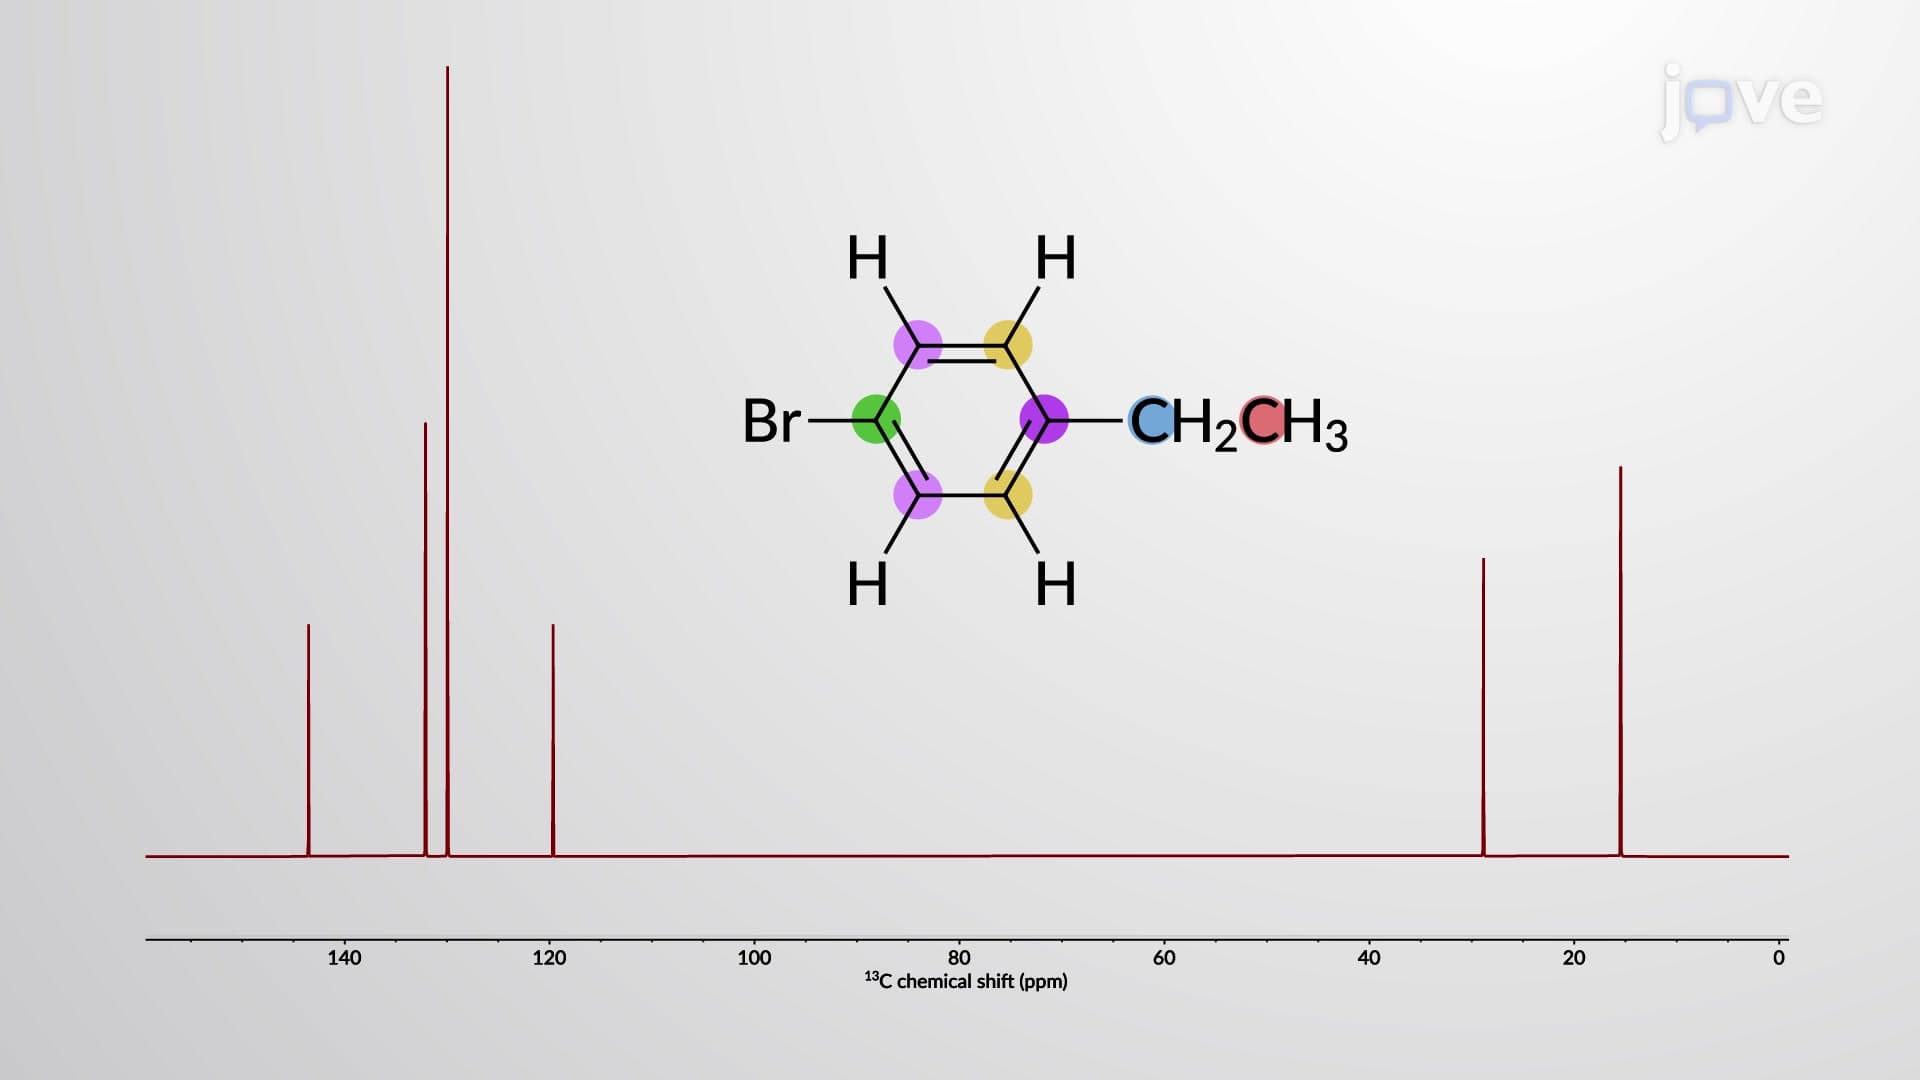
NMR Spectroscopy of Benzene Derivatives
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 01:22
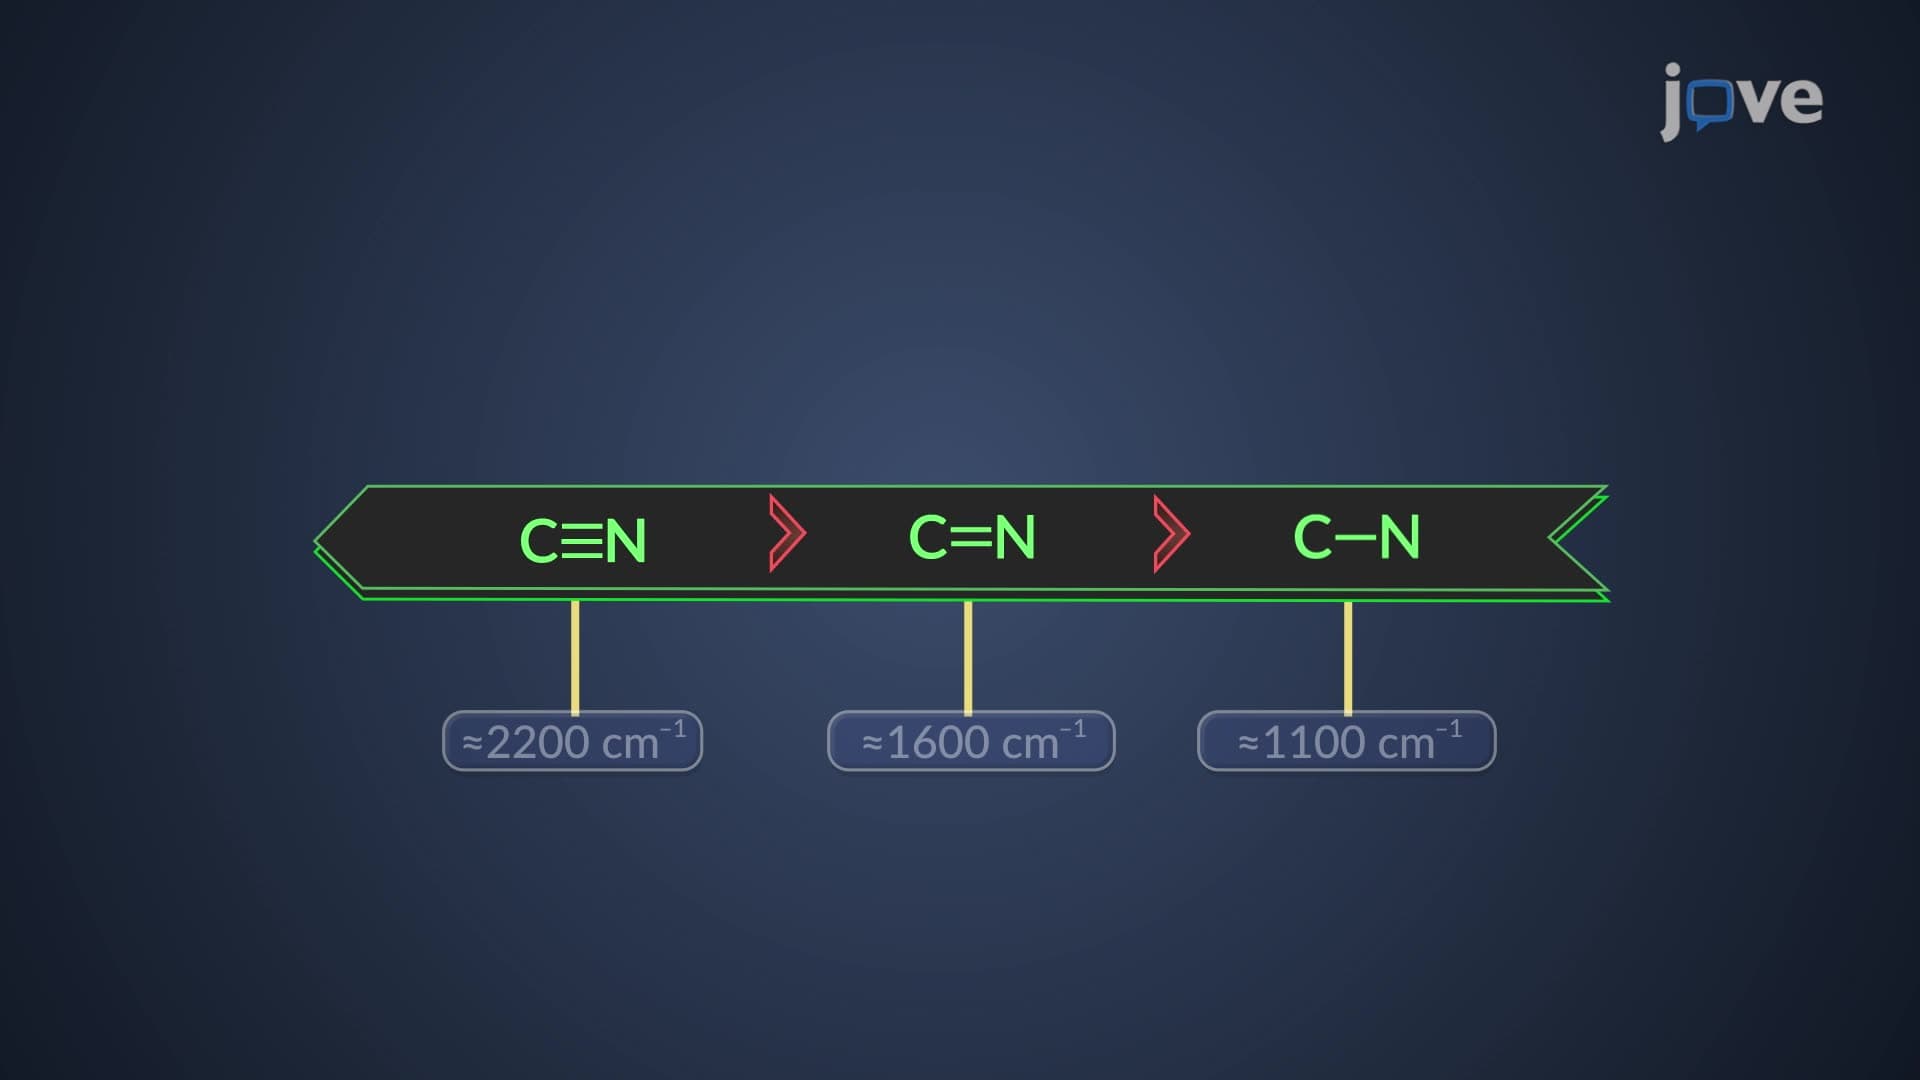
IR Frequency Region: Alkyne and Nitrile Stretching
Comparing the stretching vibrational frequency of C≡C triple bonds with that of double and single bonds, it is evident that C≡C triple bonds exhibit a higher stretching frequency than C=C double and C–C single bonds. Similarly, the C≡N triple bond exhibits higher stretching absorption than the C=N...
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds