Related Experiment Video
Updated: Aug 22, 2026

Rapid Viscoelastic Characterization of Airway Mucus Using a Benchtop Rheometer
Published on: April 21, 2022
Dornase alpha and exhaled NO in cystic fibrosis
Hartmut Grasemann1, Hildegard Lax, Jennifer W Treseler
1Department of Pediatrics, University of Essen, 45122, Germany. hartmutg@hotmail.com

Abstract:
Nitric oxide (NO) that is produced within the airways can be measured in the exhaled air. Concentrations of exhaled NO (FENO) are decreased in cystic fibrosis (CF) and, in cross sectional studies, have been shown to be even lower in patients with more advanced pulmonary disease. This may result from retention and metabolisation of NO within viscous airway secretions. Treatment with recombinant human DNase I (dornase alpha) modifies the rheological properties of airway secretions and thereby improves pulmonary function even in young and apparently healthy patients with CF. We studied FENO and pulmonary function in children with CF with little clinical evidence of lung morbidity in a two-year randomized double-blind placebo-controlled study with nebulized dornase alpha. Mean age at enrollment was 8 years (range 6 to 11 years), mean forced vital capacity (FVC) was 112% (range 86 to 133%), and mean forced expiratory volume in one second (FEV1)was 109% (range 88 to 128%) of predicted values. In five of six (83%) of the dornase alpha treated patients, FENO changed in parallel to changes in pulmonary function tests while no such correlation was observed in any of the eight patients receiving placebo. This difference between treatment groups was statistically significant for both FVC (P = 0.026, Wilcoxon-test) and FEV1 (P = 0.042). These data suggest that FENO may serve as a surrogate measure for evaluating the effectiveness of interventions that affect airway clearance in CF.
More Related Videos
 06:57
The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
 08:00
Generation of Human Nasal Epithelial Cell Spheroids for Individualized Cystic Fibrosis Transmembrane Conductance Regulator Study
Published on: April 11, 2018
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:05
Breathing
 01:23
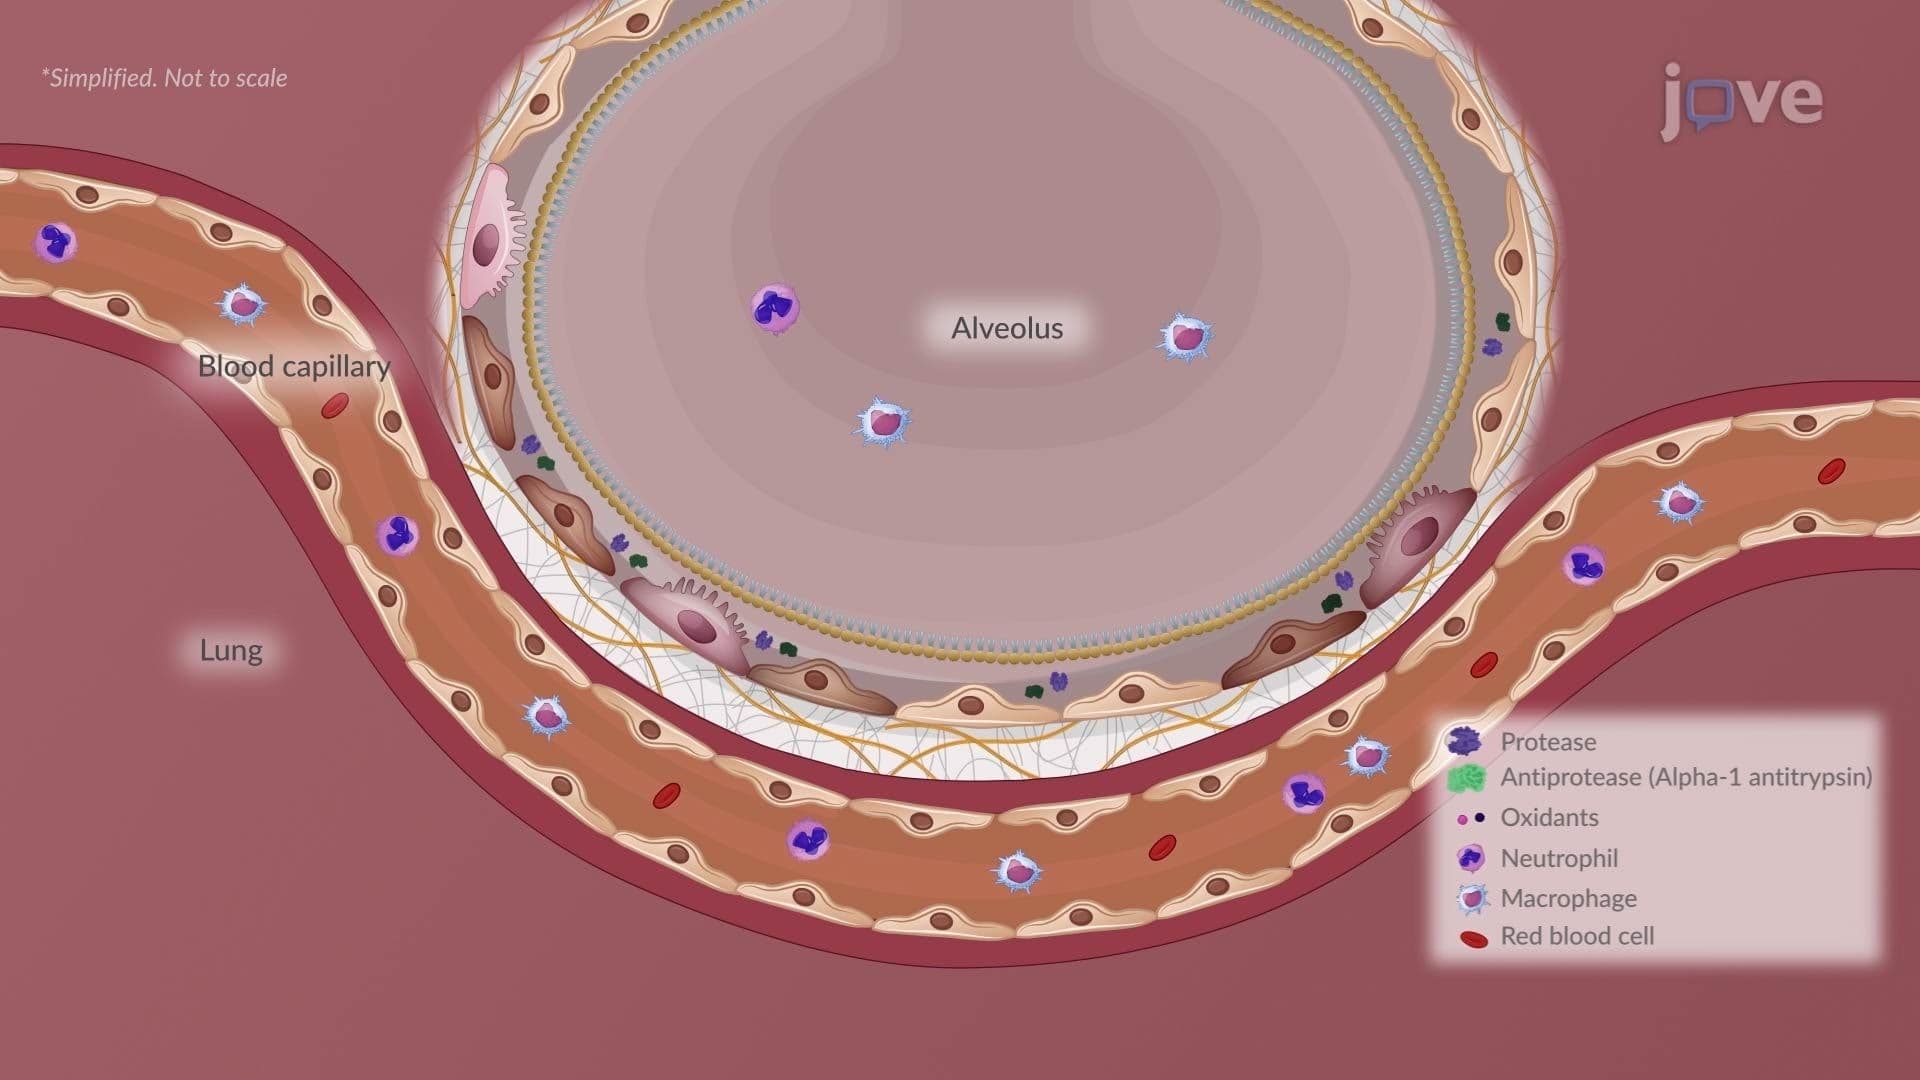
Chronic Obstructive Pulmonary Disease II: Emphysema
 01:17
Pulmonary Cycle: Exhalation
 02:32
Exon Recombination
Exon shuffling follows “splice frame rules.” Each exon has three reading...