Synthetic stitching with silicon: geminal alkylation-hydroxylation of alkynyl carbonyl compounds
Related Experiment Video
Updated: Jul 6, 2026
Optimization of the Ugi Reaction Using Parallel Synthesis and Automated Liquid Handling
Published on: November 11, 2008
Barry M Trost1, Zachary T Ball
1Department of Chemistry, Stanford University, Stanford, California 94305-5080, USA. bmtrost@stanford.edu

Abstract:
A new strategy for the synthesis of beta-carbonyl-substituted tertiary alcohols from alpha,beta-alkynyl ketones and esters has been demonstrated. A silicon tether is used to internally deliver an alkyl group, which, combined with a C-Si to C-O transformation, can regio- and diastereoselectively "stitch" together geminal C-C and C-O bonds at the beta-position of an electron-withdrawing group. Regioselective alkyne hydrosilylation by a trans addition process provides clean access to trisubstituted vinylsilanes. Subsequent one-pot fluoride-induced C-C bond formation and oxidation of the resulting tertiary silane, a type of silane not normally reactive to such conditions, affords the desired products. The utility of neighboring ketone and carboxylate groups in promoting the oxidation of these highly hindered tertiary alcohols, an observation that may affect synthetic design of routes depending on such oxidations, is demonstrated. Good diastereoselection (>10:1) is observed for substrates bearing gamma-alkoxy stereocenters.
Related Concept Videos
 00:51
Base-Promoted α-Halogenation of Aldehydes and Ketones
 01:08
Base-Catalyzed Aldol Addition Reaction
 01:09
Conjugate Addition to α,β-Unsaturated Carbonyl Compounds
 00:54
Radical Formation: Homolysis
 01:10
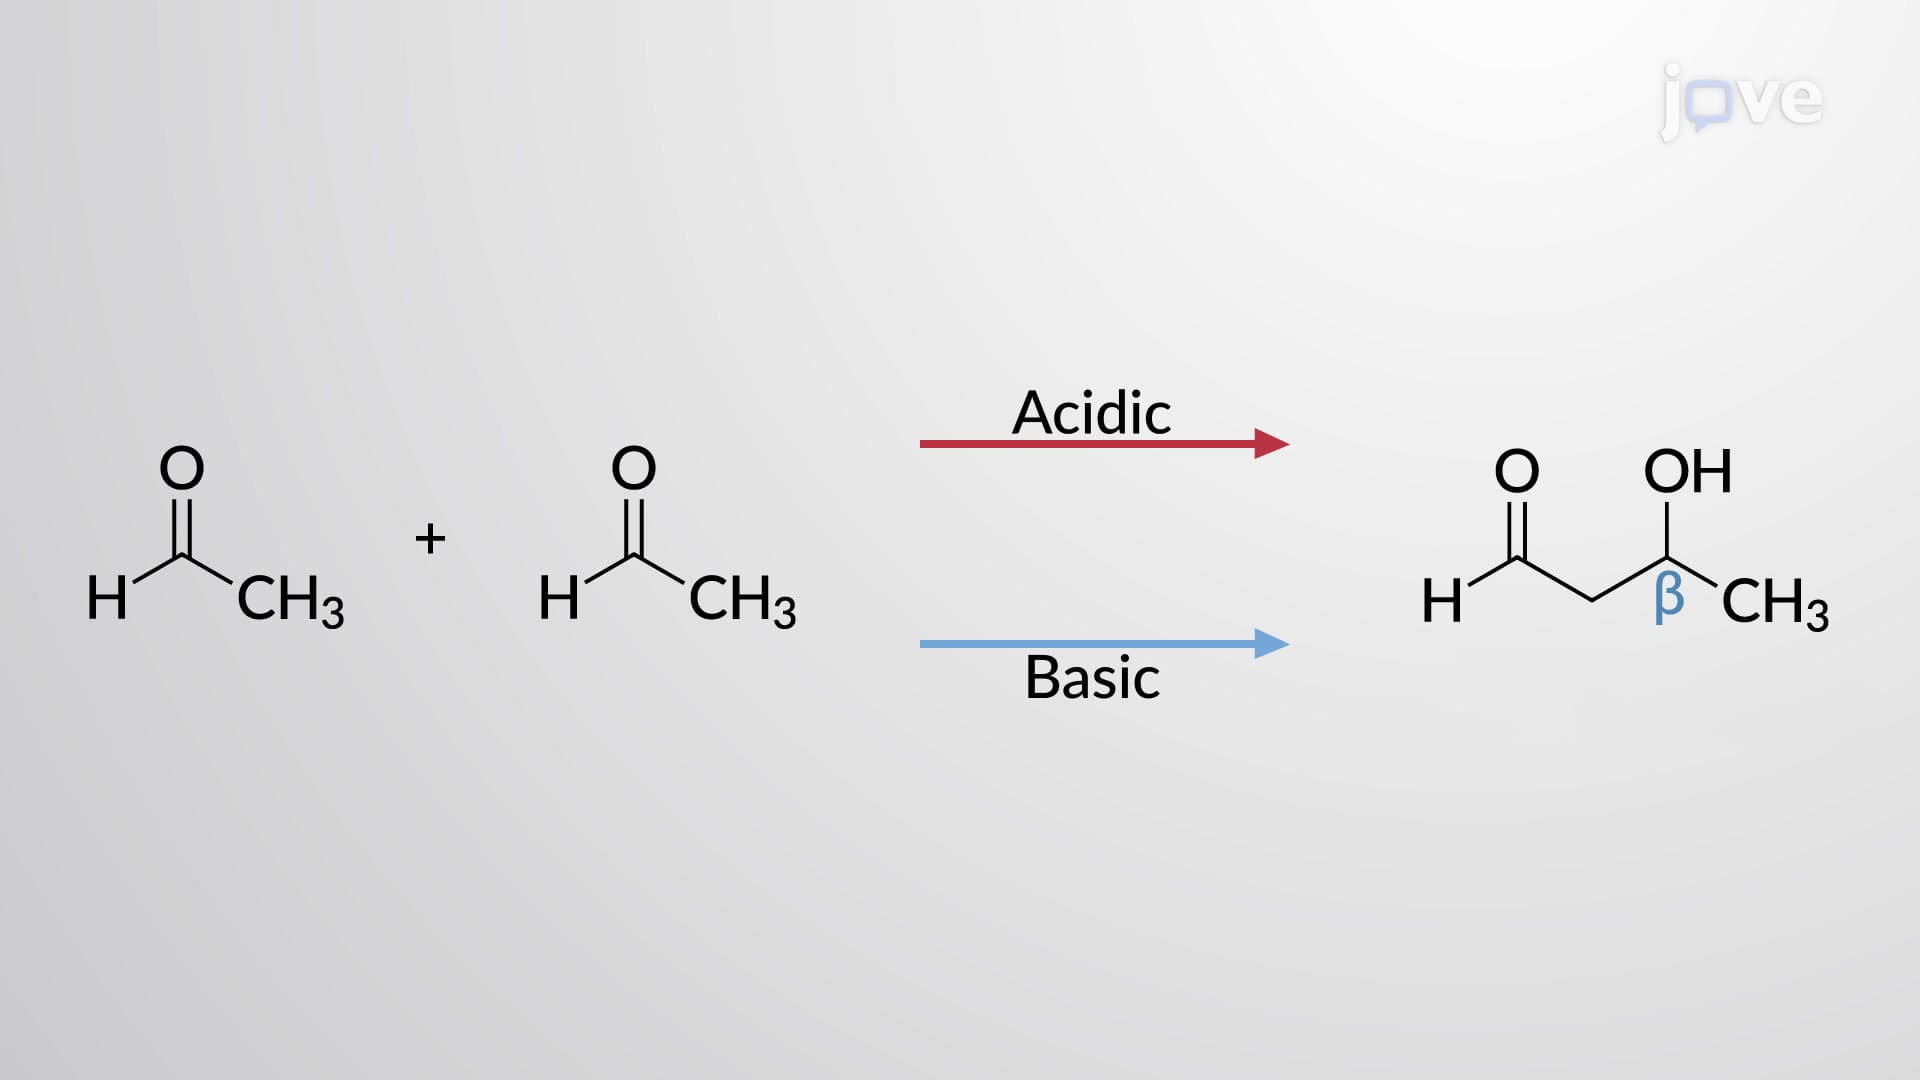
C–C Bond Formation: Aldol Condensation Overview
 01:27
Conjugate Addition (1,4-Addition) vs Direct Addition (1,2-Addition)
Conjugate addition results in a thermodynamically stable product. The reaction retains the stronger C=O bond at the expense of the weaker C=C π bond. The process is slow as the β carbon is less electrophilic than the carbonyl carbon.
Direct addition products are formed faster owing to...