An efficient molecular docking using conformational space annealing
Related Experiment Video
Updated: Aug 21, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Kyoungrim Lee1, Cezary Czaplewski, Seung-Yeon Kim
1School of Computational Sciences, Korea Institute for Advanced Study, 207-43 Cheongnyangni 2-dong, Dongdaemun-gu, Seoul, South Korea.

Abstract:
Molecular docking falls into the general category of global optimization problems because its main purpose is to find the most stable complex consisting of a receptor and its ligand. Conformational space annealing (CSA), a powerful global optimization method, is incorporated with the Tinker molecular modeling package to perform molecular docking simulations of six receptor-ligand complexes (3PTB, 1ULB, 2CPP, 1STP, 3CPA, and 1PPH) from the Protein Data Bank. In parallel, Monte Carlo with the minimization (MCM) method is also incorporated into the Tinker package for comparison. The energy function, consisting of electrostatic interactions, van der Waals interactions, and torsional energy terms, is calculated using the AMBER94 all-atom empirical force field. Rigid docking simulations for all six complexes and flexible docking simulations for three complexes (1STP, 3CPA, and 1PPH) are carried out using the CSA and the MCM methods. The simulation results show that the docking procedures using the CSA method generally find the most stable complexes as well as the native-like complexes more efficiently and accurately than those using the MCM, demonstrating that CSA is a promising search method for molecular docking problems.
More Related Videos
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
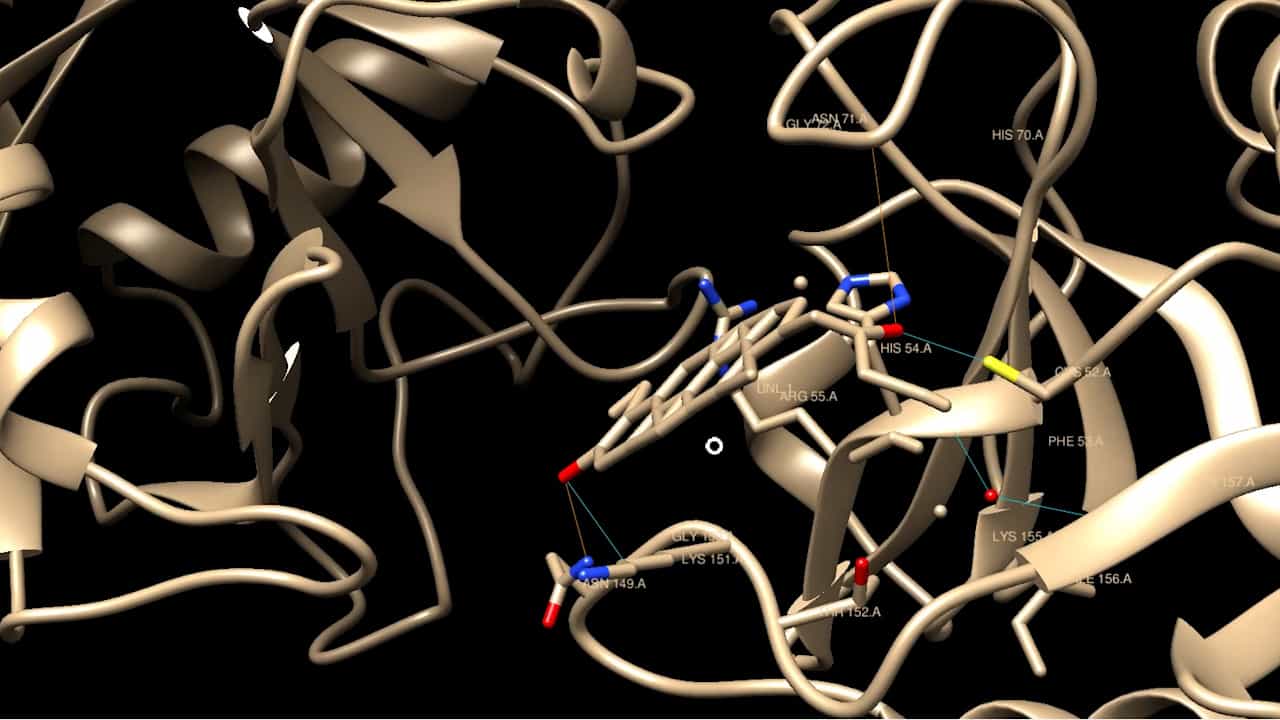 10:01
Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...