Related Experiment Video
Updated: Jul 17, 2026
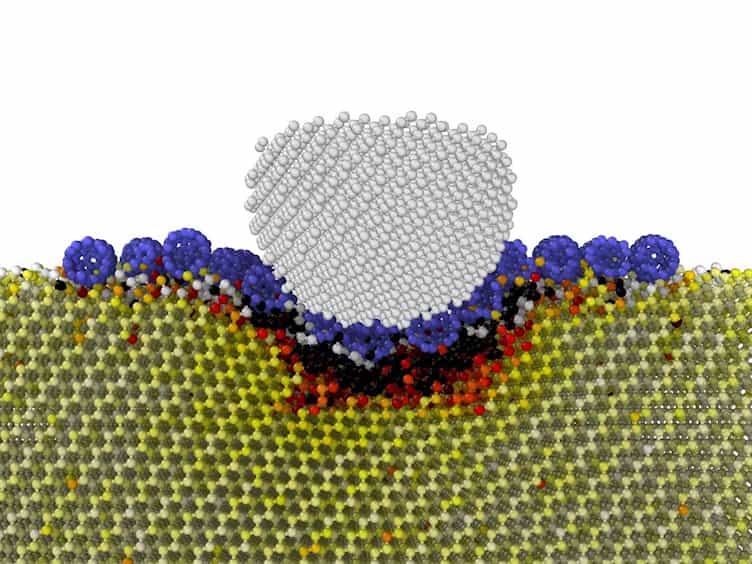

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Preparation and characterization of pi-stacking quinodimethane oligothiophenes. Predicting semiconductor behavior and
Daron E Janzen1, Michael W Burand, Paul C Ewbank
1Department of Chemistry, University of Minnesota, Minneapolis, Minnesota 55455, USA.

Abstract:
A series of new quinodimethane-substituted terthiophene and quaterthiophene oligomers has been investigated for comparison with a previously studied quinoid oligothiophene that has demonstrated high mobilities and ambipolar transport behavior in thin-film transistor devices. Each new quinoidal thiophene derivative shows a reversible one-electron oxidation between 0.85 and 1.32 V, a quasi-reversible one-electron second oxidation between 1.37 and 1.96 V, and a reversible two-electron reduction between -0.05 and -0.23 V. The solution UV-vis-NIR spectrum of each compound is dominated by an intense (epsilon congruent with 100 000 M(-1) cm(-1)) low energy pi-pi transition that has a lambda(max) ranging between 648 and 790 nm. All X-ray crystal structures exhibit very planar quinoidal backbones and short intermolecular pi-stacking distances (3.335-3.492 A). Structures exhibit a single pi-stacking distance with parallel cofacial stacking (sulfur atoms of equivalent rings pointed in the same direction) or with alternating distances and antiparallel cofacial stacking (sulfur atoms of equivalent rings pointed in the opposite direction). Examples of the layered and herringbone-packing motifs are observed for both the parallel and the antiparallel cofacial stacking. Analysis of the X-ray structures and molecular orbital calculations indicates that all of these compounds have one-dimensional electronic band structures as a result of the pi-stacking. For structures with a unique pi-stacking distance, a simple geometric overlap parameter calculated from the shape of the molecule and the slip from perfect registry in the pi-stack correlates well with the transfer integrals (t) calculated using molecular orbital theory. The calculated valence (633 meV) and conduction (834 meV) bandwidths for a quinoid quaterthiophene structure are similar to those calculated for the benchmark pentacene and indicate that both hole and electron mobilities could be significant.
More Related Videos

Related Concept Videos
 02:27
Predicting Molecular Geometry
 02:22
Structures of Solids
 02:42
Ionic Crystal Structures
Most monatomic ions behave as charged spheres, and their attraction for ions of opposite charge is the same in every direction. Consequently, stable structures for ionic compounds result (1) when ions of one charge are surrounded by as many ions as possible of the opposite...
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability. Many...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:18
Unit Cells