Related Experiment Video
Updated: Aug 3, 2026

Spatial Separation of Molecular Conformers and Clusters
Published on: January 9, 2014
Magnitude and orientation dependence of intermolecular interaction of perfluoropropane dimer studied by high-level ab
Seiji Tsuzuki1, Tadafumi Uchimaru, Masuhiro Mikami
1National Institute of Advanced Industrial Science and Technology (AIST), Tsukuba, Ibaraki 305-8568, Japan. s.tsuzuki@aist.go.jp

Abstract:
Intermolecular interaction energies of 12 orientations of C(3)F(8) dimers were calculated with electron correlation correction by the second-order Møller-Plesset perturbation method. The antiparallel C(2h) dimer has the largest interaction energy (-1.45 kcal/mol). Electron correlation correction increases the attraction considerably. Electrostatic energy is not large. Dispersion is mainly responsible for the attraction. Orientation dependence of the interaction energy of the C(3)F(8) dimer is substantially smaller than that of the C(3)H(8) dimer. The calculated interaction energy of the C(3)F(8) dimer at the potential minimum is 78% of that of the C(3)H(8) dimer (-1.85 kcal/mol), whereas the interaction energies of the CF(4) and C(2)F(6) dimers are larger than those of the CH(4) and C(2)H(6) dimers. The intermolecular separation in the C(3)F(8) dimer at the potential minimum is substantially larger than that in the C(3)H(8) dimer. The larger intermolecular separation due to the steric repulsion between fluorine atoms is the cause of the smaller interaction energy of the C(3)F(8) dimer at the potential minimum. The calculated intermolecular interaction energy potentials of the C(3)F(8) dimers using an all atom model OPLS-AA (OPLS all atom model) force field and a united atom model force field were compared with the ab initio calculations. Although the two force fields well reproduces the experimental vapor and liquid properties of perfluoroalkenes, the comparison shows that the united atom model underestimates the potential depth and orientation dependence of the interaction energy. The potentials obtained by the OPLS-AA force field are close to those obtained by the ab initio calculations.
More Related Videos
 09:43
Interfacial Molecular-level Structures of Polymers and Biomacromolecules Revealed via Sum Frequency Generation Vibrational Spectroscopy
Published on: August 13, 2019
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Related Concept Videos
 03:37
Molecular Shape and Polarity
 03:24
Hybridization of Atomic Orbitals I
 02:34
Comparing Intermolecular Forces: Melting Point, Boiling Point, and Miscibility
Temporary attractive forces like dispersion are present in all molecules, whether they are polar or nonpolar. They...
 02:56
Intermolecular Forces and Physical Properties
 02:18
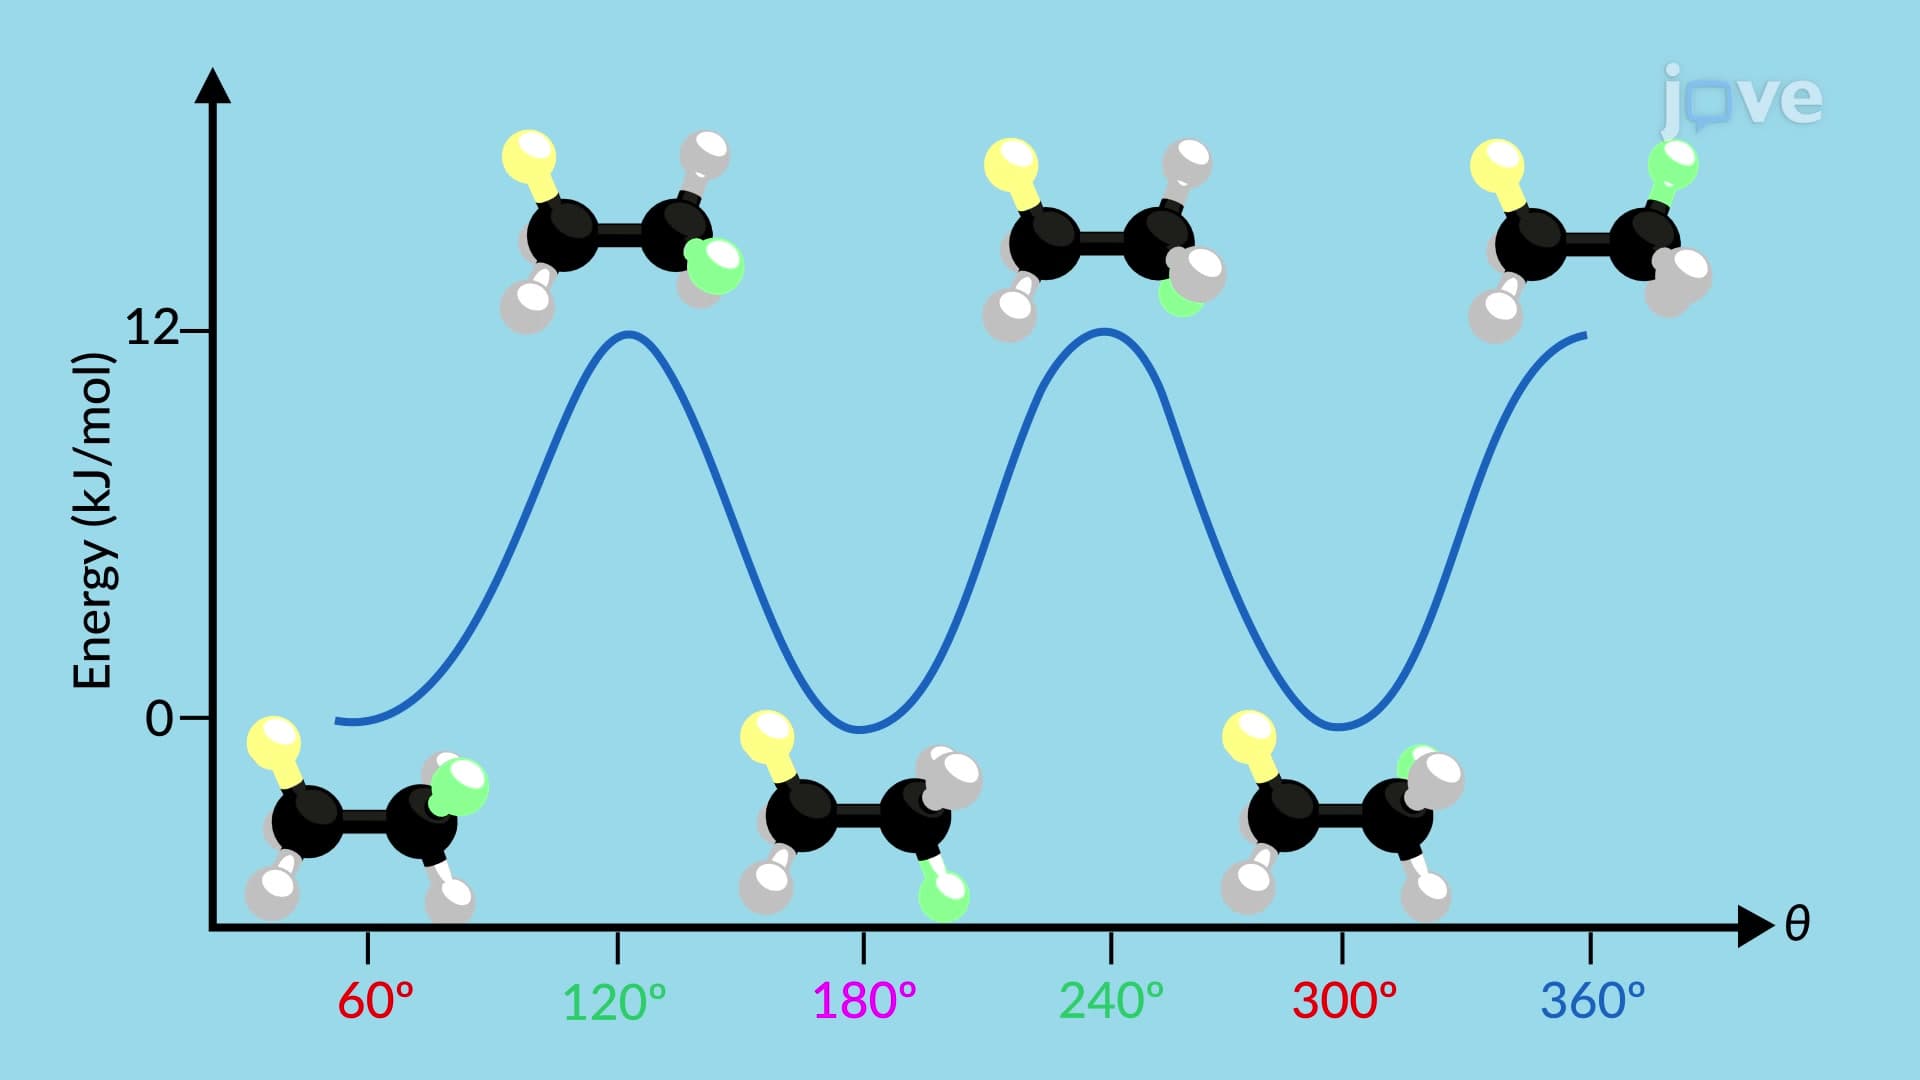
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered ethane, the...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...